Related Experiment Video
Updated: Dec 5, 2025
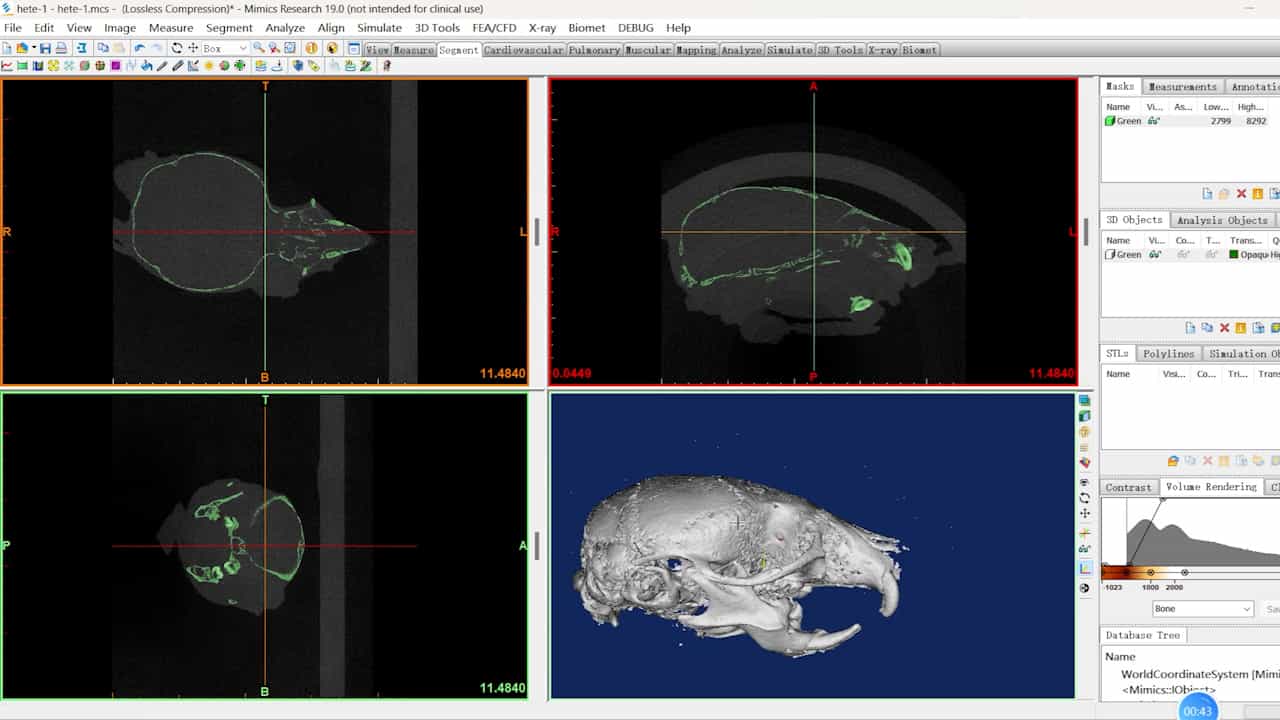

Analysis of Craniomaxillofacial Malformations in Mice Using Three-dimensional Microcomputed Tomography
Published on: January 17, 2025
Immunodeficiency in a patient with microcephalic osteodysplastic primordial dwarfism type I as compared to Roifman
Hidetoshi Hagiwara1, Hiroshi Matsumoto1, Kenji Uematsu1
1Department of Pediatrics, National Defense Medical College Hospital, 3-2 Namiki, Tokorozawa, Saitama 359-8513, Japan.

Background:
Microcephalic osteodysplastic primordial dwarfism type I (MOPD I, also known as Taybi-Linder syndrome) is a rare genetic disorder associated with severe intrauterine growth retardation, short stature, microcephaly, brain anomalies, stunted limbs, and early mortality. RNU4ATAC, the gene responsible for this disorder, does not encode a protein but instead the U4atac small nuclear RNA (snRNA), a crucial component of the minor spliceosome. Roifman syndrome is an allelic disorder of MOPD I that is characterized by immunodeficiency complications.
Case Report:
The patient described herein is an 18-year-old woman exhibiting congenital dwarfism and microcephaly with structural brain anomaly. She suffered human herpesvirus 6 (HHV-6)-associated acute necrotizing encephalopathy at the age of one, thereafter resulting in severe psychomotor disabilities. Genetic analysis using gene microarray and whole-exome sequencing could not identify the cause of her congenital anomalies. However, Sanger sequencing revealed a compound heterozygous mutation within RNU4ATAC (NR_023343.1:n.[50G > A];[55G > A]). Immunological findings showed decreases in total lymphocytes, CD4+ T cells, and T cell regenerative activity. Furthermore, antibodies against varicella-zoster, rubella, measles, mumps, and influenza were very low or negative despite having received vaccinations for these viruses. HHV-6 IgG antibodies were also undetected.
Discussion:
The patient here exhibited a marked MOPD I phenotype complicated by various immunodeficiencies. Previous studies have not demonstrated immunodeficiency comorbidities within MOPD I subjects, but this report suggests an evident immunodeficiency in MOPD I. Patients with MOPD I should be treated with one of the immunodeficiency syndromes.
Insights
Microcephalic osteodysplastic primordial dwarfism type I (MOPD I) is a rare genetic disorder. This case study reveals MOPD I can present with significant immunodeficiency, suggesting patients need treatment for both conditions.
Area of Science:
- Genetics
- Immunology
- Rare Diseases
Background:
- Microcephalic osteodysplastic primordial dwarfism type I (MOPD I), or Taybi-Linder syndrome, is a rare genetic disorder.
- It is caused by mutations in the RNU4ATAC gene, which affects the minor spliceosome.
- Roifman syndrome is an allelic disorder of MOPD I with immunodeficiency.
Observation:
- An 18-year-old woman with congenital dwarfism, microcephaly, and brain anomalies presented with severe psychomotor disabilities after HHV-6 encephalitis.
- Genetic analysis identified compound heterozygous mutations in RNU4ATAC.
- Immunological evaluation revealed decreased lymphocytes, T cells, and antibody responses to vaccinations.
Findings:
- The patient exhibited a severe MOPD I phenotype.
- Significant immunodeficiencies, including low antibody titers and reduced T cell activity, were observed.
- The RNU4ATAC mutations are linked to both MOPD I and immunodeficiency.
Implications:
- This case suggests that immunodeficiency may be an underrecognized comorbidity in MOPD I.
- Patients diagnosed with MOPD I should be evaluated for and potentially treated for associated immunodeficiency syndromes.
- Further research is needed to understand the link between RNU4ATAC mutations and immune system dysfunction.