Related Experiment Video
Updated: Dec 5, 2025

Generation, Amplification, and Titration of Recombinant Respiratory Syncytial Viruses
Published on: April 4, 2019
Towards a unified classification for human respiratory syncytial virus genotypes
Kaat Ramaekers1, Annabel Rector1, Lize Cuypers1,2
1KU Leuven, Department of Microbiology, Immunology and Transplantation, Rega Institute for Medical Research, Laboratory of Clinical and Epidemiological Virology, Herestraat 49 box 1040, BE-3000 Leuven, Belgium.

Abstract:
Since the first human respiratory syncytial virus (HRSV) genotype classification in 1998, inconsistent conclusions have been drawn regarding the criteria that define HRSV genotypes and their nomenclature, challenging data comparisons between research groups. In this study, we aim to unify the field of HRSV genotype classification by reviewing the different methods that have been used in the past to define HRSV genotypes and by proposing a new classification procedure, based on well-established phylogenetic methods. All available complete HRSV genomes (>12,000 bp) were downloaded from GenBank and divided into the two subgroups: HRSV-A and HRSV-B. From whole-genome alignments, the regions that correspond to the open reading frame of the glycoprotein G and the second hypervariable region (HVR2) of the ectodomain were extracted. In the resulting partial alignments, the phylogenetic signal within each fragment was assessed. Maximum likelihood phylogenetic trees were reconstructed using the complete genome alignments. Patristic distances were calculated between all pairs of tips in the phylogenetic tree and summarized as a density plot in order to determine a cutoff value at the lowest point following the major distance peak. Our data show that neither the HVR2 fragment nor the G gene contains sufficient phylogenetic signal to perform reliable phylogenetic reconstruction. Therefore, whole-genome alignments were used to determine HRSV genotypes. We define a genotype using the following criteria: a bootstrap support of 70 per cent for the respective clade and a maximum patristic distance between all members of the clade of ≤0.018 substitutions per site for HRSV-A or ≤0.026 substitutions per site for HRSV-B. By applying this definition, we distinguish twenty-three genotypes within subtype HRSV-A and six genotypes within subtype HRSV-B. Applying the genotype criteria on subsampled data sets confirmed the robustness of the method.
More Related Videos
 09:01
An In vitro Model to Study Immune Responses of Human Peripheral Blood Mononuclear Cells to Human Respiratory Syncytial Virus Infection
Published on: December 10, 2013
 09:14
An Improved and High Throughput Respiratory Syncytial Virus RSV Micro-neutralization Assay
Published on: January 26, 2019
Related Concept Videos
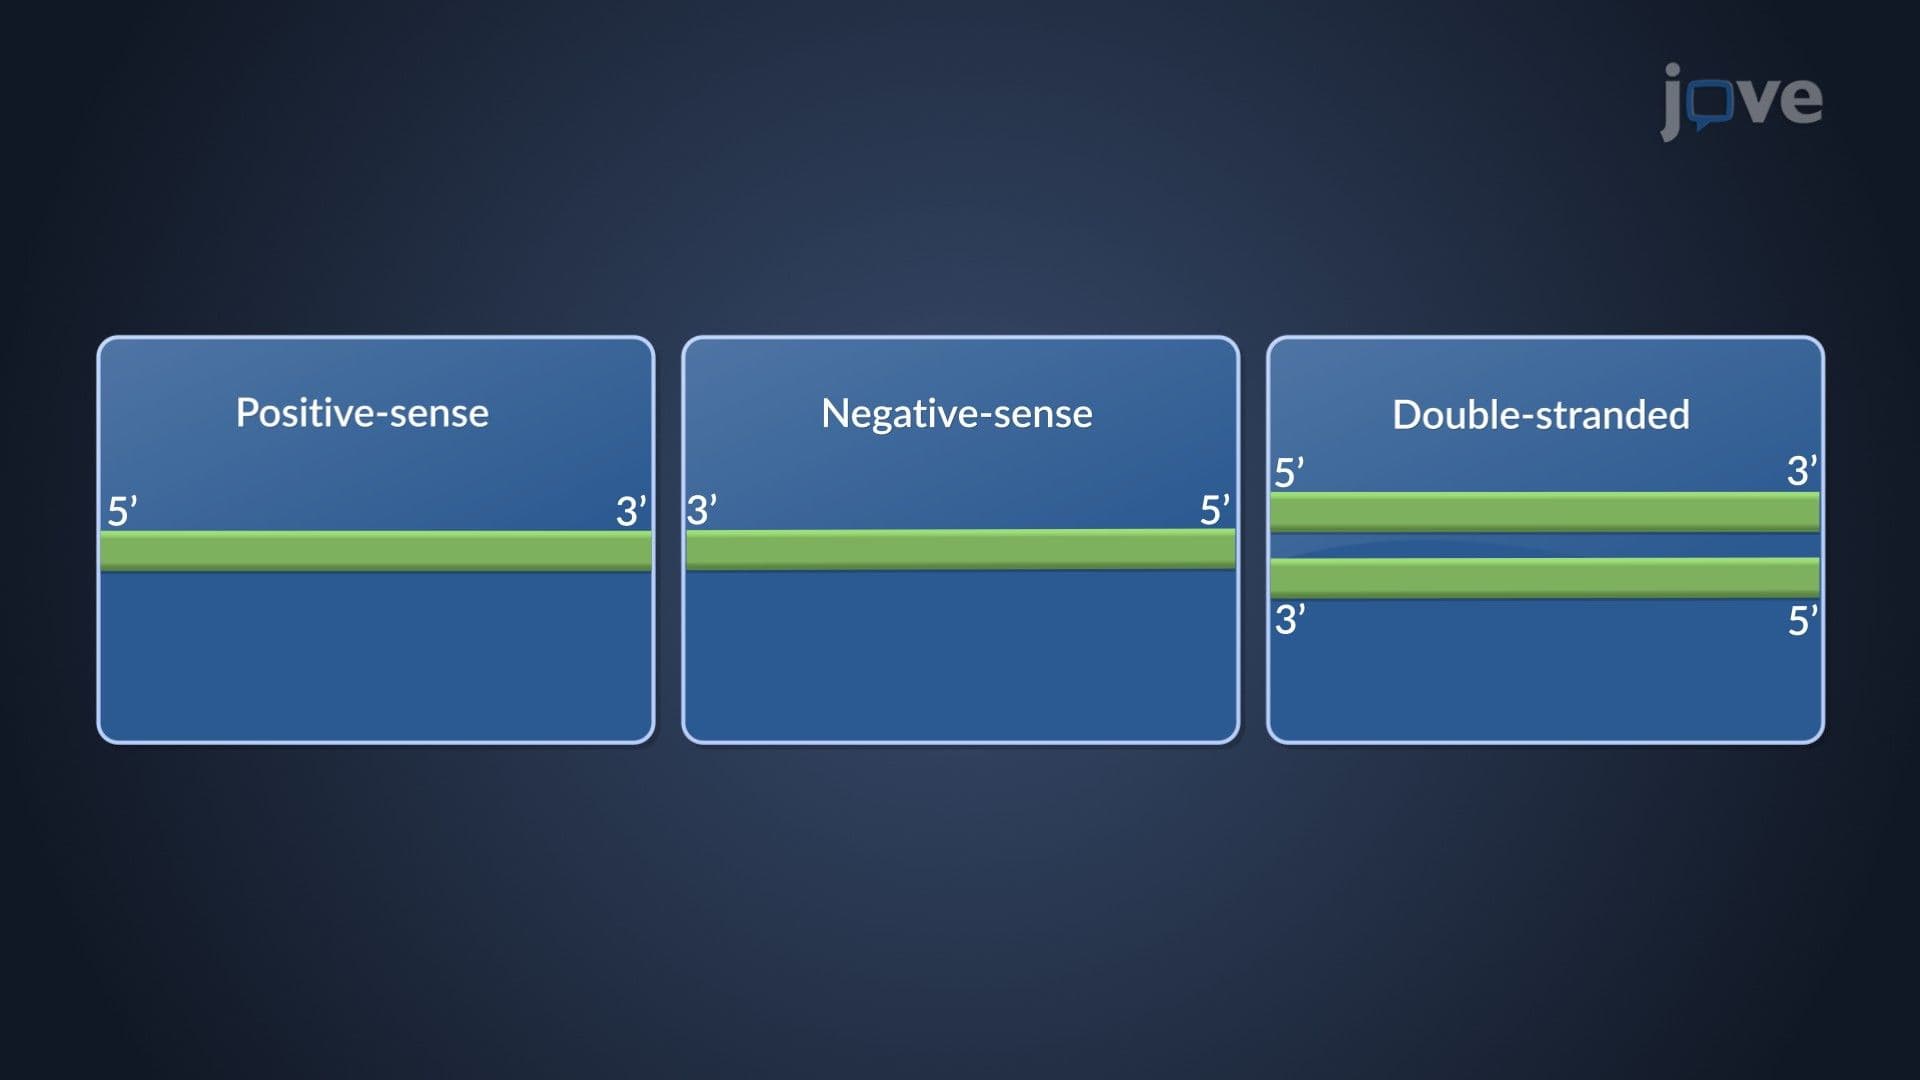 01:29
Viruses with RNA Genomes
 01:31
Common Respiratory Disorders
Upper respiratory disorders impact the airways above the vocal cords, encompassing areas like the nose, sinuses, and throat. Various conditions fall under this category, including the common cold and allergic rhinitis. These disorders can stem from several causes,...
 01:26
Asthma-II: Pathophysiology and Classification
Additionally, environmental and genetic factors play crucial roles in determining an individual's susceptibility to asthma and the severity of their condition.
Critical processes in asthma pathophysiology include:
 01:15
Upper Respiratory Drugs: First and Second-Generation Antihistamines
Histamine binds to specific receptor sites, known as H1 receptors, on tissue cells, triggering inflammation and swelling. Antihistamines combat these effects by competing with histamine for these receptor sites. By...
 00:57
Viral Recombination
 01:16
Drugs Used in Upper Respiratory Disorders: Overview
Antihistamines (e.g., Benadryl) block histamines from binding. Histamines are chemicals released during an allergic reaction in the body. As a...