Inactivation of the mitochondrial protease Afg3l2 results in severely diminished respiratory chain activity and
Gautam Pareek1, Leo J Pallanck1
1Department of Genome Sciences, University of Washington, Seattle, United States of America.

Abstract:
The m-AAA proteases play a critical role in the proteostasis of inner mitochondrial membrane proteins, and mutations in the genes encoding these proteases cause severe incurable neurological diseases. To further explore the biological role of the m-AAA proteases and the pathological consequences of their deficiency, we used a genetic approach in the fruit fly Drosophila melanogaster to inactivate the ATPase family gene 3-like 2 (AFG3L2) gene, which encodes a critical component of the m-AAA proteases. We found that null alleles of Drosophila AFG3L2 die early in development, but partial inactivation of AFG3L2 using RNAi allowed survival to the late pupal and adult stages of development. Flies with partial inactivation of AFG3L2 exhibited behavioral defects, neurodegeneration, accumulation of unfolded mitochondrial proteins, and diminished respiratory chain (RC) activity. Further work revealed that the reduced RC activity was primarily a consequence of severely diminished mitochondrial transcription and translation. These defects were accompanied by activation of the mitochondrial unfolded protein response (mito-UPR) and autophagy. Overexpression of mito-UPR components partially rescued the AFG3L2-deficient phenotypes, indicating that protein aggregation partly accounts for the defects of AFG3L2-deficient animals. Our work suggests that strategies designed to activate mitochondrial stress pathways and mitochondrial gene expression could be therapeutic in the diseases caused by mutations in AFG3L2.
Insights
Defects in AFG3L2, crucial for mitochondrial protein quality control, cause neurodegeneration. Activating mitochondrial stress responses may offer therapeutic strategies for these incurable neurological diseases.
Area of Science:
- Mitochondrial biology
- Neurogenetics
- Molecular medicine
Background:
- The m-AAA proteases are essential for maintaining mitochondrial protein homeostasis.
- Mutations in genes encoding m-AAA proteases lead to severe, untreatable neurological disorders.
Purpose of the Study:
- To investigate the biological functions of m-AAA proteases and the pathology of their deficiency.
- To explore the role of the ATPase family gene 3-like 2 (AFG3L2) in mitochondrial proteostasis and neurological disease.
Main Methods:
- Utilized a genetic approach in Drosophila melanogaster to inactivate the AFG3L2 gene.
- Employed RNA interference (RNAi) for partial gene inactivation.
- Assessed developmental viability, neurodegeneration, mitochondrial protein aggregation, respiratory chain activity, mitochondrial transcription/translation, and the mitochondrial unfolded protein response (mito-UPR).
Main Results:
- Complete AFG3L2 inactivation resulted in early developmental lethality.
- Partial AFG3L2 inactivation led to adult flies with behavioral deficits, neurodegeneration, and accumulated unfolded mitochondrial proteins.
- Reduced respiratory chain activity was linked to impaired mitochondrial transcription and translation.
- Mitochondrial unfolded protein response (mito-UPR) and autophagy were activated.
- Overexpression of mito-UPR components partially rescued phenotypes, suggesting protein aggregation contributes to pathology.
Conclusions:
- AFG3L2 deficiency severely impacts mitochondrial function and leads to neurodegeneration.
- Mitochondrial stress pathways, including mito-UPR, are activated in response to AFG3L2 loss.
- Therapeutic strategies targeting mitochondrial stress pathways and gene expression show promise for AFG3L2-related neurological diseases.
More Related Videos
 06:53
Visualization of Mitochondrial Respiratory Function using Cytochrome C Oxidase / Succinate Dehydrogenase COX/SDH Double-labeling Histochemistry
Published on: November 23, 2011
 09:40
Phosphorus-31 Magnetic Resonance Spectroscopy: A Tool for Measuring In Vivo Mitochondrial Oxidative Phosphorylation Capacity in Human Skeletal Muscle
Published on: January 19, 2017
Related Concept Videos
 01:48
ATP Synthase: Mechanism
 01:30
The Electron Transport Chain
Inhibitors of the electron transport chain
Rotenone, a widely used pesticide, prevents electron transfer from Fe-S cluster to ubiquinone or Q...
 01:39
Mitochondrial Precursor Proteins
Most of the mitochondrial...
 01:46
Electron Transport Chain: Complex I and II
ROS generation is regulated and maintained at moderate levels necessary...
 02:59
Animal Mitochondrial Genetics
 01:19
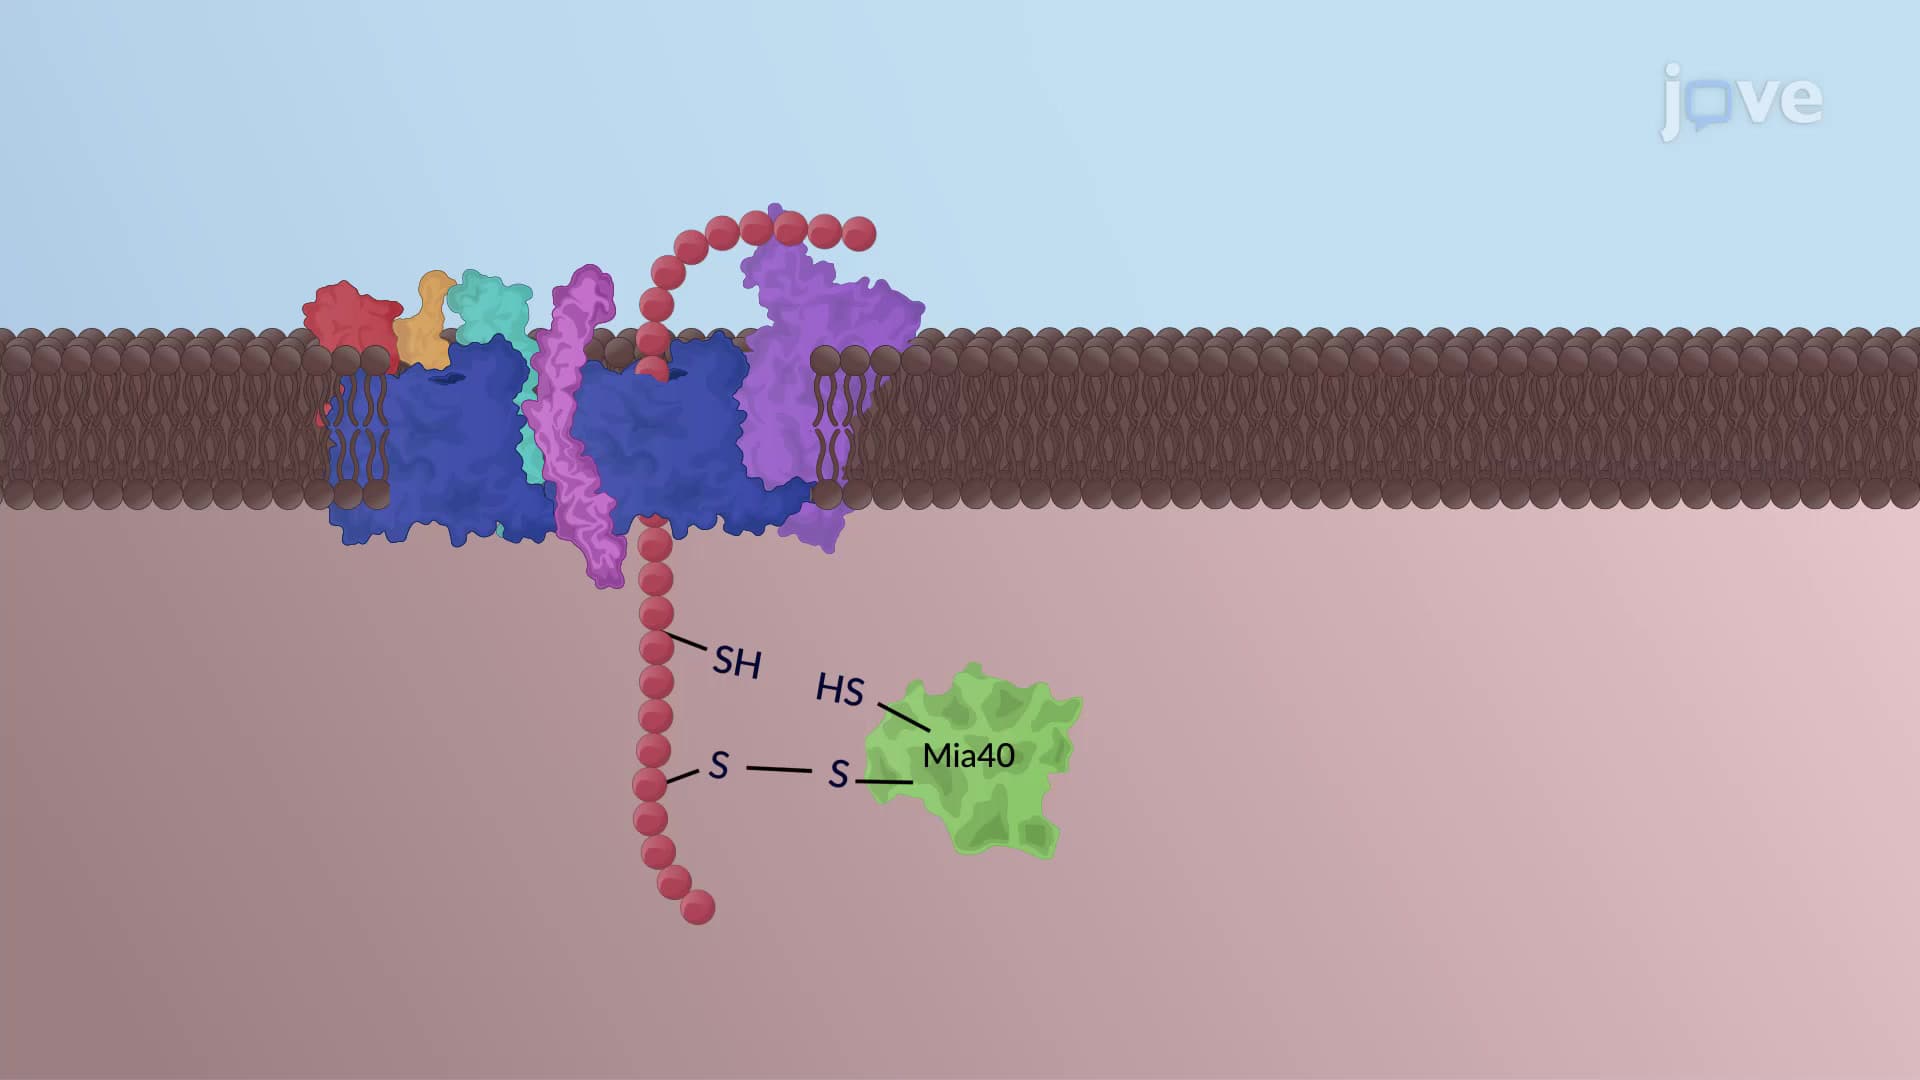
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...