Related Experiment Video
Updated: Dec 5, 2025


Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Binding Mode Exploration of B1 Receptor Antagonists' by the Use of Molecular Dynamics and Docking Simulation-How
Marica Gemei1, Carmine Talarico1, Laura Brandolini1
1Dompé Farmaceutici SpA, via Campo di Pile, 67100 L'Aquila, Italy.

Abstract:
The kinin B1 receptor plays a critical role in the chronic phase of pain and inflammation. The development of B1 antagonists peaked in recent years but almost all promising molecules failed in clinical trials. Little is known about these molecules' mechanisms of action and additional information will be necessary to exploit the potential of the B1 receptor. With the aim of contributing to the available knowledge of the pharmacology of B1 receptors, we designed and characterized a novel class of allosteric non-peptidic inhibitors with peculiar binding characteristics. Here, we report the binding mode analysis and pharmacological characterization of a new allosteric B1 antagonist, DFL20656. We analyzed the binding of DFL20656 by single point mutagenesis and radioligand binding assays and we further characterized its pharmacology in terms of IC50, B1 receptor internalization and in vivo activity in comparison with different known B1 antagonists. We highlighted how different binding modes of DFL20656 and a Merck compound (compound 14) within the same molecular pocket can affect the biological and pharmacological properties of B1 inhibitors. DFL20656, by its peculiar binding mode, involving tight interactions with N114, efficiently induced B1 receptor internalization and evoked a long-lasting effect in an in vivo model of neuropathic pain. The pharmacological characterization of different B1 antagonists highlighted the effects of their binding modes on activity, receptor occupancy and internalization. Our results suggest that part of the failure of most B1 inhibitors could be ascribed to a lack of knowledge about target function and engagement.
More Related Videos
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
 08:15
Author Spotlight: Network Pharmacology and Molecular Docking to Decipher the Action of Jiawei Shengjiang San Against Diabetic Kidney Disease
Published on: May 10, 2024
Related Concept Videos
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with...
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:29
Drug-Receptor Interactions
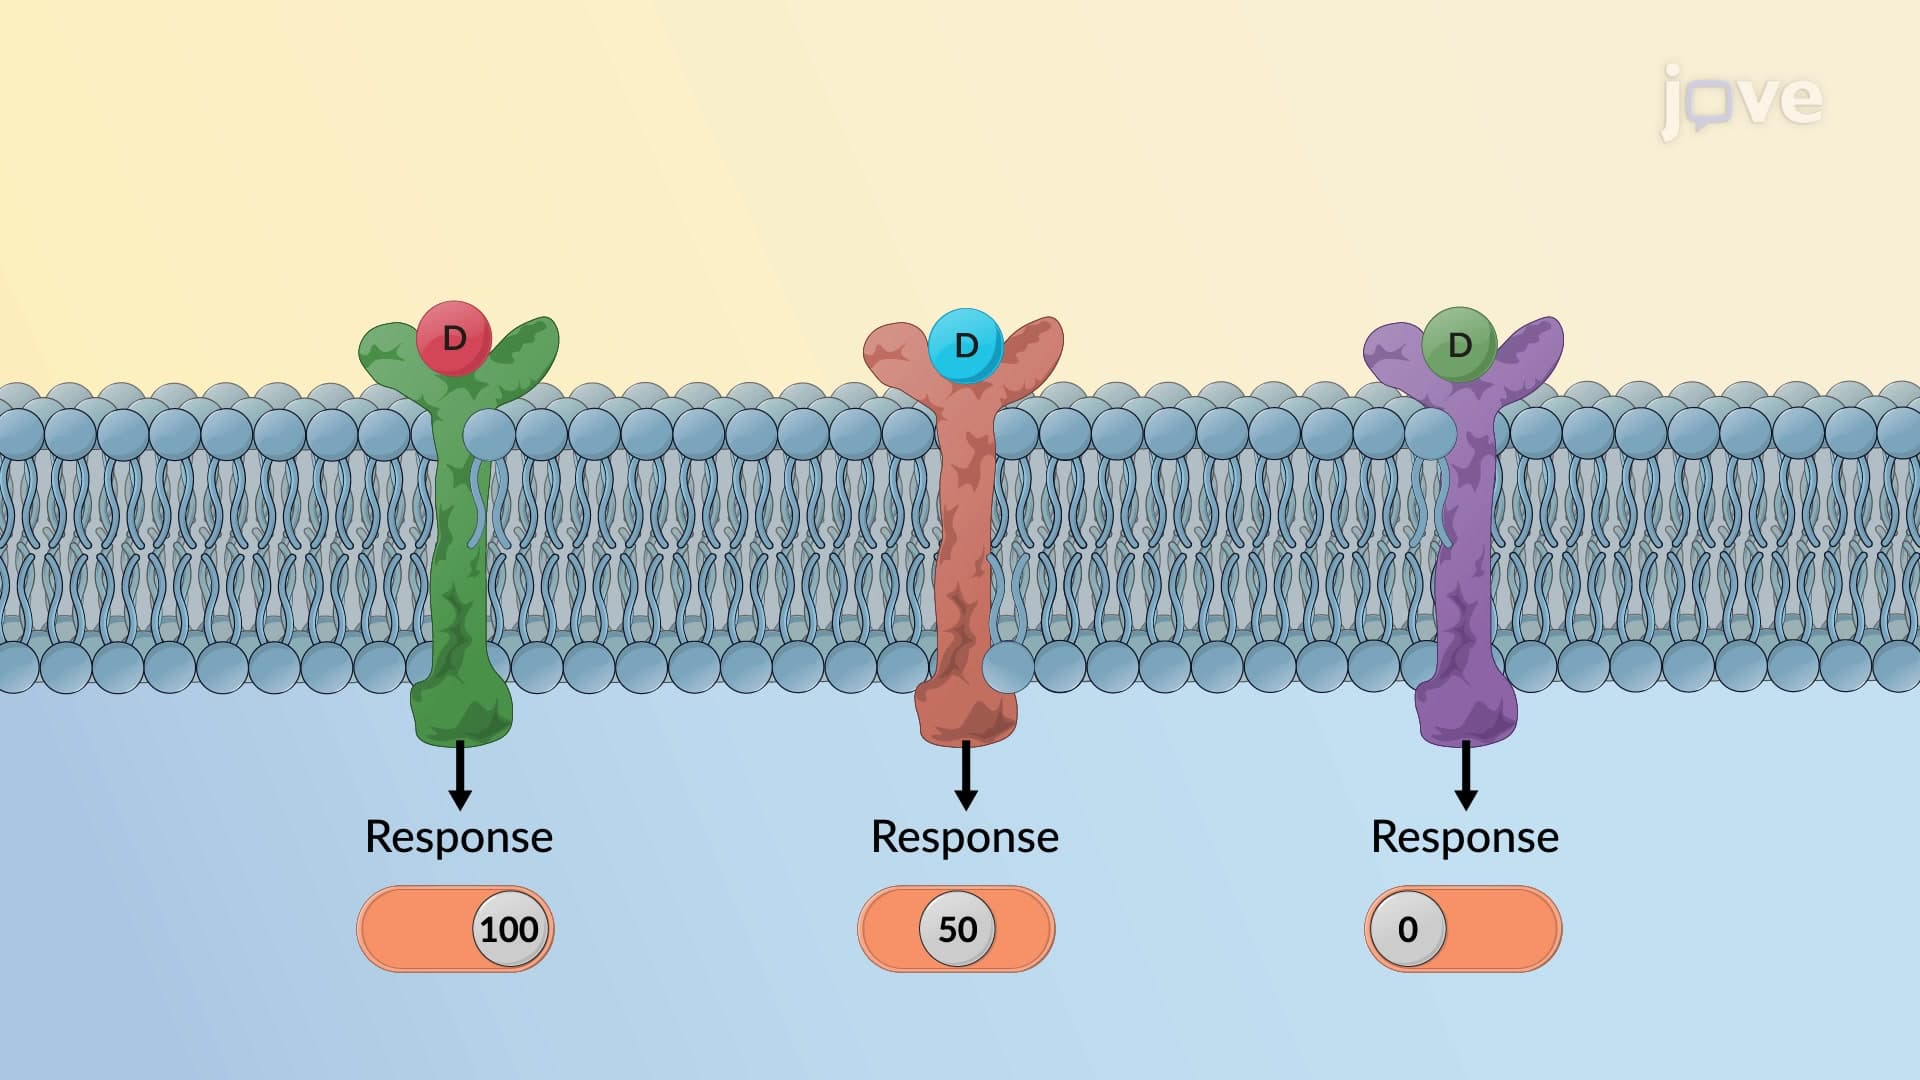
Several parameters, such as the drug's affinity for its receptor and its efficacy, which is its ability to activate the receptor, determine the drug's effect on the tissue....
 02:18
The Equilibrium Binding Constant and Binding Strength
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
02:40Ligand Binding Sites