Related Experiment Video
Updated: Dec 1, 2025
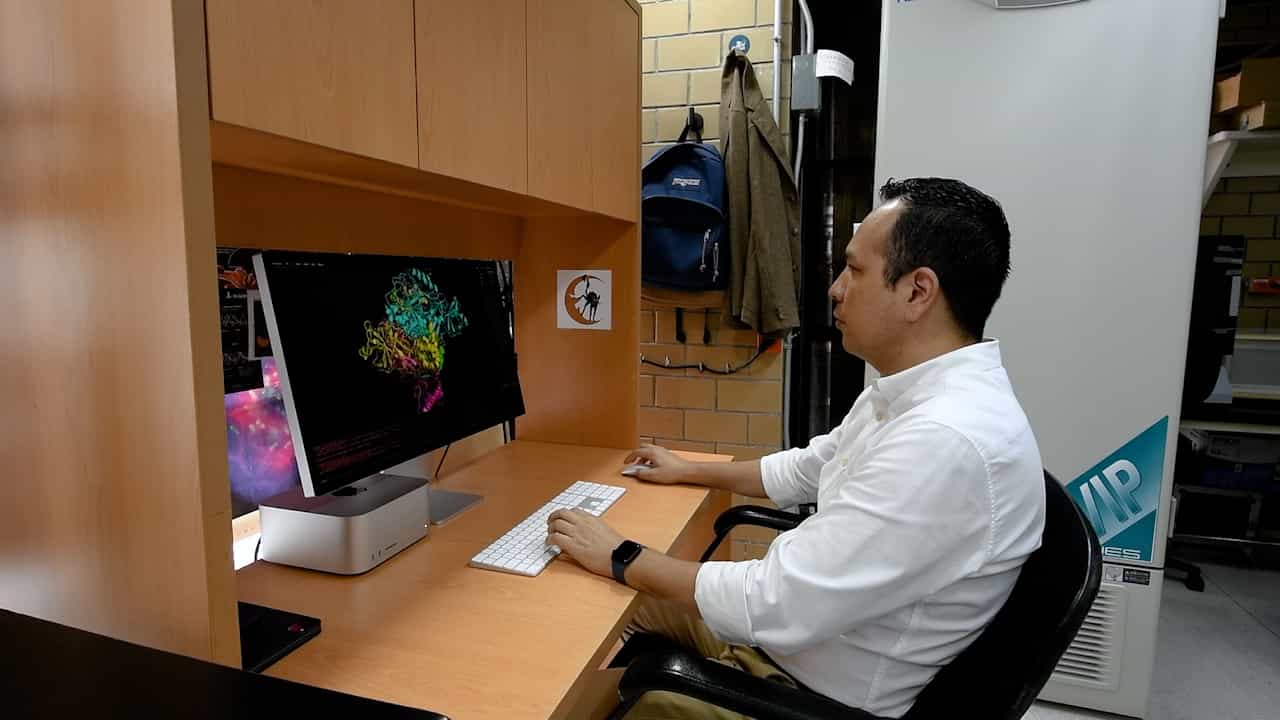
Pharmacophore Modeling for Targets with Extensive Ligand Libraries: A Case Study on SARS-CoV-2 Mpro
Published on: September 26, 2025
Drugs Repurposing Using QSAR, Docking and Molecular Dynamics for Possible Inhibitors of the SARS-CoV-2 Mpro Protease
Eduardo Tejera1,2, Cristian R Munteanu3,4, Andrés López-Cortés5,6
1Grupo de Bio-Quimioinformática, Universidad de Las Américas, Quito 170513, Ecuador.

Abstract:
Wuhan, China was the epicenter of the first zoonotic transmission of the severe acute respiratory syndrome coronavirus clade 2 (SARS-CoV-2) in December 2019 and it is the causative agent of the novel human coronavirus disease 2019 (COVID-19). Almost from the beginning of the COVID-19 outbreak several attempts were made to predict possible drugs capable of inhibiting the virus replication. In the present work a drug repurposing study is performed to identify potential SARS-CoV-2 protease inhibitors. We created a Quantitative Structure-Activity Relationship (QSAR) model based on a machine learning strategy using hundreds of inhibitor molecules of the main protease (Mpro) of the SARS-CoV coronavirus. The QSAR model was used for virtual screening of a large list of drugs from the DrugBank database. The best 20 candidates were then evaluated in-silico against the Mpro of SARS-CoV-2 by using docking and molecular dynamics analyses. Docking was done by using the Gold software, and the free energies of binding were predicted with the MM-PBSA method as implemented in AMBER. Our results indicate that levothyroxine, amobarbital and ABP-700 are the best potential inhibitors of the SARS-CoV-2 virus through their binding to the Mpro enzyme. Five other compounds showed also a negative but small free energy of binding: nikethamide, nifurtimox, rebimastat, apomine and rebastinib.
More Related Videos
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
