Related Experiment Video
Updated: Dec 1, 2025

Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Machine Learning Prediction of Nine Molecular Properties Based on the SMILES Representation of the QM9
Gabriel A Pinheiro1, Johnatan Mucelini2, Marinalva D Soares3
1Associate Laboratory for Computing and Applied Mathematics, National Institute for Space Research, PO BOX 515, 12227-010, São José dos Campos, SP, Brazil.

Abstract:
Machine learning (ML) models can potentially accelerate the discovery of tailored materials by learning a function that maps chemical compounds into their respective target properties. In this realm, a crucial step is encoding the molecular systems into the ML model, in which the molecular representation plays a crucial role. Most of the representations are based on the use of atomic coordinates (structure); however, it can increase ML training and predictions' computational cost. Herein, we investigate the impact of choosing free-coordinate descriptors based on the Simplified Molecular Input Line Entry System (SMILES) representation, which can substantially reduce the ML predictions' computational cost. Therefore, we evaluate a feed-forward neural network (FNN) model's prediction performance over five feature selection methods and nine ground-state properties (including energetic, electronic, and thermodynamic properties) from a public data set composed of ∼130k organic molecules. Our best results reached a mean absolute error, close to chemical accuracy, of ∼0.05 eV for the atomization energies (internal energy at 0 K, internal energy at 298.15 K, enthalpy at 298.15 K, and free energy at 298.15 K). Moreover, for the atomization energies, the results obtained an out-of-sample error nine times less than the same FNN model trained with the Coulomb matrix, a traditional coordinate-based descriptor. Furthermore, our results showed how limited the model's accuracy is by employing such low computational cost representation that carries less information about the molecular structure than the most state-of-the-art methods.


Related Concept Videos
 02:27
Predicting Molecular Geometry
 02:35
Molecular Orbital Theory I
 02:36
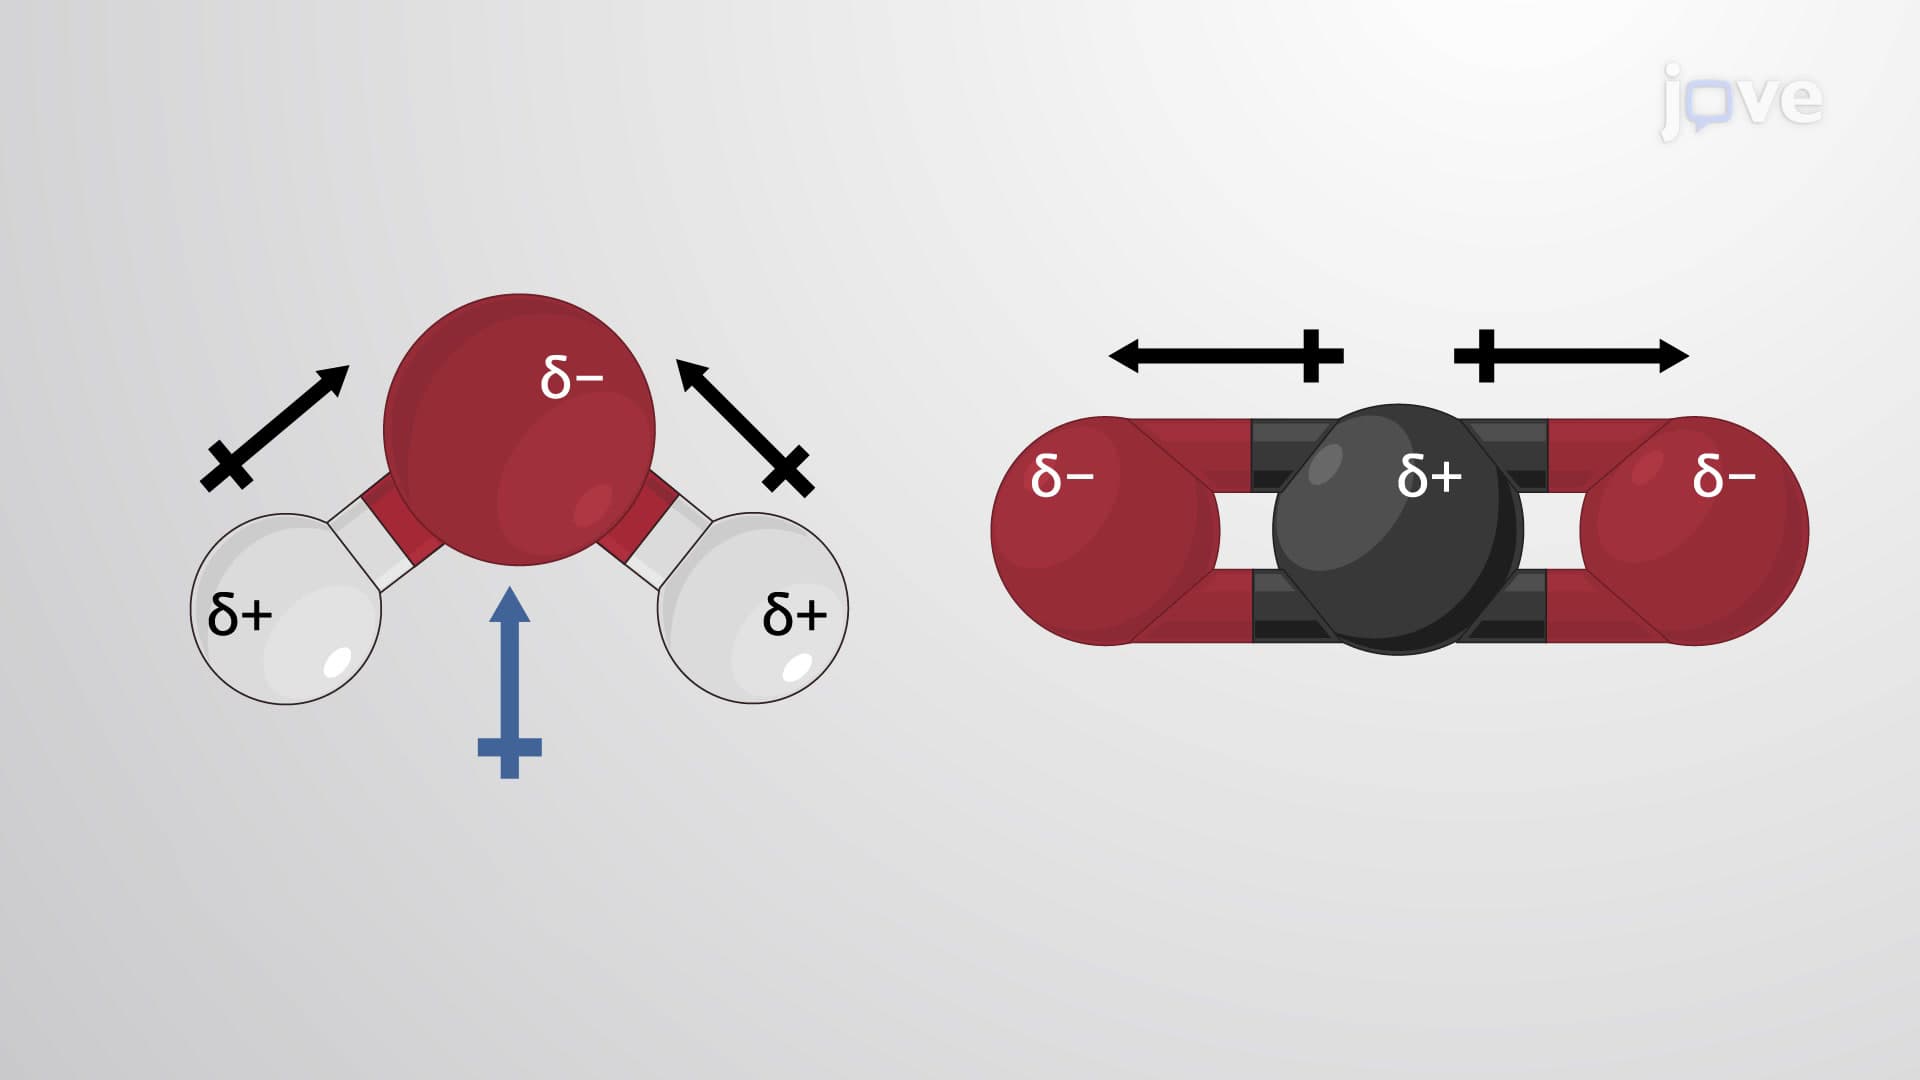
Molecular Geometry and Dipole Moments
 02:00
Molecular Models
 02:27
Predicting Products: SN1 vs. SN2
With increased substitution on the alkyl halide,...
 02:24
Predicting Reaction Outcomes