Related Experiment Video
Updated: Nov 30, 2025
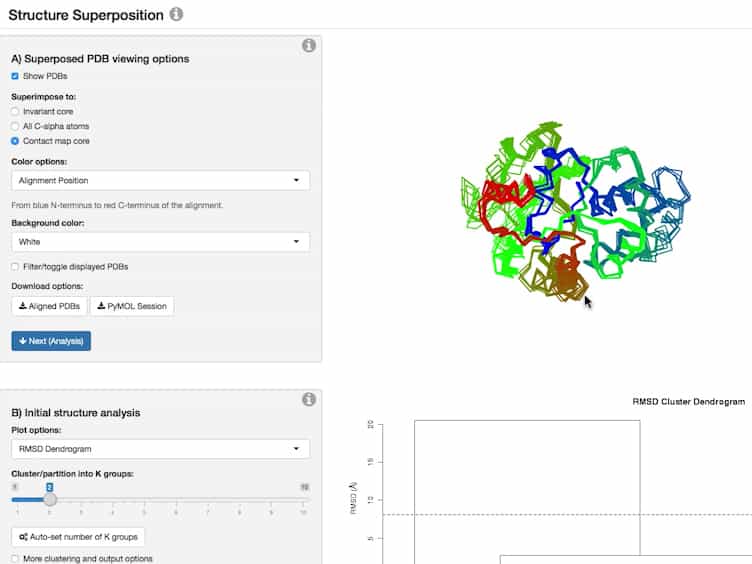

Investigating Protein Sequence-structure-dynamics Relationships with Bio3D-web
Published on: July 16, 2017
Validation of DBFOLD: An efficient algorithm for computing folding pathways of complex proteins
Amir Bitran1,2, William M Jacobs3, Eugene Shakhnovich1
1Department of Chemistry and Chemical Biology, Harvard University, Cambridge, Massachusetts, United States of America.

Abstract:
Atomistic simulations can provide valuable, experimentally-verifiable insights into protein folding mechanisms, but existing ab initio simulation methods are restricted to only the smallest proteins due to severe computational speed limits. The folding of larger proteins has been studied using native-centric potential functions, but such models omit the potentially crucial role of non-native interactions. Here, we present an algorithm, entitled DBFOLD, which can predict folding pathways for a wide range of proteins while accounting for the effects of non-native contacts. In addition, DBFOLD can predict the relative rates of different transitions within a protein's folding pathway. To accomplish this, rather than directly simulating folding, our method combines equilibrium Monte-Carlo simulations, which deploy enhanced sampling, with unfolding simulations at high temperatures. We show that under certain conditions, trajectories from these two types of simulations can be jointly analyzed to compute unknown folding rates from detailed balance. This requires inferring free energies from the equilibrium simulations, and extrapolating transition rates from the unfolding simulations to lower, physiologically-reasonable temperatures at which the native state is marginally stable. As a proof of principle, we show that our method can accurately predict folding pathways and Monte-Carlo rates for the well-characterized Streptococcal protein G. We then show that our method significantly reduces the amount of computation time required to compute the folding pathways of large, misfolding-prone proteins that lie beyond the reach of existing direct simulation. Our algorithm, which is available online, can generate detailed atomistic models of protein folding mechanisms while shedding light on the role of non-native intermediates which may crucially affect organismal fitness and are frequently implicated in disease.


Related Concept Videos
 01:22
Protein Folding
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
 01:29
Protein Folding Quality Check in the RER
 03:00
Molecular Chaperones and Protein Folding
The...
03:00Molecular Chaperones and Protein Folding
 02:26
Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to...