Related Experiment Video
Updated: Nov 29, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
A new release of MOPAC incorporating the INDO/S semiempirical model with CI excited states
1Department of Chemistry, Brandeis University, Waltham, Massachusetts, USA.

Abstract:
The semiempirical INDO/S Hamiltonian is incorporated into a new release of MOPAC2016. The MOPAC2016 software package has long been at the forefront of semiempirical quantum chemical methods (SEQMs) for small molecules, proteins, and solids and until this release has included only NDDO-type SEQMs. The new code enables the calculation of excited states using the INDO/S Hamiltonian combined with a configuration interaction (CI) approach using single excitations (CIS), single and double excitations (CISD), or multiple reference determinants (MRCI) where reference determinants are generated using a complete active space (CAS) approach. The capacity to perform excited-state calculations beyond the CIS level makes INDO/CI one of the few low-cost computational methods capable of accurately modeling states with substantial double-excitation character. Solvent corrections to the ground-state and excited-state energies can be computed using the COSMO implicit solvent model, incorporating state-specific corrections to the excited states based on the solvent refractive index. This code produces physically reasonable electronic structures, absorption spectra, and solvatochromic shifts at low computational costs for systems up to hundreds of atoms, and for both organic molecules and metal clusters.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
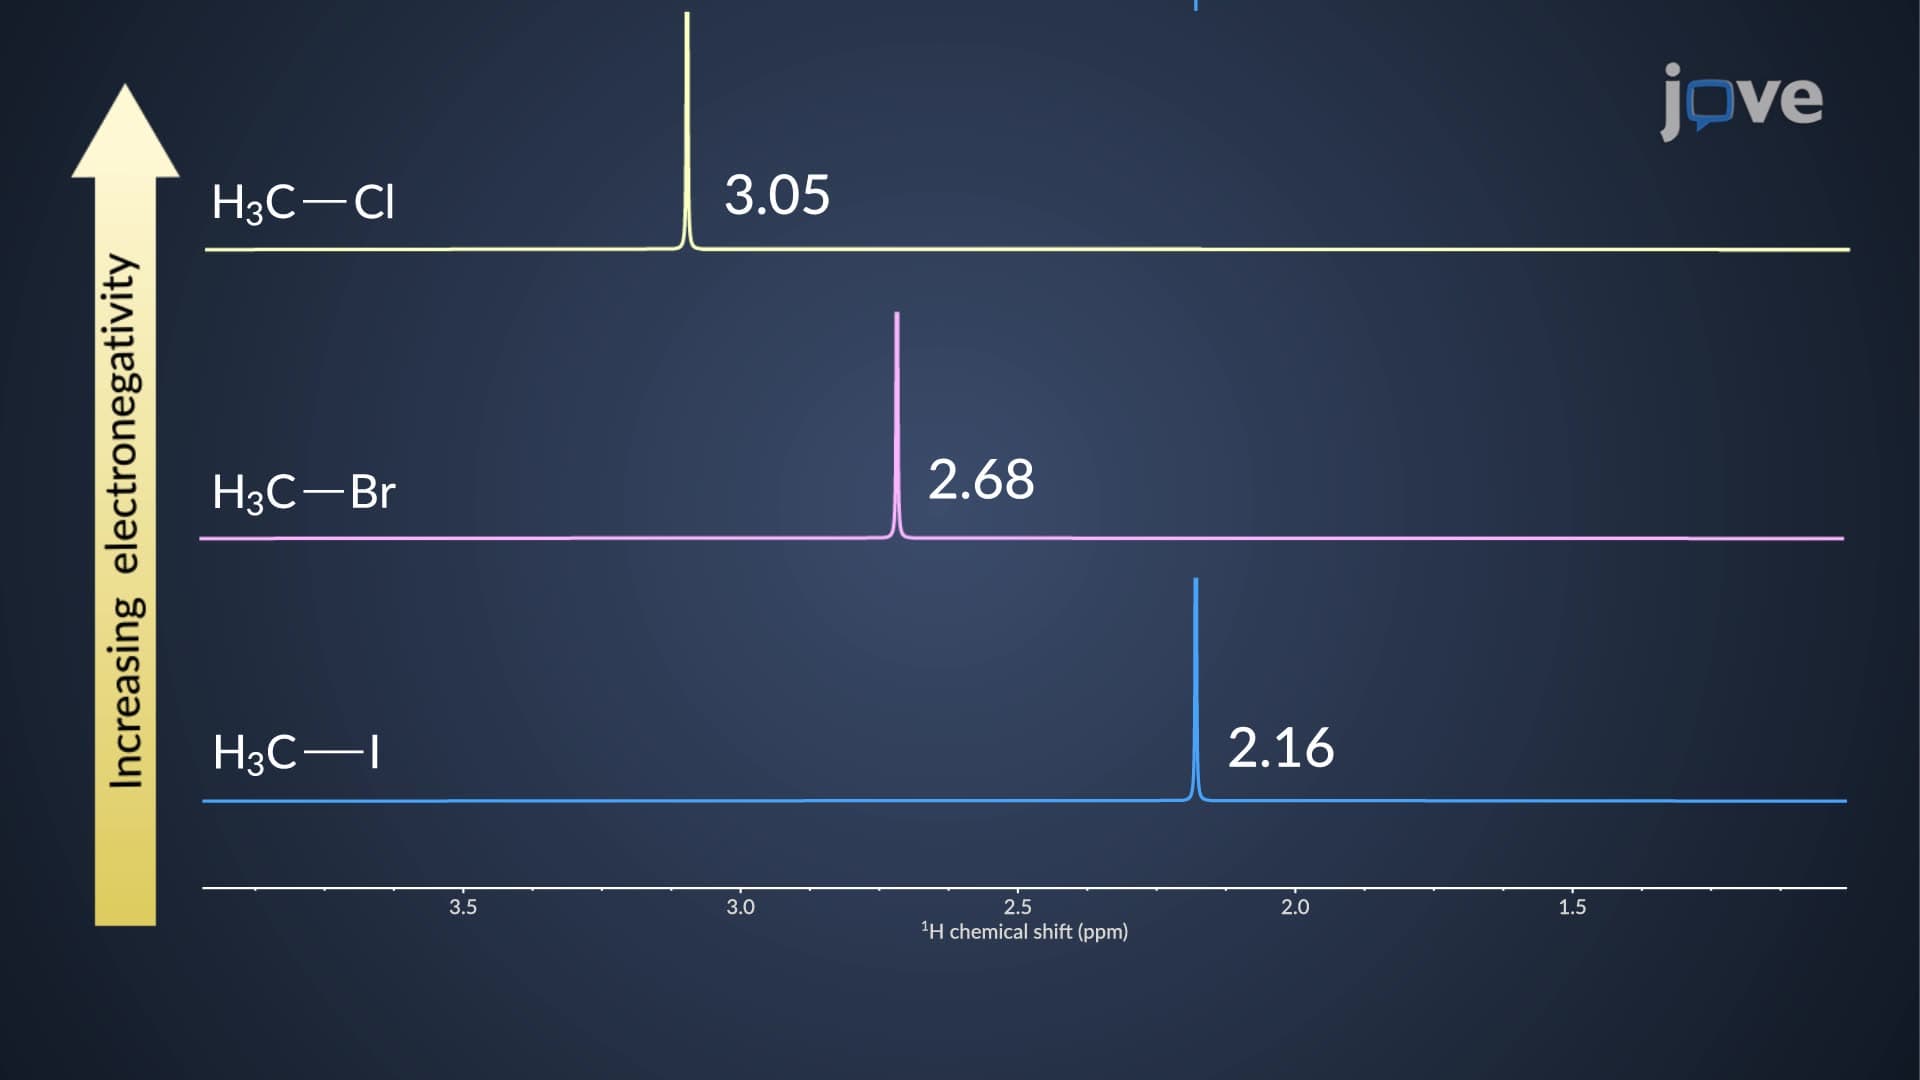 01:27
Inductive Effects on Chemical Shift: Overview
 02:35
Molecular Orbital Theory I
 01:24
One-Compartment Open Model: Wagner-Nelson and Loo Riegelman Method for ka Estimation
On...
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...
 03:51
Molecular Orbital Theory II
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...