Related Experiment Video
Updated: Nov 27, 2025

Mass Spectrometry and Luminogenic-based Approaches to Characterize Phase I Metabolic Competency of In Vitro Cell Cultures
Published on: March 28, 2017
Deciphering Reaction Determinants of Altered-Activity CYP2D6 Variants by Well-Tempered Metadynamics Simulation and
Charleen G Don1, Martin Smieško1
1Computational Pharmacy Group, Department of Pharmaceutical Sciences, University of Basel, Klingelbergstrasse 50, 4056 Basel, Switzerland.

Abstract:
The xenobiotic metabolizing enzyme CYP2D6 is the P450 cytochrome family member with the highest rate of polymorphism. This causes changes in the enzyme activity and specificity, which can ultimately lead to adverse reactions during drug treatment. To avoid or lower CYP-related toxicity risks, prediction of the most likely positions within a molecule where a metabolic reaction might occur is paramount. In order to obtain accurate predictions, it is crucial to understand all phenomena within the active site of the enzyme that contribute to an efficient substrate recognition and the subsequent catalytic reaction together with their relative weight within the overall thermodynamic context. This study aims to define the weight of the driving forces upon the C-H bond activation within CYP2D6 wild-type and a clinically relevant allelic variant with increased activity (CYP2D6*53) featuring two amino acid mutations in close vicinity of the heme. First, we investigated the steric and electrostatic complementarity of the substrate bufuralol using well-tempered metadynamics simulations with the aim to obtain the free energy profiles for each site of metabolism (SoM) within the different active sites. Second, the stereoelectronic complementarity was determined for each SoM within the two different active-site environments. Relying on the well-tempered metadynamics simulation energy profiles of each SoM, we identified the binding mode that was closest to the preferred transition-state geometry for efficient C-H bond activation. The binding modes were then used as starting structures for the quantum mechanics/molecular mechanics calculations performed to quantify the corresponding activation barriers. Our results show the relevance of the steric component in orienting the SoM in an energetically accessible position toward the heme. However, the corresponding intrinsic reactivity and electronic complementarity within the active site must be accurately evaluated in order to obtain a meaningful reaction prediction, from which the predominant SoM can be determined. The F120I mutation lowered the activation barrier for the major site and one of the minor SoMs. However, it had an impact neither on the CYP2D6 enantioselectivity preference of the oxidation reaction nor on the stereoselectivity from the substrate point of view.
More Related Videos
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
 05:51
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
Related Concept Videos
 01:17
Drug Metabolism: Phase I Reactions
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:19
Phase I Reactions: Oxidation of Aliphatic and Aromatic Carbon-Containing Systems
Oxidation reactions are fundamental in aromatic carbon-containing systems. An example is the hydroxylation of phenobarbital, a process that transforms it into...
 01:34
Woodward–Hoffmann Selection Rules and Microscopic Reversibility
 02:35
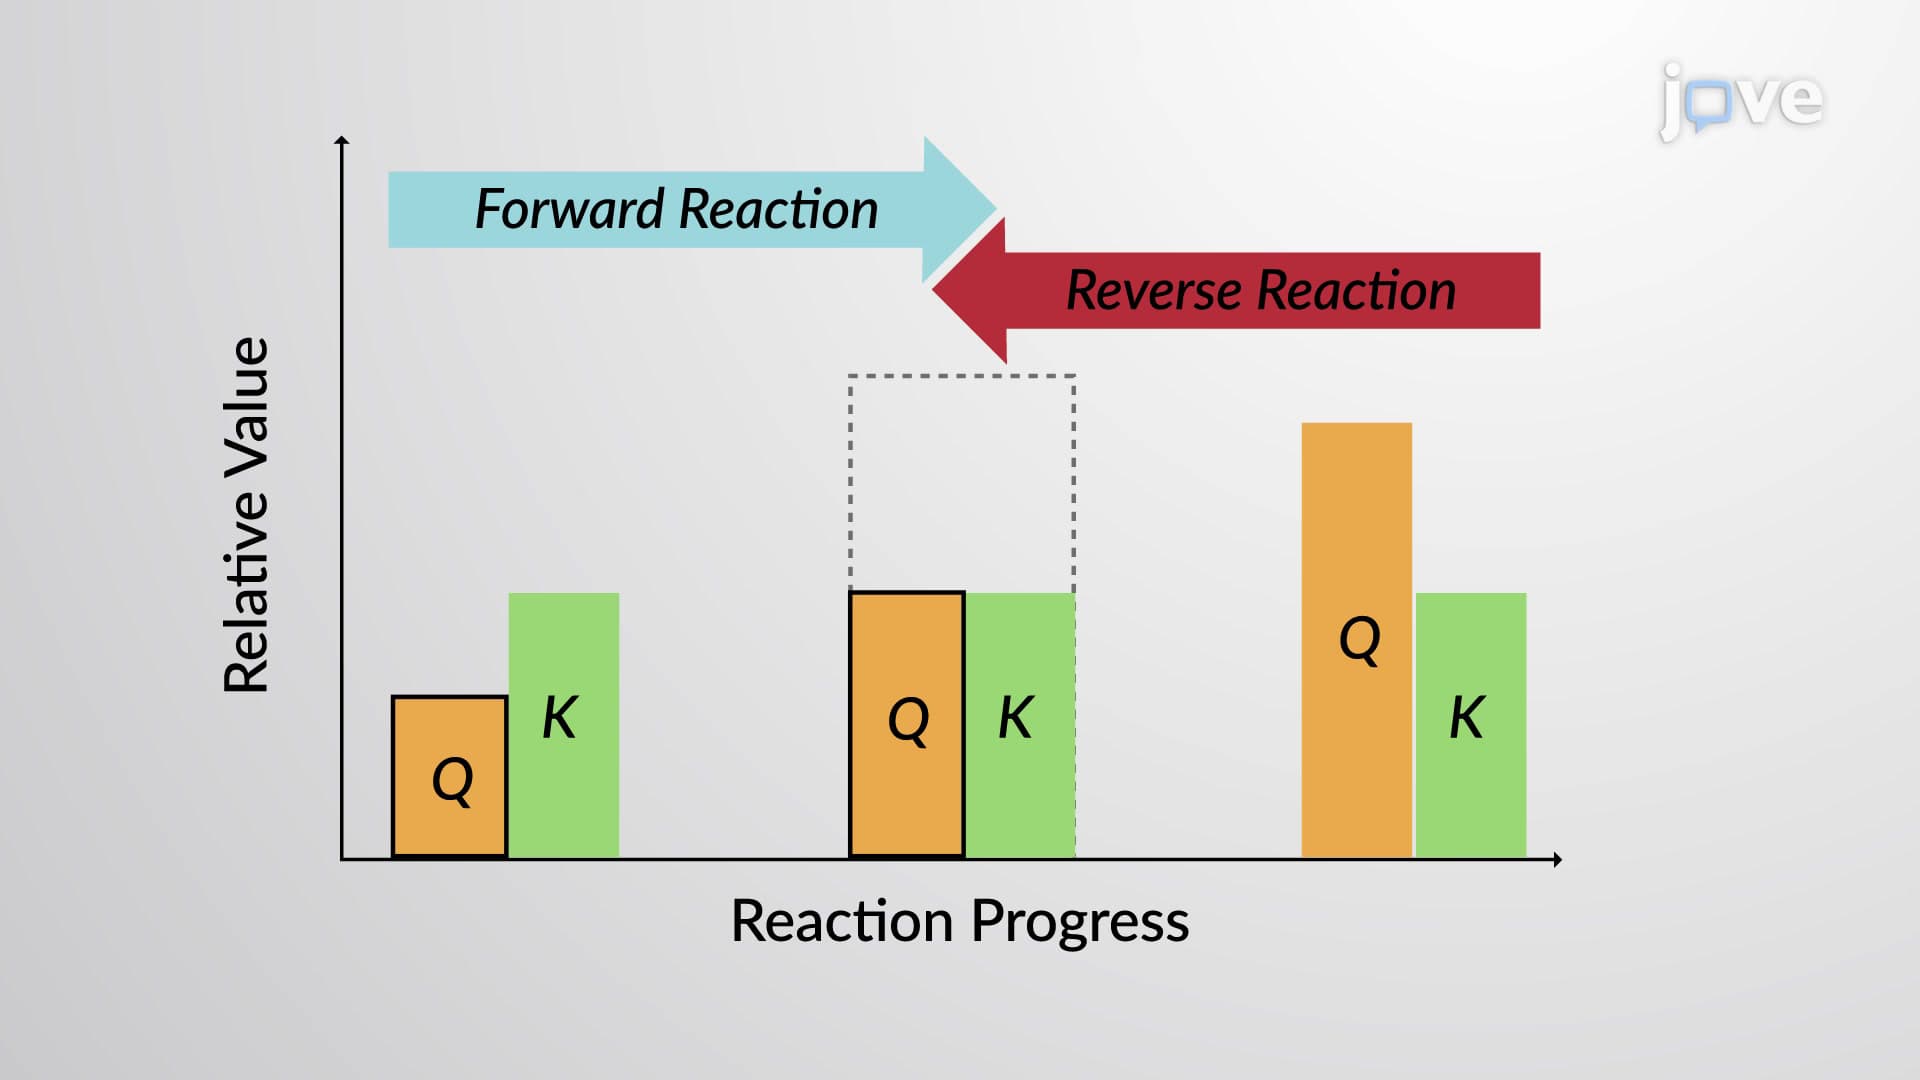
Reaction Quotient
 01:14
Drug Metabolism: Phase II Reactions