Related Experiment Video
Updated: Nov 26, 2025


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
London Dispersion Corrections to Density Functional Theory for Transition Metals Based on Fitting to Experimental
Hao Yang1, Tao Cheng1, William A Goddard2
1Institute of Functional Nano and Soft Materials (FUNSOM), Soochow University, Suzhou 215123, China.

Abstract:
Standard implementations of generalized gradient approximation (GGA)-based density functional theory (DFT) describe well strongly bound molecules and solids but fail to describe long-range London dispersion or van der Waals (vdW) attraction interactions that are important in molecular crystals and two-dimensional solids. To provide accurate values for the vdW distance and energies for the metals Cu, Ag, Au, Ni, Pd, and Pt, we determined empirical vdW corrections to Perdew, Burke, and Ernzerhof (PBE) DFT by fitting the experimental adsorption enthalpies measured by temperature-programmed desorption (TPD) from benzene monolayers by Campbell and co-workers ( J. Phys. Chem. C 2016, 120, 25161-25172). Benzene physisorbed to these metals without chemical reaction; therefore, we consider the bonding to be vdW. We use the low gradient form for the vdW corrections, EvdW-LG = -C6LG/[R6 + RvdwLG6] with just two parameters per atom (C6LG and RvdwLG). This LG form leads to negligible changes in bond distances and angles, so adjusting the parameters should not sacrifice accuracy for the bonding interactions. We demonstrate that the parameters fitted to benzene also describe well the physisorption enthalpies for other hydrocarbons (naphthalene, cyclohexane, methane, ethane, and propane) on Pt. We also report low gradient vdW correction parameters for the noble gases that fit the equilibrium lattice parameter and heat of vaporization of the crystals.
More Related Videos


Related Concept Videos
 03:51
Molecular Orbital Theory II
 01:10
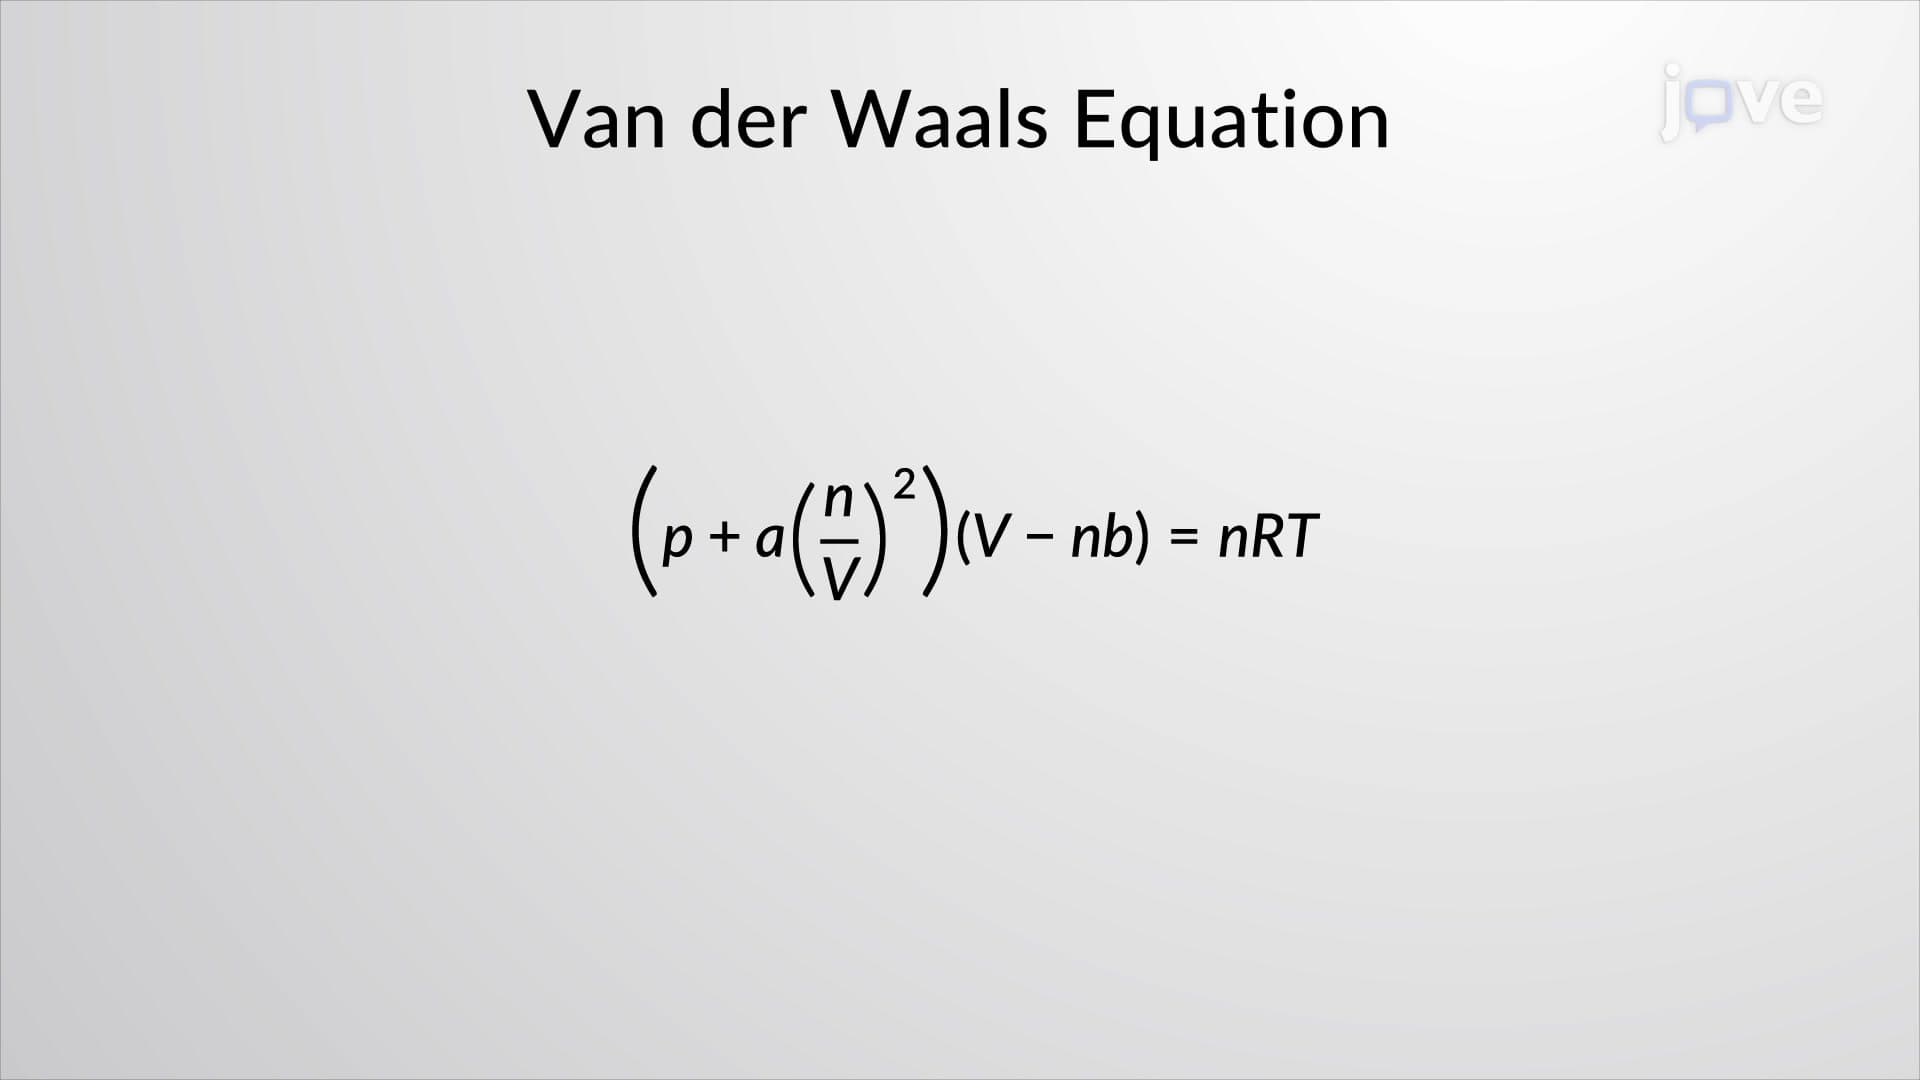
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:18
Structure of Benzene: Molecular Orbital Model
 01:24
Van der Waals Interactions
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 01:34
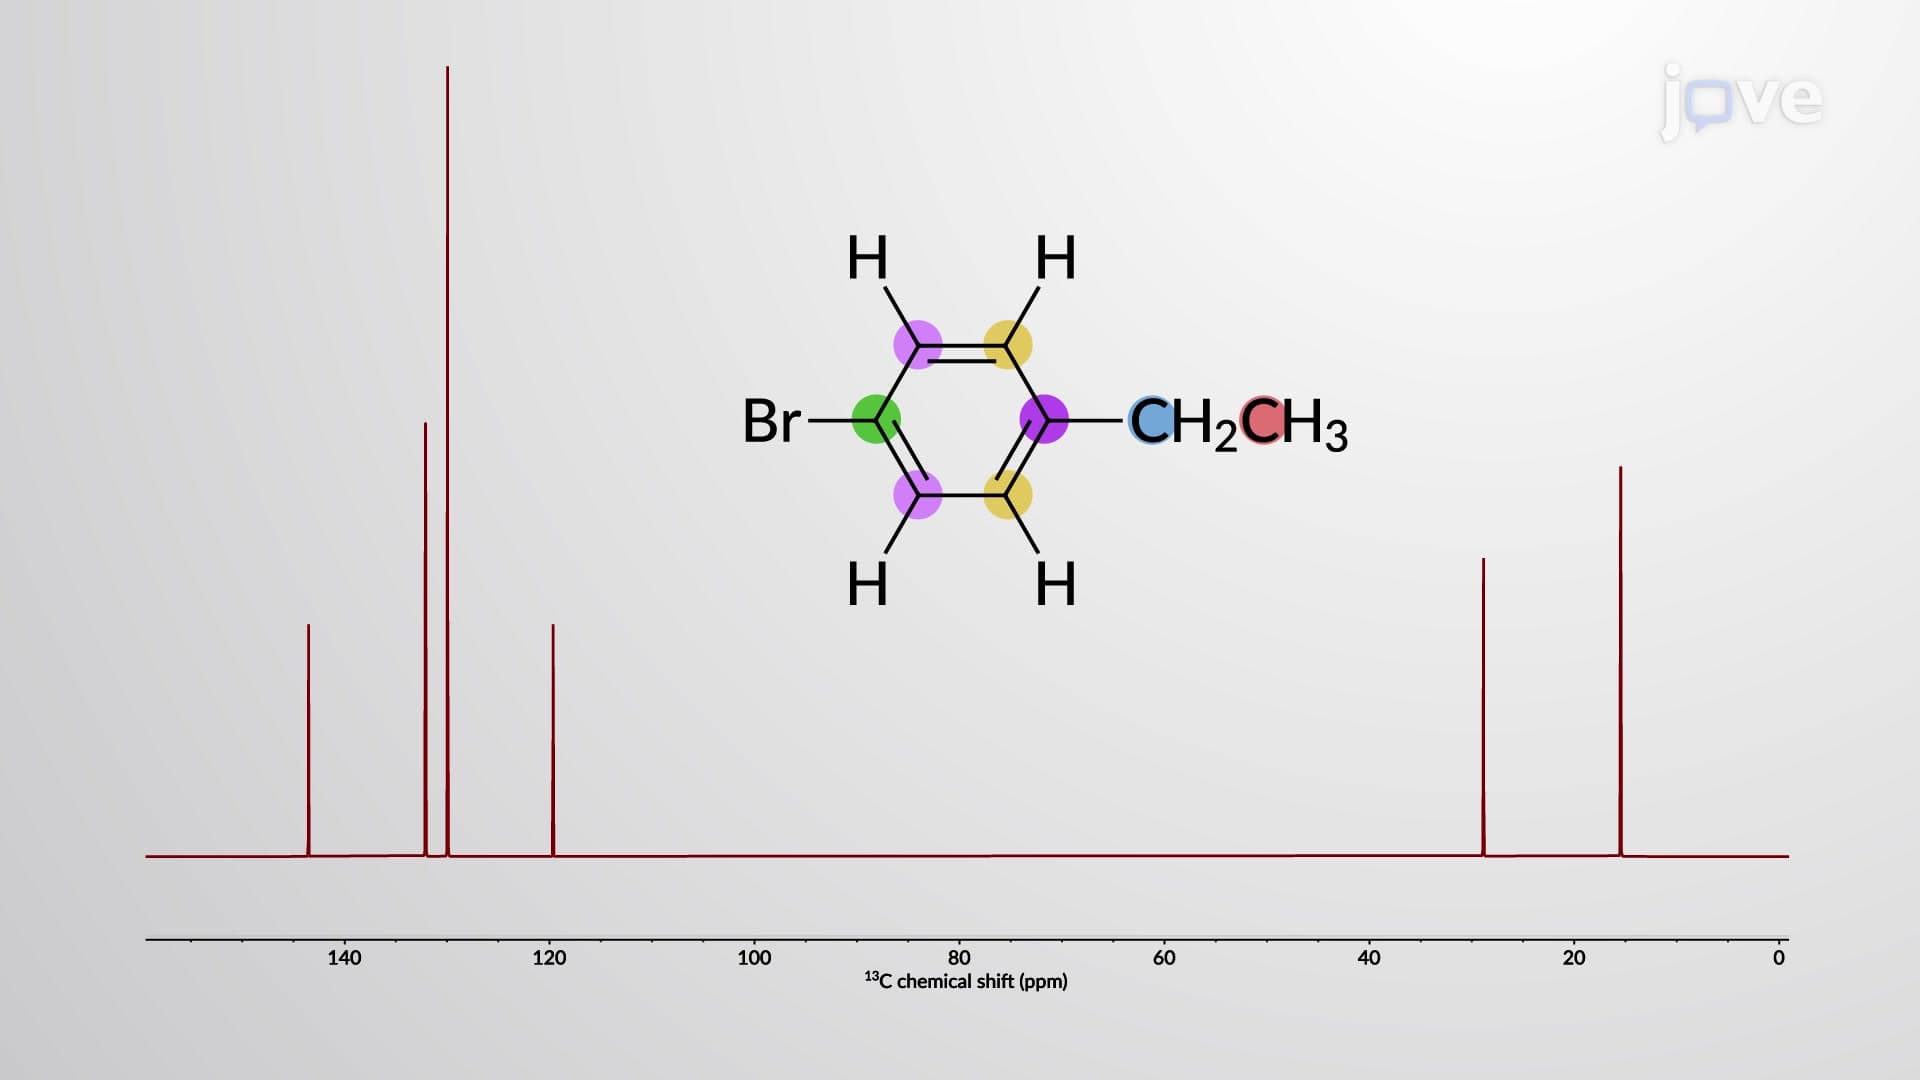
NMR Spectroscopy of Benzene Derivatives