Related Experiment Video
Updated: Nov 26, 2025

Rapid Isolation of the Mitoribosome from HEK Cells
Published on: October 4, 2018
The Diseased Mitoribosome
Alberto Ferrari1, Samuel Del'Olio1,2, Antoni Barrientos1,3
1Department of Neurology, University of Miami Miller School of Medicine, FL, USA.

Abstract:
Mitochondria control life and death in eukaryotic cells. Harboring a unique circular genome, a by-product of an ancient endosymbiotic event, mitochondria maintains a specialized and evolutionary divergent protein synthesis machinery, the mitoribosome. Mitoribosome biogenesis depends on elements encoded in both the mitochondrial genome (the RNA components) and the nuclear genome (all ribosomal proteins and assembly factors). Recent cryo-EM structures of mammalian mitoribosomes have illuminated their composition and provided hints regarding their assembly and elusive mitochondrial translation mechanisms. A growing body of literature involves the mitoribosome in inherited primary mitochondrial disorders. Mutations in genes encoding mitoribosomal RNAs, proteins, and assembly factors impede mitoribosome biogenesis, causing protein synthesis defects that lead to respiratory chain failure and mitochondrial disorders such as encephalo- and cardiomyopathy, deafness, neuropathy, and developmental delays. In this article, we review the current fundamental understanding of mitoribosome assembly and function, and the clinical landscape of mitochondrial disorders driven by mutations in mitoribosome components and assembly factors, to portray how basic and clinical studies combined help us better understand both mitochondrial biology and medicine.
Insights
Mitochondria
Area of Science:
- Cell Biology
- Genetics
- Biochemistry
Background:
- Mitochondria possess a unique genome and protein synthesis machinery called the mitoribosome.
- Mitoribosome biogenesis requires nuclear and mitochondrial genetic elements.
- Recent cryo-electron microscopy (cryo-EM) studies reveal mitoribosome structure and function.
Purpose of the Study:
- To review current knowledge of mitoribosome assembly and function.
- To explore the clinical aspects of mitochondrial disorders linked to mitoribosome defects.
- To highlight the synergy between basic research and clinical studies in mitochondrial medicine.
Main Methods:
- Literature review of fundamental mitoribosome research.
- Analysis of clinical data on mitochondrial disorders.
- Integration of structural biology findings (cryo-EM) with genetic and clinical information.
Main Results:
- Mitoribosome dysfunction due to mutations causes severe inherited mitochondrial disorders.
- Defects in mitoribosome biogenesis lead to protein synthesis impairment and respiratory chain failure.
- Specific disorders include encephalomyocardiopathy, deafness, neuropathy, and developmental delays.
Conclusions:
- Understanding mitoribosome assembly and function is crucial for diagnosing and treating mitochondrial diseases.
- Mutations in mitoribosome components provide insights into mitochondrial biology and disease mechanisms.
- Combined basic and clinical research advances both mitochondrial medicine and fundamental biology.
Related Concept Videos
 01:31
Translation
Translation Produces the Building Blocks of Life
Proteins are...
01:31Translation
Translation is the process of synthesizing proteins from the genetic information carried by messenger RNA (mRNA). Following transcription, it constitutes the final step in the expression of genes. This process is carried out by ribosomes, complexes of protein and specialized RNA molecules. Ribosomes, transfer RNA (tRNA), and other proteins produce a chain of amino acids—the polypeptide—as the end product of translation.
Translation Produces the Building Blocks of...
 01:19
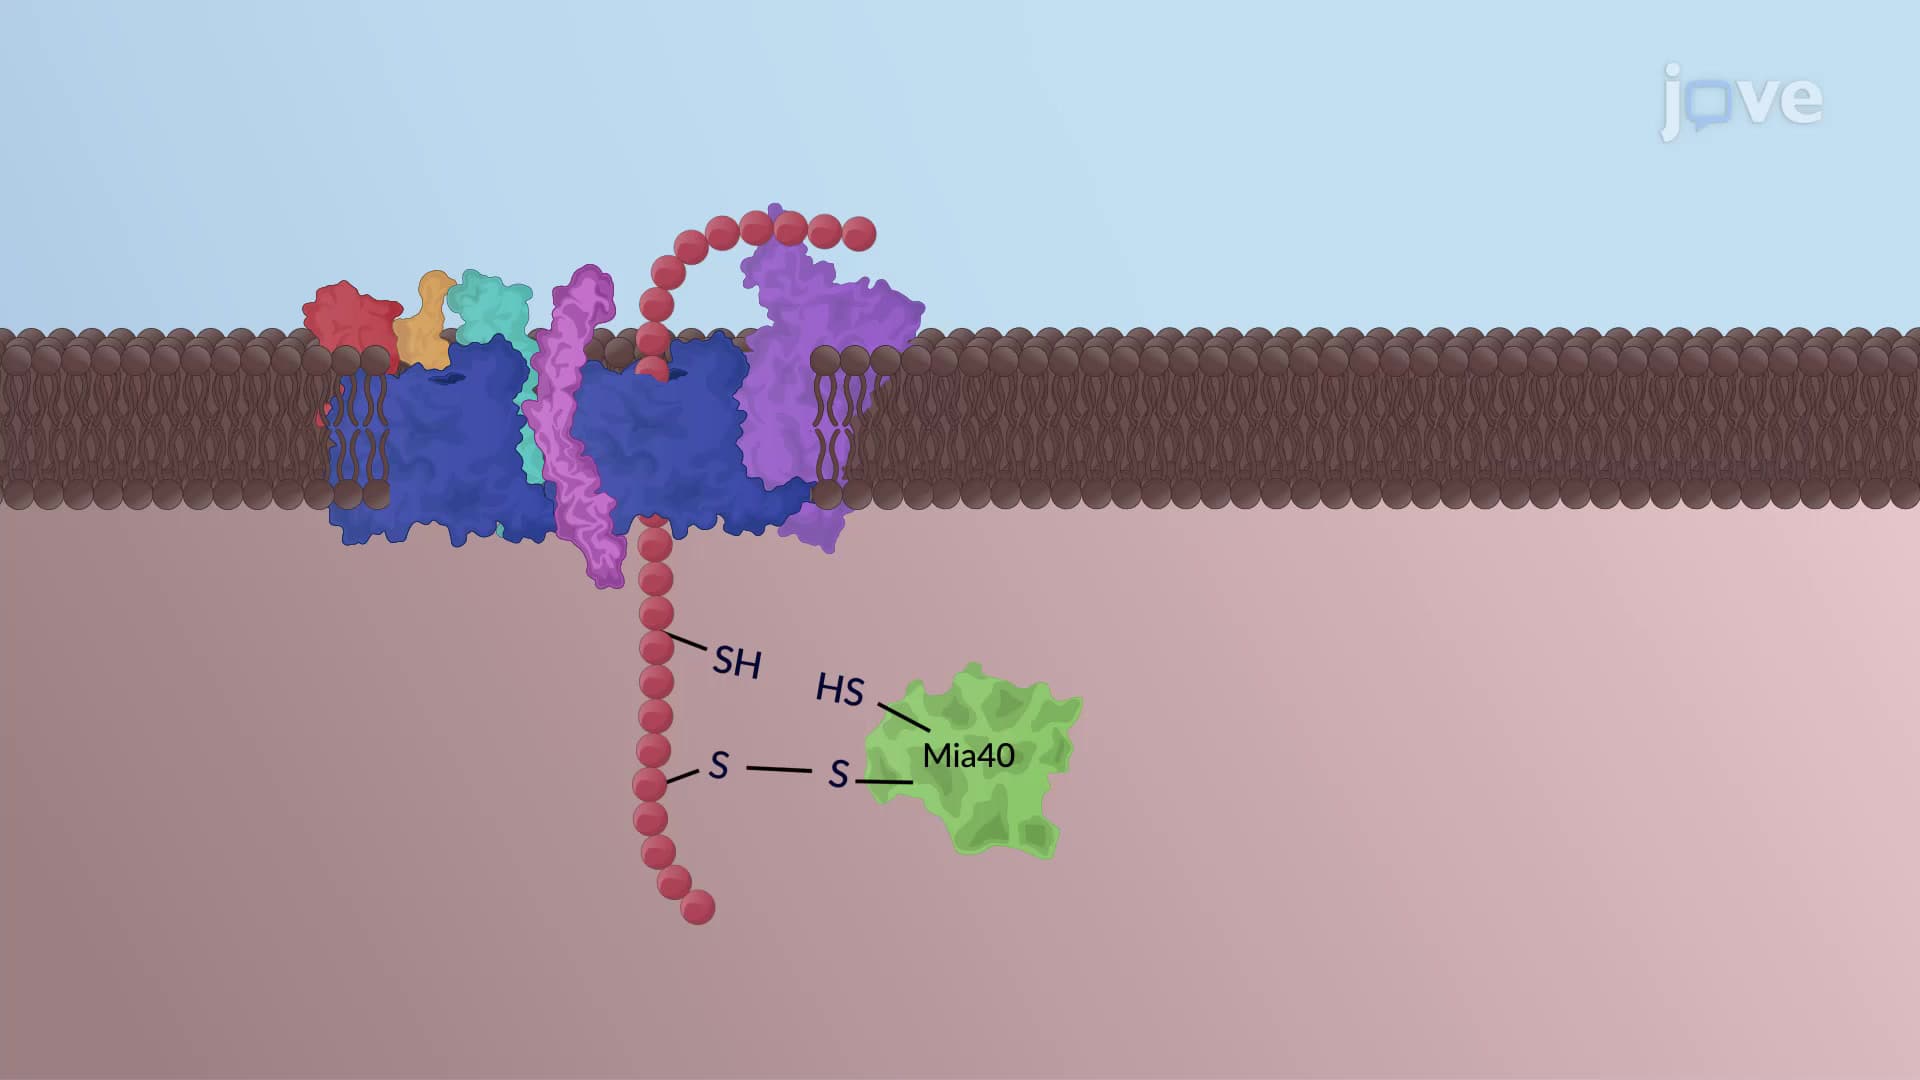
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
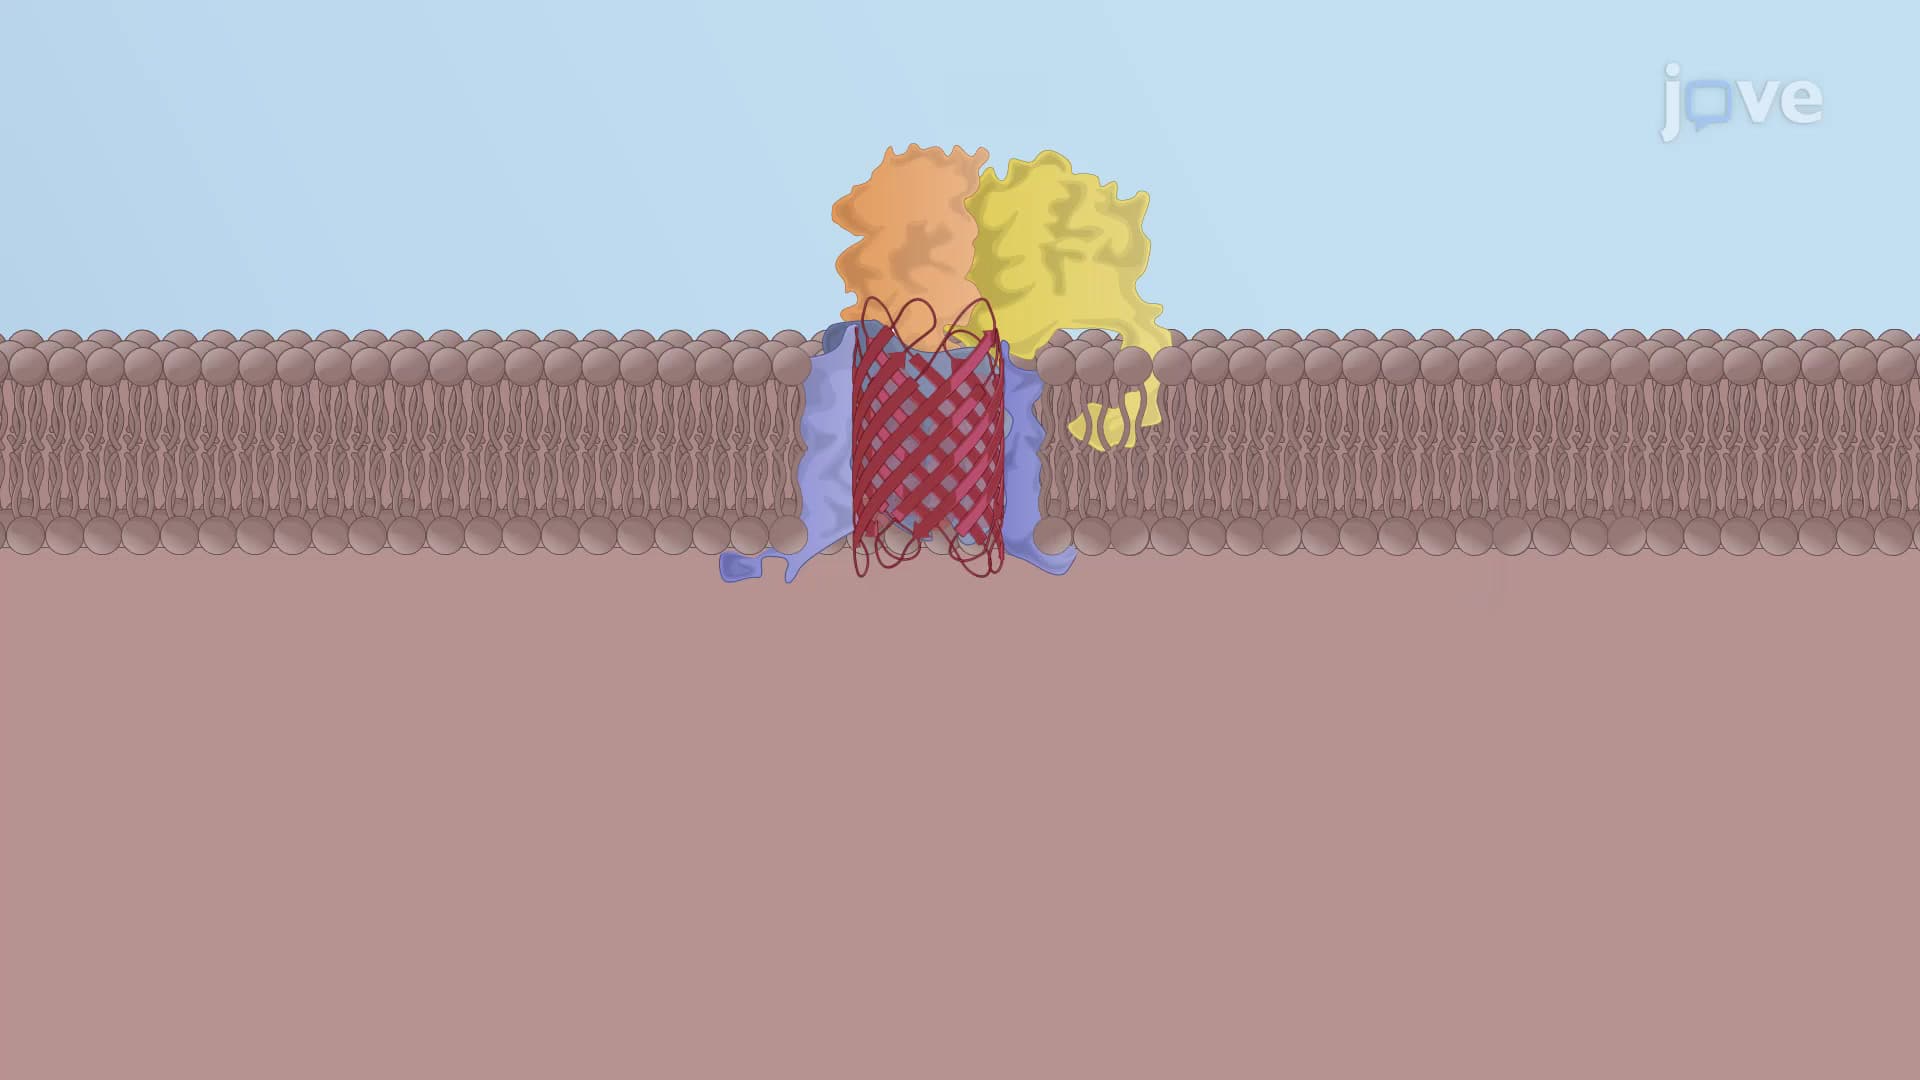 01:12
Porin Insertion in the Outer Mitochondrial Membrane
Three models describe the assembly of porins by the SAM complex and their insertion into the outer membrane. Model 1 suggests that porins are assembled outside the SAM channel as the...
 01:48
ATP Synthase: Mechanism
 01:32
Export of Misfolded Proteins out of the ER