Related Experiment Video
Updated: Nov 23, 2025

An In Vitro Assay to Detect tRNA-Isopentenyl Transferase Activity
Published on: October 8, 2018
Disease-associated mutations in mitochondrial precursor tRNAs affect binding, m1R9 methylation, and tRNA processing
Agnes Karasik1, Catherine A Wilhelm2, Carol A Fierke3,4
1Department of Biochemistry and Molecular Biology, Uniformed Services University of the Health Sciences, Bethesda, Maryland 20814, USA.

Abstract:
Mitochondrial diseases linked to mutations in mitochondrial (mt) tRNA sequences are common. However, the contributions of these tRNA mutations to the development of diseases is mostly unknown. Mutations may affect interactions with (mt)tRNA maturation enzymes or protein synthesis machinery leading to mitochondrial dysfunction. In human mitochondria, in most cases the first step of tRNA processing is the removal of the 5' leader of precursor tRNAs (pre-tRNA) catalyzed by the three-component enzyme, mtRNase P. Additionally, one component of mtRNase P, mitochondrial RNase P protein 1 (MRPP1), catalyzes methylation of the R9 base in pre-tRNAs. Despite the central role of 5' end processing in mitochondrial tRNA maturation, the link between mtRNase P and diseases is mostly unexplored. Here, we investigate how 11 different human disease-linked mutations in (mt)pre-tRNAIle, (mt)pre-tRNALeu(UUR), and (mt)pre-tRNAMet affect the activities of mtRNase P. We find that several mutations weaken the pre-tRNA binding affinity (K s are approximately two- to sixfold higher than that of wild-type), while the majority of mutations decrease 5' end processing and methylation activity catalyzed by mtRNase P (up to ∼55% and 90% reduction, respectively). Furthermore, all of the investigated mutations in (mt)pre-tRNALeu(UUR) alter the tRNA fold which contributes to the partial loss of function of mtRNase P. Overall, these results reveal an etiological link between early steps of (mt)tRNA-substrate processing and mitochondrial disease.
Insights
Mitochondrial tRNA mutations impair processing by mtRNase P, impacting protein synthesis and causing disease. This study reveals how specific mutations affect tRNA maturation, linking early processing steps to mitochondrial dysfunction.
Area of Science:
- Mitochondrial biology
- Molecular genetics
- Biochemistry
Background:
- Mitochondrial diseases are often caused by mutations in mitochondrial tRNA genes, but their precise contribution to disease is unclear.
- Mitochondrial tRNA processing involves enzymes like mtRNase P, which removes the 5' leader sequence and methylates specific bases.
- The role of mtRNase P in disease pathogenesis remains largely unexplored.
Purpose of the Study:
- To investigate the impact of 11 disease-linked mutations in mitochondrial precursor tRNAs (pre-tRNAs) on mtRNase P activity.
- To determine how these mutations affect pre-tRNA binding, 5' end processing, and methylation.
- To explore the link between altered tRNA processing and mitochondrial dysfunction.
Main Methods:
- In vitro assays were used to assess the binding affinity of mtRNase P to mutated pre-tRNAs.
- Enzyme activity assays measured the 5' end processing and methylation capabilities of mtRNase P with mutant substrates.
- Structural analysis was performed on mutated mitochondrial tRNAs to assess their fold.
Main Results:
- Several mutations significantly weakened the binding affinity of mtRNase P to pre-tRNAs.
- The majority of mutations reduced the 5' end processing and methylation activities of mtRNase P by up to 55% and 90%, respectively.
- Mutations in mitochondrial pre-tRNALeu(UUR) altered tRNA folding, contributing to reduced mtRNase P function.
Conclusions:
- Disease-linked mutations in mitochondrial pre-tRNAs disrupt their processing by mtRNase P.
- Impaired tRNA maturation, including altered binding, reduced enzymatic activity, and structural changes, contributes to mitochondrial dysfunction.
- This study establishes an etiological link between early mitochondrial tRNA processing defects and the development of mitochondrial diseases.
Related Concept Videos
 01:39
Mitochondrial Precursor Proteins
Most of the mitochondrial...
 01:31
Translation
Translation Produces the Building Blocks of Life
Proteins are...
01:31Translation
Translation is the process of synthesizing proteins from the genetic information carried by messenger RNA (mRNA). Following transcription, it constitutes the final step in the expression of genes. This process is carried out by ribosomes, complexes of protein and specialized RNA molecules. Ribosomes, transfer RNA (tRNA), and other proteins produce a chain of amino acids—the polypeptide—as the end product of translation.
Translation Produces the Building Blocks of...
 01:19
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
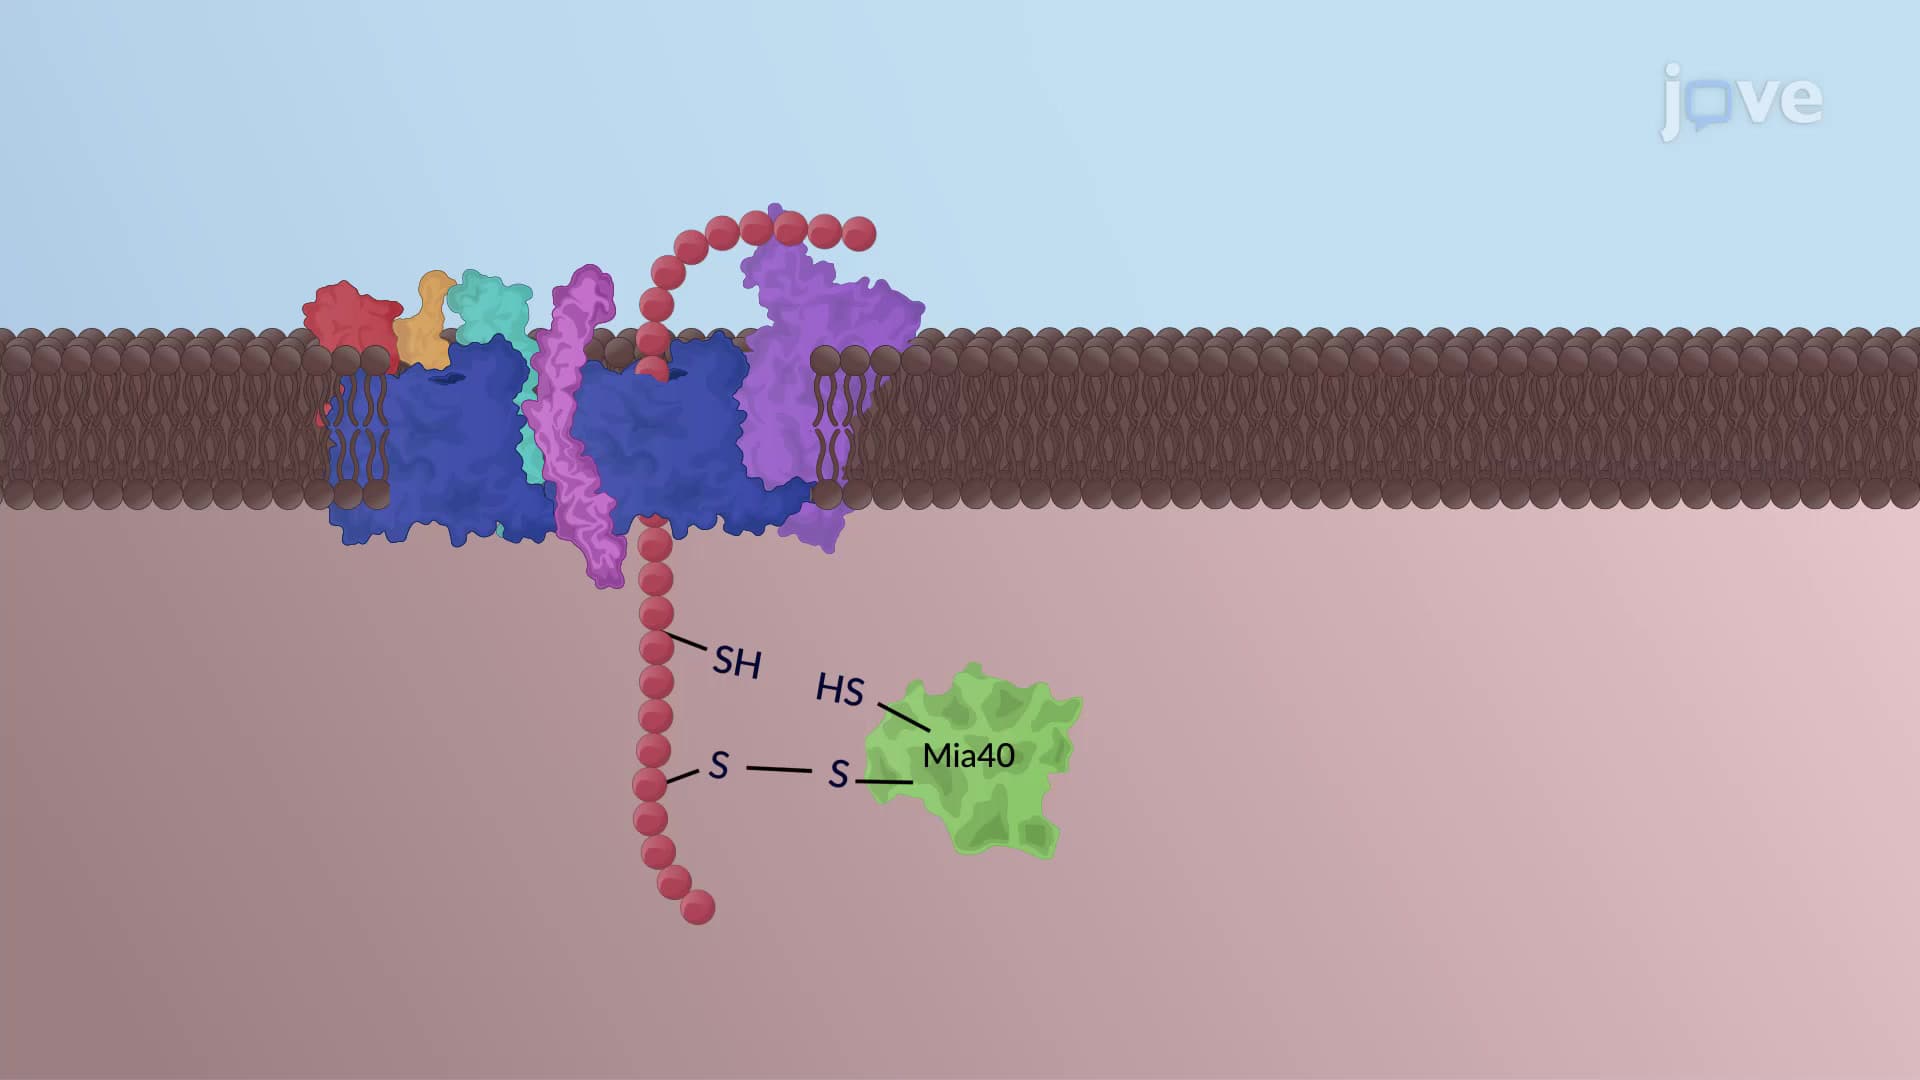
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:39
Mitochondrial Protein Sorting
Most of these mitochondrial proteins are encoded by the nucleus and imported to the mitochondria as unfolded or loosely folded precursors. Mitochondrial precursors...
 01:48
ATP Synthase: Mechanism