Related Experiment Video
Updated: Nov 23, 2025

Screening of Coatings for an All-Solid-State Battery Using In Situ Transmission Electron Microscopy
Published on: January 20, 2023
Atomistic Insights into Lithium Storage Mechanisms in Anatase, Rutile, and Amorphous TiO2 Electrodes
Jodie A Yuwono1, Patrick Burr2, Conor Galvin2
1School of Photovoltaic and Renewable Energy Engineering, UNSW Sydney, Kensington, Sydney, New South Wales 2052, Australia.

Abstract:
Density functional theory calculations were used to investigate the phase transformations of LiTiO2 (at 0 ≤ x ≤ 1), solid-state Li+ diffusion, and interfacial charge-transfer reactions in both crystalline and amorphous forms of TiO2. It is shown that in contrast to crystalline TiO2 polymorphs, the energy barrier to Li+ diffusion in amorphous TiO2 decreases with increasing mole fraction of Li+ due to the changes of chemical species pair interactions following the progressive filling of low-energy Li+ trapping sites. Sites with longer Li-Ti and Li-O interactions exhibit lower Li+ insertion energies and higher migration energy barriers. Due to its disordered atomic arrangement and increasing Li+ diffusivity at higher mole fractions, amorphous TiO2 exhibits both surface and bulk storage mechanisms. The results suggest that nanostructuring of crystalline TiO2 can increase both the rate and capacity because the capacity dependence on the bulk storage mechanism is minimized and replaced with the surface storage mechanism. These insights into Li+ storage mechanisms in different forms of TiO2 can guide the fabrication of TiO2 electrodes to maximize the capacity and rate performance in the future.
Related Concept Videos
 02:42
Ionic Crystal Structures
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 04:02
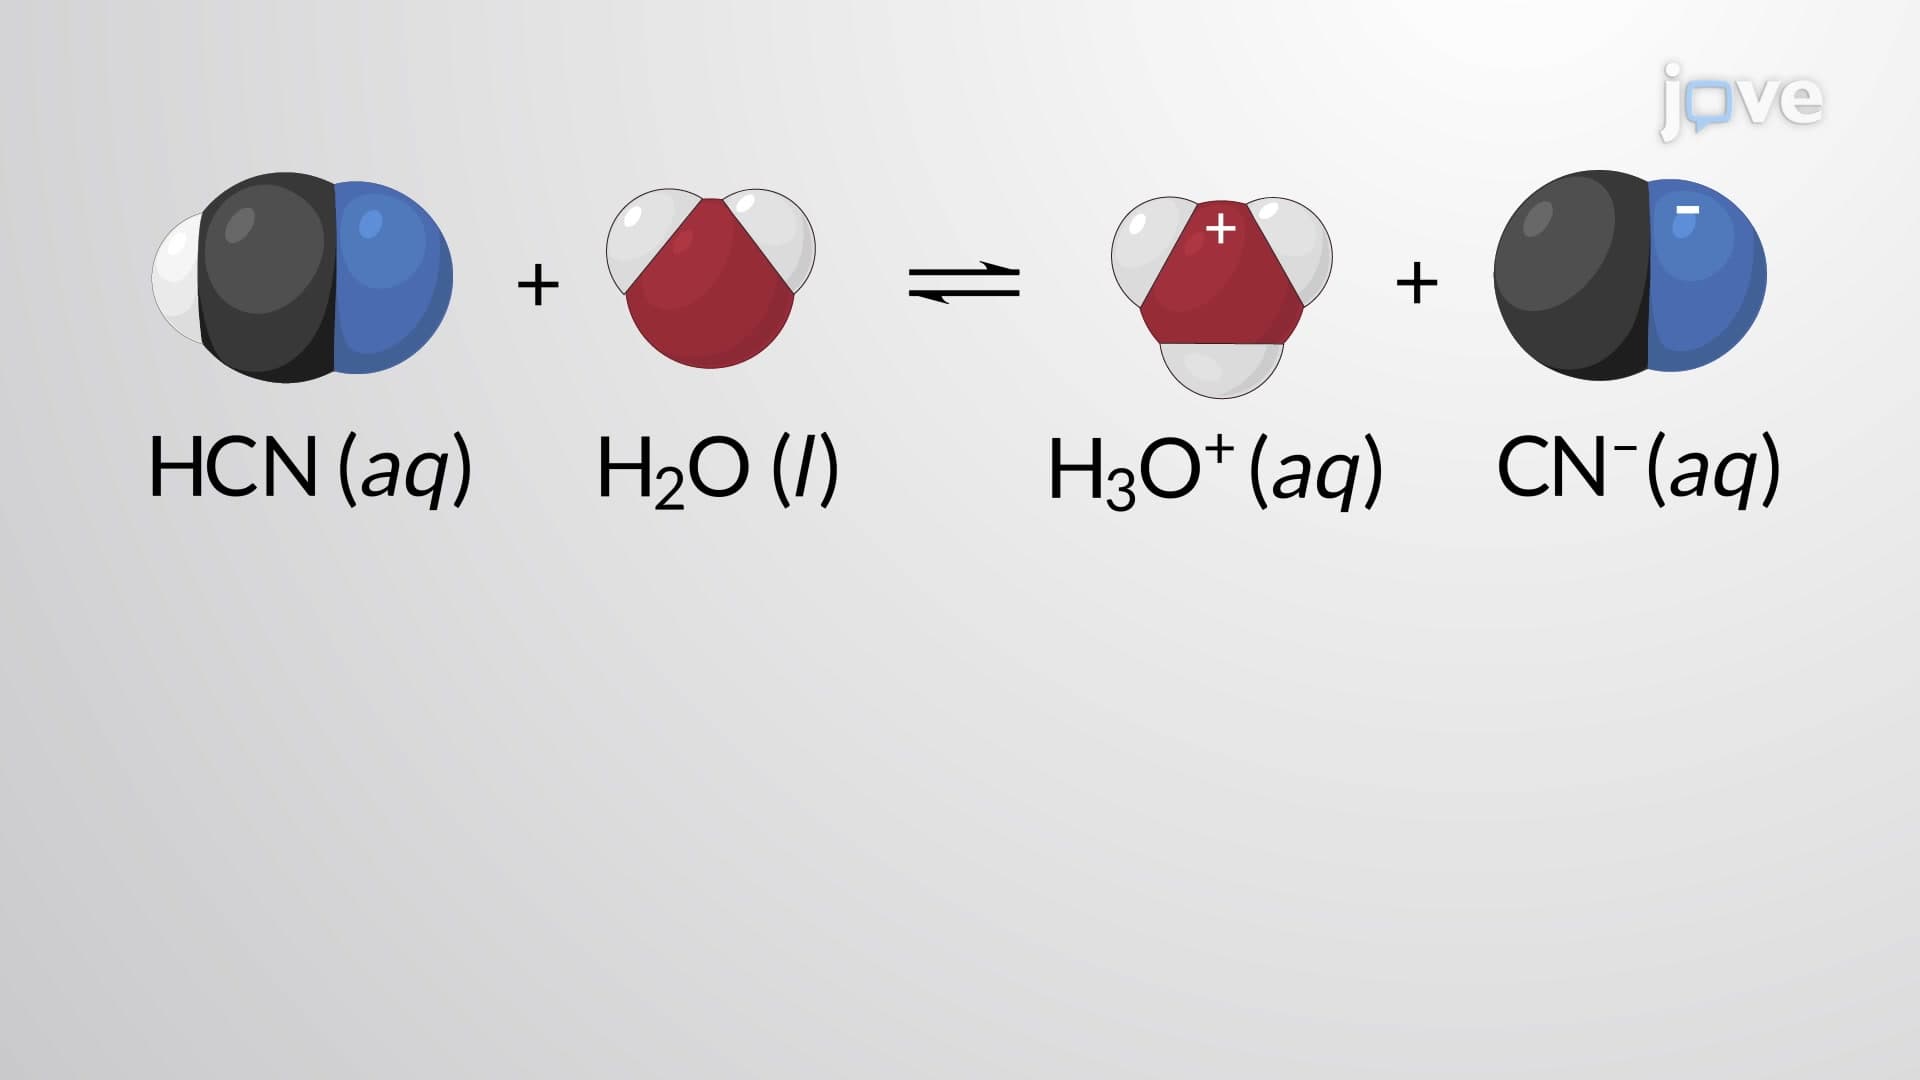
Weak Acid Solutions