Related Experiment Video
Updated: Nov 23, 2025

Structure and Coordination Determination of Peptide-metal Complexes Using 1D and 2D 1H NMR
Published on: December 16, 2013
Mapping Conformational Space of All 8000 Tripeptides by Quantum Chemical Methods: What Strain Is Affordable within
Martin Culka1, Tadeáš Kalvoda1, Ondrej Gutten1
1Institute of Organic Chemistry and Biochemistry of the Czech Academy of Sciences, Flemingovo náměstí 2, 166 10 Praha 6, Czech Republic.

Abstract:
To gain more insight into the physicochemical aspects of a protein structure from the first principles, conformational space of all 8000 "capped" tripeptides (i.e., N-Ac-X1X2X3-NH-CH3, where X is one of the 20 natural amino acids) was investigated computationally. An enormous dataset (denoted P-CONF_1.6M and containing close to 1 600 000 conformers in total) has been obtained by employing a composite protocol combining density functional theory, semiempirical quantum mechanics (SQM), and state-of-the-art solvation methods with 1000 K molecular dynamics (MD) used to generate initial structures (200 snapshots for each tripeptide). This allowed us to present the first rigorous QM-based glimpse at the vast conformational space spanned by small protein fragments. The same computational procedure was repeated for tripeptide fragments taken from the SCOPe database of three-dimensional protein folds, by restraining them to their geometry in a protein. Such complementary data allowed us to compare the distribution of conformational strain energies of unrestrained tripeptidic fragments "in solvent" with those in existing protein chains. Besides providing a rigorous (ab initio) proof of a few well-known concepts and hypotheses concerning protein structures, such as the distribution of (φ, ψ) angles in Ramachandran plots, we have made several observations that came as a certain surprise: (1) distribution of conformational energies does not significantly differ between the "unbiased/unrestrained" conformers obtained from MD sampling in solvent and the biased conformers, i.e., those of a given tripeptide obtained from protein structures; (2) conformational (strain) energy window up to ∼20 to 25 kcal·mol-1 is readily available to tripeptide fragments within the context of a protein chain; (3) overpopulation in certain regions of Ramachandran plot was observed for the unbiased conformers. Last but not least, the massive dataset of accurate (DFT-D3//COSMO-RS) conformational (free) energies of ∼1.6 M peptide conformers, P-CONF_1.6M, obtained throughout this work may serve as excellent dataset for calibrating and benchmarking of popular force fields.
Related Concept Videos
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
 01:22
Protein Folding
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal...
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 02:29
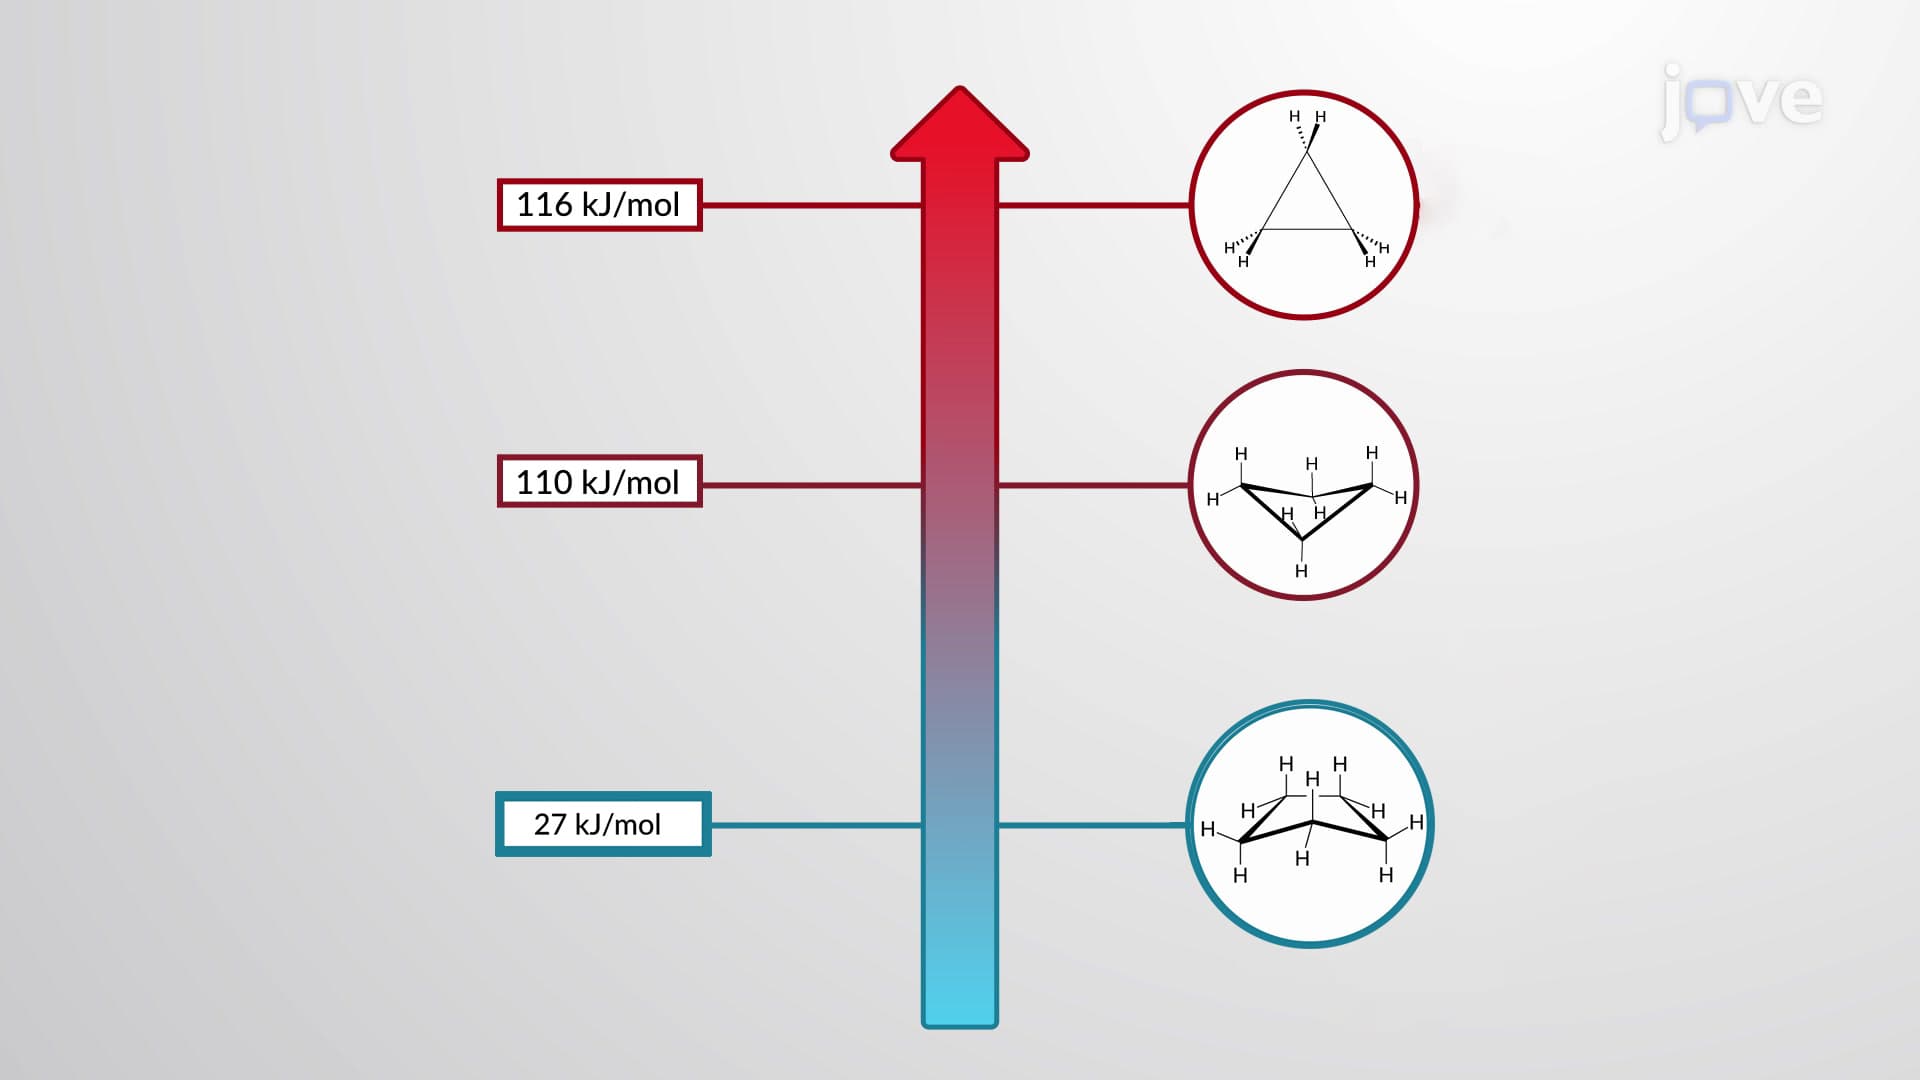
Conformations of Cycloalkanes
 03:00
Molecular Chaperones and Protein Folding
The...