Related Experiment Video
Updated: Nov 23, 2025
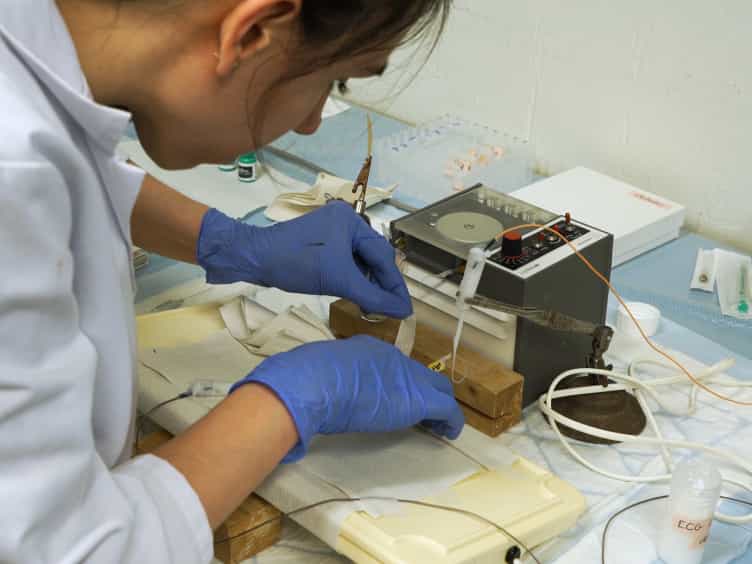

Nasal Potential Difference to Quantify Trans-epithelial Ion Transport in Mice
Published on: July 4, 2018
Synergy between Readthrough and Nonsense Mediated Decay Inhibition in a Murine Model of Cystic Fibrosis Nonsense
Daniel R McHugh1, Calvin U Cotton2,3, Craig A Hodges1,2
1Department of Genetics and Genome Sciences, Case Western Reserve University, 10900 Euclid Ave, Cleveland, OH 44106, USA.

Abstract:
Many heritable genetic disorders arise from nonsense mutations, which generate premature termination codons (PTCs) in transcribed mRNA. PTCs ablate protein synthesis by prematurely terminating the translation of mutant mRNA, as well as reducing mutant mRNA quantity through targeted degradation by nonsense-mediated decay (NMD) mechanisms. Therapeutic strategies for nonsense mutations include facilitating ribosomal readthrough of the PTC and/or inhibiting NMD to restore protein function. However, the efficacy of combining readthrough agents and NMD inhibitors has not been thoroughly explored. In this study, we examined combinations of known NMD inhibitors and readthrough agents using functional analysis of the CFTR protein in primary cells from a mouse model carrying a G542X nonsense mutation in Cftr. We observed synergy between an inhibitor of the NMD component SMG-1 (SMG1i) and the readthrough agents G418, gentamicin, and paromomycin, but did not observe synergy with readthrough caused by amikacin, tobramycin, PTC124, escin, or amlexanox. These results indicate that treatment with NMD inhibitors can increase the quantity of functional protein following readthrough, and that combining NMD inhibitors and readthrough agents represents a potential therapeutic option for treating nonsense mutations.
Insights
Combining nonsense-mediated decay (NMD) inhibitors with readthrough agents shows promise for treating genetic disorders caused by premature termination codons (PTCs). This synergistic approach can restore functional protein levels, offering a new therapeutic avenue.
Area of Science:
- Genetics
- Molecular Biology
- Pharmacology
Background:
- Heritable genetic disorders often stem from nonsense mutations, creating premature termination codons (PTCs).
- PTCs disrupt protein synthesis and trigger mRNA degradation via nonsense-mediated decay (NMD).
- Current therapies aim to bypass PTCs through ribosomal readthrough or inhibit NMD.
Purpose of the Study:
- To investigate the synergistic efficacy of combining NMD inhibitors with various readthrough agents.
- To assess the restoration of functional CFTR protein in a mouse model with a G542X nonsense mutation.
Main Methods:
- Functional analysis of CFTR protein in primary cells from a G542X mutant mouse model.
- Testing combinations of known NMD inhibitors (SMG1i) with multiple readthrough agents (G418, gentamicin, paromomycin, amikacin, tobramycin, PTC124, escin, amlexanox).
Main Results:
- Synergistic effects were observed between SMG1i and readthrough agents G418, gentamicin, and paromomycin.
- No synergy was found with amikacin, tobramycin, PTC124, escin, or amlexanox.
- NMD inhibition enhanced functional protein restoration following ribosomal readthrough.
Conclusions:
- Combining NMD inhibitors with specific readthrough agents can increase functional protein quantity.
- This combination therapy represents a potential strategy for treating genetic disorders caused by nonsense mutations.
More Related Videos

Related Concept Videos
 02:27
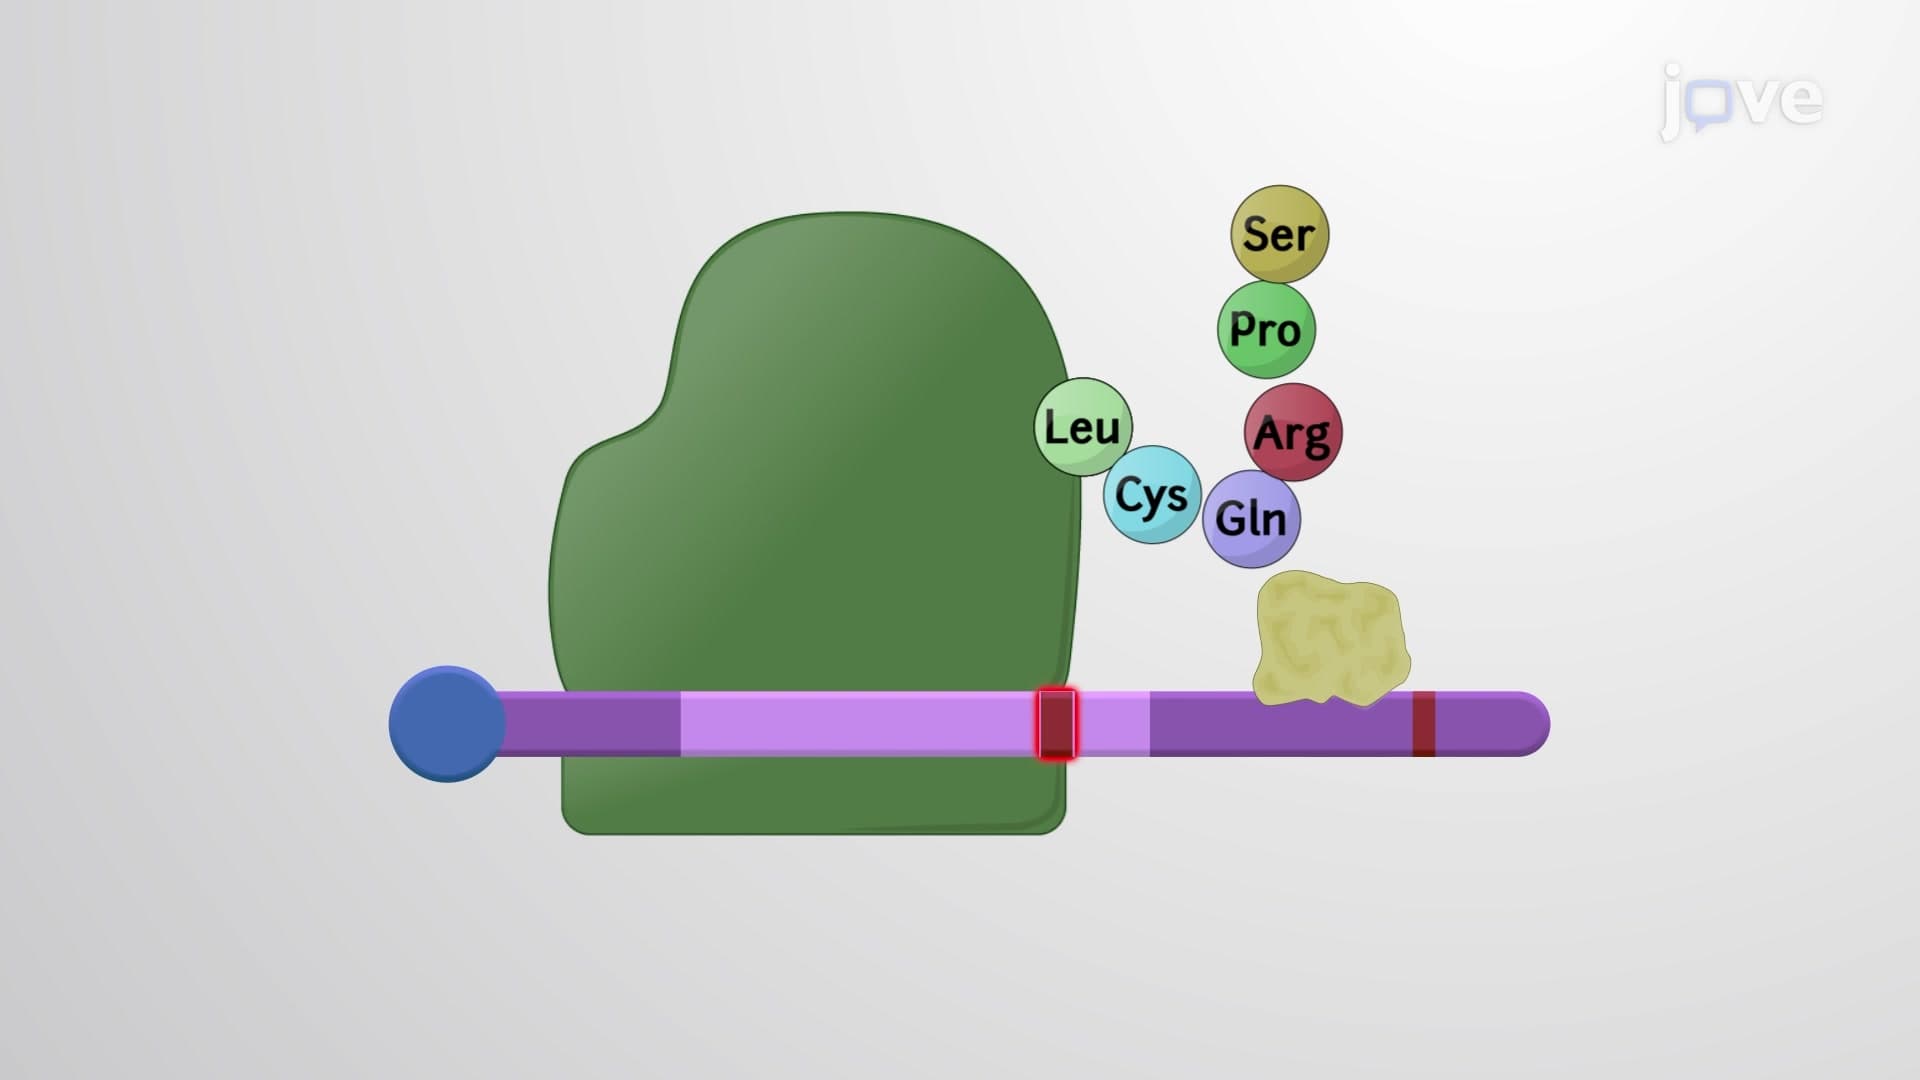
Nonsense-mediated mRNA Decay
Usually, Upf3 binds to an Exon Junction Complex (EJC) at mRNA splice sites. If a ribosome fully translates the mRNA,...
02:27Nonsense-mediated mRNA Decay
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 02:31
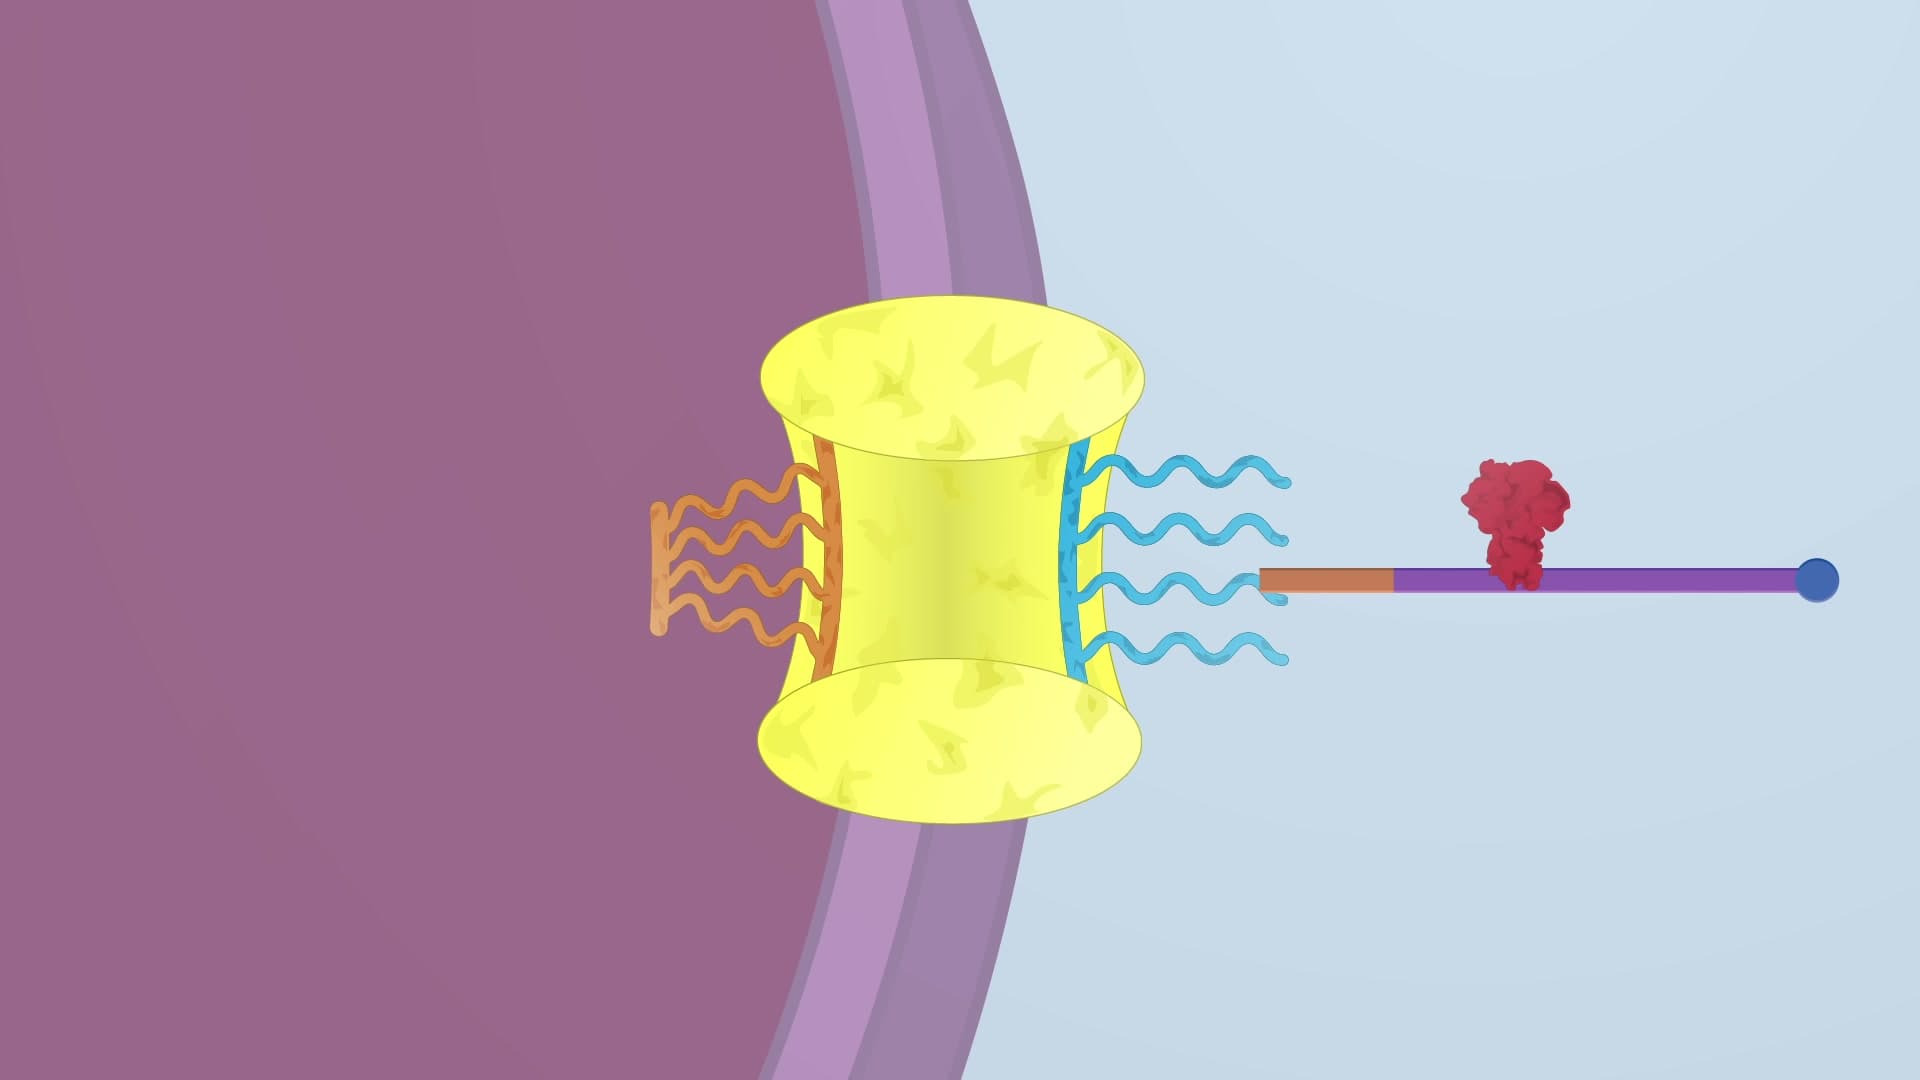
Nuclear Export of mRNA
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic...