A Simple Range-Separated Double-Hybrid Density Functional Theory for Excited States
Related Experiment Video
Updated: Nov 22, 2025
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
1Department of Physical Chemistry and Materials Science, Budapest University of Technology and Economics, P.O. Box 91, Budapest, H-1521, Hungary.

Abstract:
A simple and robust range-separated (RS) double-hybrid (DH) time-dependent density functional approach is presented for the accurate calculation of excitation energies of molecules within the Tamm-Dancoff approximation. The scheme can be considered as an excited-state extension of the ansatz proposed by Toulouse and co-workers [J. Chem. Phys. 2018, 148, 164105], which is based on the two-parameter decomposition of the Coulomb potential, for which both the exchange and correlation contributions are range-separated. A flexible and efficient implementation of the new scheme is also presented, which facilitates its extension to any combination of exchange and correlation functionals. The performance of the new approximation is tested for singlet excitations on several benchmark compilations and thoroughly compared to that of representative DH, RS hybrid, and RS DH functionals. The one-electron basis set dependence and computation times are also assessed. Our results show that the new approach improves on standard DHs in most cases, and it can provide a more robust and accurate alternative. In addition, on average, it noticeably surpasses the existing RS hybrid and RS DH functionals.

Related Concept Videos
 03:24
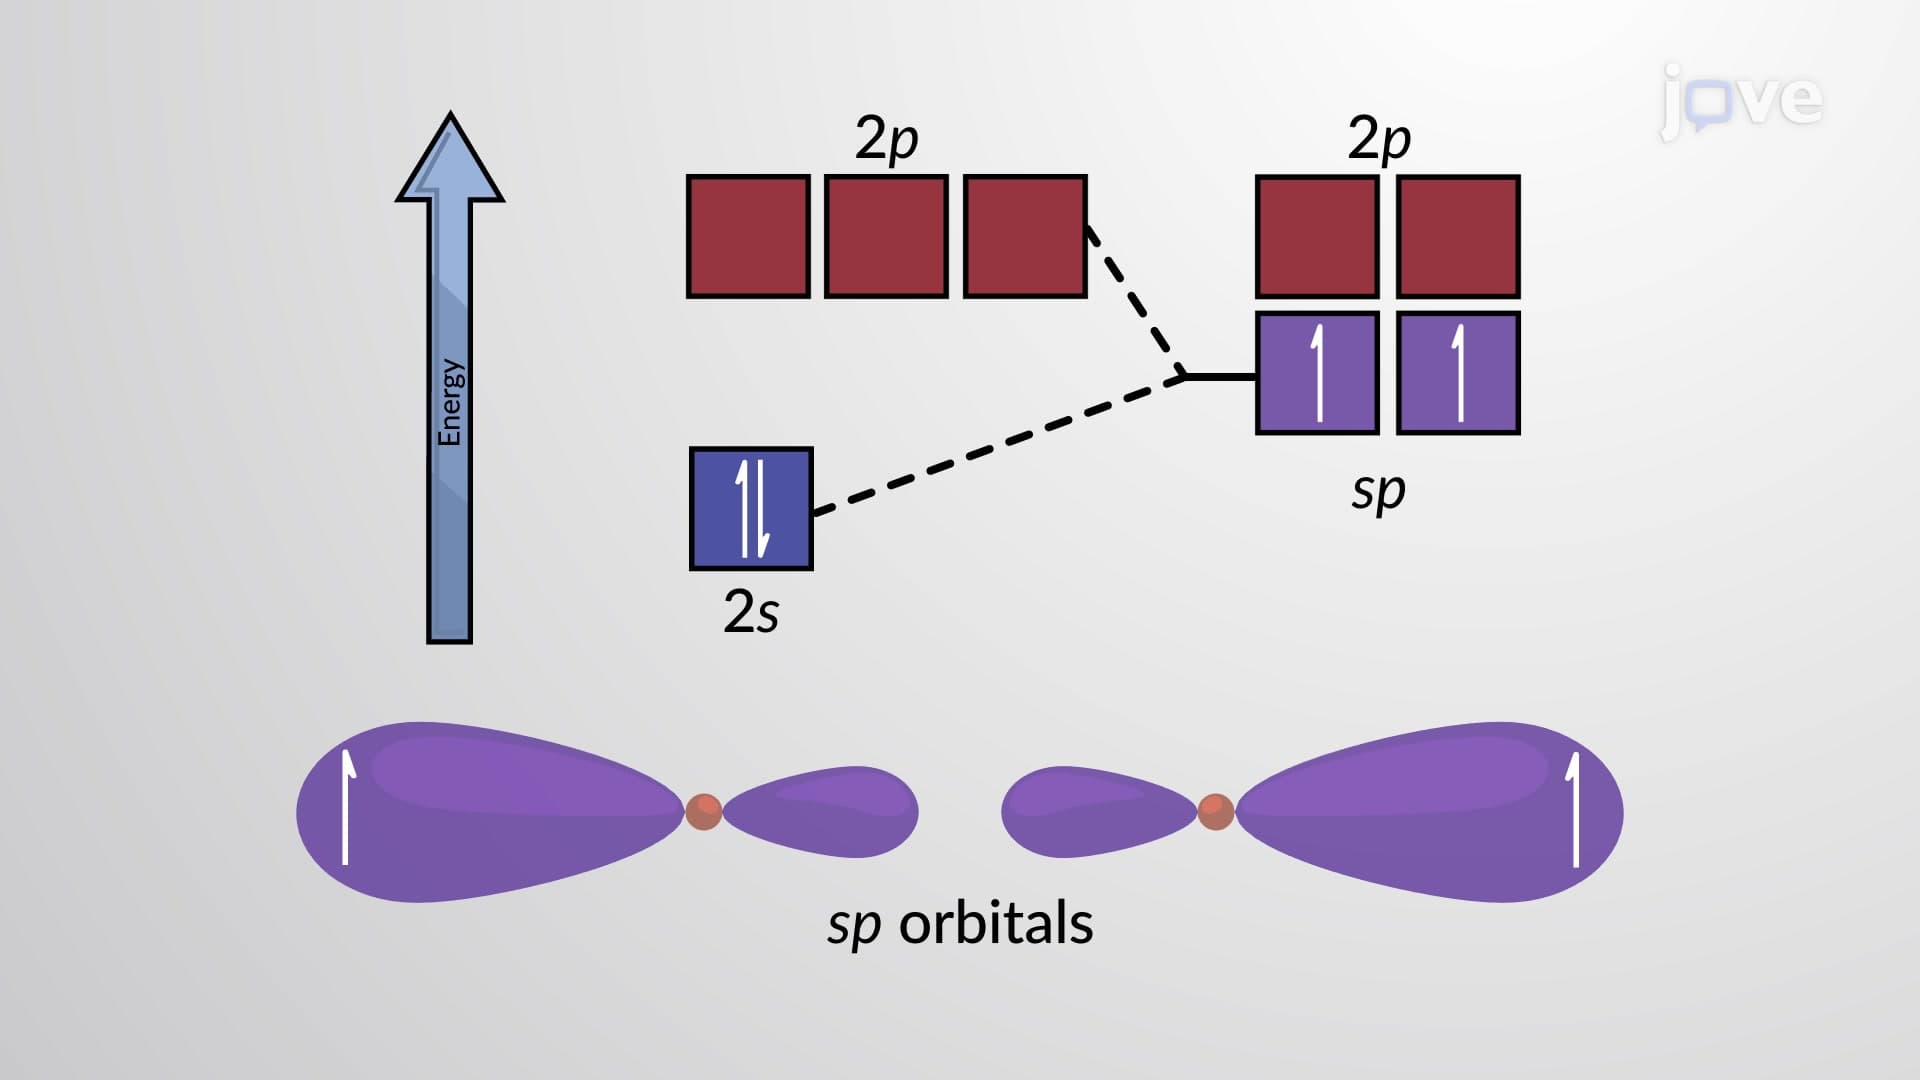
Hybridization of Atomic Orbitals I
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 03:51
Molecular Orbital Theory II
 03:35
Hybridization of Atomic Orbitals II
 02:35
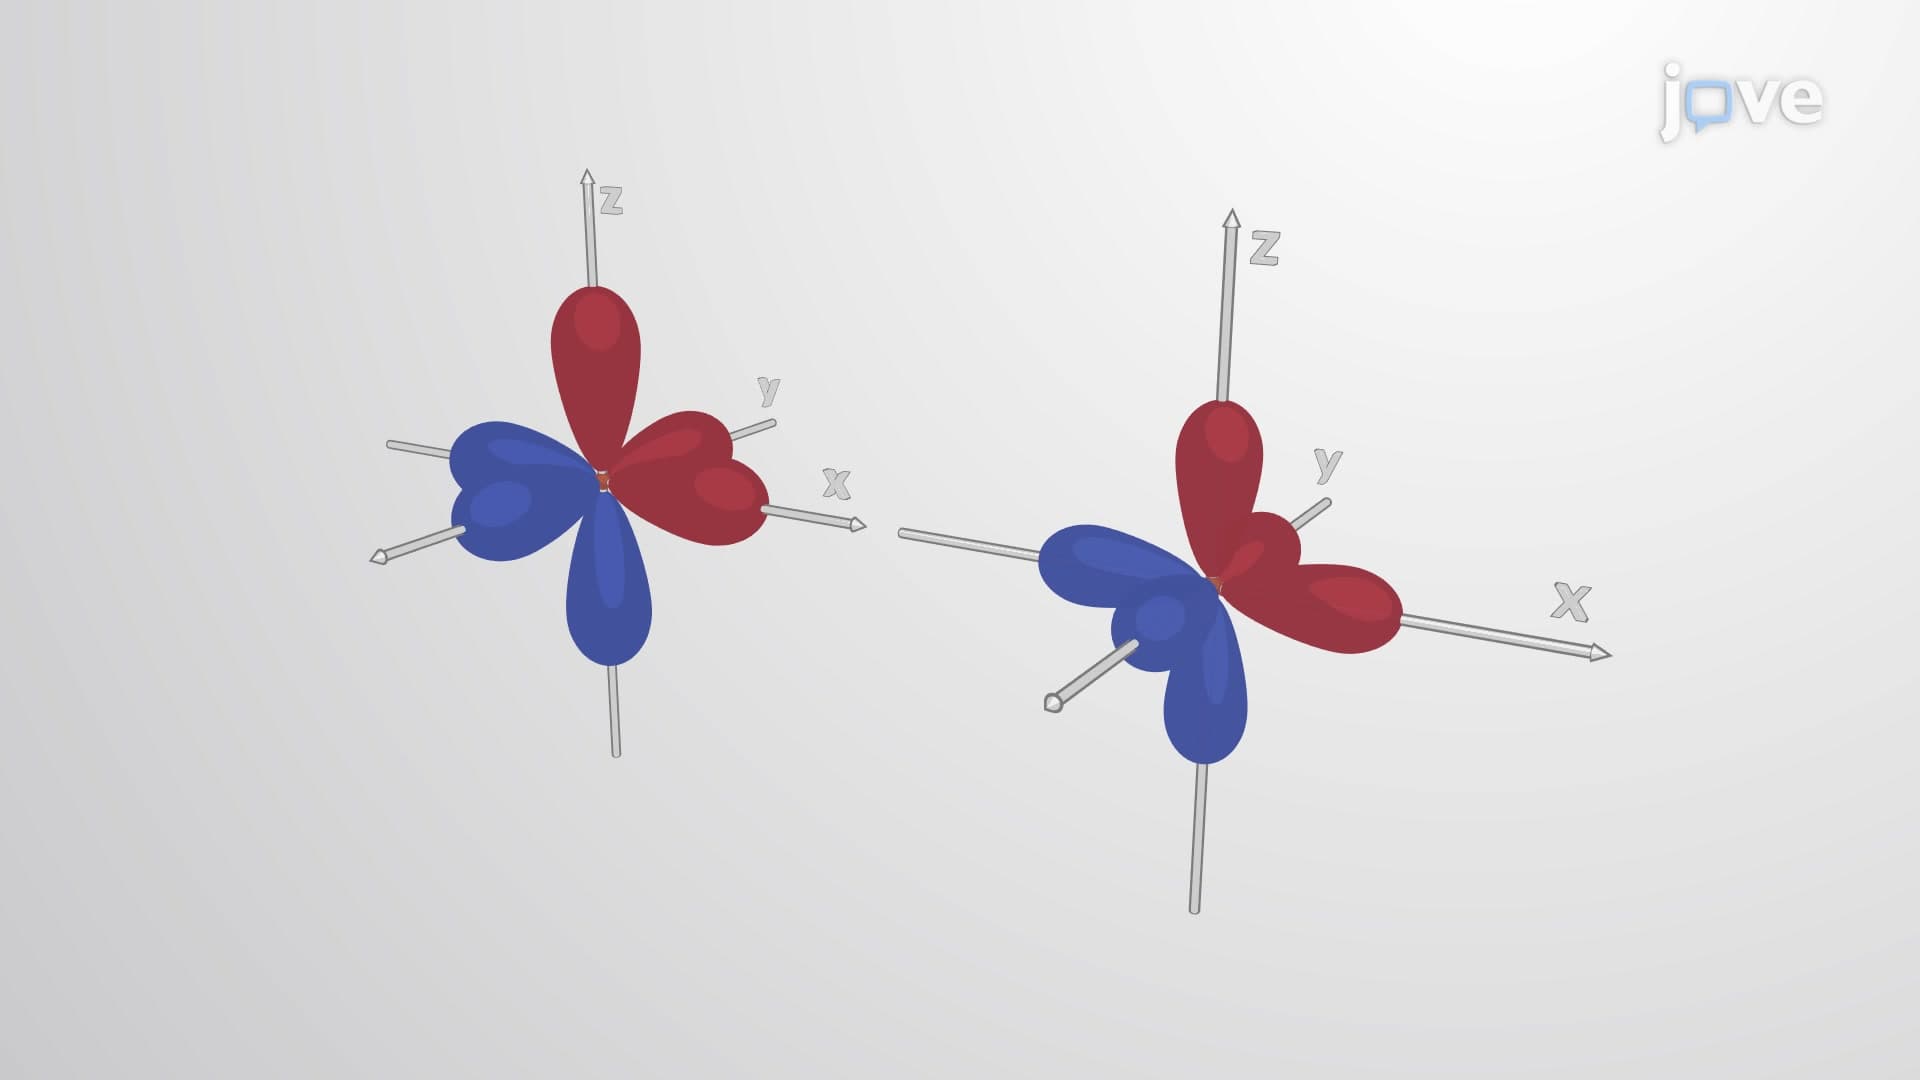
Molecular Orbital Theory I
 02:16
Resonance and Hybrid Structures
Resonance Structures and Resonance Hybrids
The Lewis structure of a nitrite anion (NO2−) may actually be drawn in two different ways, distinguished by the locations of the N–O and N=O bonds.