Related Experiment Video
Updated: Nov 22, 2025
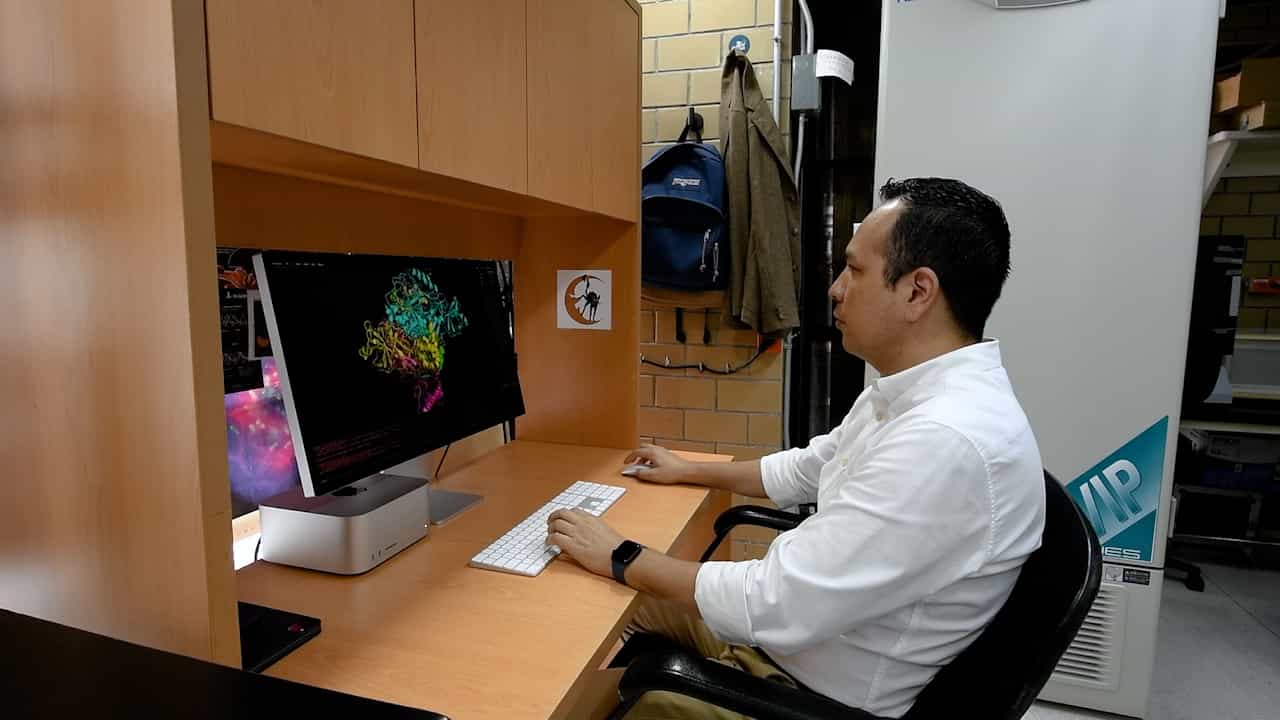

Pharmacophore Modeling for Targets with Extensive Ligand Libraries: A Case Study on SARS-CoV-2 Mpro
Published on: September 26, 2025
Identification of potential SARS-CoV-2 Mpro inhibitors integrating molecular docking and water thermodynamics
M Elizabeth Sobhia1, Ketan Ghosh1, Srikanth Sivangula1
1National Institute of Pharmaceutical Education and Research (NIPER), SAS Nagar, Punjab, India.

Abstract:
The COVID-19 pandemic is an ongoing global health emergency caused by a newly discovered coronavirus SARS-CoV-2. The entire scientific community across the globe is working diligently to tackle this unprecedented challenge. In silico studies have played a crucial role in the current situation by expediting the process of identification of novel potential chemotypes targeting the viral receptors. In this study, we have made efforts to identify molecules that can potentially inhibit the SARS-CoV-2 main protease (Mpro) using the high-resolution crystal structure of SARS-CoV-2 Mpro. The SARS-CoV-2 Mpro has a large flexible binding pocket that can accommodate various chemically diverse ligands but a complete occupation of the binding cavity is necessary for efficient inhibition and stability. We augmented glide three-tier molecular docking protocol with water thermodynamics to screen molecules obtained from three different compound libraries. The diverse hits obtained through docking studies were scored against generated WaterMap to enrich the quality of results. Five molecules were selected from each compound library on the basis of scores and protein-ligand complementarity. Further MD simulations on the proposed molecules affirm the stability of these molecules in the complex. MM-GBSA results and intermolecular hydrogen bond analysis also confirm the thermodynamic stability of proposed molecules. This study also presumably steers the structure determination of many ligand-main protease complexes using x-ray diffraction methods.Communicated by Ramaswamy H. Sarma.
More Related Videos
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Related Concept Videos
 02:04
Protein-protein Interfaces
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...