Related Experiment Video
Updated: Nov 22, 2025
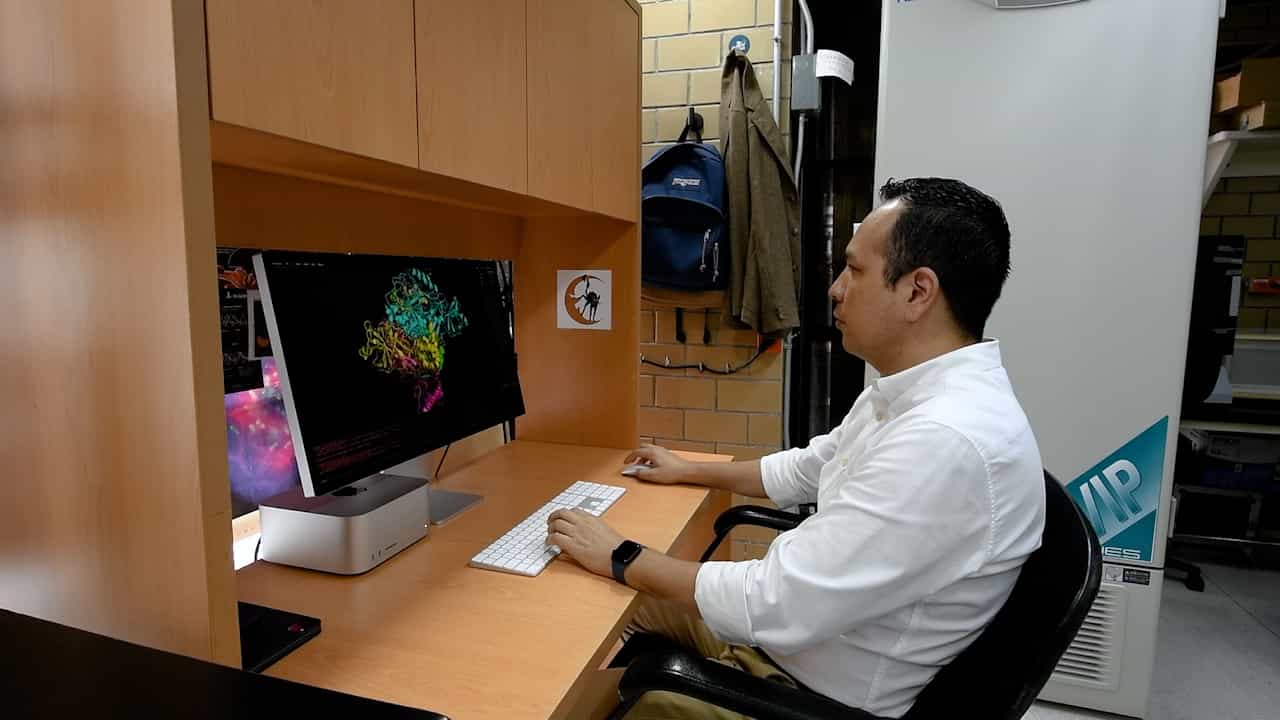

Pharmacophore Modeling for Targets with Extensive Ligand Libraries: A Case Study on SARS-CoV-2 Mpro
Published on: September 26, 2025
Computational Prediction of Potential Inhibitors of the Main Protease of SARS-CoV-2
Renata Abel1, María Paredes Ramos2, Qiaofeng Chen1
1Institute of Physiology, Charité-University Medicine Berlin, Berlin, Germany.

Insights
Computational methods identified potential inhibitors for SARS-CoV-2, the virus causing COVID-19. This research screened numerous compounds, including natural products and existing drugs, to find treatments for the ongoing pandemic.
Area of Science:
- Computational chemistry
- Drug discovery
- Virology
Background:
- The COVID-19 pandemic, caused by SARS-CoV-2, poses a significant global health risk, especially for individuals with comorbidities.
- Lack of approved treatments necessitates rapid identification of potential therapeutic agents.
Purpose of the Study:
- To develop and apply a computational screening methodology for identifying inhibitors of the SARS-CoV-2 main protease.
- To discover potential drug candidates from natural product and drug repurposing databases.
Main Methods:
- Employed ligand- and structure-based virtual screening against natural product databases (Super Natural II, Traditional Chinese Medicine) and drug databases.
- Utilized molecular docking, molecular dynamics simulations, and computational analysis of toxicity and cytochrome inhibition profiles.
- Screened approximately 360,000 compounds, with 80 selected for detailed evaluation and 12 for further analysis.
Main Results:
- Identified 12 promising candidate compounds from four distinct datasets after extensive computational analysis.
- Evaluated the toxicity and cytochrome inhibition profiles of the selected compounds.
Conclusions:
- The study successfully developed a computational approach to identify potential SARS-CoV-2 inhibitors.
- The identified candidate compounds warrant further experimental investigation for their therapeutic potential against COVID-19.
Abstract:
The rapidly developing pandemic, known as coronavirus disease 2019 (COVID-19) and caused by the severe acute respiratory syndrome coronavirus 2 (SARS-CoV-2), has recently spread across 213 countries and territories. This pandemic is a dire public health threat-particularly for those suffering from hypertension, cardiovascular diseases, pulmonary diseases, or diabetes; without approved treatments, it is likely to persist or recur. To facilitate the rapid discovery of inhibitors with clinical potential, we have applied ligand- and structure-based computational approaches to develop a virtual screening methodology that allows us to predict potential inhibitors. In this work, virtual screening was performed against two natural products databases, Super Natural II and Traditional Chinese Medicine. Additionally, we have used an integrated drug repurposing approach to computationally identify potential inhibitors of the main protease of SARS-CoV-2 in databases of drugs (both approved and withdrawn). Roughly 360,000 compounds were screened using various molecular fingerprints and molecular docking methods; of these, 80 docked compounds were evaluated in detail, and the 12 best hits from four datasets were further inspected via molecular dynamics simulations. Finally, toxicity and cytochrome inhibition profiles were computationally analyzed for the selected candidate compounds.
Related Concept Videos
 02:04
Protein-protein Interfaces
 02:50
Conjugated Proteins
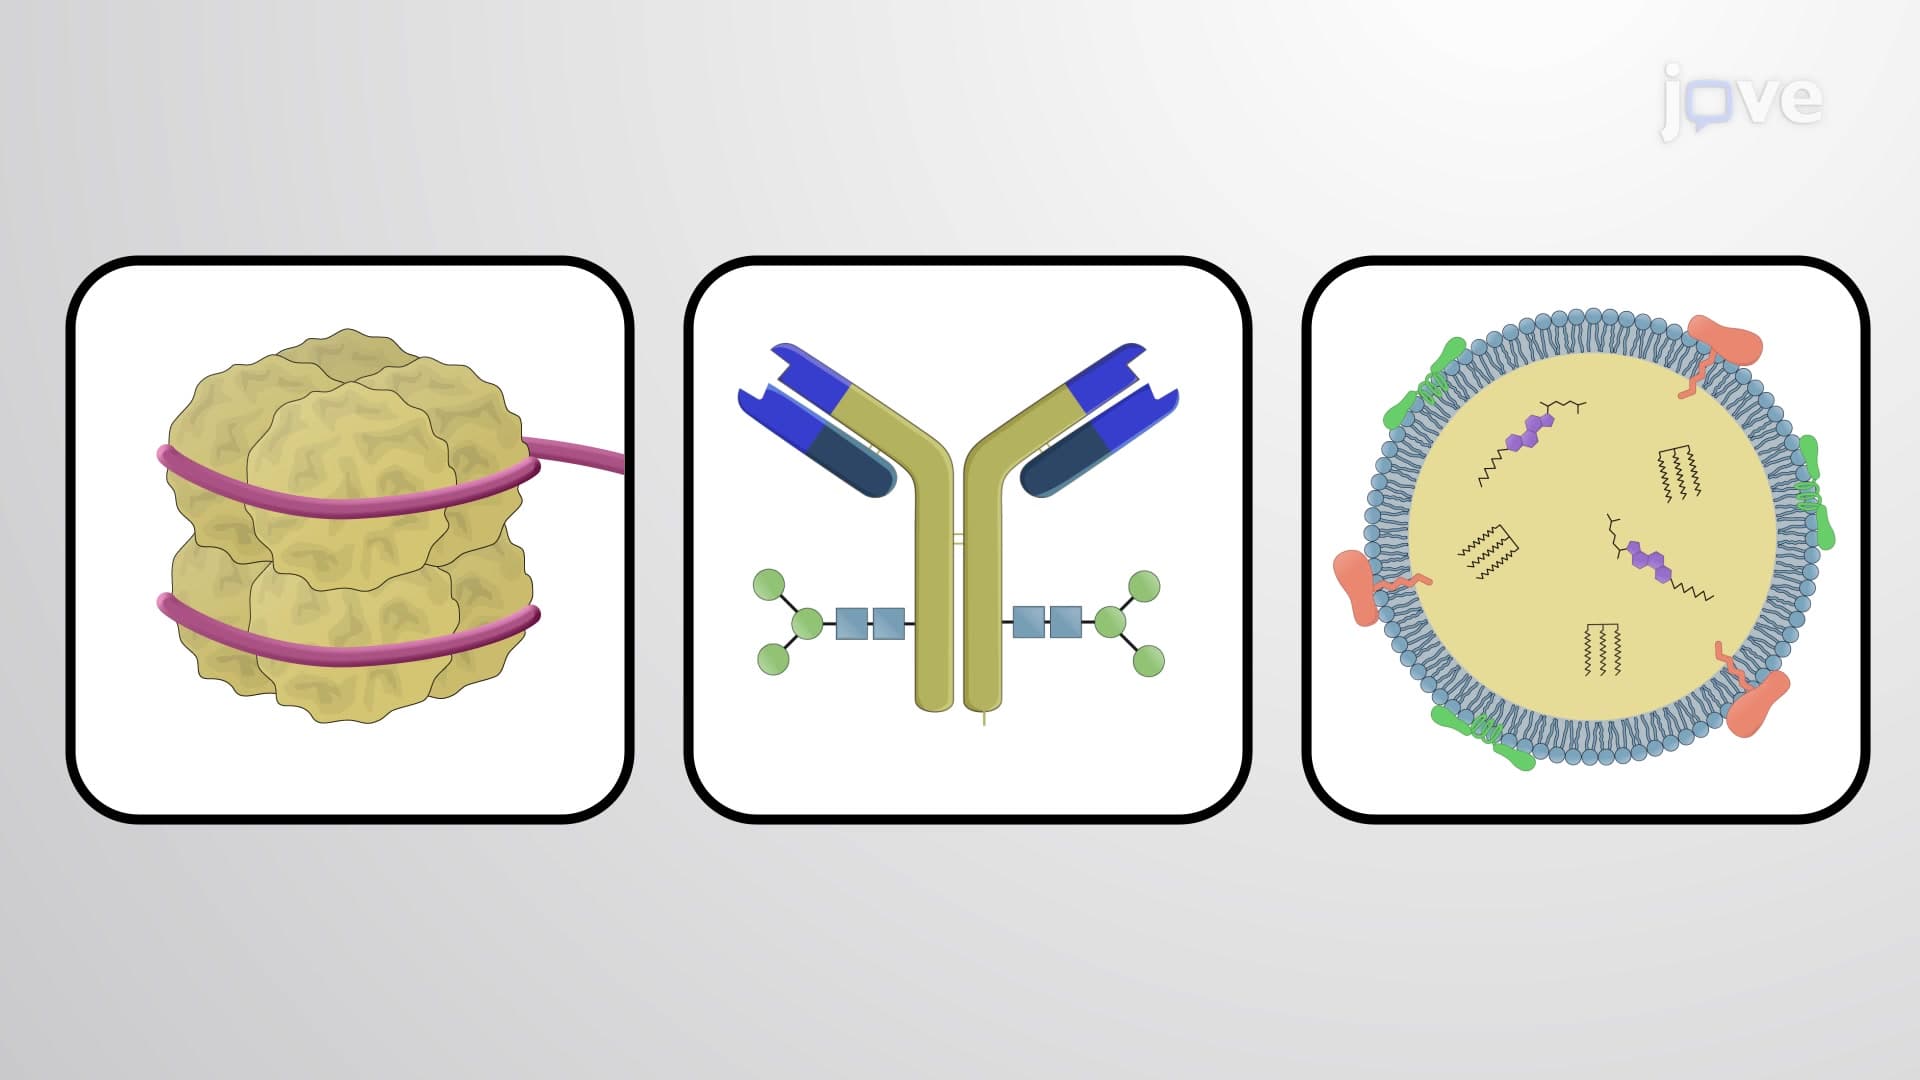
Nucleoproteins are protein complexes that contain nucleic acids, categorized as deoxyribonucleoproteins (DNPs) or ribonucleoproteins (RNPs) respectively. The nucleosome is a typical example of a DNP where nuclear DNA is associated with histone proteins. The major antigen for the Covid-19 virus SARS-CoV is an RNP that is critical...
 02:24
Predicting Reaction Outcomes