Born-Oppenheimer molecular dynamics simulations on structures of high-density and low-density water: a comparison of the SCAN meta-GGA and PBE GGA functionals
Related Experiment Video
Updated: Nov 21, 2025

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Mengli Li1, Lu Chen, Lirong Gui
1School of Physics and Optoelectronic Engineering, Ludong University, Yantai 264025, P. R. China. gzhao19800209@126.com.

Abstract:
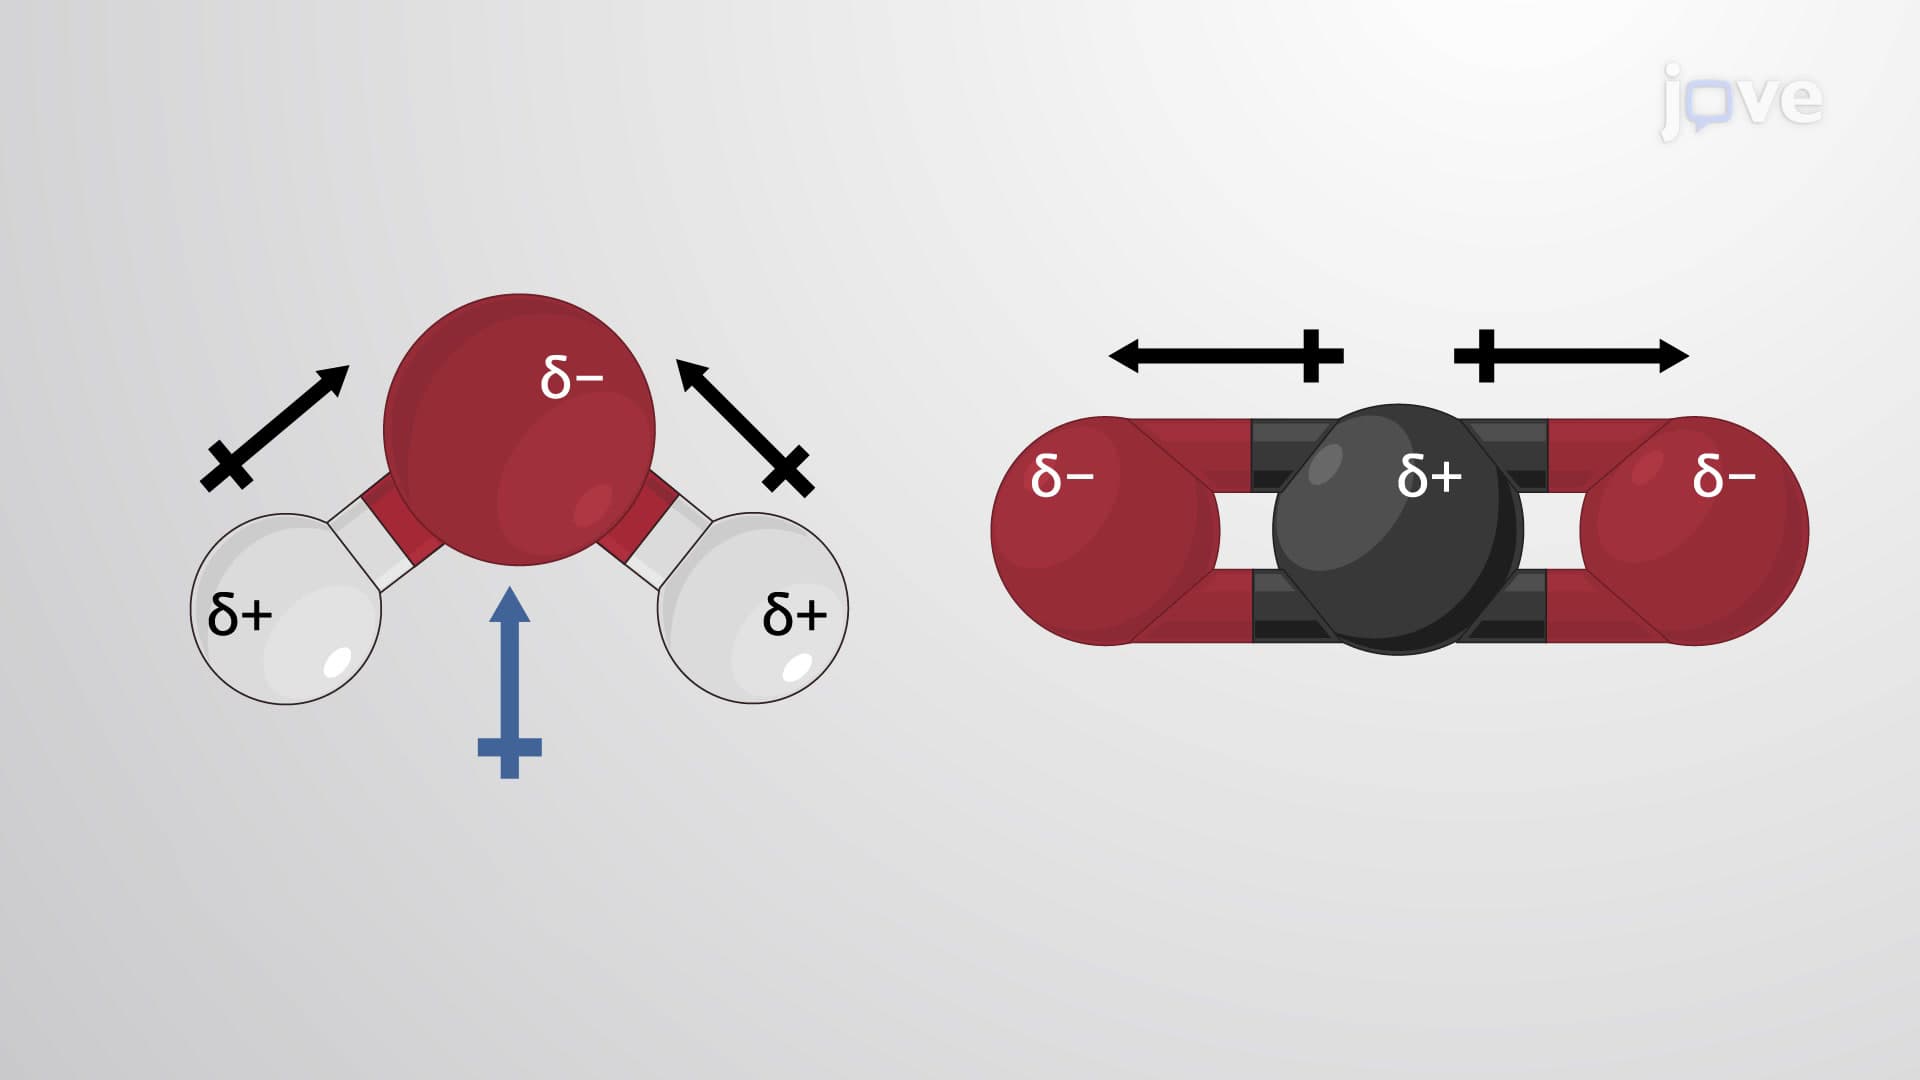
Using Born-Oppenheimer ab initio molecular dynamics (BOAIMD) simulations, the high-density water (HDW) and low-density water (LDW) structures based on SCAN meta-GGA are compared with those based on PBE GGA. Compared with Car-Parrinello ab initio molecular dynamics (CPAIMD) simulations, BOAIMD simulations can produce more accurate results because no fictitious electron mass is introduced. At each state point, our simulations continue for 100 ps after the system reached equilibrium, which is the longest ab initio simulations of liquid water reported so far and can ensure an accurate statistical average. The influence of the size effect and nuclear quantum effect on structure is not considered in the present work, but only that of two different functionals on the structure is discussed. It is found that, in HDW, just as shown using CPAIMD simulations, the SCAN-based hydrogen-bonds (HBs) are more flexible than the PBE-based ones, which makes the structure based on SCAN obviously closer to the experimental results than that based on PBE. However, it is not the case in LDW, and the difference between the results based on these two functionals is very small.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 15:05
Deciphering the Structural Effects of Activating EGFR Somatic Mutations with Molecular Dynamics Simulation
Published on: May 20, 2020