Related Experiment Video
Updated: Nov 20, 2025


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Predicting metal-metal interactions. II. Accelerating generalized schemes through physical insights
Tej S Choksi1, Verena Streibel1, Frank Abild-Pedersen2
1SUNCAT Center for Interface Science and Catalysis, Department of Chemical Engineering, Stanford University, 443 Via Ortega, Stanford, California 94305, USA.

Abstract:
Operando-computational frameworks that integrate descriptors for catalyst stability within catalyst screening paradigms enable predictions of rates and selectivity on chemically faithful representations of nanoparticles under reaction conditions. These catalyst stability descriptors can be efficiently predicted by density functional theory (DFT)-based models. The alloy stability model, for example, predicts the stability of metal atoms in nanoparticles with site-by-site resolution. Herein, we use physical insights to present accelerated approaches of parameterizing this recently introduced alloy-stability model. These accelerated approaches meld quadratic functions for the energy of metal atoms in terms of the coordination number with linear correlations between model parameters and the cohesive energies of bulk metals. By interpolating across both the coordination number and chemical space, these accelerated approaches shrink the training set size for 12 fcc p- and d-block metals from 204 to as few as 24 DFT calculated total energies without sacrificing the accuracy of our model. We validate the accelerated approaches by predicting adsorption energies of metal atoms on extended surfaces and 147 atom cuboctahedral nanoparticles with mean absolute errors of 0.10 eV and 0.24 eV, respectively. This efficiency boost will enable a rapid and exhaustive exploration of the vast material space of transition metal alloys for catalytic applications.
More Related Videos
 06:53
Author Spotlight: Magnetometric Characterization of Intermediates in the Solid-State Electrochemistry of Redox-Active Metal-Organic Frameworks
Published on: June 9, 2023
 11:04
Ion Mobility-Mass Spectrometry Techniques for Determining the Structure and Mechanisms of Metal Ion Recognition and Redox Activity of Metal Binding Oligopeptides
Published on: September 7, 2019
Related Concept Videos
 02:32
Bonding in Metals
 02:51
Metal-Ligand Bonds
In these complexes, transition metals form coordinate covalent bonds, a kind of Lewis acid-base interaction in which both of the electrons in the bond are contributed by a donor (Lewis base) to an electron acceptor (Lewis acid). The Lewis acid in...
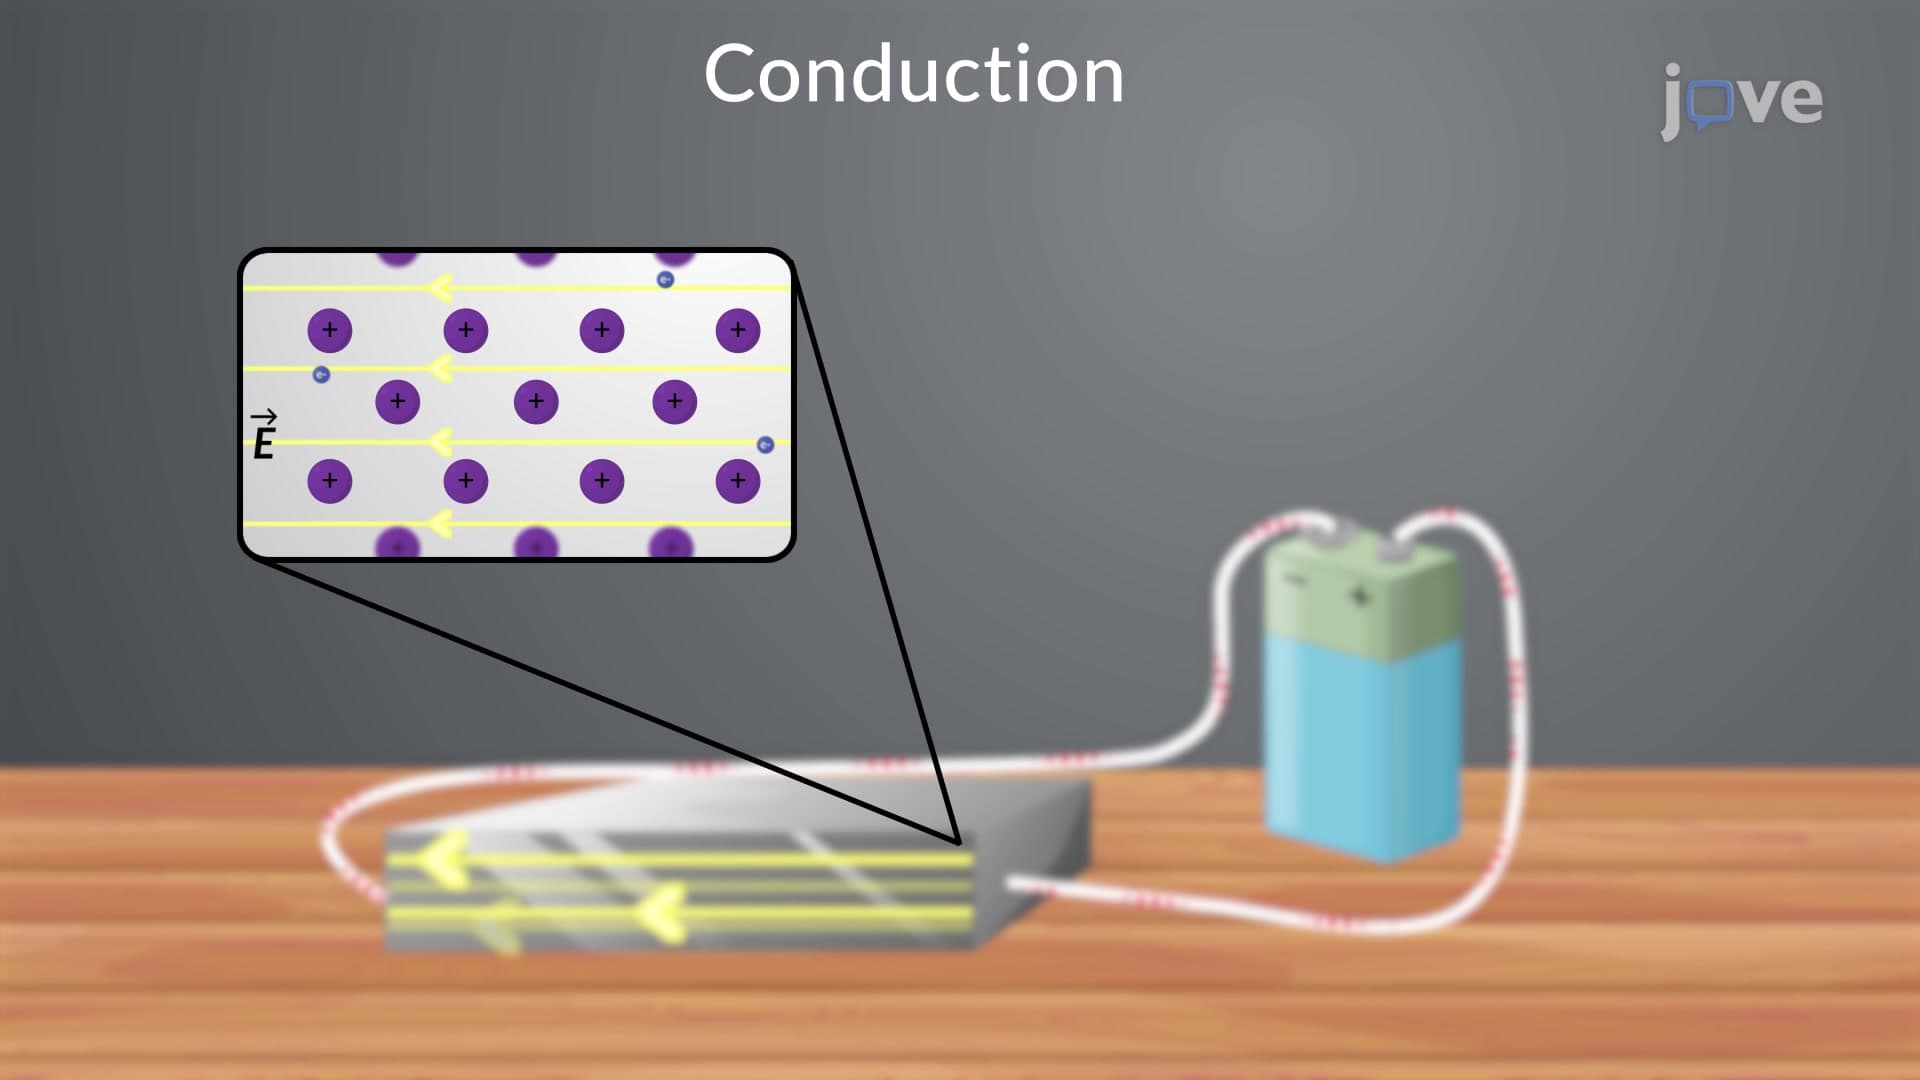 01:17
Theory of Metallic Conduction
In this theory, Newton's second law of motion is used to determine the acceleration of an electron in the presence of an applied electric field. Then, its velocity is expressed via this acceleration.
An electron moves through the crystal, containing positive ions,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:37
Metallic Solids
All metallic solids exhibit high thermal and electrical conductivity, metallic luster, and malleability....
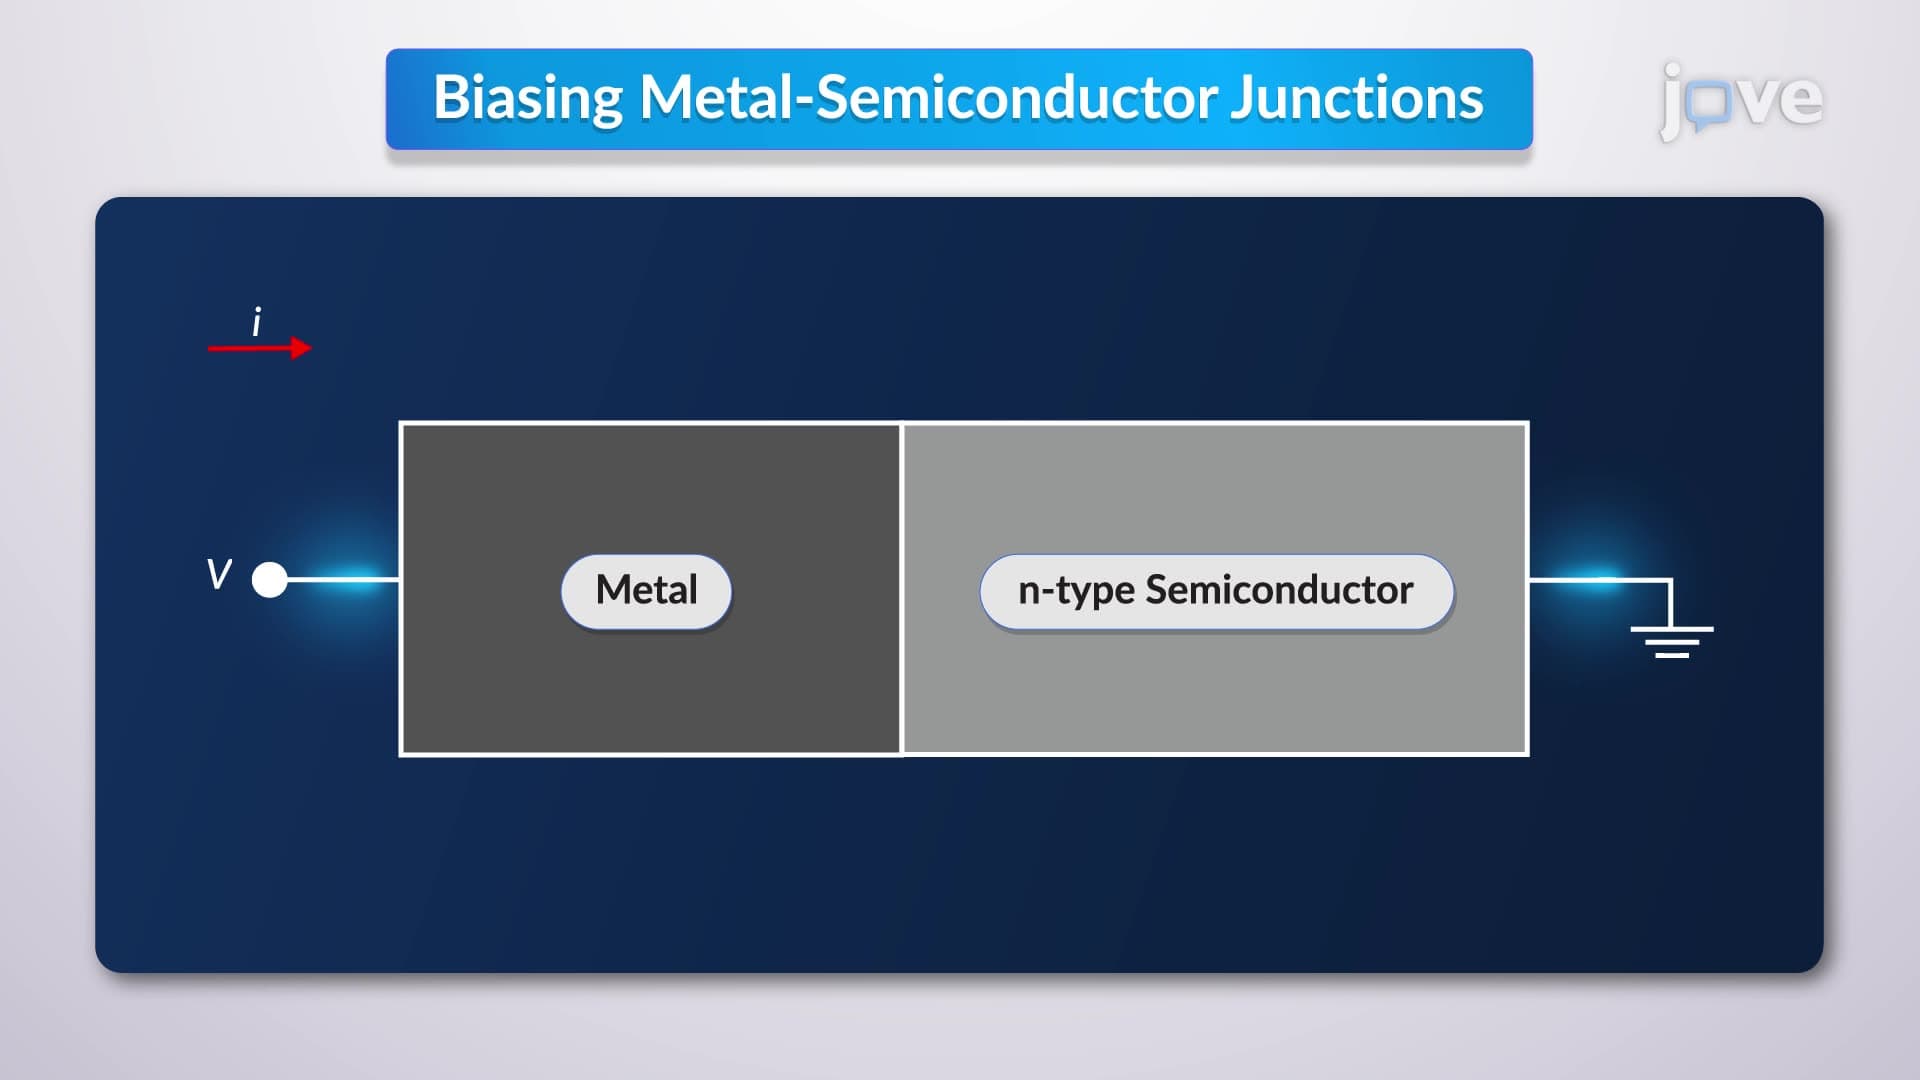 01:27
Biasing of Metal-Semiconductor Junctions
In Schottky junctions, where the semiconductor is n-type, applying a positive voltage to the metal relative to the semiconductor reduces its Fermi...