Related Experiment Video
Updated: Nov 20, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Long-range-corrected multiconfiguration density functional with the on-top pair density
Michał Hapka1, Ewa Pastorczak1, Agnieszka Krzemińska1
1Institute of Physics, Lodz University of Technology, ul. Wolczanska 219, 90-924 Lodz, Poland.

Abstract:
We propose a multiconfiguration density functional combining a short-range density functional approximation with a novel long-range correction for dynamic correlation effects. The correction is derived from the adiabatic connection formalism so that the resulting functional requires access only to one- and two-electron reduced density matrices of the system. In practice, the functional is formulated for wavefunctions of the complete active space (CAS) type and the short-range density functional part is made dependent on the on-top pair density via auxiliary spin densities. The latter allows for reducing the self-interaction and the static correlation errors without breaking the spin symmetry. We study the properties and the performance of the non-self-consistent variant of the method, termed lrAC0-postCAS. Numerical demonstration on a set of dissociation energy curves and excitation energies shows that lrAC0-postCAS provides accuracy comparable with more computationally expensive ab initio rivals.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 01:10
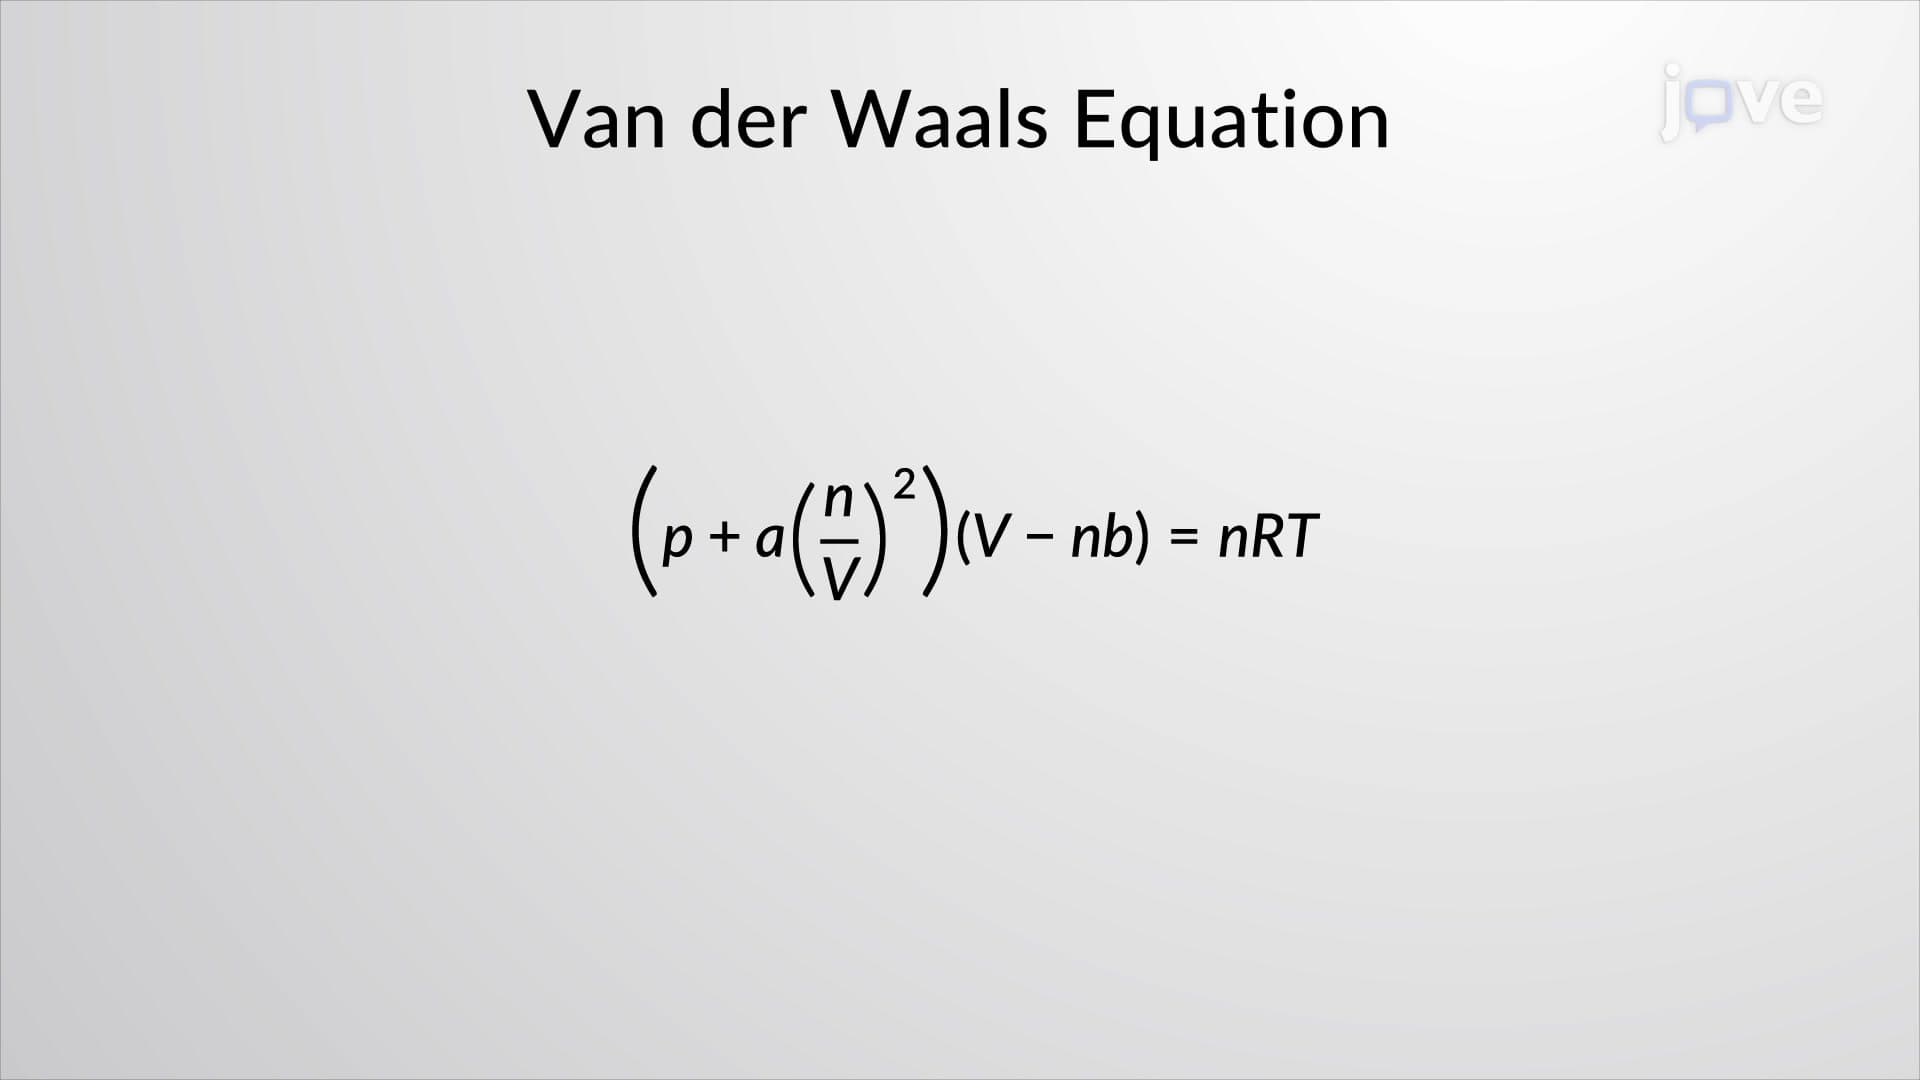
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 02:40
MO Theory and Covalent Bonding
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:24
Van der Waals Interactions
 03:51
Molecular Orbital Theory II
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...