Related Experiment Video
Updated: Nov 19, 2025

Author Spotlight: Unveiling Oxidative Phosphorylation System Dynamics and Mitochondrial Roles in Health and Disease
Published on: May 3, 2024
Structural basis for a complex I mutation that blocks pathological ROS production
Zhan Yin1, Nils Burger1, Duvaraka Kula-Alwar2
1MRC Mitochondrial Biology Unit, University of Cambridge, Cambridge Biomedical Campus, Cambridge, UK.

Abstract:
Mitochondrial complex I is central to the pathological reactive oxygen species (ROS) production that underlies cardiac ischemia-reperfusion (IR) injury. ND6-P25L mice are homoplasmic for a disease-causing mtDNA point mutation encoding the P25L substitution in the ND6 subunit of complex I. The cryo-EM structure of ND6-P25L complex I revealed subtle structural changes that facilitate rapid conversion to the "deactive" state, usually formed only after prolonged inactivity. Despite its tendency to adopt the "deactive" state, the mutant complex is fully active for NADH oxidation, but cannot generate ROS by reverse electron transfer (RET). ND6-P25L mitochondria function normally, except for their lack of RET ROS production, and ND6-P25L mice are protected against cardiac IR injury in vivo. Thus, this single point mutation in complex I, which does not affect oxidative phosphorylation but renders the complex unable to catalyse RET, demonstrates the pathological role of ROS production by RET during IR injury.
Insights
Mitochondrial complex I mutations can prevent reactive oxygen species (ROS) production during cardiac injury. This study shows a specific mutation protects mice from ischemia-reperfusion injury by blocking ROS generation.
Area of Science:
- Biochemistry
- Cardiovascular Biology
- Mitochondrial Medicine
Background:
- Mitochondrial complex I is implicated in cardiac ischemia-reperfusion (IR) injury via reactive oxygen species (ROS) production.
- A specific point mutation (P25L) in the ND6 subunit of complex I alters its structure and function.
Purpose of the Study:
- To investigate the structural and functional consequences of the ND6-P25L mutation in mitochondrial complex I.
- To determine the role of reverse electron transfer (RET)-mediated ROS production in cardiac IR injury.
Main Methods:
- Cryo-electron microscopy (cryo-EM) to determine the structure of the mutant complex I.
- Biochemical assays to assess NADH oxidation and ROS production.
- In vivo studies using ND6-P25L mice subjected to cardiac IR injury.
Main Results:
- The ND6-P25L mutation causes subtle structural changes in complex I, promoting a rapid transition to the 'deactive' state.
- Mutant complex I remains fully active for NADH oxidation but cannot produce ROS via RET.
- ND6-P25L mice exhibit normal mitochondrial function but are protected against cardiac IR injury due to the absence of RET ROS.
Conclusions:
- A single point mutation in mitochondrial complex I that abolishes RET ROS production protects against cardiac IR injury.
- This highlights the pathological role of RET-derived ROS in IR injury.
- Targeting RET in complex I may offer a therapeutic strategy for cardiac protection.
More Related Videos
Related Concept Videos
 01:46
Electron Transport Chain: Complex I and II
ROS generation is regulated and maintained at moderate levels necessary...
 01:30
The Electron Transport Chain
Inhibitors of the electron transport chain
Rotenone, a widely used pesticide, prevents electron transfer from Fe-S cluster to ubiquinone or Q...
 01:41
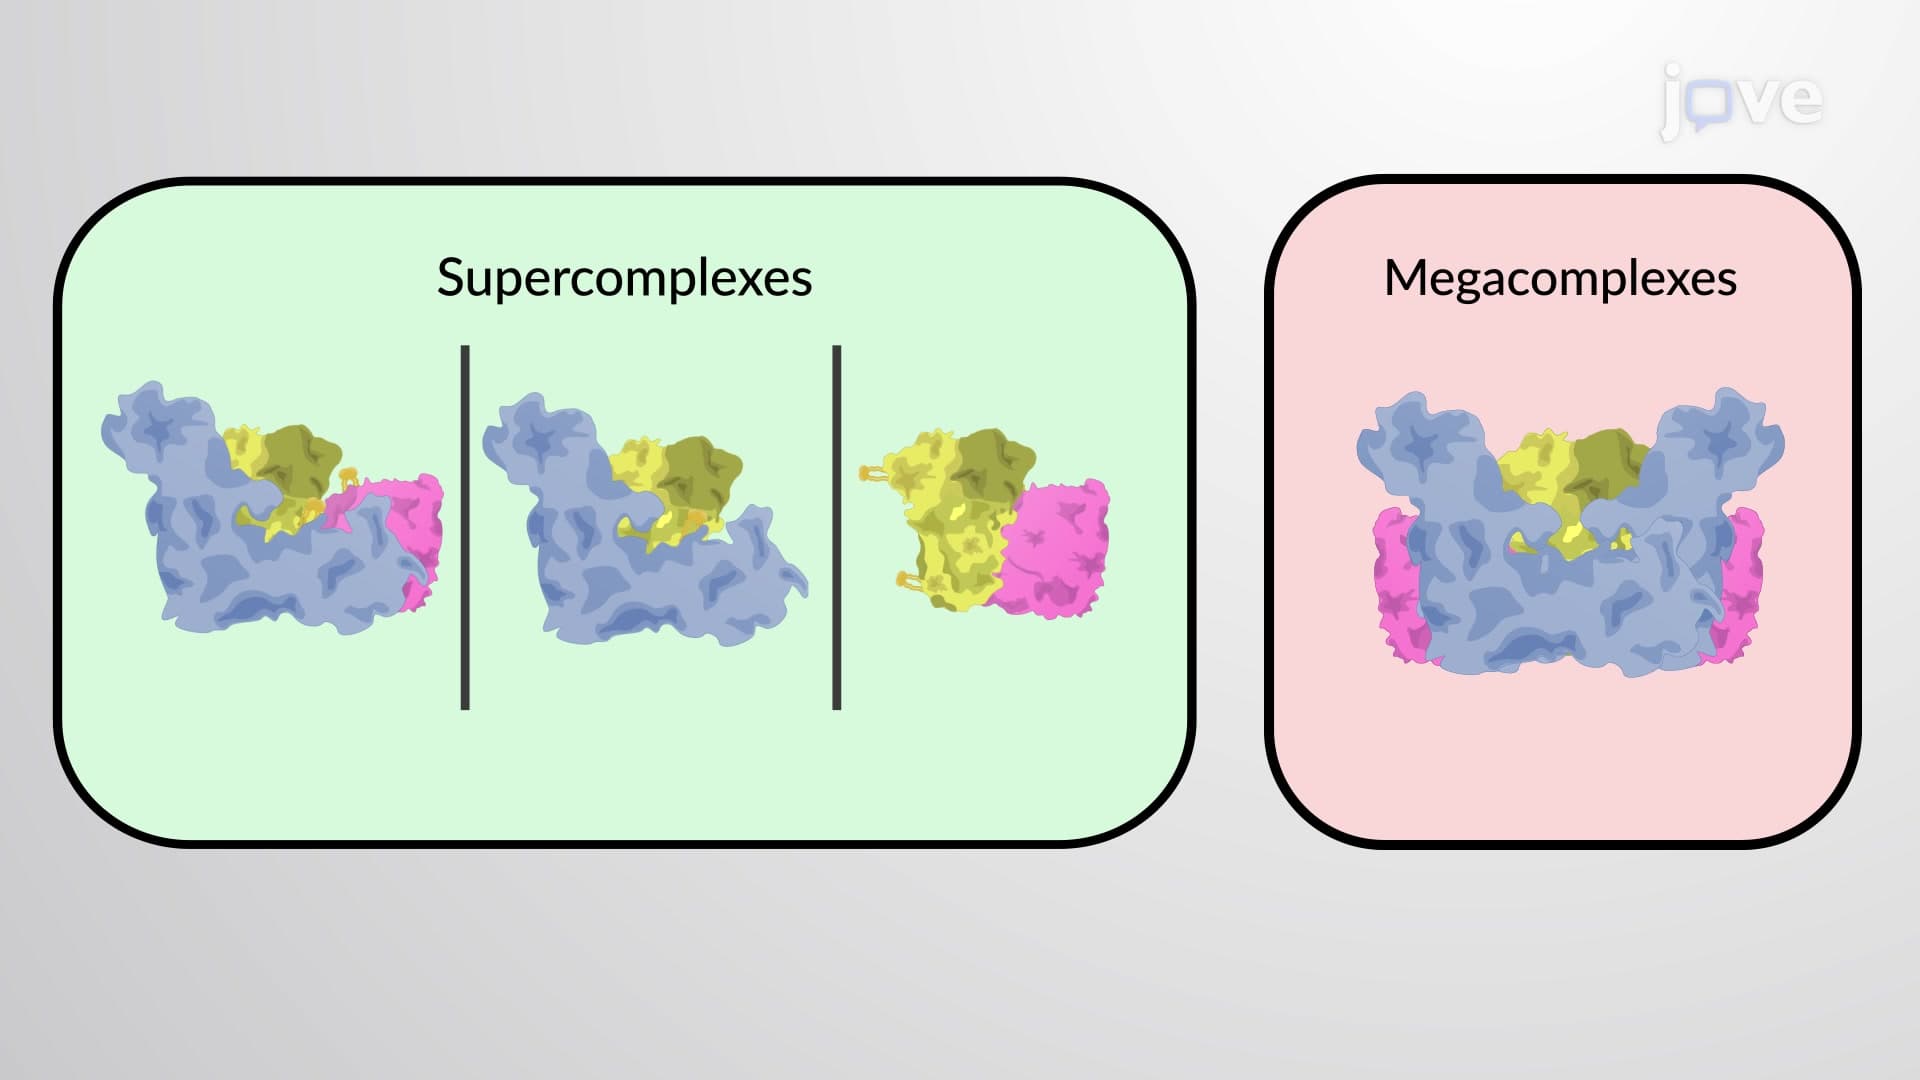
The Supercomplexes in the Crista Membrane
 01:48
ATP Synthase: Mechanism
 01:43
Electron Transport Chain: Complex III and IV
 01:19
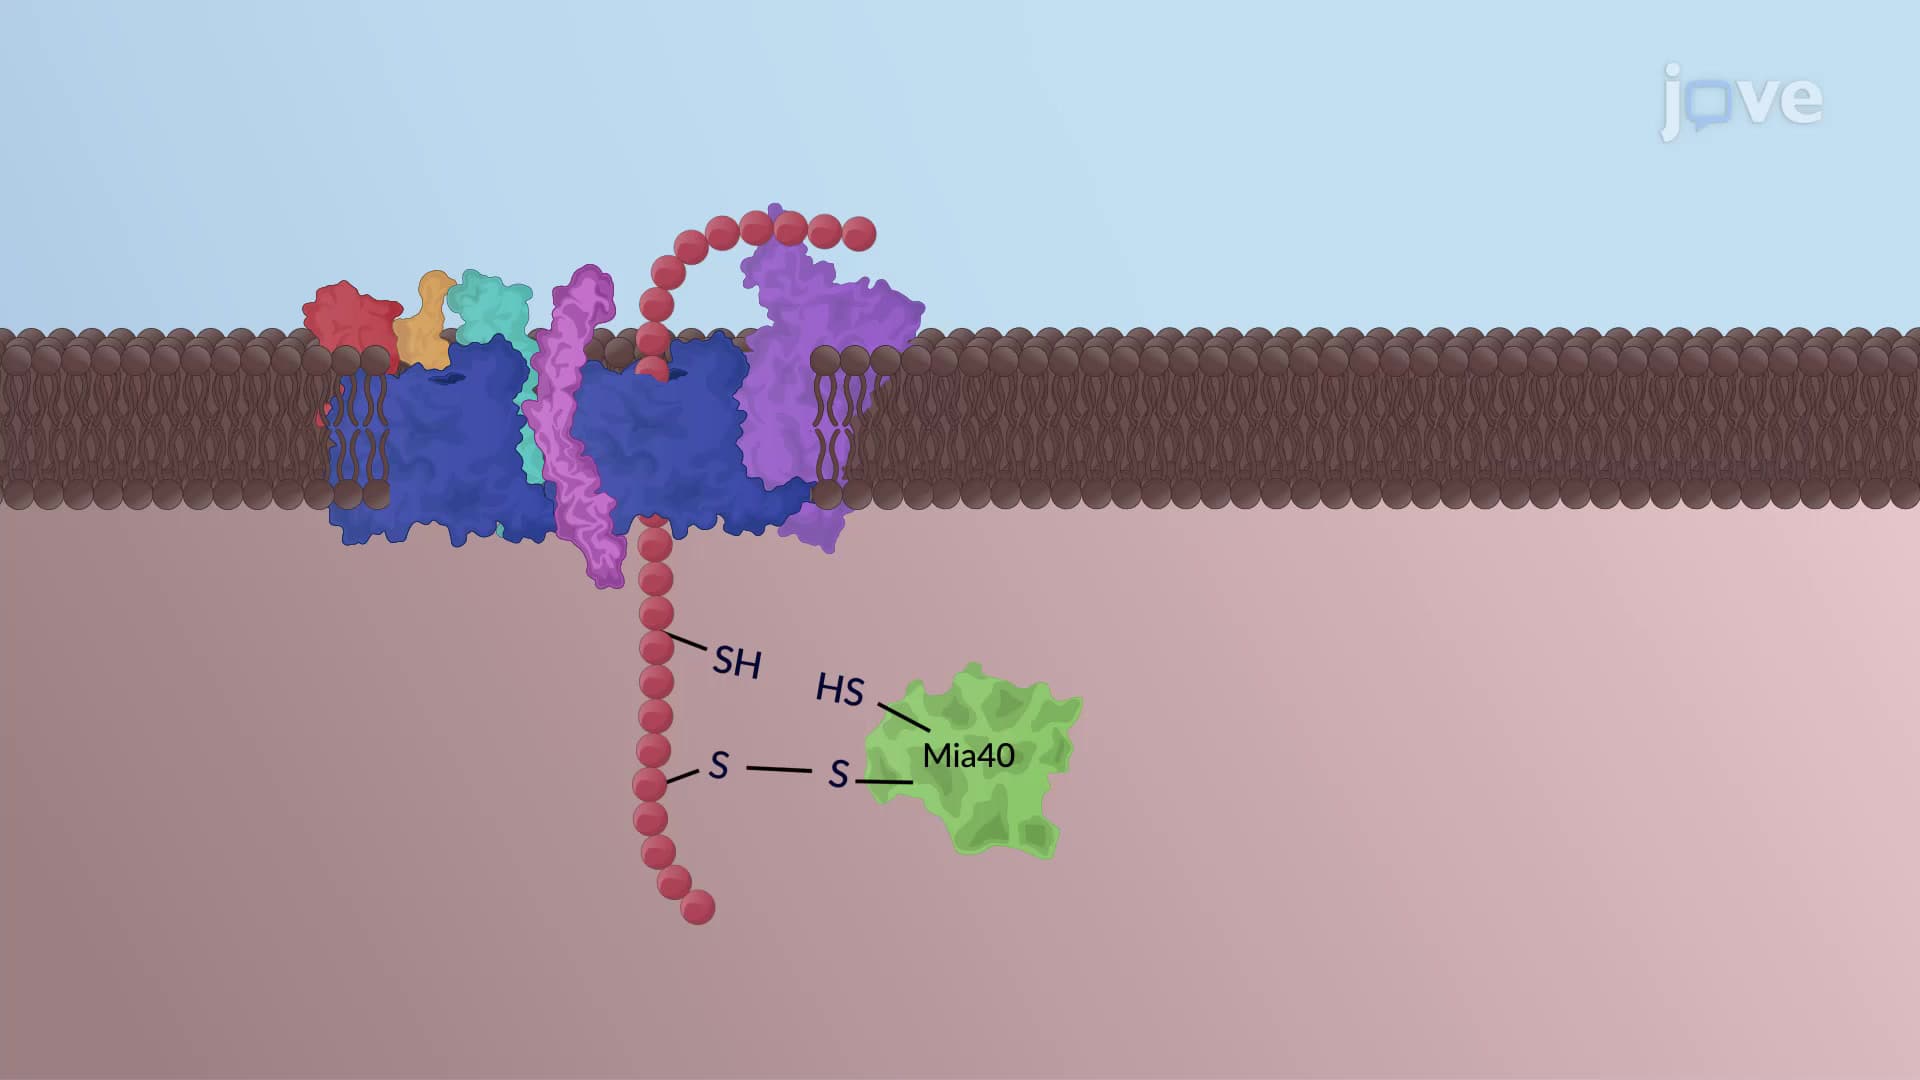
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...