Related Experiment Video
Updated: Nov 18, 2025

Characterization of Biological Absorption Spectra Spanning the Visible to the Short-Wave Infrared
Published on: January 10, 2025
XABOOM: An X-ray Absorption Benchmark of Organic Molecules Based on Carbon, Nitrogen, and Oxygen 1s → π* Transitions
Thomas Fransson1,2, Iulia E Brumboiu3,4, Marta L Vidal5
1Interdisciplinary Center for Scientific Computing, Ruprecht-Karls University, Im Neuenheimer Feld 205, 69120 Heidelberg, Germany.

Abstract:
The performance of several standard and popular approaches for calculating X-ray absorption spectra at the carbon, nitrogen, and oxygen K-edges of 40 primarily organic molecules up to the size of guanine has been evaluated, focusing on the low-energy and intense 1s → π* transitions. Using results obtained with CVS-ADC(2)-x and fc-CVS-EOM-CCSD as benchmark references, we investigate the performance of CC2, ADC(2), ADC(3/2), and commonly adopted density functional theory (DFT)-based approaches. Here, focus is on precision rather than on accuracy of transition energies and intensities-in other words, we target relative energies and intensities and the spread thereof, rather than absolute values. The use of exchange-correlation functionals tailored for time-dependent DFT calculations of core excitations leads to error spreads similar to those seen for more standard functionals, despite yielding superior absolute energies. Long-range corrected functionals are shown to perform particularly well compared to our reference data, showing error spreads in energy and intensity of 0.2-0.3 eV and ∼10%, respectively, as compared to 0.3-0.6 eV and ∼20% for a typical pure hybrid. In comparing intensities, state mixing can complicate matters, and techniques to avoid this issue are discussed. Furthermore, the influence of basis sets in high-level ab initio calculations is investigated, showing that reasonably accurate results are obtained with the use of 6-311++G**. We name this benchmark suite as XABOOM (X-ray absorption benchmark of organic molecules) and provide molecular structures and ground-state self-consistent field energies and spectroscopic data. We believe that it provides a good assessment of electronic structure theory methods for calculating X-ray absorption spectra and will become useful for future developments in this field.
More Related Videos
 10:03
Proton Transfer and Protein Conformation Dynamics in Photosensitive Proteins by Time-resolved Step-scan Fourier-transform Infrared Spectroscopy
Published on: June 27, 2014
 09:40
Measurement and Analysis of Atomic Hydrogen and Diatomic Molecular AlO, C2, CN, and TiO Spectra Following Laser-induced Optical Breakdown
Published on: February 14, 2014
Related Concept Videos
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 01:14
Molecular Spectroscopy: Absorption and Emission
 01:26
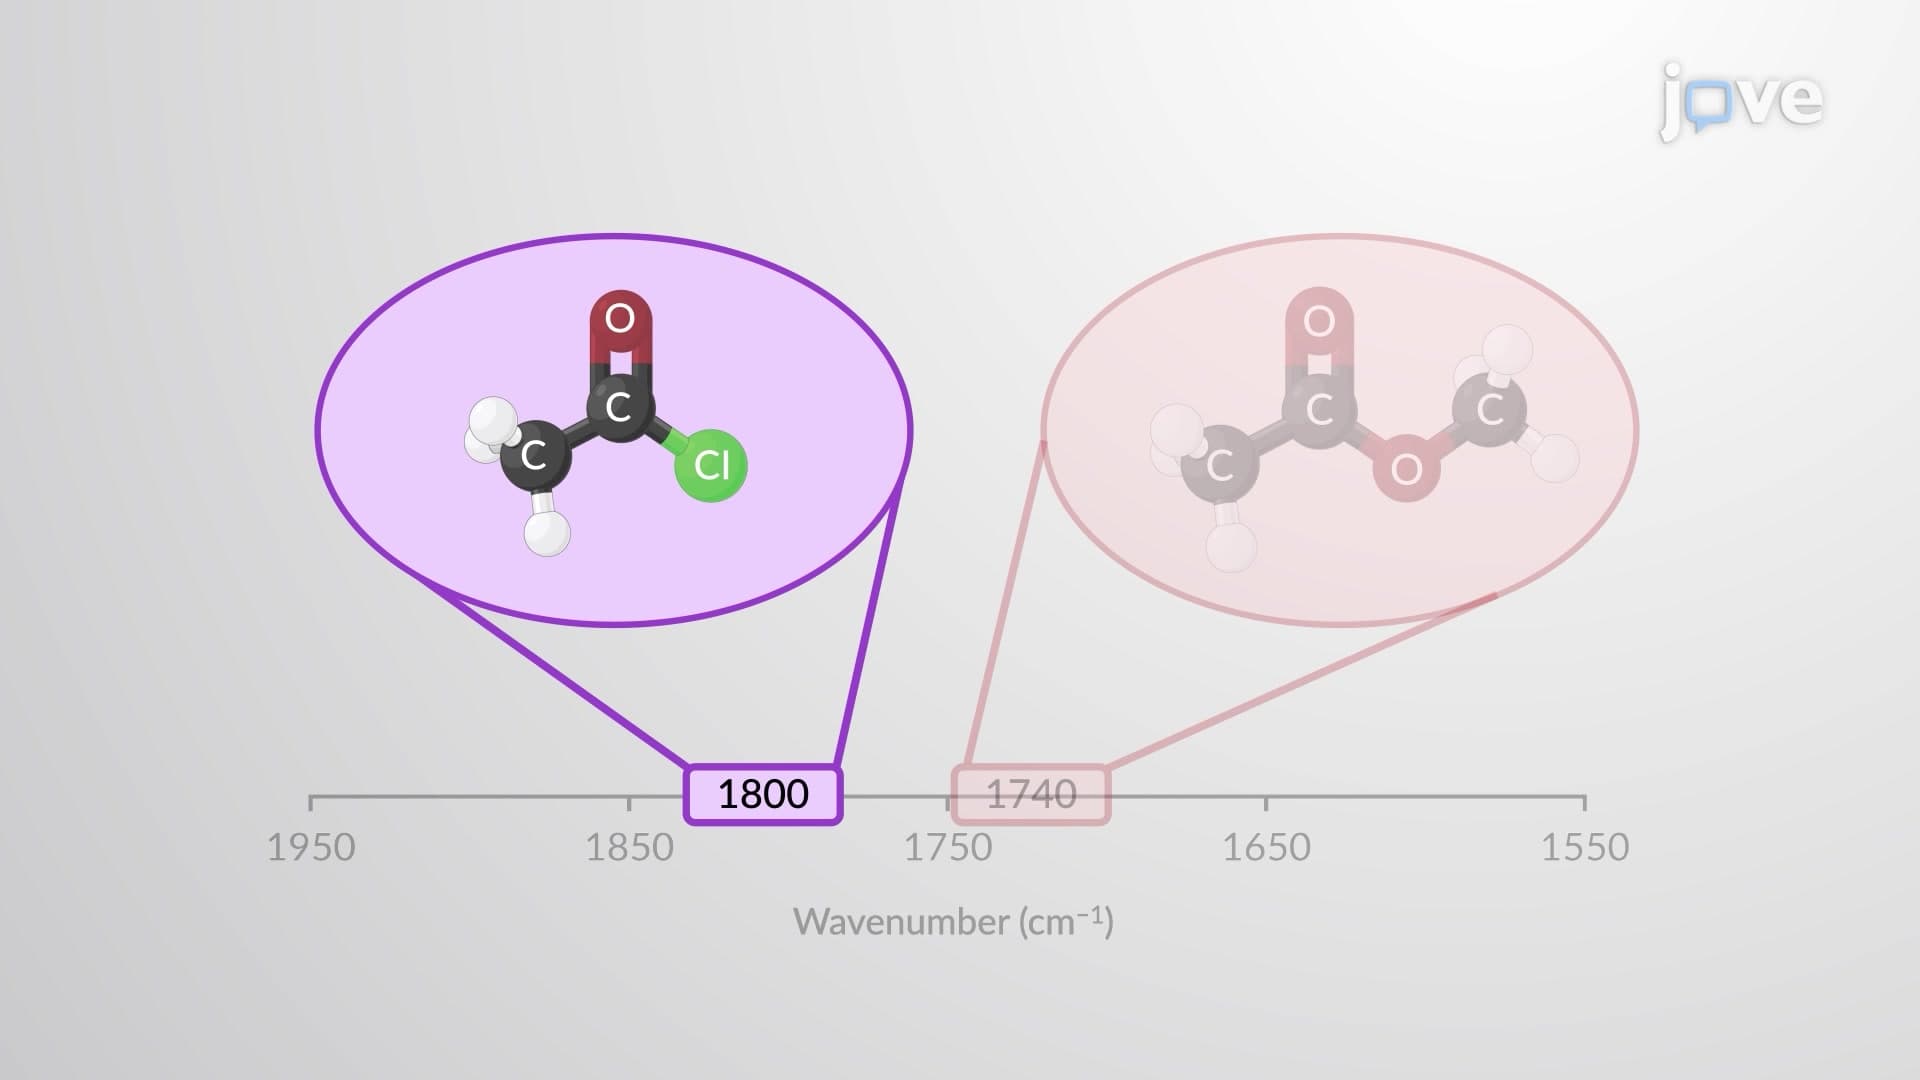
Spectroscopy of Carboxylic Acid Derivatives
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...
 01:14
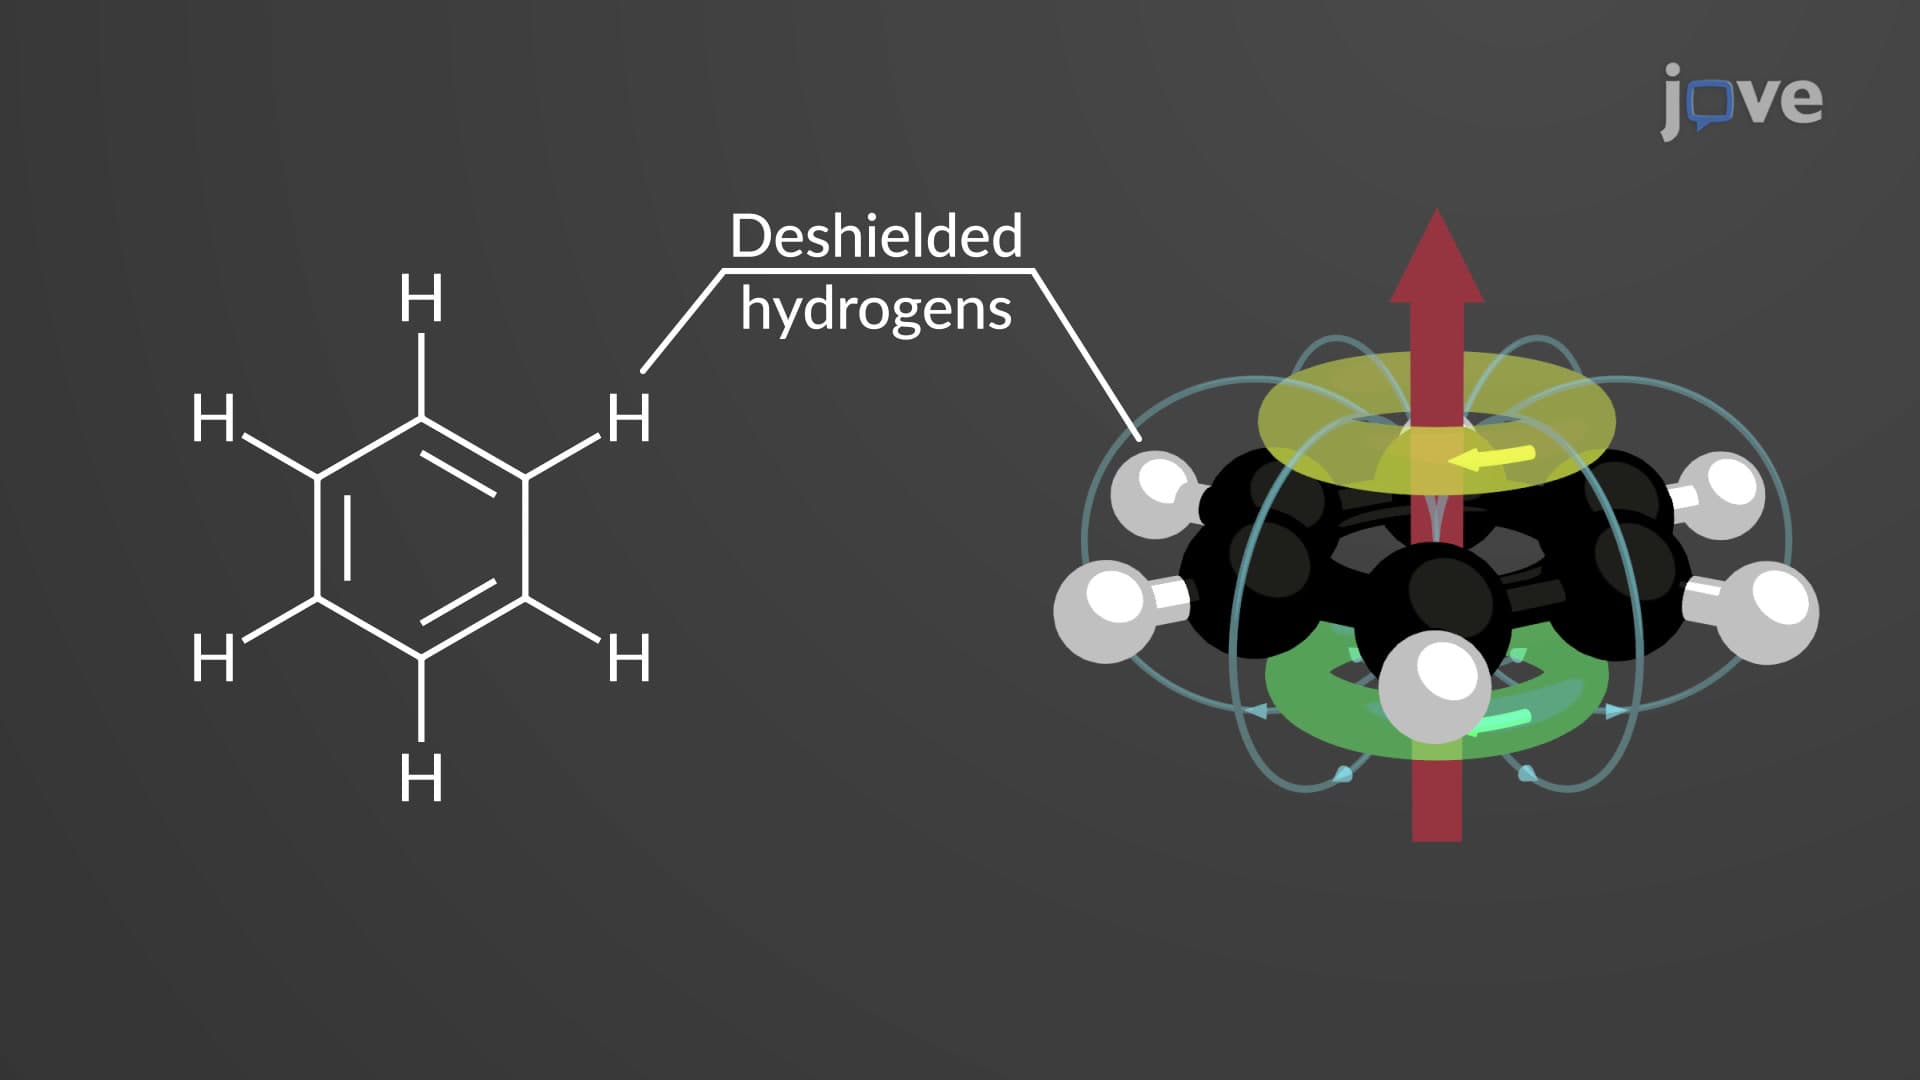
NMR Spectroscopy of Aromatic Compounds
 01:27
Atomic Absorption Spectroscopy: Overview
When irradiated by EMR of a particular wavelength, these...