Related Experiment Video
Updated: Nov 17, 2025

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Accelerated Computation of Free Energy Profile at Ab Initio Quantum Mechanical/Molecular Mechanics Accuracy via a
Jia-Ning Wang1, Wei Liu1, Pengfei Li1
1State Key Laboratory of Precision Spectroscopy, School of Physics and Electronic Science, East China Normal University, Shanghai 200062, China.

Abstract:
Although quantum mechanical/molecular mechanics (QM/MM) methods are now routinely applied to the studies of chemical reactions in condensed phases and enzymatic reactions, they may experience technical difficulties when the reactive region is varying over time. For instance, when the solvent molecules are directly participating in the reaction, the exchange of water molecules between the QM and MM regions may occur on a time scale comparable to the reaction time. To cope with this situation, several adaptive QM/MM schemes have been proposed. However, these methods either add significantly to the computational cost or introduce artificial restraints to the system. In this work, we developed a novel adaptive QM/MM scheme and applied it to the study of a nucleophilic addition reaction. In this scheme, the configuration sampling was performed with a small QM region (without solvent molecules), and the thermodynamic properties under another potential energy function with a larger QM region (with a certain number of solvent molecules and/or different levels of QM theory) are computed via extrapolation using the reference-potential method. Our simulation results show that this adaptive QM/MM scheme is numerically stable, at least for the case studied in this work. Furthermore, this method also offers an inexpensive way to examine the convergence of the QM/MM calculation with respect to the size of the QM region.
Related Concept Videos
 01:26
Thermodynamic Potentials
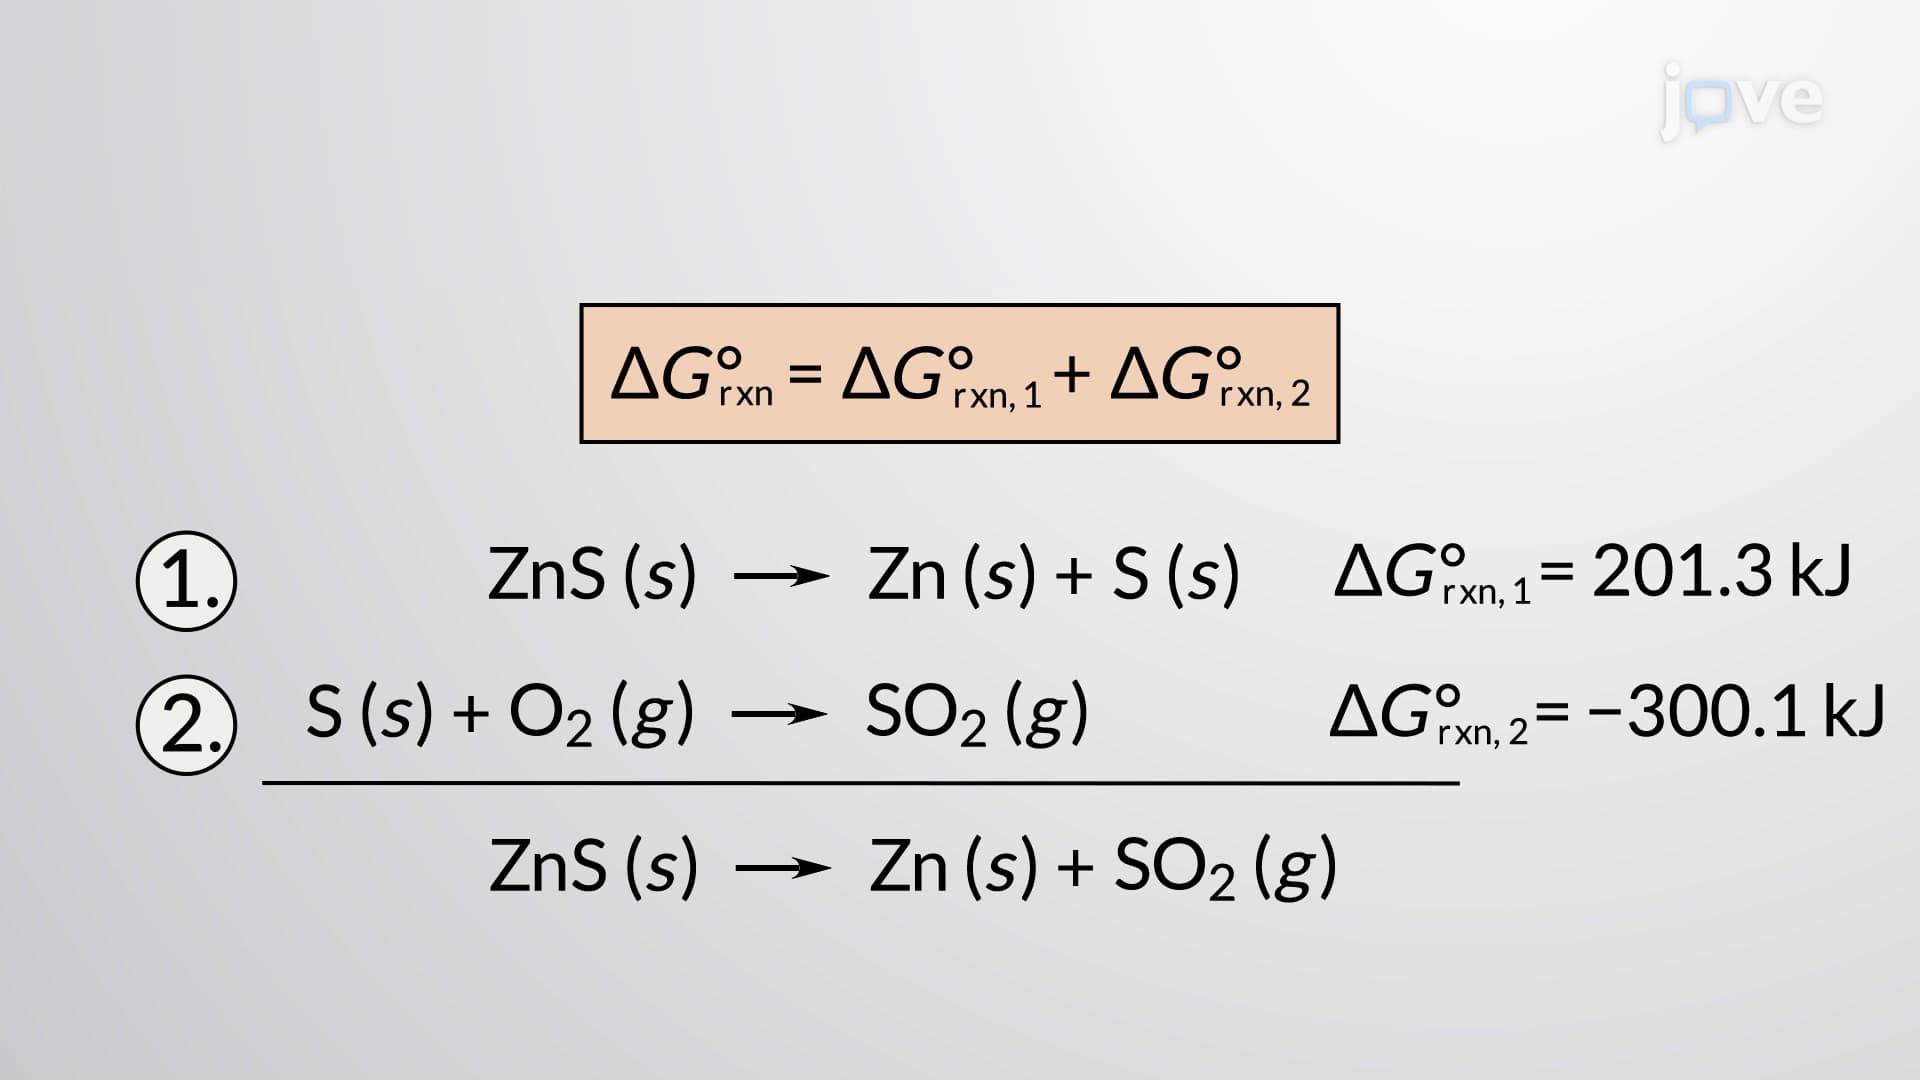 02:49
Calculating Standard Free Energy Changes
 01:16
Potential-Energy Criterion for Equilibrium
 03:25
Free Energy Changes for Nonstandard States
 01:10
Thermodynamics: Chemical Potential and Activity
The thermodynamic equilibrium constant is more accurately defined in terms of activity rather than concentration.
 01:10
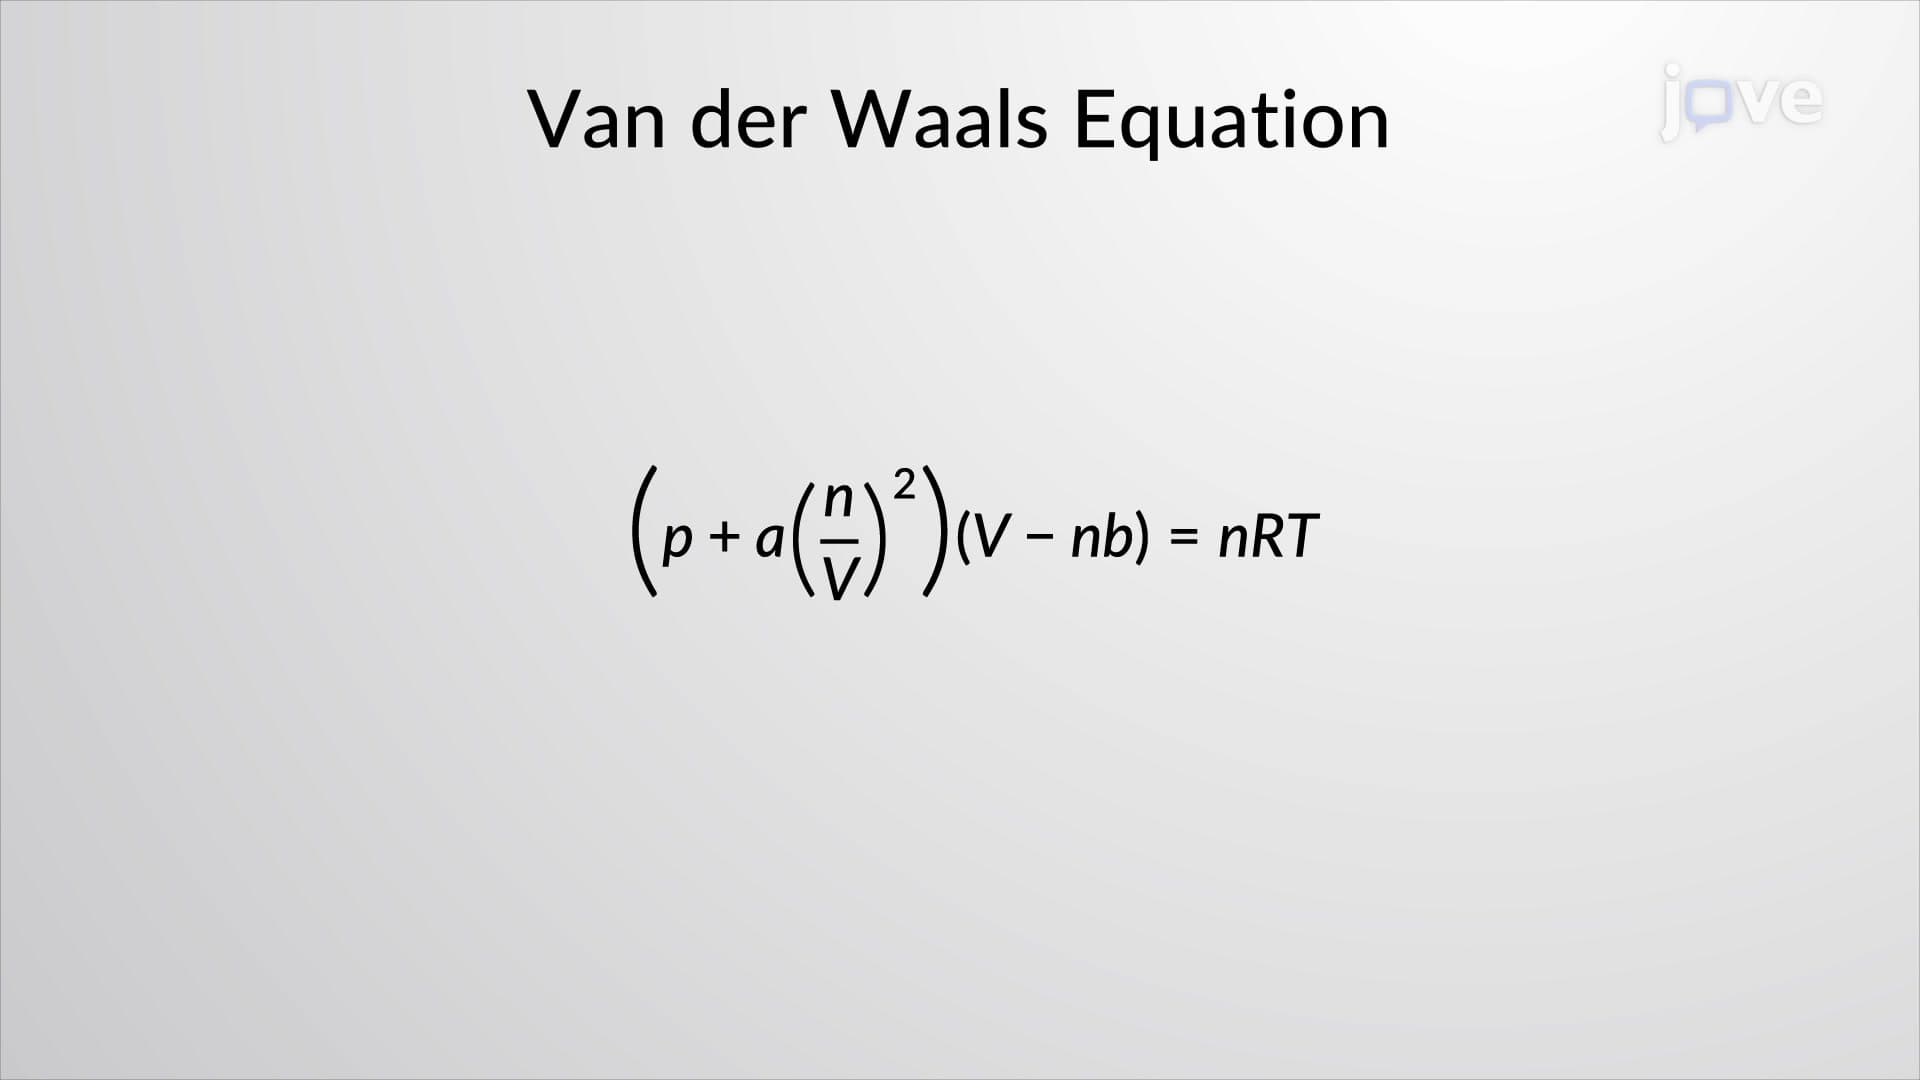
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...