Related Experiment Video
Updated: Nov 17, 2025

Analyzing Mitochondrial Transport and Morphology in Human Induced Pluripotent Stem Cell-Derived Neurons in Hereditary Spastic Paraplegia
Published on: February 9, 2020
Genomic variants causing mitochondrial dysfunction are common in hereditary lower motor neuron disease
Natalie Keller1,2,3, Cem Paketci4, Janine Altmueller5
1Institute of Human Genetics and Institute of Genetics, University of Cologne, Cologne, Germany.

Abstract:
Hereditary lower motor neuron diseases (LMND) other than 5q-spinal muscular atrophy (5q-SMA) can be classified according to affected muscle groups. Proximal and distal forms of non-5q-SMA represent a clinically and genetically heterogeneous spectrum characterized by significant overlaps with axonal forms of Charcot-Marie-Tooth (CMT) disease. A consensus for the best approach to molecular diagnosis needs to be reached, especially in light of continuous novel gene discovery and falling costs of next-generation sequencing (NGS). We performed exome sequencing (ES) in 41 families presenting with non-5q-SMA or axonal CMT, 25 of which had undergone a previous negative neuromuscular disease (NMD) gene panel analysis. The total diagnostic yield of ES was 41%. Diagnostic success in the cohort with a previous NMD-panel analysis was significantly extended by ES, primarily due to novel gene associated-phenotypes and uncharacteristic phenotypic presentations. We recommend early ES for individuals with hereditary LMND presenting uncharacteristic or significantly overlapping features. As mitochondrial dysfunction was the underlying pathomechanism in 47% of the solved individuals, we highlight the sensitivity of the anterior horn cell and peripheral nerve to mitochondrial imbalance as well as the necessity to screen for mitochondrial disorders in individuals presenting predominant lower motor neuron symptoms.
More Related Videos
 07:24
Genotyping Single Nucleotide Polymorphisms in the Mitochondrial Genome by Pyrosequencing
Published on: February 10, 2023
 06:53
Visualization of Mitochondrial Respiratory Function using Cytochrome C Oxidase / Succinate Dehydrogenase COX/SDH Double-labeling Histochemistry
Published on: November 23, 2011
Related Concept Videos
 02:59
Animal Mitochondrial Genetics
 01:48
ATP Synthase: Mechanism
 01:31
Translation
Translation Produces the Building Blocks of Life
Proteins are...
01:31Translation
Translation is the process of synthesizing proteins from the genetic information carried by messenger RNA (mRNA). Following transcription, it constitutes the final step in the expression of genes. This process is carried out by ribosomes, complexes of protein and specialized RNA molecules. Ribosomes, transfer RNA (tRNA), and other proteins produce a chain of amino acids—the polypeptide—as the end product of translation.
Translation Produces the Building Blocks of...
 01:39
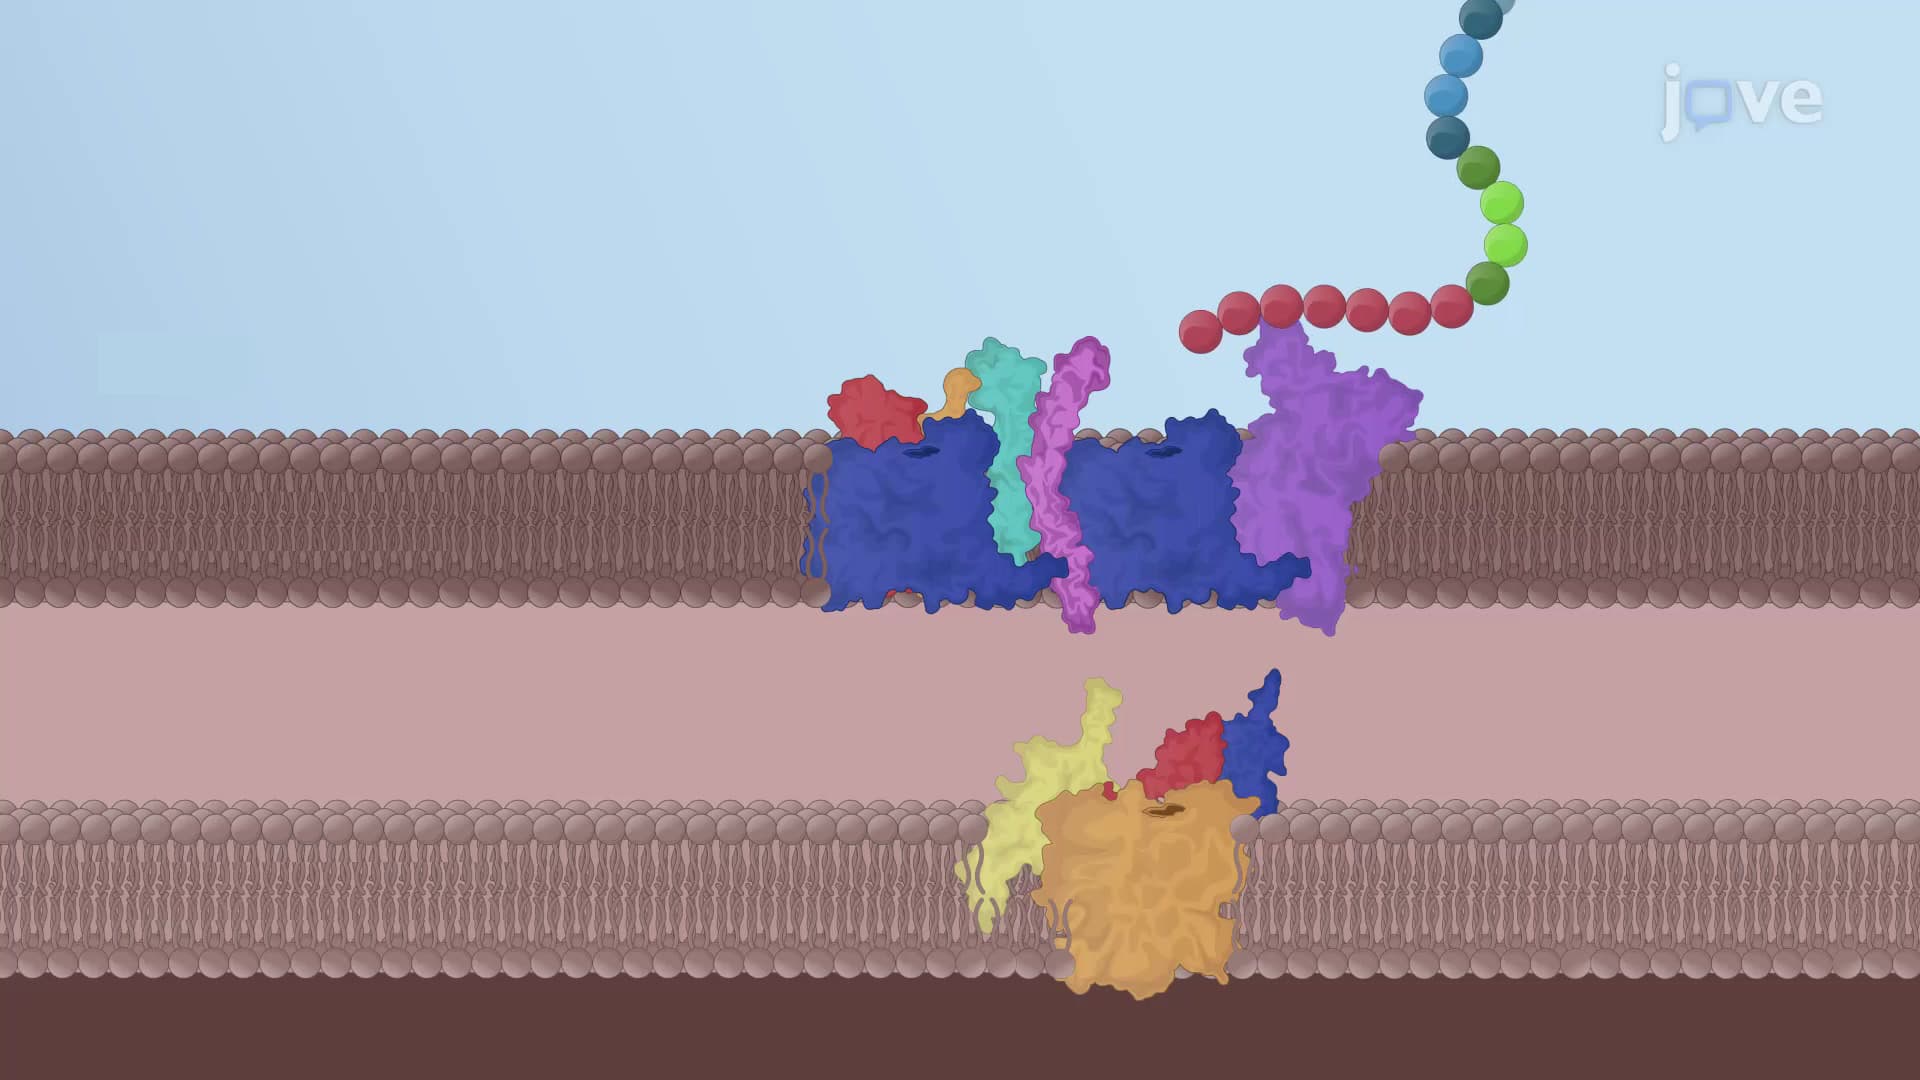
Mitochondrial Precursor Proteins
Most of the mitochondrial...
 01:19
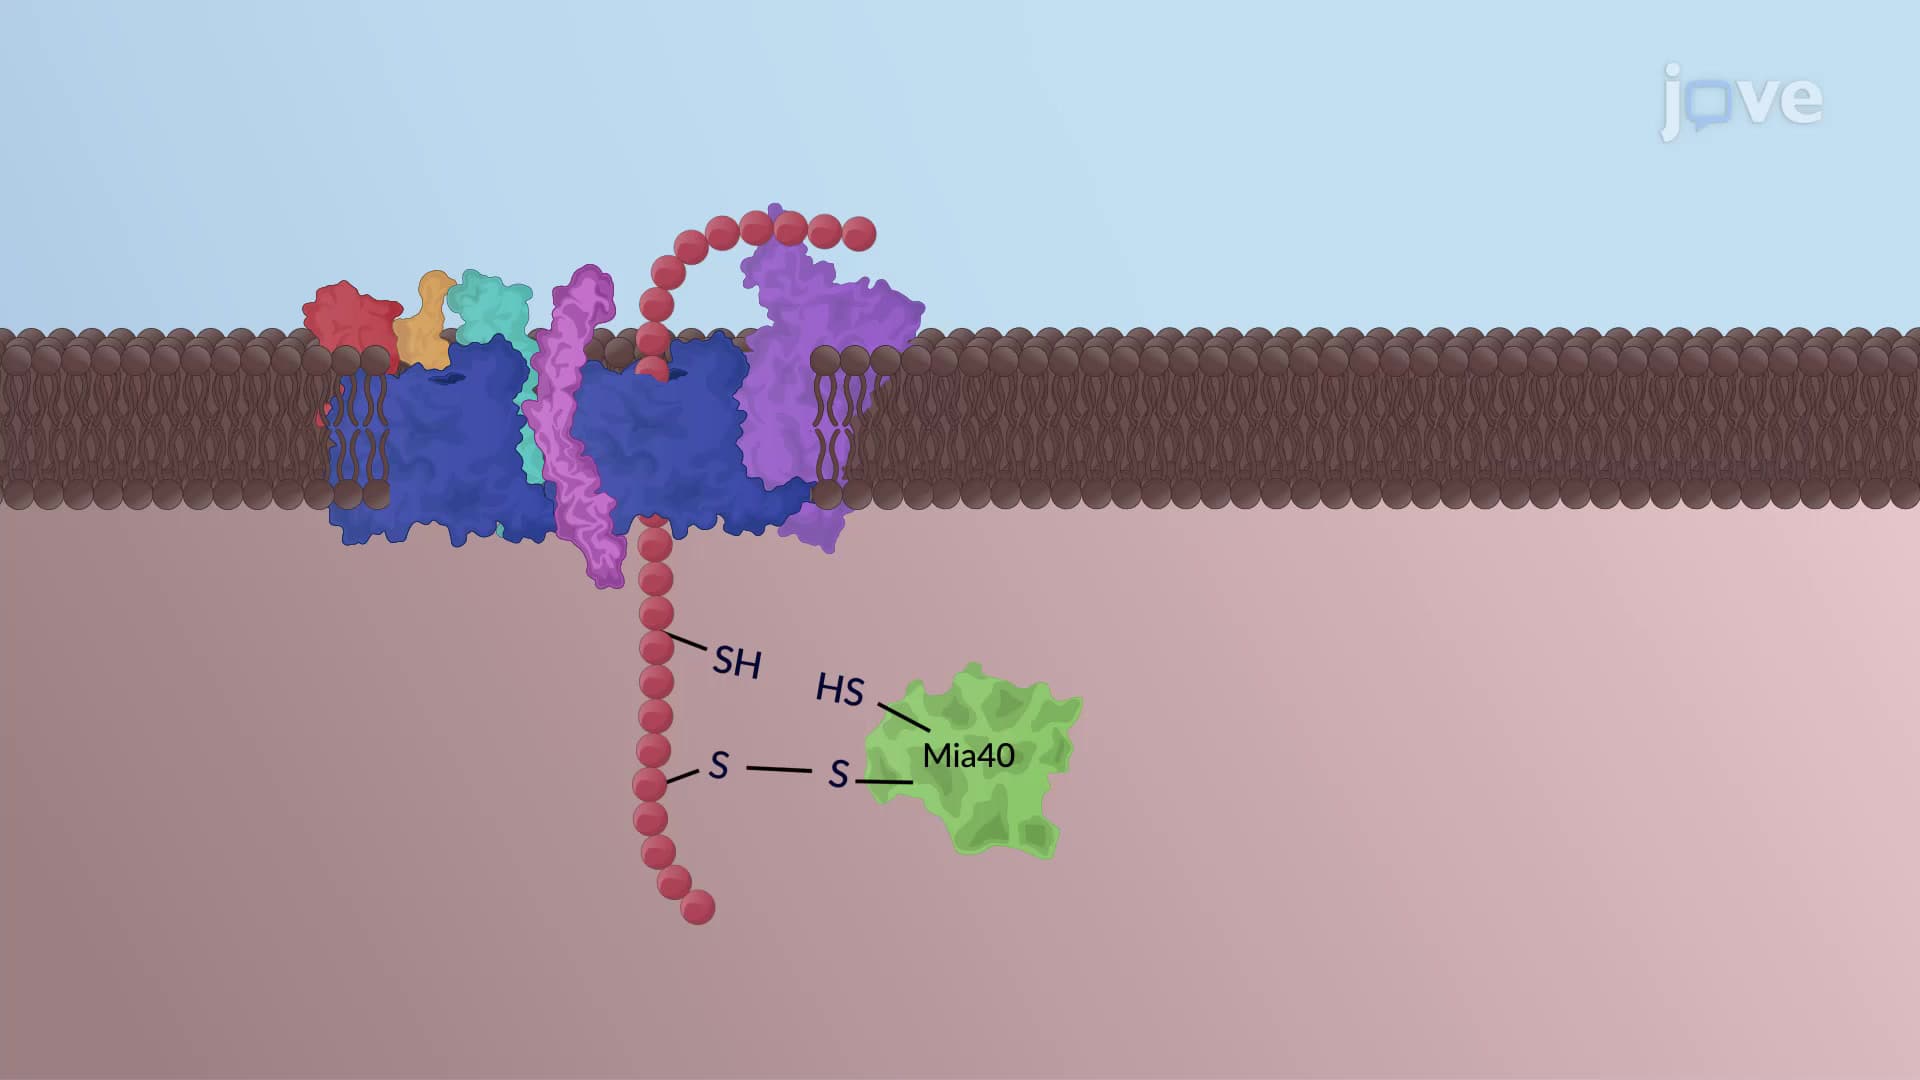
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...