Related Experiment Video
Updated: Nov 16, 2025

Detection of Rare Genomic Variants from Pooled Sequencing Using SPLINTER
Published on: June 23, 2012
Efficient and flexible Integration of variant characteristics in rare variant association studies using integrated
Hana Susak1,2,3, Laura Serra-Saurina4,5,6, German Demidov1,7,8
1Centre for Genomic Regulation (CRG), The Barcelona Institute of Science and Technology, Barcelona, Spain.

Abstract:
Rare variants are thought to play an important role in the etiology of complex diseases and may explain a significant fraction of the missing heritability in genetic disease studies. Next-generation sequencing facilitates the association of rare variants in coding or regulatory regions with complex diseases in large cohorts at genome-wide scale. However, rare variant association studies (RVAS) still lack power when cohorts are small to medium-sized and if genetic variation explains a small fraction of phenotypic variance. Here we present a novel Bayesian rare variant Association Test using Integrated Nested Laplace Approximation (BATI). Unlike existing RVAS tests, BATI allows integration of individual or variant-specific features as covariates, while efficiently performing inference based on full model estimation. We demonstrate that BATI outperforms established RVAS methods on realistic, semi-synthetic whole-exome sequencing cohorts, especially when using meaningful biological context, such as functional annotation. We show that BATI achieves power above 70% in scenarios in which competing tests fail to identify risk genes, e.g. when risk variants in sum explain less than 0.5% of phenotypic variance. We have integrated BATI, together with five existing RVAS tests in the 'Rare Variant Genome Wide Association Study' (rvGWAS) framework for data analyzed by whole-exome or whole genome sequencing. rvGWAS supports rare variant association for genes or any other biological unit such as promoters, while allowing the analysis of essential functionalities like quality control or filtering. Applying rvGWAS to a Chronic Lymphocytic Leukemia study we identified eight candidate predisposition genes, including EHMT2 and COPS7A.
More Related Videos
 11:35
Screening for Functional Non-coding Genetic Variants Using Electrophoretic Mobility Shift Assay EMSA and DNA-affinity Precipitation Assay DAPA
Published on: August 21, 2016
 09:34
Targeted Next-generation Sequencing and Bioinformatics Pipeline to Evaluate Genetic Determinants of Constitutional Disease
Published on: April 4, 2018
Related Concept Videos
 02:26
Comparing Copy Number Variations and SNPs
Copy number variations or CNVs are the structural variations that cover more than 1kb of DNA sequence. The single nucleotide polymorphism (SNP), on the other hand, is a single nucleotide change or a point mutation that is found in more than 1%...
 01:05
Single Nucleotide Polymorphisms-SNPs
 01:11
Genome-wide Association Studies-GWAS
GWAS does not require the identification of the target gene involved in...
 01:11
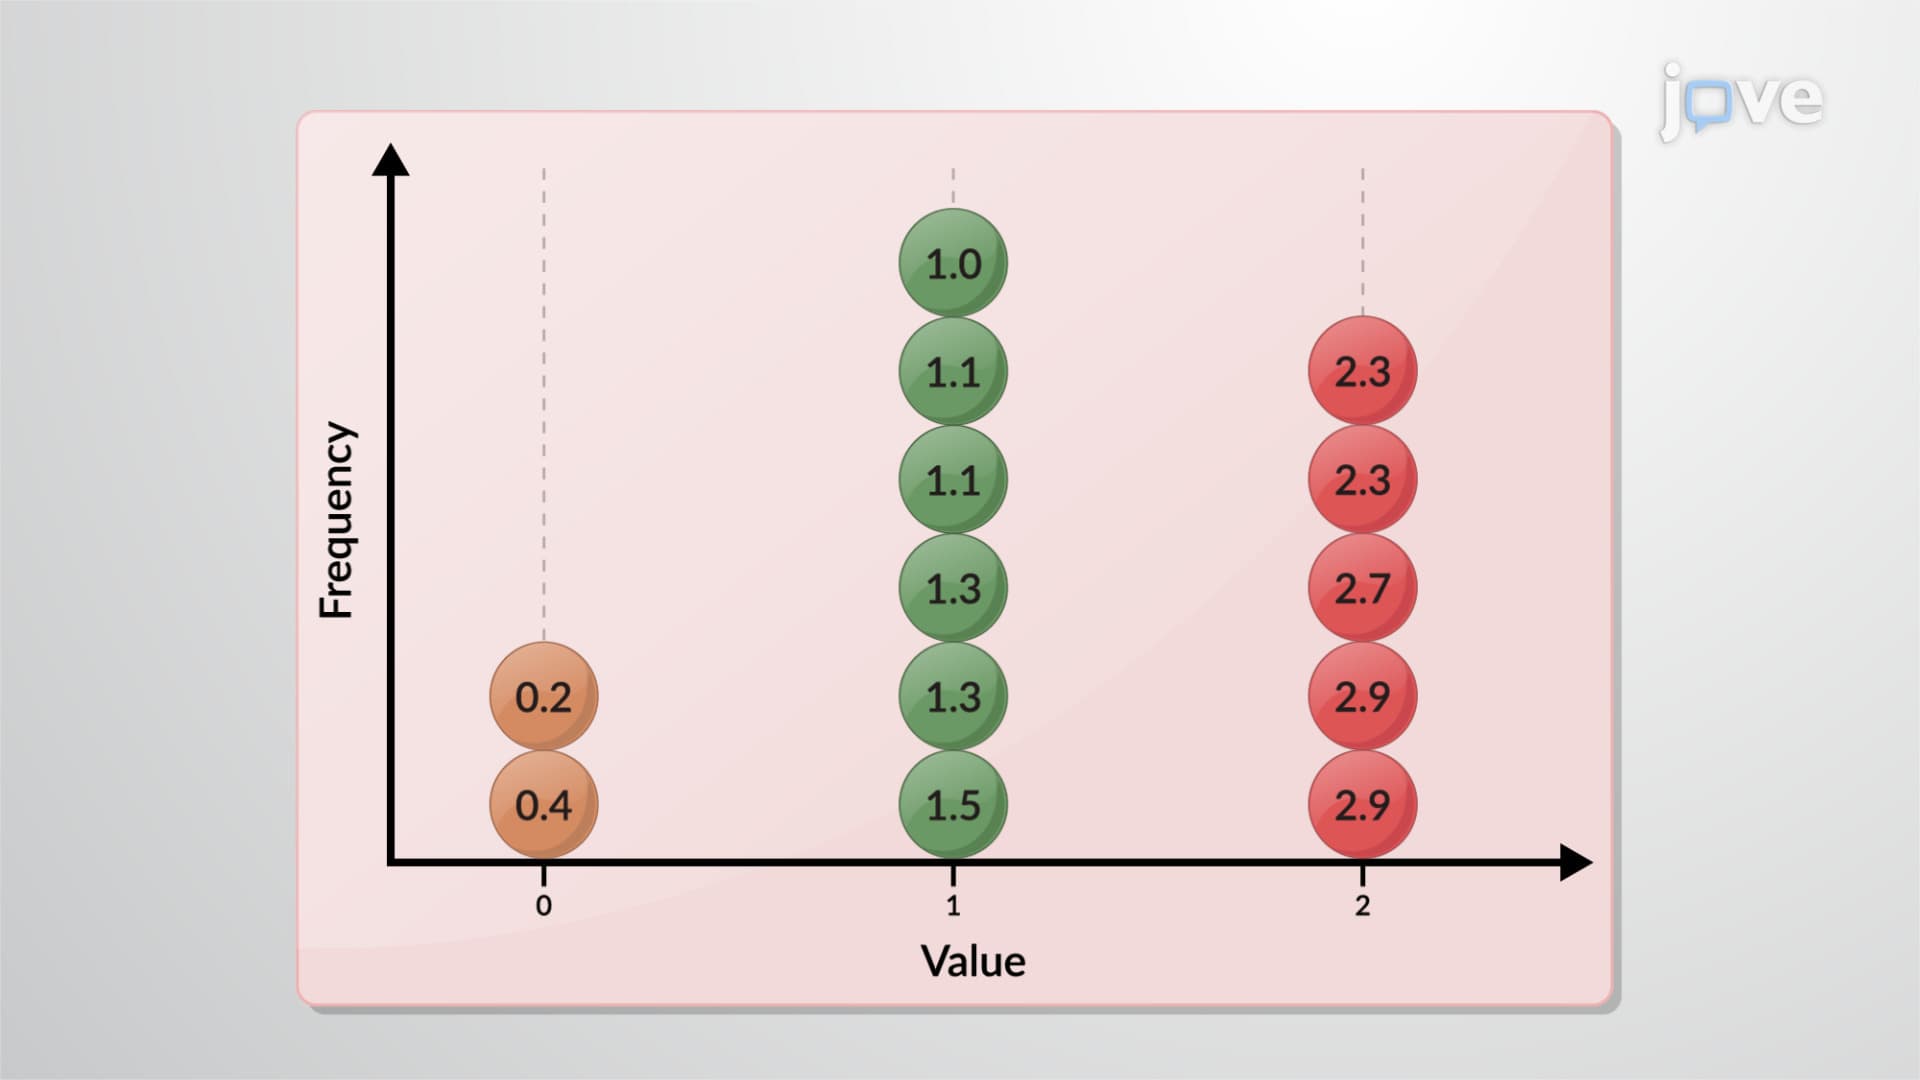
Variability: Analysis
The range is a simple measure of variability, indicating the difference between the highest and...
 01:25
Genetic Variation
Genes exist in different versions called alleles,...