Related Experiment Video
Updated: Nov 14, 2025

Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Dynamics and kinetics of the Si(1D) + H2/D2 reactions on a new global ab initio potential energy surface
Jianwei Cao1, Yanan Wu2, Haitao Ma1
1Beijing National Laboratory for Molecular Sciences, Institute of Chemistry, Chinese Academy of Sciences, Beijing 100190, China. bian@iccas.ac.cn.

Abstract:
Recent studies on the exothermic complex-forming reactions have improved our understanding of complex-forming reactions greatly, however, so far a similar level of study on endothermic ones has been rather limited. In this work, the endothermic complex-forming reaction Si(1D) + H2 → SiH + H and its deuterated isotopic variant are investigated by quantum dynamics (QD) and ring polymer molecular dynamics (RPMD) calculations on a new global ab initio potential energy surface (PES) for the ground electronic state, which is constructed based on 8996 symmetry unique points computed at the icMRCI+Q/aug-cc-pVQZ level. The PES reproduces our ab initio data very well in the dynamically important regions, on which the ro-vibrational energy levels of SiH2 are calculated and general good agreement with experiment is obtained. The integral cross sections and product angular and state distributions are computed in a wide range of collision energies, and interesting dynamics behaviors are revealed. The rate coefficients are also investigated, which display an exponential rise from 2.09 × 10⁻²⁰ to 6.00 × 10⁻¹² cm3 s-1 for the Si(1D) + H2 reaction as the temperature increases from 300 to 1500 K, in contrast to the nearly temperature-independent behavior of exothermic complex-forming reactions. In addition, the applicability of the RPMD approach is demonstrated, and the kinetic isotope effect is investigated, the ratio of which decreases from 7.89 (300 K) to 1.70 (1500 K). The effects of tunneling and initial rotational excitation are also discussed.
Related Concept Videos
 03:51
Molecular Orbital Theory II
 01:28
Stability of Conjugated Dienes
A comparison of the enthalpies of hydrogenation of dienes reveals that conjugated dienes release less heat on hydrogenation, rendering them more stable than their nonconjugated analogs.
 01:23
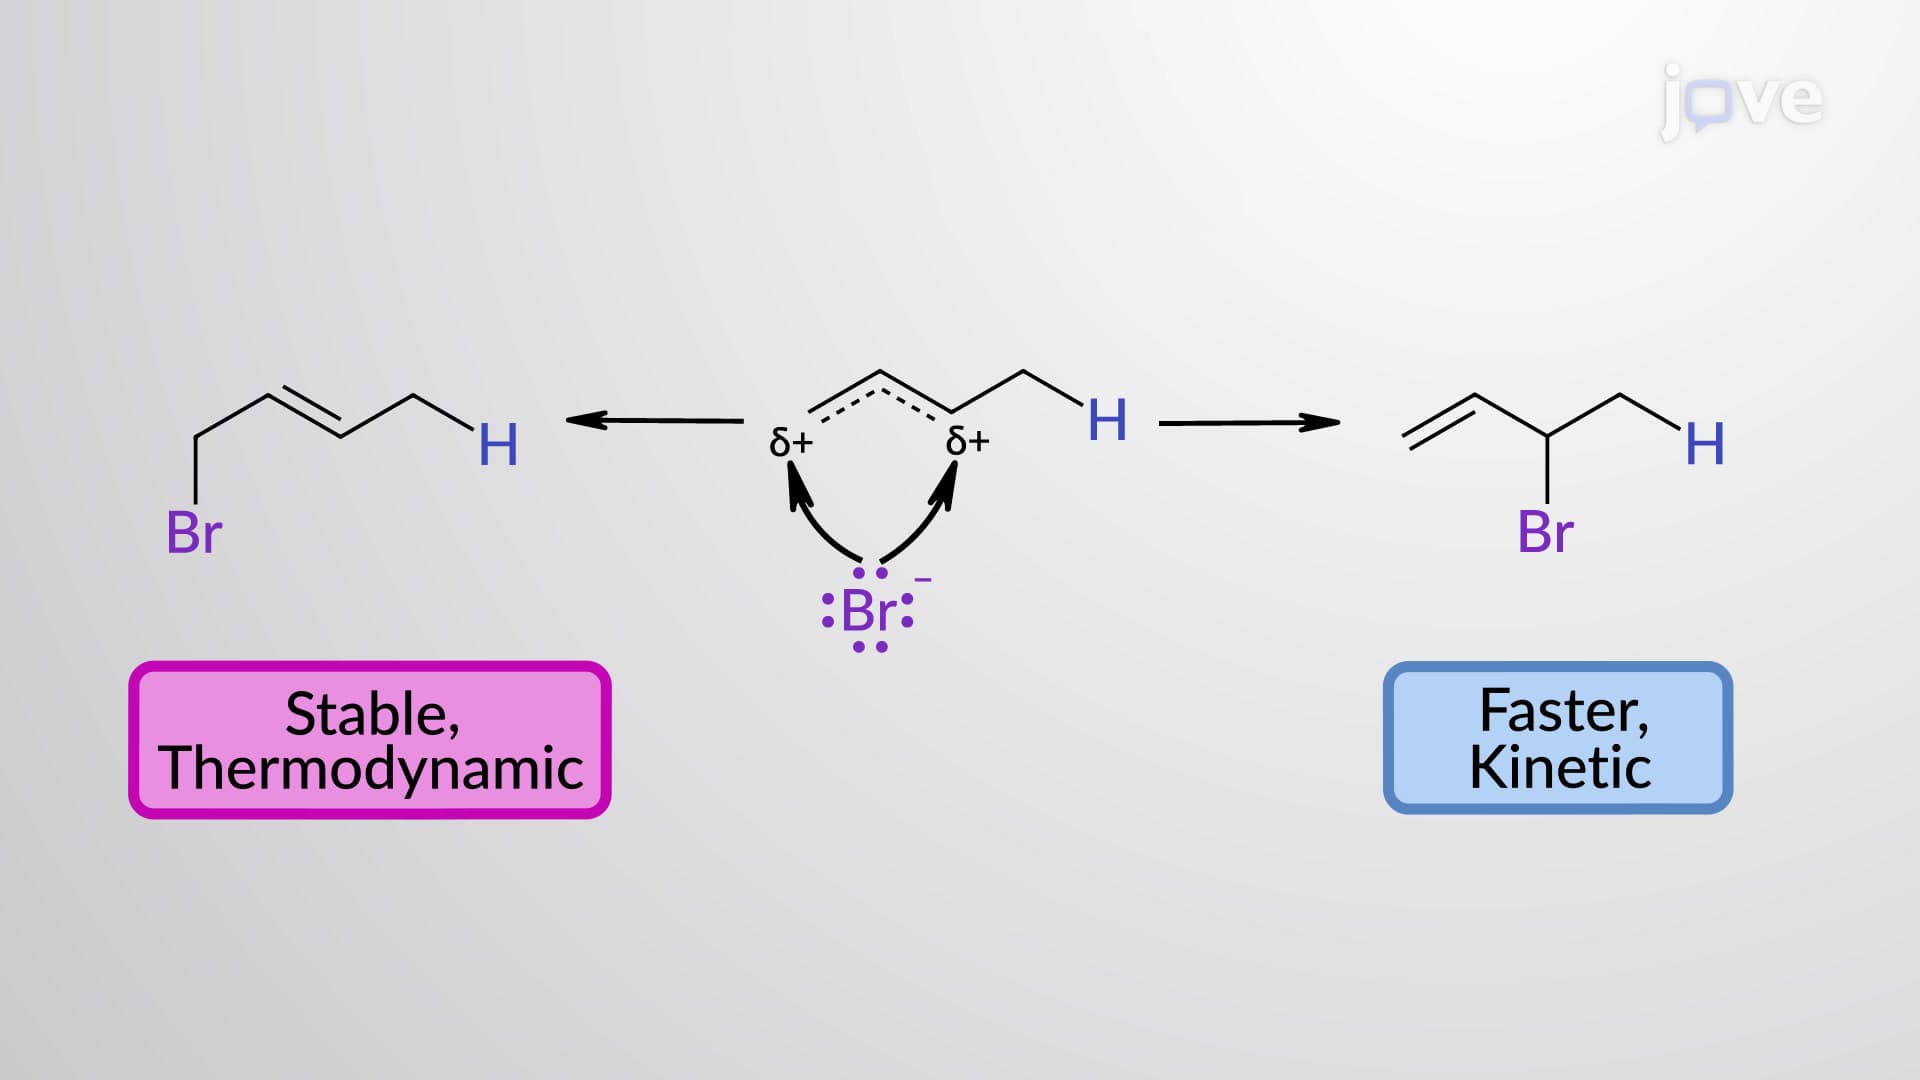
Electrophilic Addition of HX to 1,3-Butadiene: Thermodynamic vs Kinetic Control
 02:38
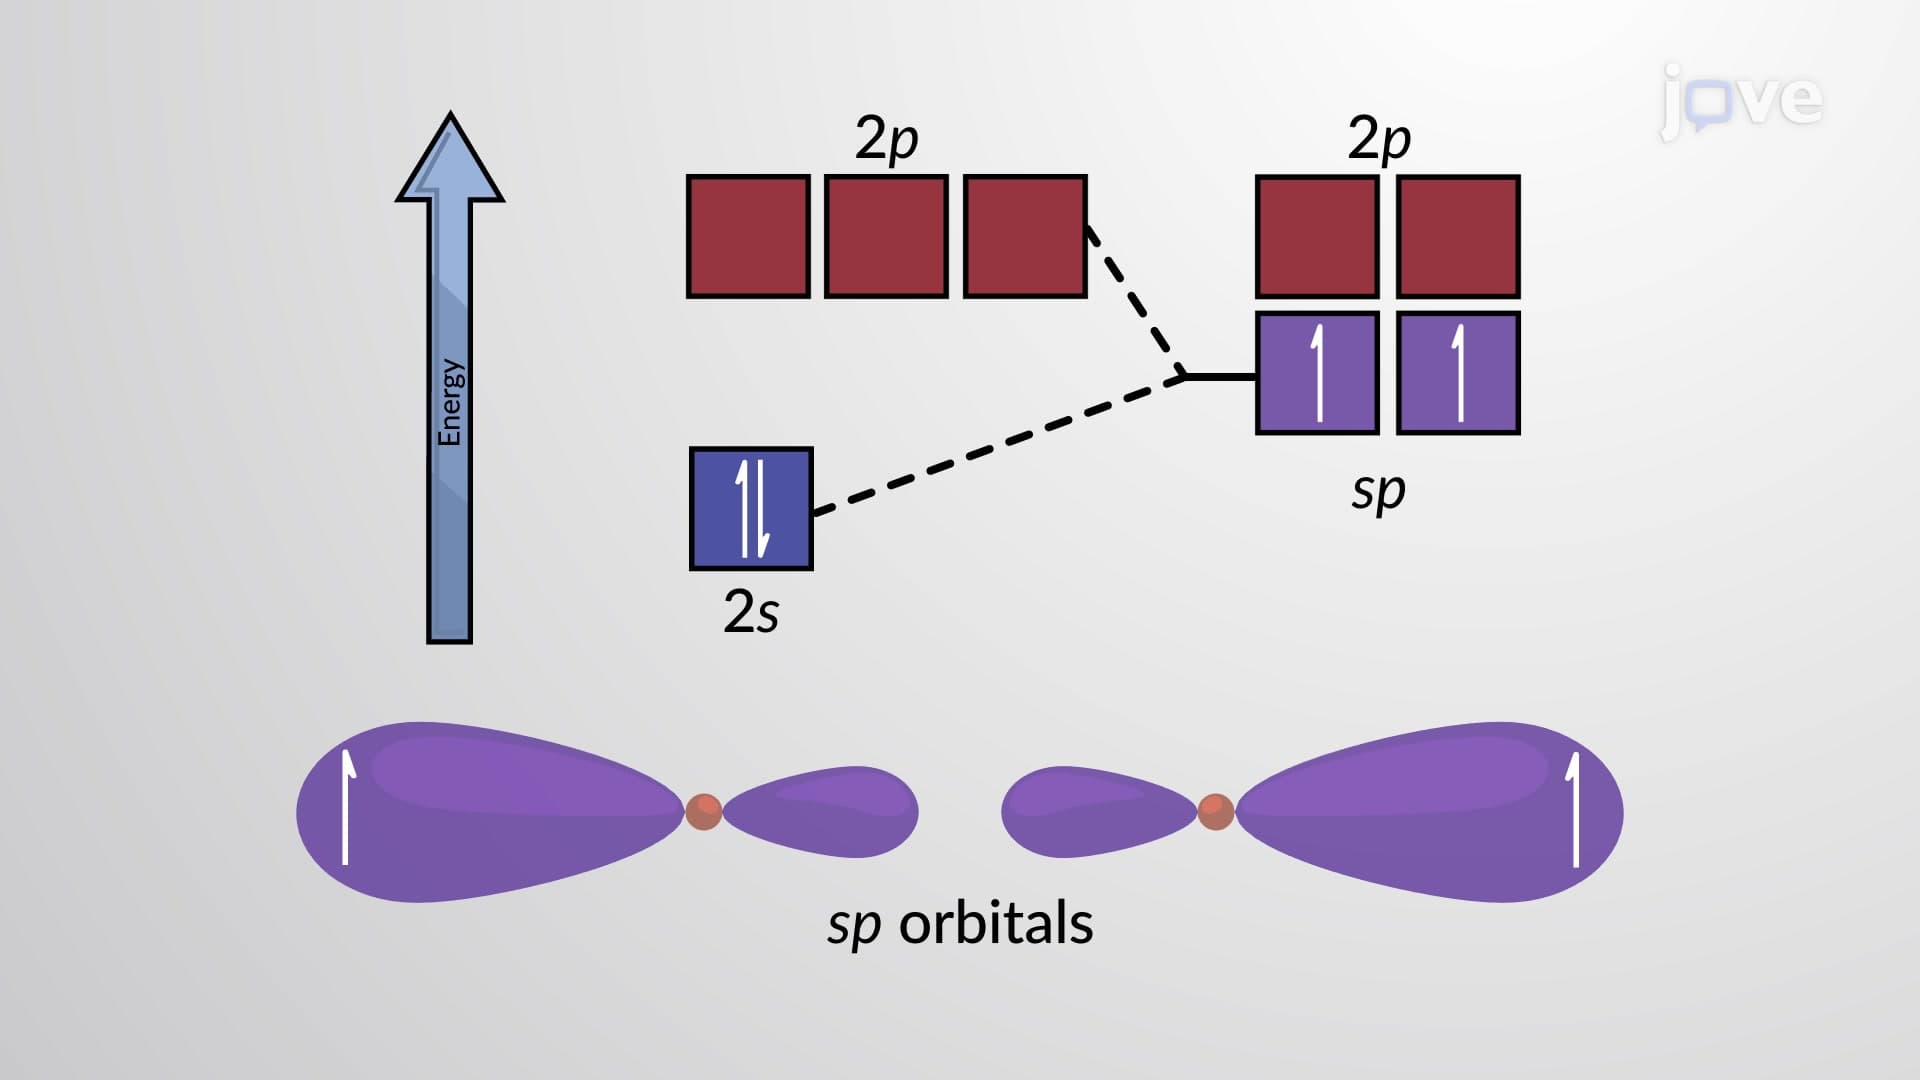
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 03:35
Hybridization of Atomic Orbitals II
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...