Related Experiment Video
Updated: Nov 12, 2025
An Intestine/Liver Microphysiological System for Drug Pharmacokinetic and Toxicological Assessment
Published on: December 3, 2020
Regulatory guidelines do not accurately predict tolvaptan and metabolite interactions at BCRP, OATP1B1, and OAT3
Susan E Shoaf1, Patricia Bricmont2, Jennifer Repella Gordon2
1Quantitative Pharmacology, Otsuka Pharmaceutical Development & Commercialization, Inc., Rockville, Maryland, USA.

Abstract:
Tolvaptan (TLV) was US Food and Drug Administration (FDA)-approved for the indication to slow kidney function decline in adults at risk of rapidly progressing autosomal dominant polycystic kidney disease in 2018. In vitro, TLV was a breast cancer resistance protein (BCRP) inhibitor, whereas the oxobutyric acid metabolite of TLV (DM-4013) was an inhibitor of organic anion transport polypeptide (OATP)1B1 and organic anion transporter (OAT)3. Based on the 2017 FDA guidance, potential for clinically relevant inhibition at these transporters was indicated for the highest TLV regimen. Consequently, two postmarketing clinical trials in healthy subjects were required. In trial 1, 5 mg rosuvastatin calcium (BCRP and OATP1B1 substrate) was administered alone, with 90 mg TLV or 48 h following 7 days of once daily 300 mg TLV (i.e., in the presence of DM-4103). In trial 2, 40 mg furosemide (OAT3 substrate) was administered alone and in presence of DM-4103. For BCRP, rosuvastatin geometric mean ratios (90% confidence intervals [CIs]) for maximum plasma concentration (Cmax ) were 1.54 (90% CI 1.26-1.88) and for area under the concentration-time curve from time 0 to the time of the last measurable concentration (AUCt ) were 1.69 (90% CI 1.34-2.14), indicating no clinically significant interaction. DM-4103 produced no clinically meaningful changes in rosuvastatin or furosemide concentrations, indicating no inhibition at OATP1B1 or OAT3. The BCRP prediction assumed the drug dose is completely soluble in 250 ml; TLV has solubility of ~0.01 g/250 ml. For OATP1B1/OAT3, if fraction unbound for plasma protein binding (PPB) is less than 1%, then 1% is assumed. DM-4103 has PPB greater than 99.8%. Use of actual drug substance solubility and unbound fraction in plasma would have produced predictions consistent with the clinical results.
Insights
Tolvaptan (TLV) did not show clinically significant drug interactions with rosuvastatin or furosemide, despite in vitro predictions. Postmarketing trials confirmed TLV’s safety profile regarding BCRP, OATP1B1, and OAT3 transporters.
Area of Science:
- Pharmacology
- Drug Metabolism and Transporter Interactions
Background:
- Tolvaptan (TLV) is FDA-approved to slow kidney function decline in autosomal dominant polycystic kidney disease.
- In vitro studies indicated TLV and its metabolite DM-4103 could inhibit breast cancer resistance protein (BCRP), organic anion transport polypeptide (OATP)1B1, and organic anion transporter (OAT)3.
Purpose of the Study:
- To evaluate potential drug-drug interactions between Tolvaptan (TLV) and substrates of BCRP, OATP1B1, and OAT3 transporters.
- To assess the clinical relevance of in vitro predictions for TLV and its metabolite DM-4103 on transporter inhibition.
Main Methods:
- Two postmarketing clinical trials in healthy subjects were conducted.
- Trial 1: Rosuvastatin (BCRP/OATP1B1 substrate) co-administered with TLV or DM-4103.
- Trial 2: Furosemide (OAT3 substrate) co-administered with DM-4103.
Main Results:
- Rosuvastatin exposure showed no clinically significant changes when co-administered with TLV or DM-4103, indicating no relevant BCRP or OATP1B1 inhibition.
- Furosemide concentrations remained unchanged in the presence of DM-4103, demonstrating no OAT3 inhibition.
- In vitro prediction discrepancies were attributed to assumptions about drug solubility and plasma protein binding.
Conclusions:
- Tolvaptan (TLV) does not appear to cause clinically significant drug interactions via BCRP, OATP1B1, or OAT3 inhibition at the studied doses.
- The clinical findings support the safety of TLV concerning these major drug transporters.
- Actual drug properties, like solubility and unbound fraction, are crucial for accurate in vitro to in vivo extrapolation of drug interactions.
More Related Videos
 16:02
Demonstration of the Sequence Alignment to Predict Across Species Susceptibility Tool for Rapid Assessment of Protein Conservation
Published on: February 10, 2023
 09:50
Technical Aspect of the Automated Synthesis and Real-Time Kinetic Evaluation of [11C]SNAP-7941
Published on: April 28, 2019
Related Concept Videos
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion,...
 01:14
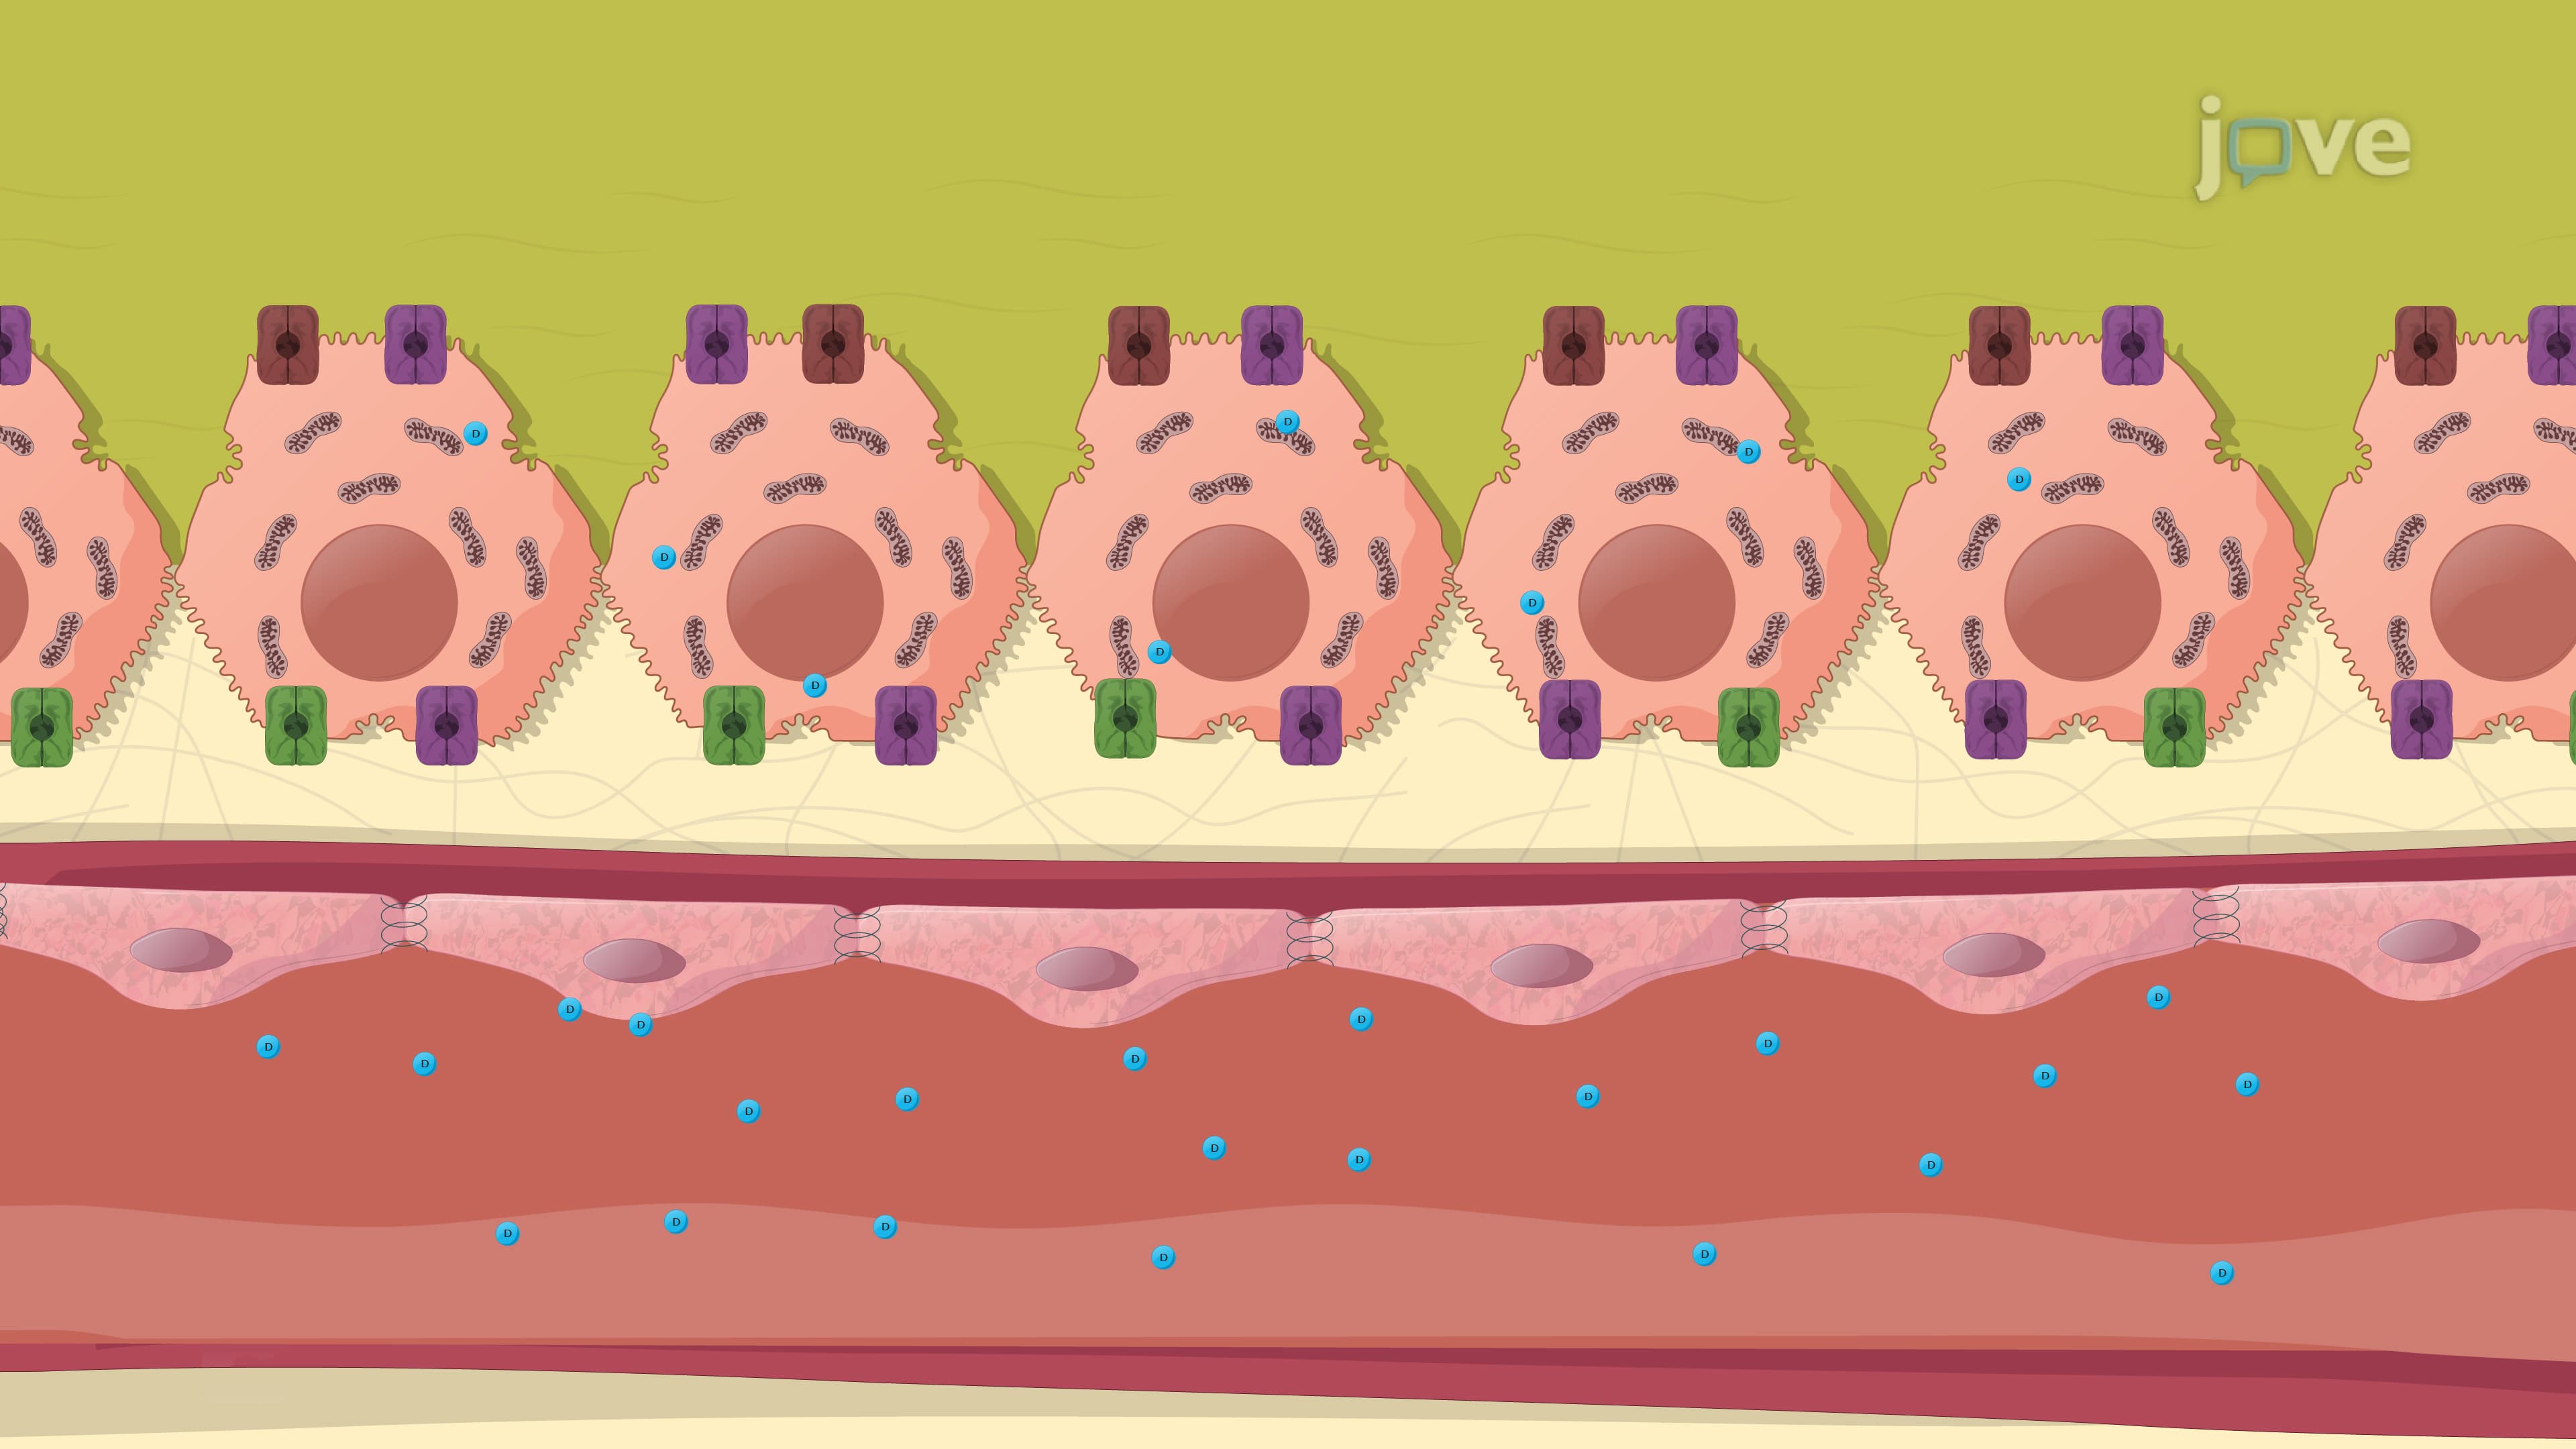
Hepatic Drug Clearance: Role of Transporters
 01:27
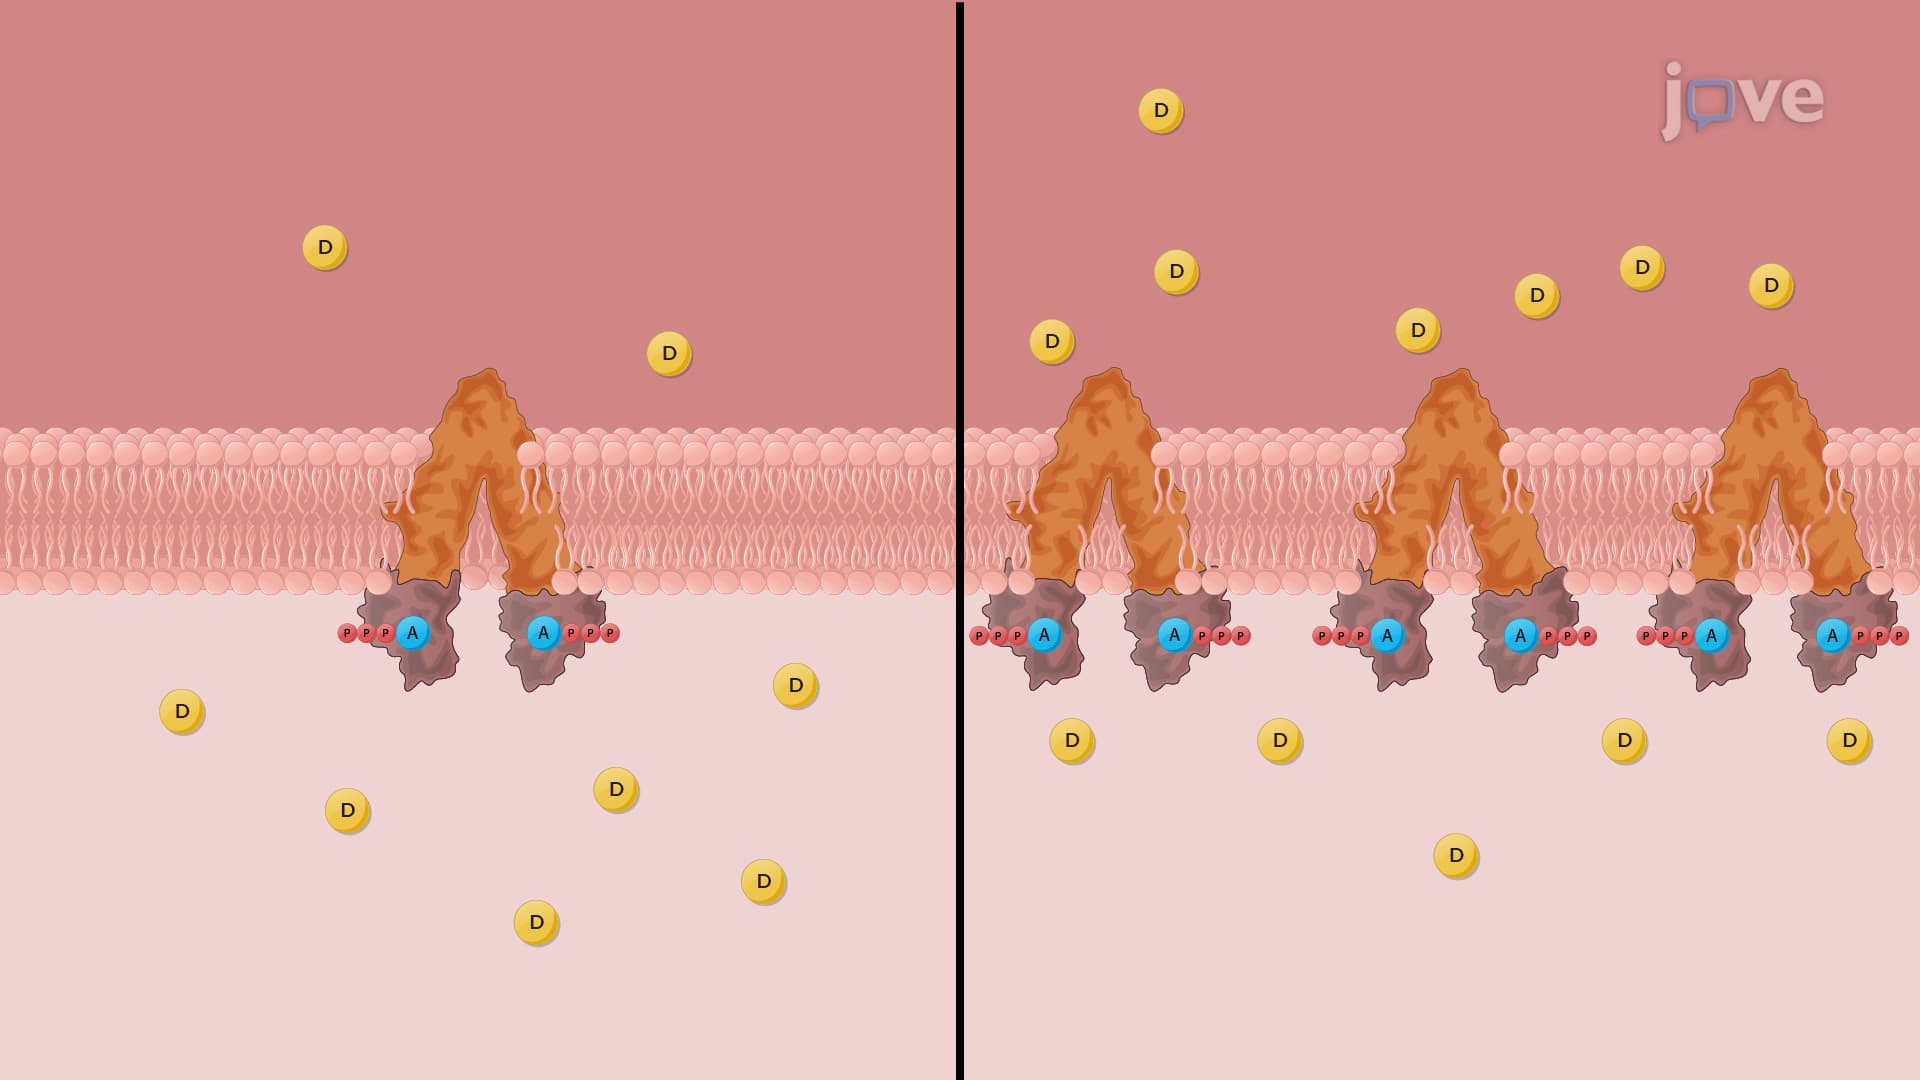
Nonlinear Pharmacokinetics: Role of Transporters
Polymorphisms occurring in drug transporters can alter...
 01:25
Pharmacokinetics: Drug–Drug Interactions
 01:09
Factors Affecting Renal Clearance: Drug Distribution and Drug Interactions
One important factor is the relationship between renal clearance and the apparent volume of distribution. Renal clearance tends to be inversely proportional to the apparent volume of distribution. Drugs with an extensive distribution volume or those...
 01:23
Nonlinear Pharmacokinetics: Dependence of Elimination Half-Life and Dose Clearance
A study on guinea pigs examined the...