Related Experiment Video
Updated: Nov 11, 2025
Pharmacophore Modeling for Targets with Extensive Ligand Libraries: A Case Study on SARS-CoV-2 Mpro
Published on: September 26, 2025
Structural insights on the interaction potential of natural leads against major protein targets of SARS-CoV-2:
Sinosh Skariyachan1, Dharshini Gopal2, Aditi G Muddebihalkar3
1Department of Microbiology, St. Pius X College Rajapuram, Kasaragod, Kerala, India.

Abstract:
Though significant efforts are in progress for developing drugs and vaccines against COVID-19, limited therapeutic agents are available currently. Thus, it is essential to undertake COVID-19 research and to identify therapeutic interventions in which computational modeling and virtual screening of lead molecules provide significant insights. The present study aimed to predict the interaction potential of natural lead molecules against prospective protein targets of SARS-CoV-2 by molecular modeling, docking, and dynamic simulation. Based on the literature survey and database search, fourteen molecular targets were selected and the three targets which lack the native structures were computationally modeled. The drug-likeliness and pharmacokinetic features of ninety-two natural molecules were predicted. Four lead molecules with ideal drug-likeliness and pharmacokinetic properties were selected and docked against fourteen targets, and their binding energies were compared with the binding energy of the interaction between Chloroquine and Hydroxychloroquine to their usual targets. The stabilities of selected docked complexes were confirmed by MD simulation and energy calculations. Four natural molecules demonstrated profound binding to most of the prioritized targets, especially, Hyoscyamine and Tamaridone to spike glycoprotein and Rotiorinol-C and Scutifoliamide-A to replicase polyprotein-1ab main protease of SARS-CoV-2 showed better binding energy, conformational and dynamic stabilities compared to the binding energy of Chloroquine and its usual target glutathione-S-transferase. The aforementioned lead molecules can be used to develop novel therapeutic agents towards the protein targets of SARS-CoV-2, and the study provides significant insight for structure-based drug development against COVID-19.
More Related Videos
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
 10:21
Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
Related Concept Videos
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 02:04
Protein-protein Interfaces
 02:50
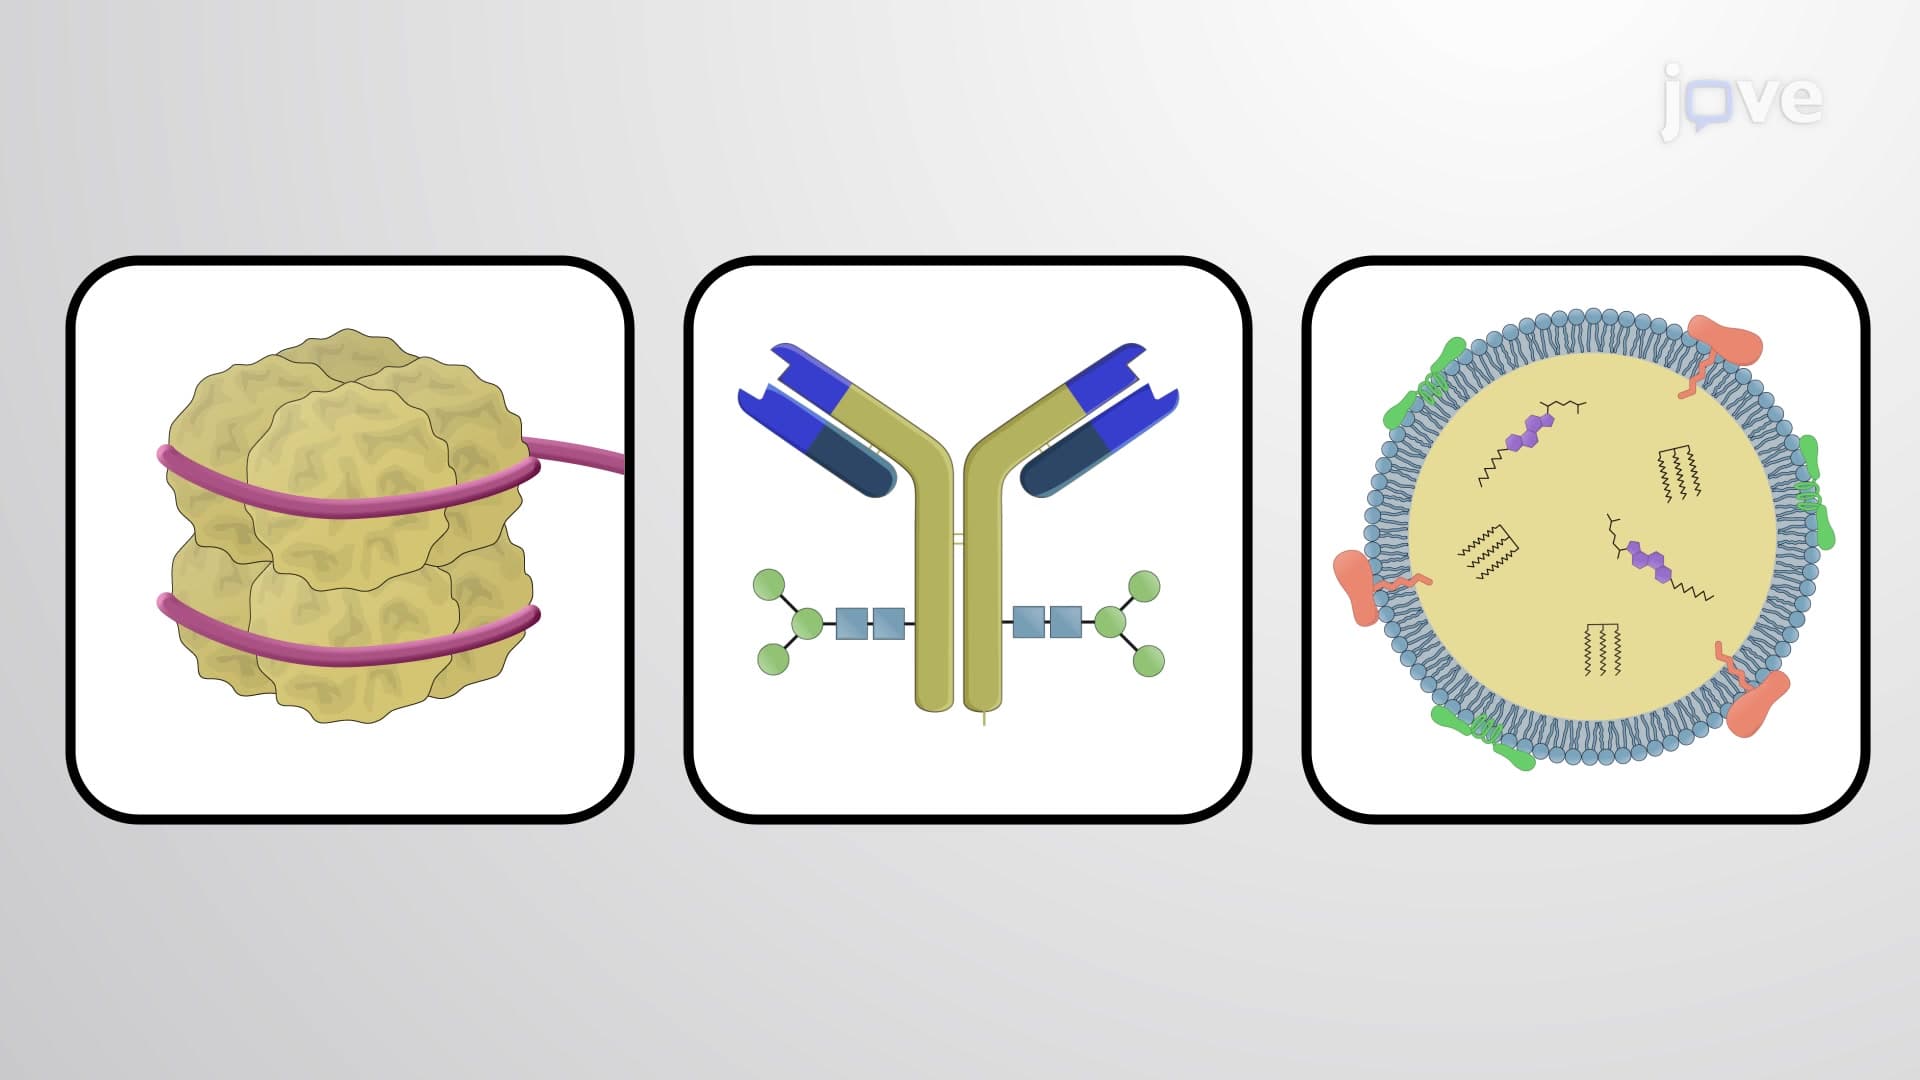
Conjugated Proteins
Nucleoproteins are protein complexes that contain nucleic acids, categorized as deoxyribonucleoproteins (DNPs) or ribonucleoproteins (RNPs) respectively. The nucleosome is a typical example of a DNP where nuclear DNA is associated with histone proteins. The major antigen for the Covid-19 virus SARS-CoV is an RNP that is critical...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 01:16
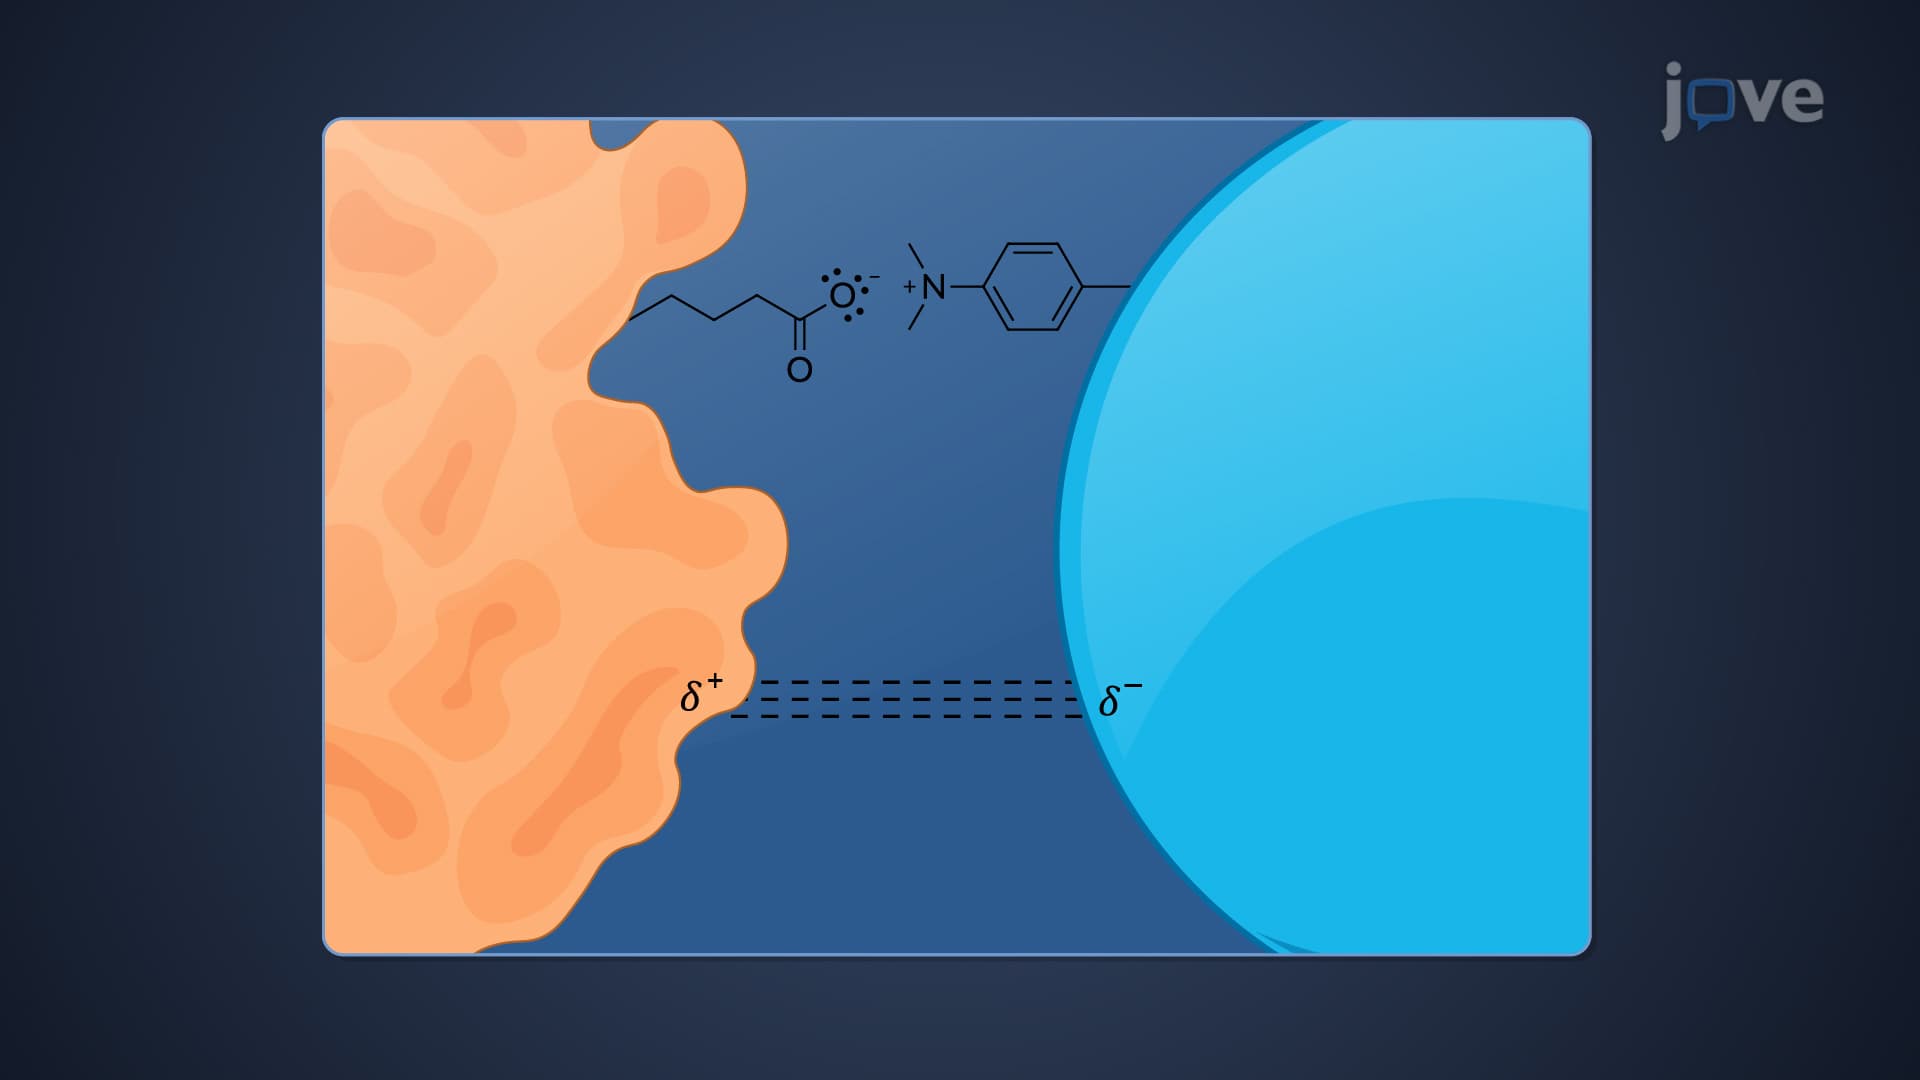
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:24
Protein Organization
The primary structure of a protein is its amino acid sequence....