Related Experiment Video
Updated: Nov 11, 2025


Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Electronic structure of ternary palladates and effect of hole doping: a valence band photoemission spectroscopic
B H Reddy1, Asif Ali1, Ravi Shankar Singh1
1Department of Physics, Indian Institute of Science Education and Research Bhopal, Bhopal Bypass Road, Bhauri, Bhopal-462 066, India.

Abstract:
We investigate the electronic structure of ternary palladatesAPd3O4(A= Sr, Ca) using valence band photoemission spectroscopy and band structure calculations. Energy positions of various features and overall width of the experimental valence band spectra are well captured by band structure calculations using hybrid functional. Band structure calculations within local density approximations lead to metallic ground state while the calculations using hybrid functional provide band gap of 0.25 eV and 0.22 eV for CaPd3O4and SrPd3O4respectively, suggesting moderate to strong electron correlation strength in these narrow band gap semiconducting palladates. High resolution spectra reveal negligibly small intensity at Fermi level,EF, for parent compounds, while hole doped SrPd3O4(by 15% Li substitution at Sr site) exhibits a Fermi cut-off suggesting metallic character in contrast to semiconducting transport. These observations reveal the importance of localization of electrons in case where the Fermi edge falls in the mobility edge.
More Related Videos
Related Concept Videos
 02:42
Valence Bond Theory
 03:26
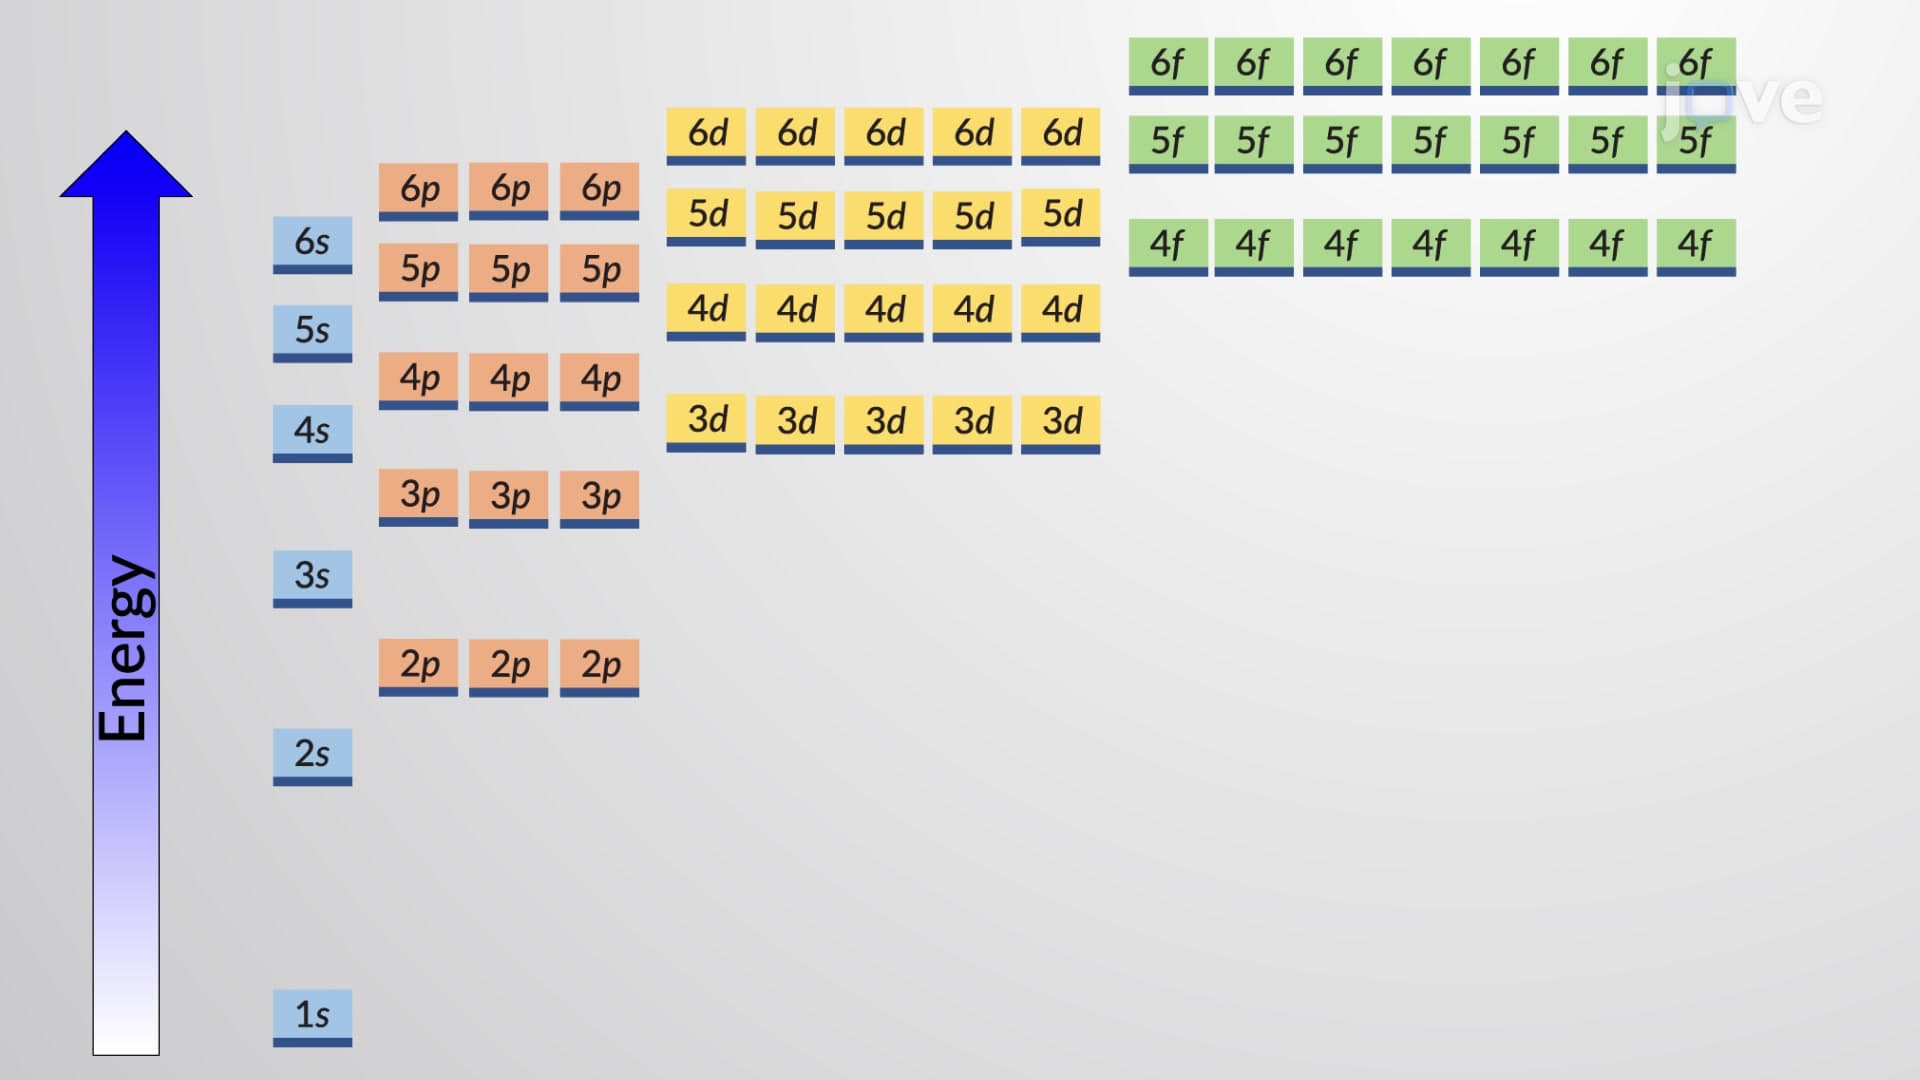
Electron Configuration of Multielectron Atoms
 01:12
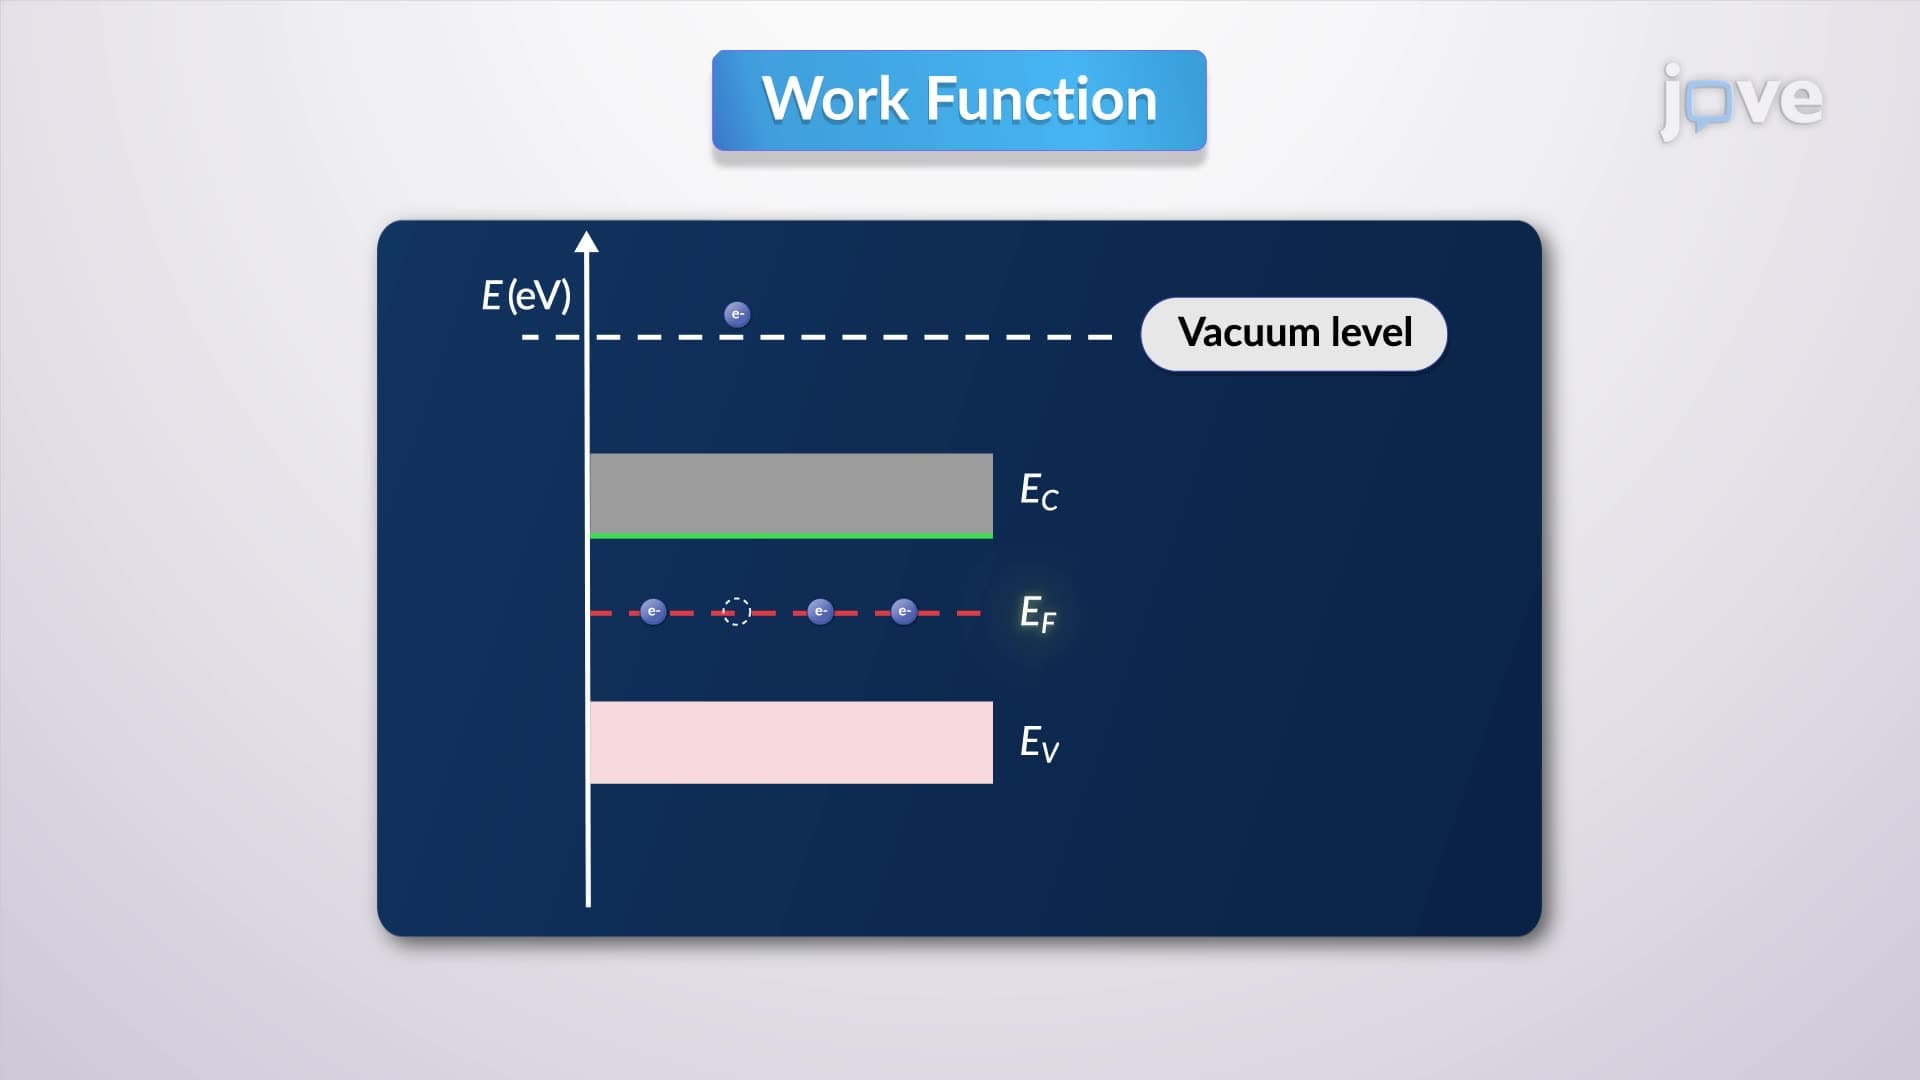
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
 02:46
Electron Configurations
The relative energies of the subshells determine the order in which atomic orbitals are filled (1s, 2s, 2p, 3s, 3p,...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...