
Non long-range corrected density functionals incorrectly describe the intensity of the CH stretching band in
Hugo Geindre1, Abdul-Rahman Allouche2, Daniel Peláez3
1Université Lille, UMR 8523 - Physique des Lasers, Atomes et Molécules, Lille, France.

Abstract:
We present a comprehensive study of the most relevant numerical aspects influencing frequencies and intensities in the infrared spectrum of isolated polycyclic aromatic hydrocarbons (PAHs) regarding the overestimate of the IR CH-stretching bands. We use naphthalene as benchmark and show the validity of our results to different members of the PAH family. Our analysis relies on widely employed density functional theory methods and second-order vibrational perturbational theory for the computation of vibrational eigenstates. We have focused on the elucidation of the origin of the systematic overestimate of the intensities in the CH-stretching region. To rule out nonfundamental numerical errors, we have initially considered the influence of the electronic basis set and various other parameters on the different stages of the vibrational analysis. In a second stage, we have benchmarked the results of different density functional theory functionals with respect to the aforementioned overestimate taken as the ratio between the most prominent features of the spectrum, the CH-bending and the CH-stretching bands. Our results unambiguously indicate that the long-range correction plays a major role in this spurious numerical issue. More specifically, this phenomenon is due to an incorrect description of the charge distribution (and hence dipole) within the symmetrically relevant CH bonds. Long-range correction specifically remedies this issue. It improves the description of the intensities in the stretching region while at the same time it does not perturb significantly the rest of the spectrum. With respect to the frequencies, we have observed an overall improvement when compared to noncorrected functionals.
More Related Videos
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:24
IR Frequency Region: X–H Stretching
 01:21
IR Absorption Frequency: Hybridization
Among the sp, sp2, and sp3 hybridized orbitals, sp orbitals have the maximum s character (50%). Consequently, the electrons are held more closely to the nucleus, resulting in stronger and shorter C–H bonds that...
 01:22
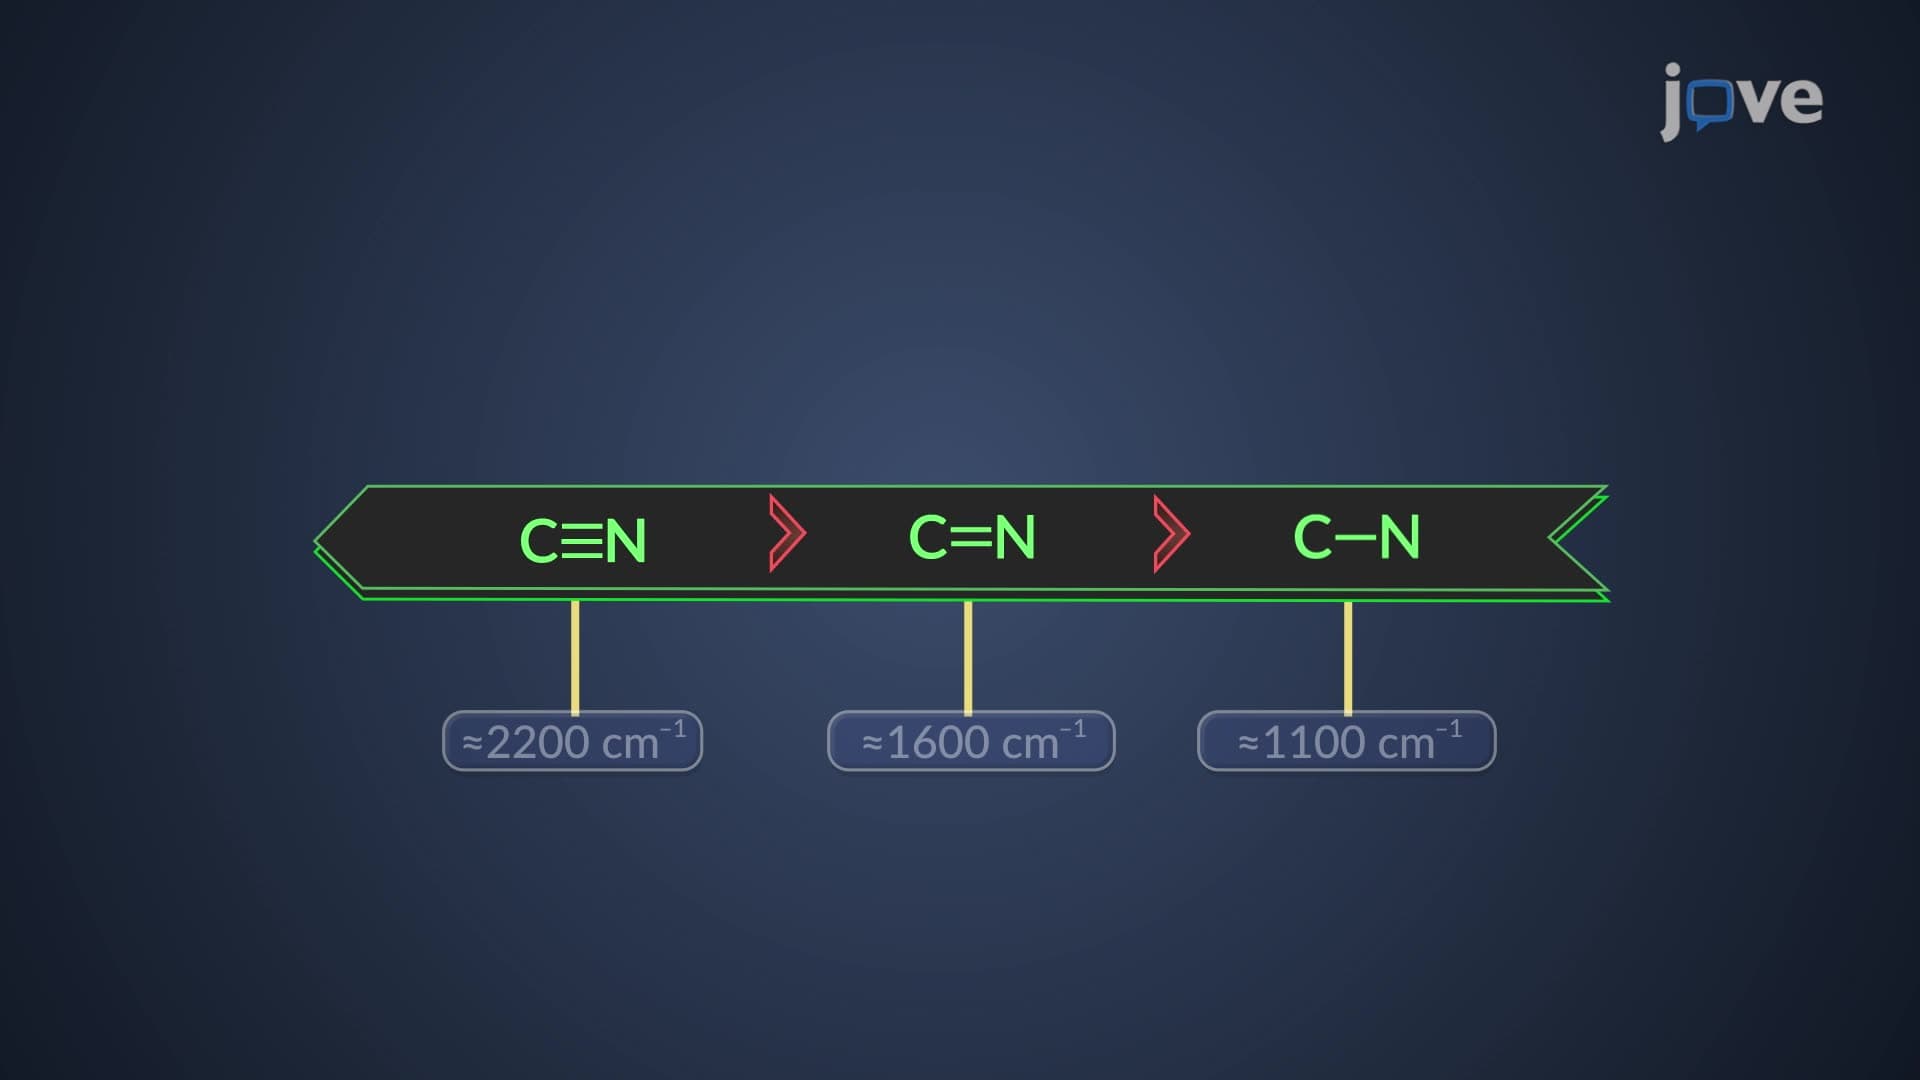
IR Frequency Region: Alkyne and Nitrile Stretching
Comparing the stretching vibrational frequency of C≡C triple bonds with that of double and single bonds, it is evident that C≡C triple bonds exhibit a higher stretching frequency than C=C double and C–C single bonds. Similarly, the C≡N triple bond...
 01:29
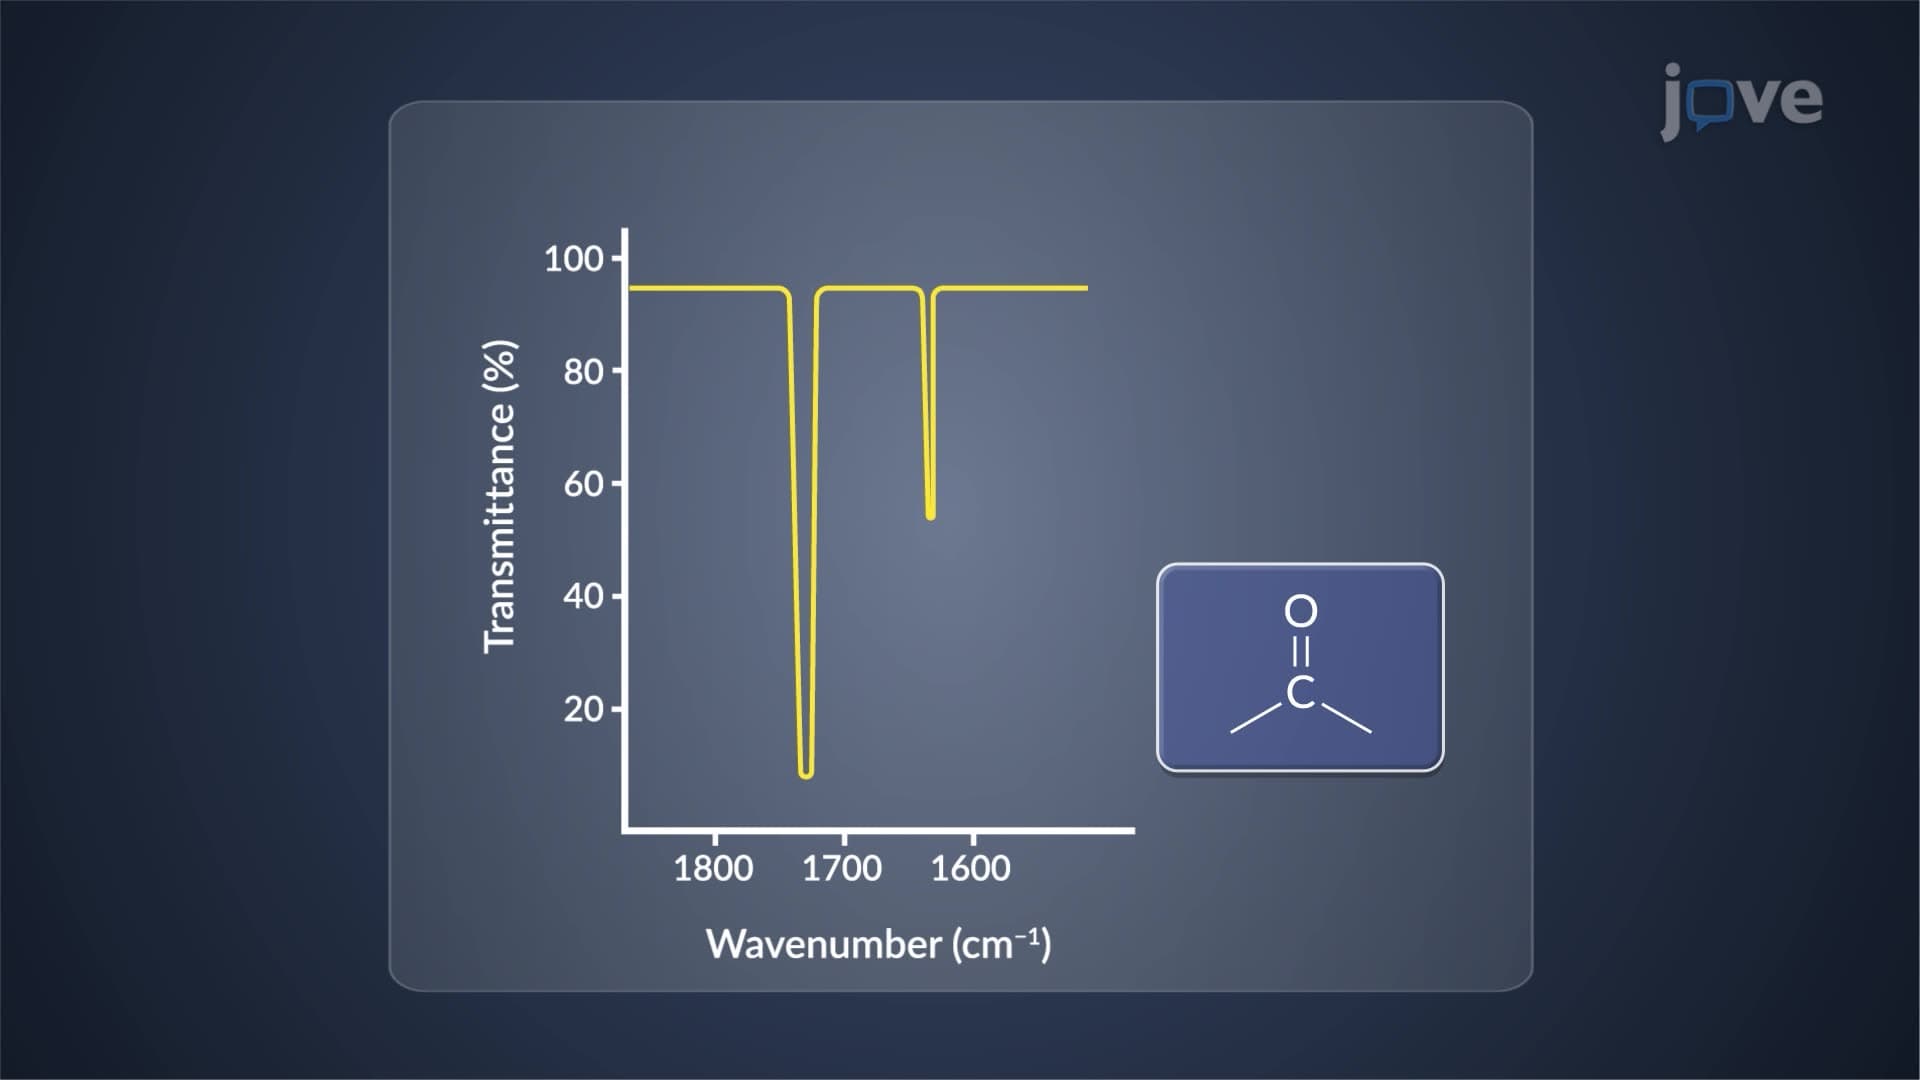
IR Frequency Region: Alkene and Carbonyl Stretching
Stretching frequencies are affected by several factors, such as resonance, inductive effects, ring strain, dipole moment, and hydrogen bonding. Consequently, the stretching frequency of the carbonyl double bond...
 01:23
IR Spectrum Peak Broadening: Hydrogen Bonding
However, the extent of hydrogen bonding influences the observed stretching frequency and band broadening. Intermolecular or intramolecular...
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...