Related Experiment Video
Updated: Nov 9, 2025

High-Pressure NMR Experiments for Detecting Protein Low-Lying Conformational States
Published on: June 29, 2021
Conformational Shifts of Stacked Heteroaromatics: Vacuum vs. Water Studied by Machine Learning
Johannes R Loeffler1, Monica L Fernández-Quintero1, Franz Waibl1
1Center of Molecular Biosciences Innsbruck, Institute of General, Inorganic and Theoretical Chemistry, University of Innsbruck, Innsbruck, Austria.

Abstract:
Stacking interactions play a crucial role in drug design, as we can find aromatic cores or scaffolds in almost any available small molecule drug. To predict optimal binding geometries and enhance stacking interactions, usually high-level quantum mechanical calculations are performed. These calculations have two major drawbacks: they are very time consuming, and solvation can only be considered using implicit solvation. Therefore, most calculations are performed in vacuum. However, recent studies have revealed a direct correlation between the desolvation penalty, vacuum stacking interactions and binding affinity, making predictions even more difficult. To overcome the drawbacks of quantum mechanical calculations, in this study we use neural networks to perform fast geometry optimizations and molecular dynamics simulations of heteroaromatics stacked with toluene in vacuum and in explicit solvation. We show that the resulting energies in vacuum are in good agreement with high-level quantum mechanical calculations. Furthermore, we show that using explicit solvation substantially influences the favored orientations of heteroaromatic rings thereby emphasizing the necessity to include solvation properties starting from the earliest phases of drug design.
More Related Videos
 14:55
Atomic Scale Structural Studies of Macromolecular Assemblies by Solid-state Nuclear Magnetic Resonance Spectroscopy
Published on: September 17, 2017
 08:49
Author Spotlight: Unveiling the Potential of VSFG Microscopy in Studying Mesoscopically Heterogeneous Self-Assembled Structures
Published on: December 1, 2023
Related Concept Videos
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
 02:29
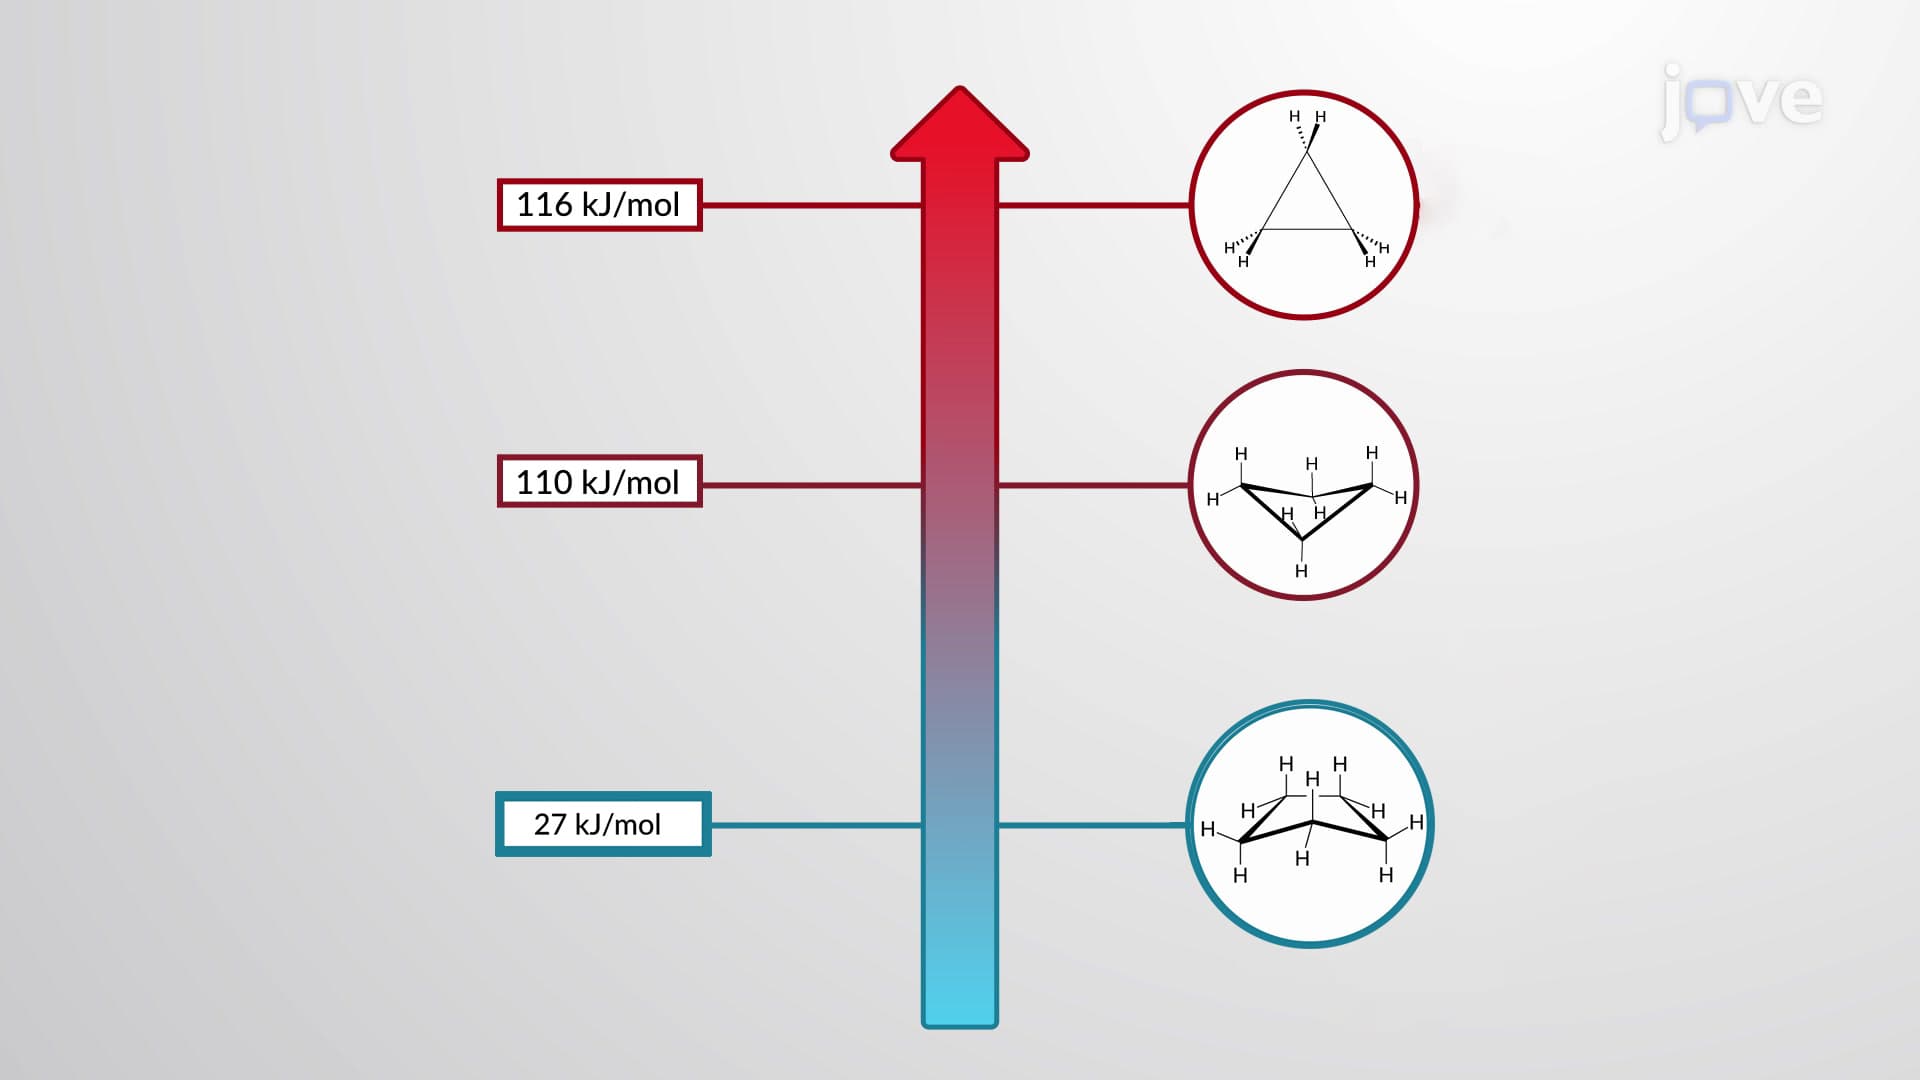
Conformations of Cycloalkanes
 01:18
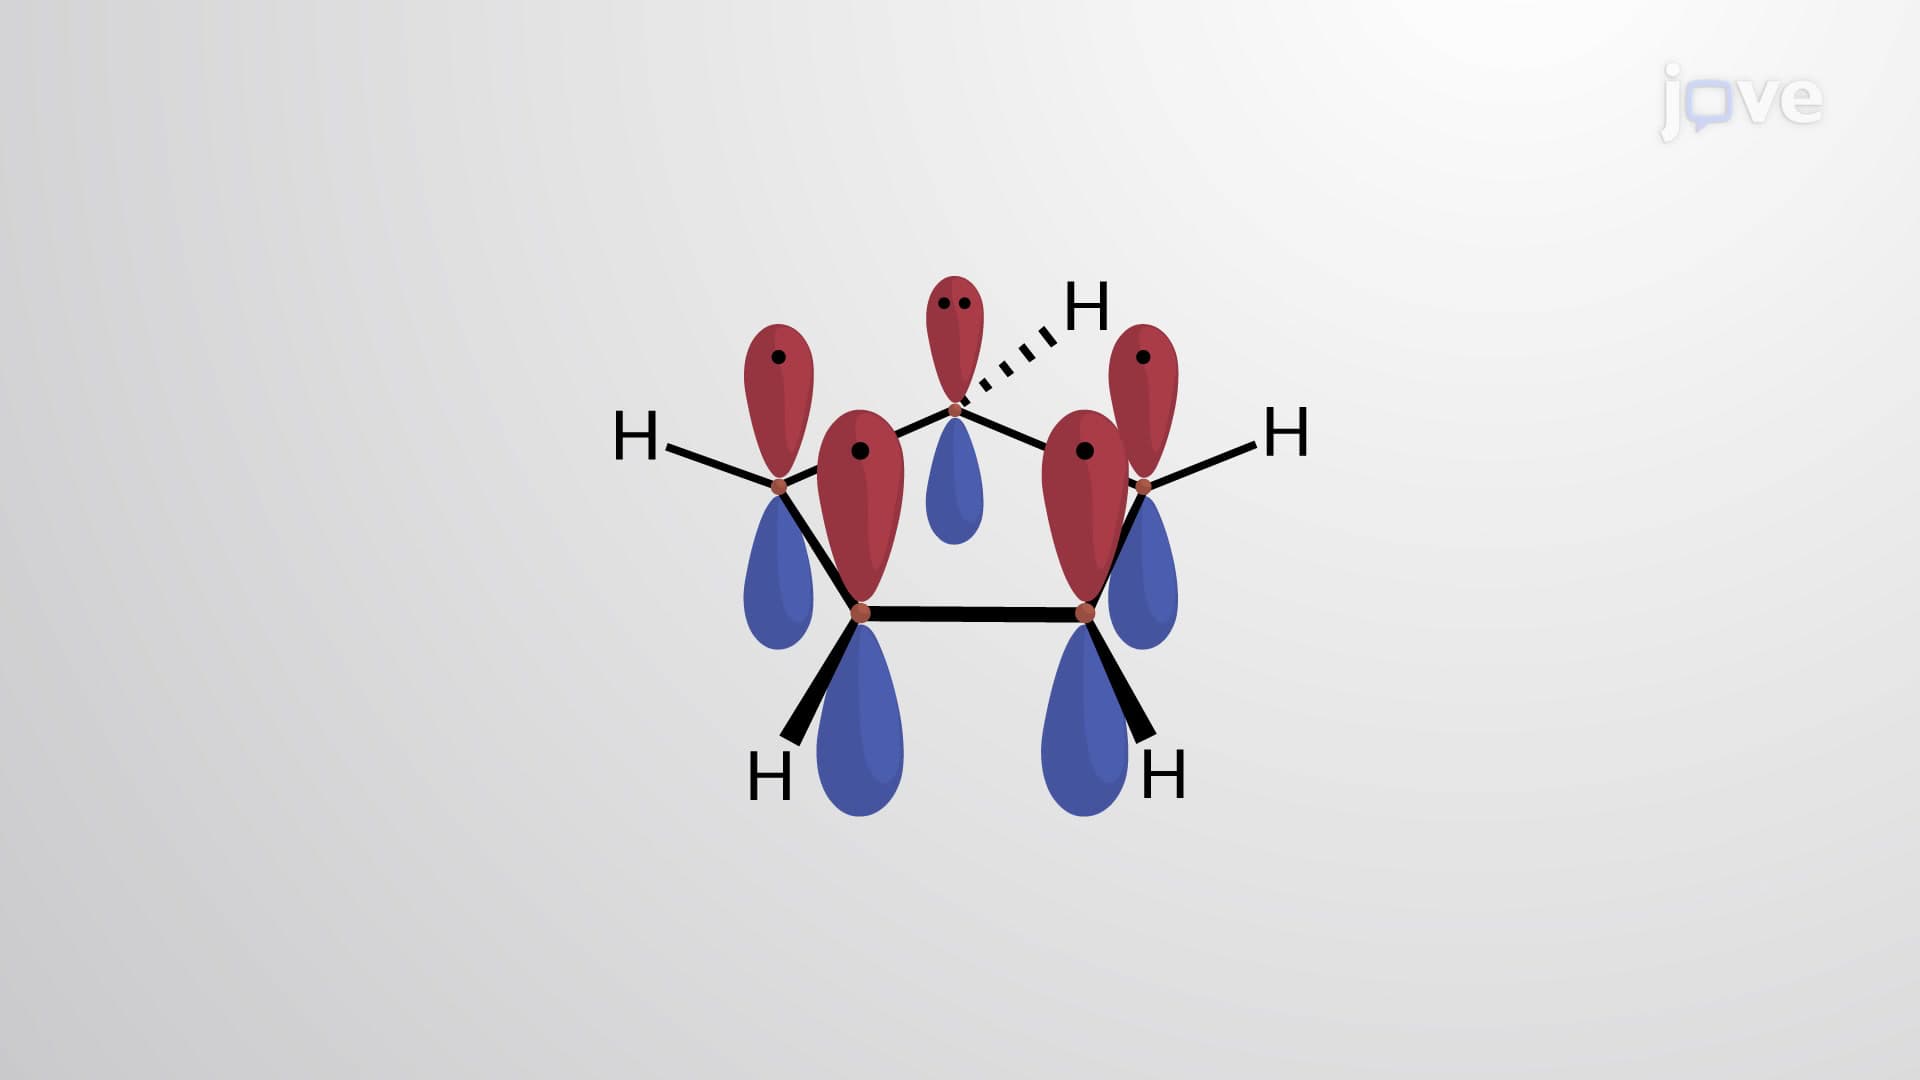
Aromatic Hydrocarbon Anions: Structural Overview
Due to the absence of continuous...
 01:14
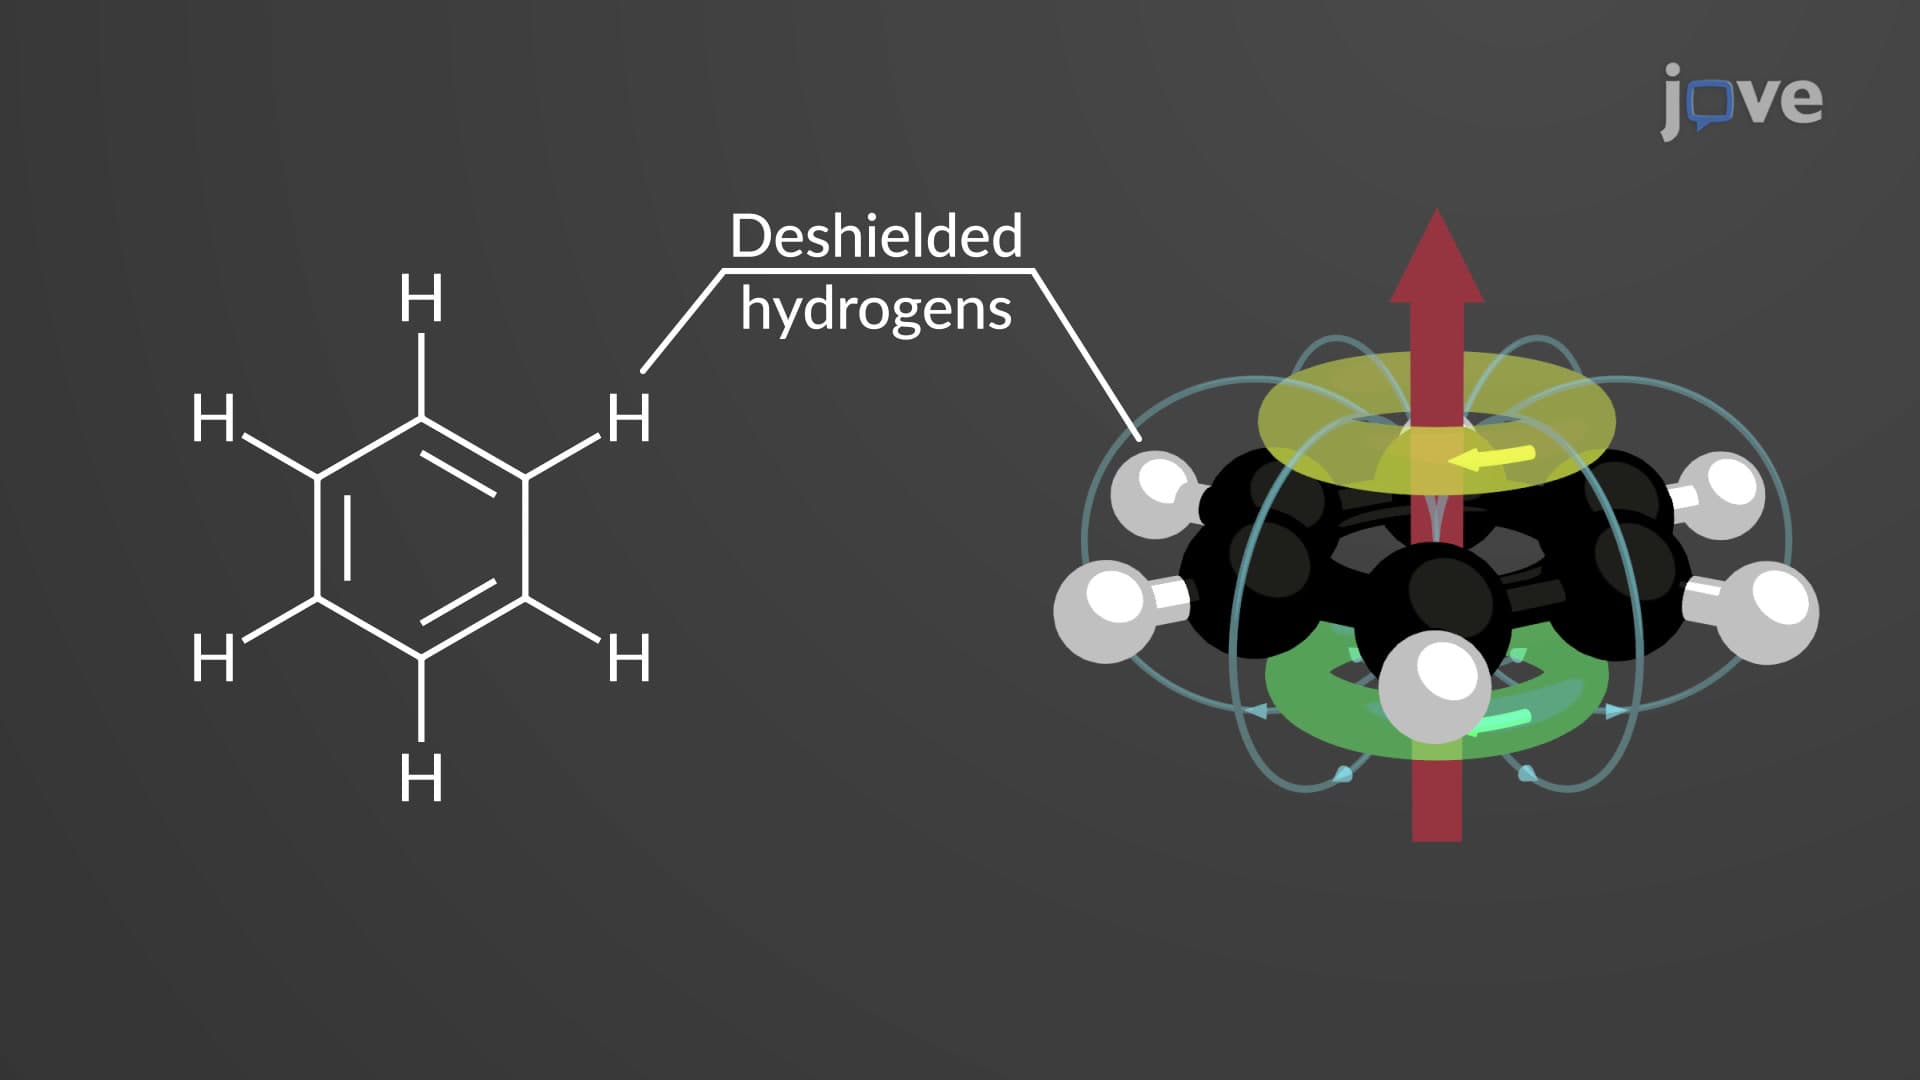
NMR Spectroscopy of Aromatic Compounds
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal...