Related Experiment Video
Updated: Nov 8, 2025

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
A test of ab initio-generated, radial intermolecular potential energy functions for five axially-symmetric,
1School of Chemistry, University of Bristol, Cantock's Close, Bristol, BS8 1TS, UK. a.c.legon@bristol.ac.uk.

Abstract:
The radial potential energy functions for five axially symmetric, hydrogen-bonded complexes BHF (B = N2, CO, PH3, HCN and NH3) have been calculated ab initio at the explicitly correlated level of theory CCSD(T)(F12c)/cc-pVTZ-F12 as a function of the hydrogen-bond distance r(ZH), where Z is the hydrogen-bond acceptor atom of B. The remaining geometric parameters were optimised at each point in the calculation. The functions so generated were then used to estimate the spectroscopic constants ωσ, ωσxσ,ασ, and the vσ = 1 ← 0 transition wavenumber associated with the intermolecular stretching mode νσ of each BHF by using two equivalent approaches. Both involved the assumption that the vibrational modes of the B and HF molecules were sufficiently stiff relative to the intermolecular stretching mode that BHF could be treated in a pseudo-diatomic approximation. One approach used derivatives of the potential evaluated at the distance r = re while the other used the potential constants obtained by non-linear regression fits of three analytical functions (Morse, Rydberg and Hulburt-Hirschfelder) to the ab initio calculated points. The two approaches would give exactly the same results if the functions were a perfect fit. The H-H function gave the best fit. The determined spectroscopic constants were found to be in reasonable agreement with the limited number available by experiment. The efficacy of the approach was tested for the diatomic molecule H35Cl by taking advantage of both an accurate RKR-type potential and an accurate set of spectroscopic constants. It was also established that the relationship De = kσ/(2a2) between two measures (De and kσ, both calculated ab initio) of the strength of the hydrogen bond in the BHF complexes (and required if the H-H function were an accurate representation of the BHF potential functions) holds to an excellent level of approximation, and supports the conclusion that this function is appropriate to represent the hydrogen bond in the complexes investigated.
More Related Videos
 06:37
Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 03:51
Molecular Orbital Theory II
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 03:24
Hybridization of Atomic Orbitals I
 01:18
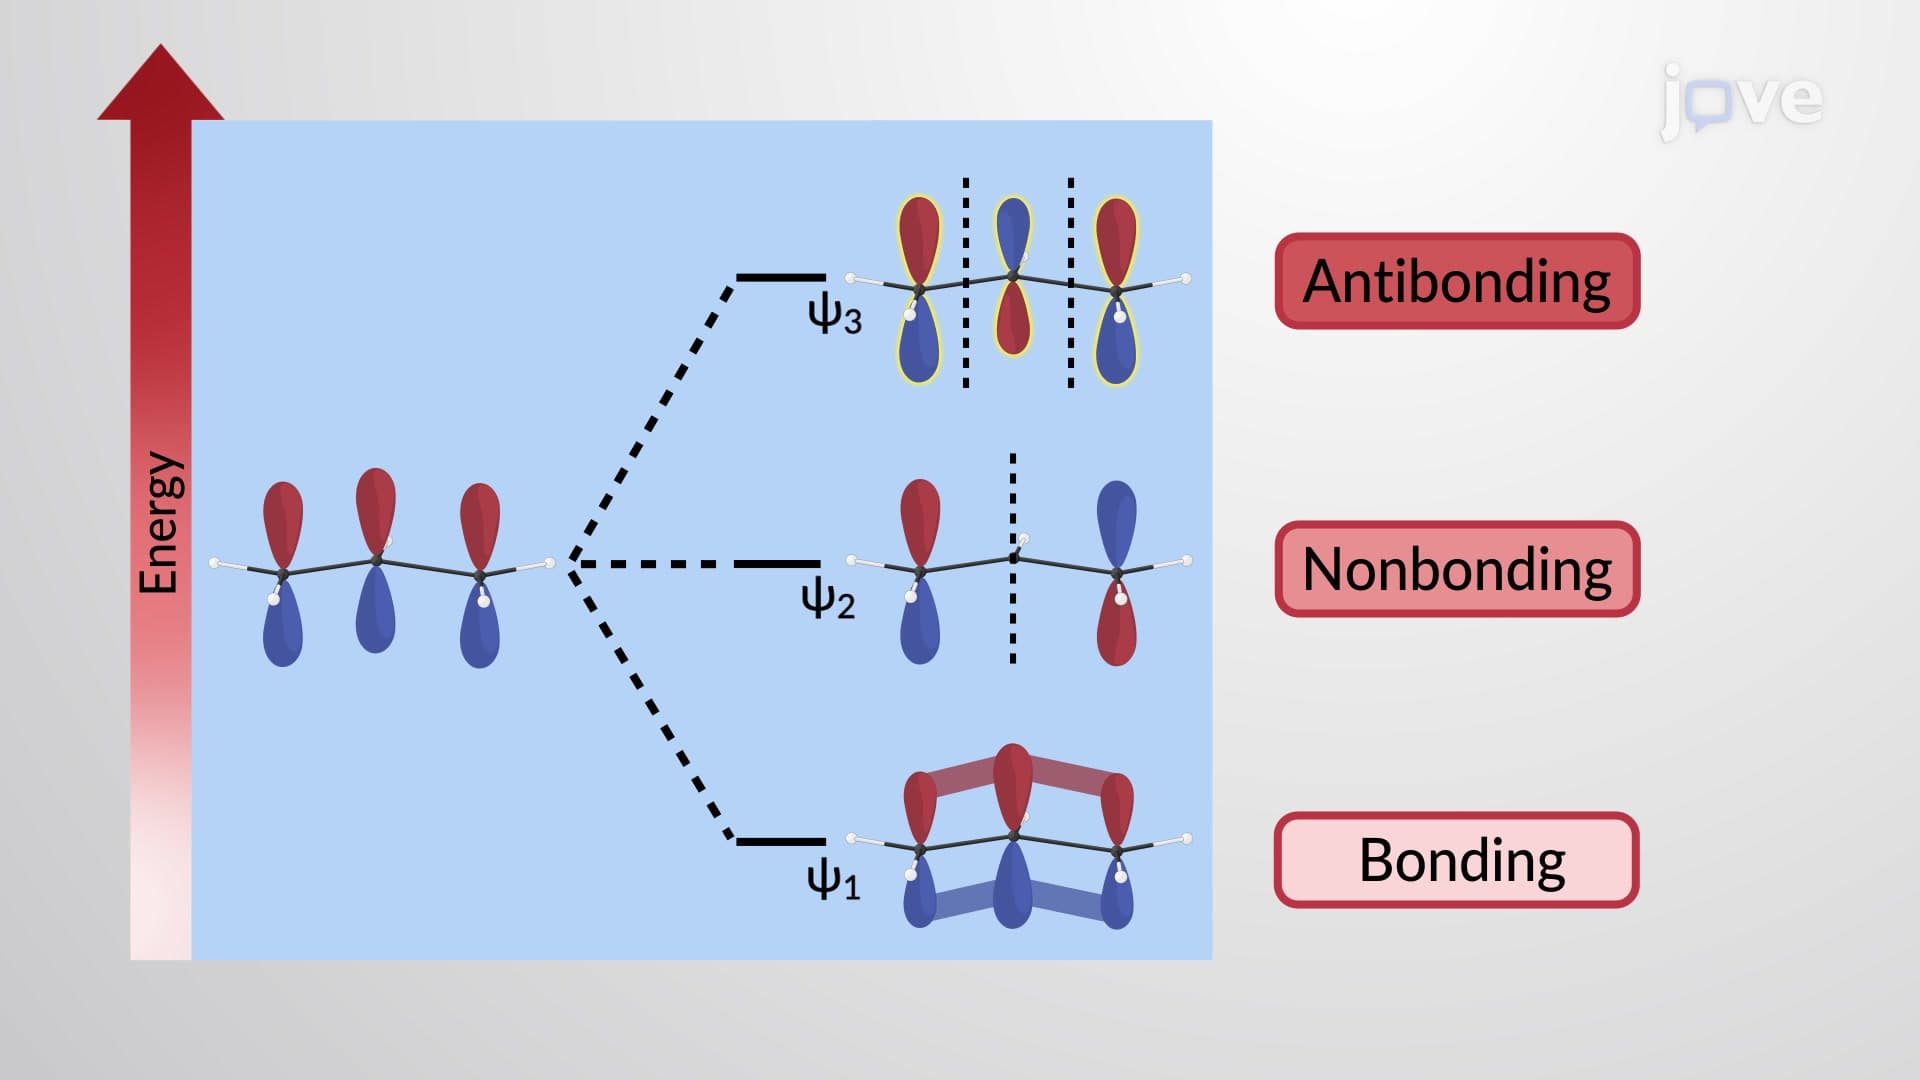
π Molecular Orbitals of the Allyl Cation and Anion
 01:04
Hydrogen Bonds
 00:26
Hydrogen Bonds
Hydrogen Bonds Control the World!
Because hydrogen has very weak electronegativity when it binds with a strongly electronegative atom, such as oxygen or nitrogen, electrons in the bond are unequally shared....