Related Experiment Video
Updated: Nov 7, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Improved predictions of thermomechanical properties of molecular crystals from energy and dispersion corrected DFT
1Department of Civil, Materials and Environmental Engineering, University of Illinois at Chicago, Chicago, Illinois 60607, USA.

Abstract:
Thermal stability and pressure-dependent changes are key to molecular crystals and their properties. The determination of their thermal properties from ab initio methods is, however, a challenging task. While the low-frequency phonon spectrum related to intermolecular vibrations remains difficult to describe, the Quasi-Harmonic Approximation (QHA) also induces for molecular crystals a significant volume deviation, which makes their thermal behavior ill-determined. To overcome these difficulties, we consider a pragmatic energy correction (EC) that has long been used for atomic crystals, and we presently report the first ever use for molecular crystals. Applying the QHA in dispersion-corrected density functional theory (DFT-D) calculations with an ab initio parameterized EC, the resulting model can simultaneously and accurately derive thermal and mechanical properties of high-explosive molecular crystals. When compared to experiments, the mean absolute percent error of previous DFT-based thermomechanical models is 12% for mechanical and 31% for thermal properties. Our model performs significantly better and reduces these uncertainties to 4.1% and 9.8%, respectively. In particular, the agreement between our model and experiments for the thermal properties is three times better. This significant improvement greatly benefits the determination of thermomechanical properties such as the Grüneisen parameter and the shock properties. The method has been successfully applied to molecular crystals showing a large diversity of weak intermolecular interactions (β-1,3,5,7-tetranitro-1,3,5,7-tetrazoctane (HMX), α-1,1-diamino-2,2-dinitroethylene (FOX-7), Triaminotrinitrobenzene (TATB), ε-Hexanitrohexaazaisowurtzitane (CL20), and Pentaerythritol tetranitrate (PETN)-I). Due to its accuracy and transferability, our model is expected to work for a large class of computationally designed molecular crystals and co-crystals, providing a basis for a predictive framework.
More Related Videos
 06:37
Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:27
Predicting Molecular Geometry
 01:16
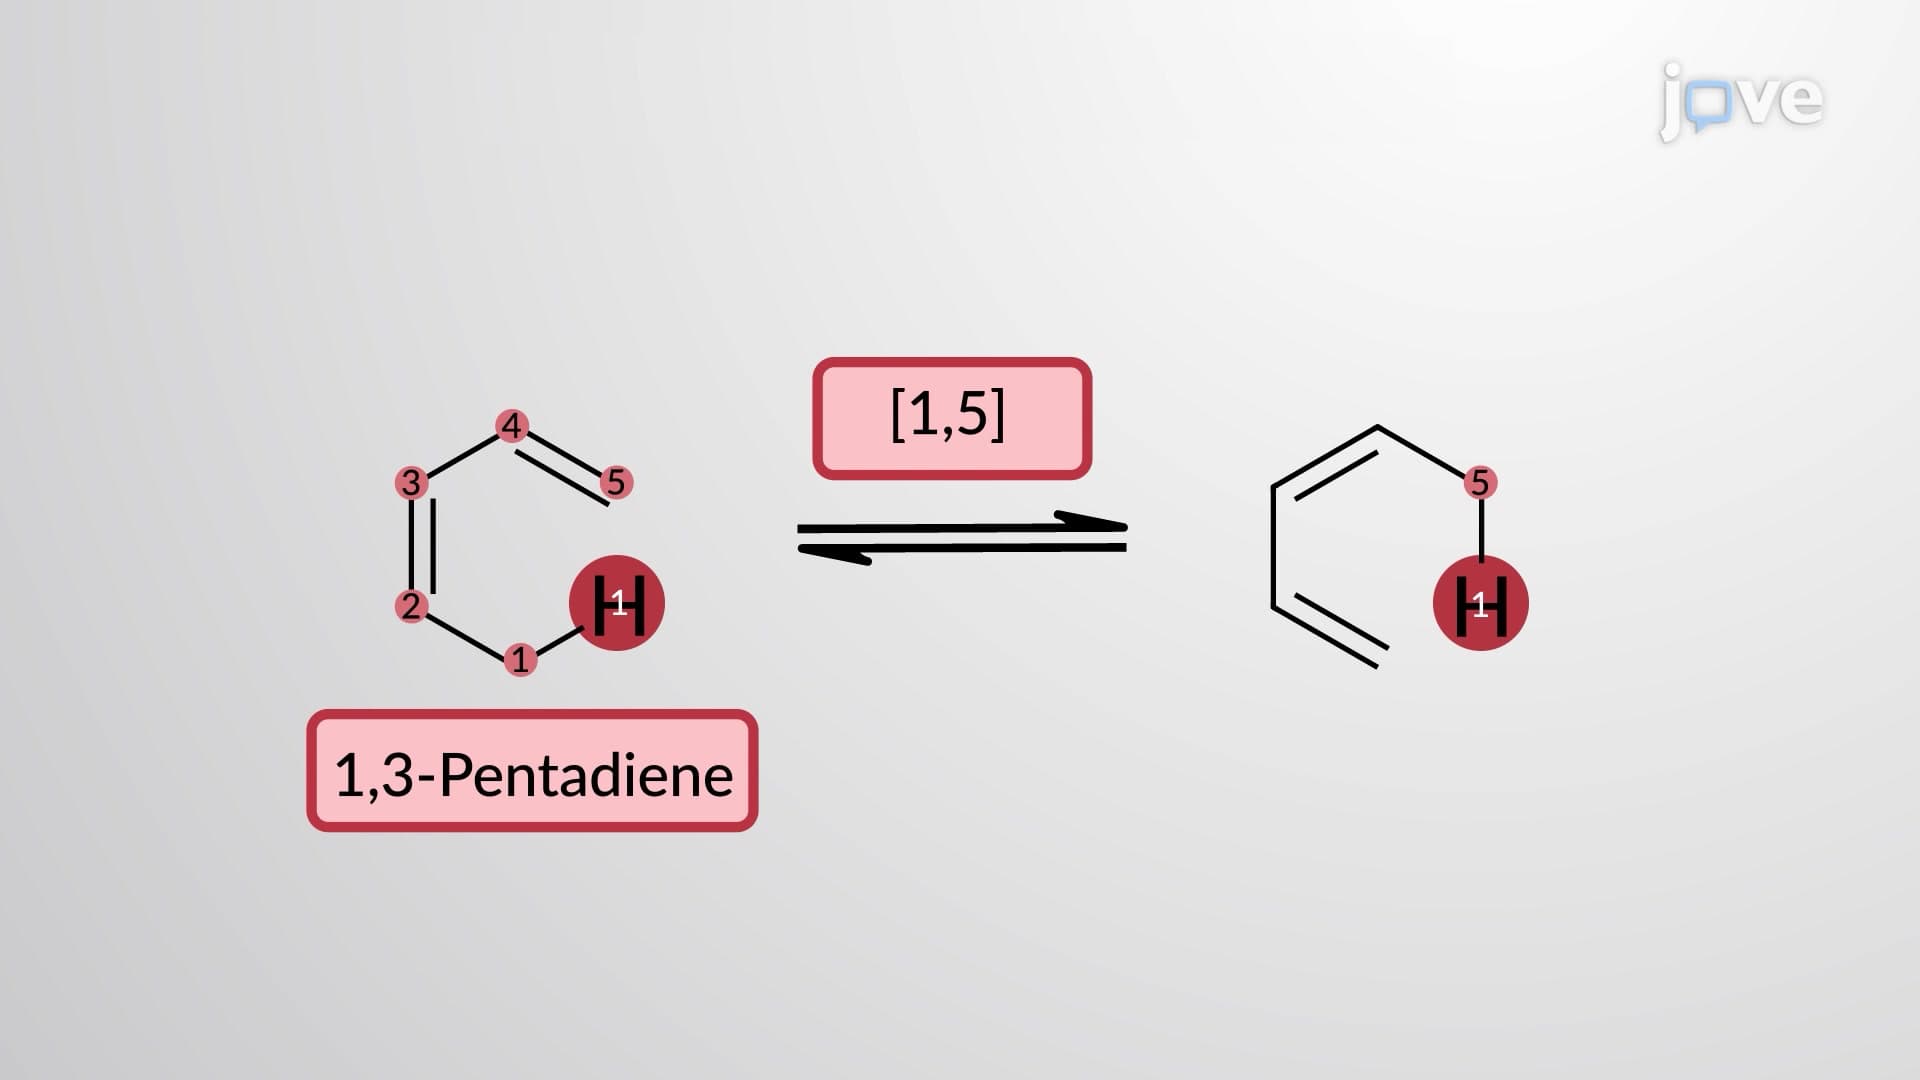
Thermal Sigmatropic Reactions: Overview
Sigmatropic shifts are classified based on an order term [i, j ], where i and j indicate the number of atoms across which each end of the σ bond migrates. Below are examples of a [3,3] sigmatropic shift in 1,5-hexadiene, referred...
 02:18
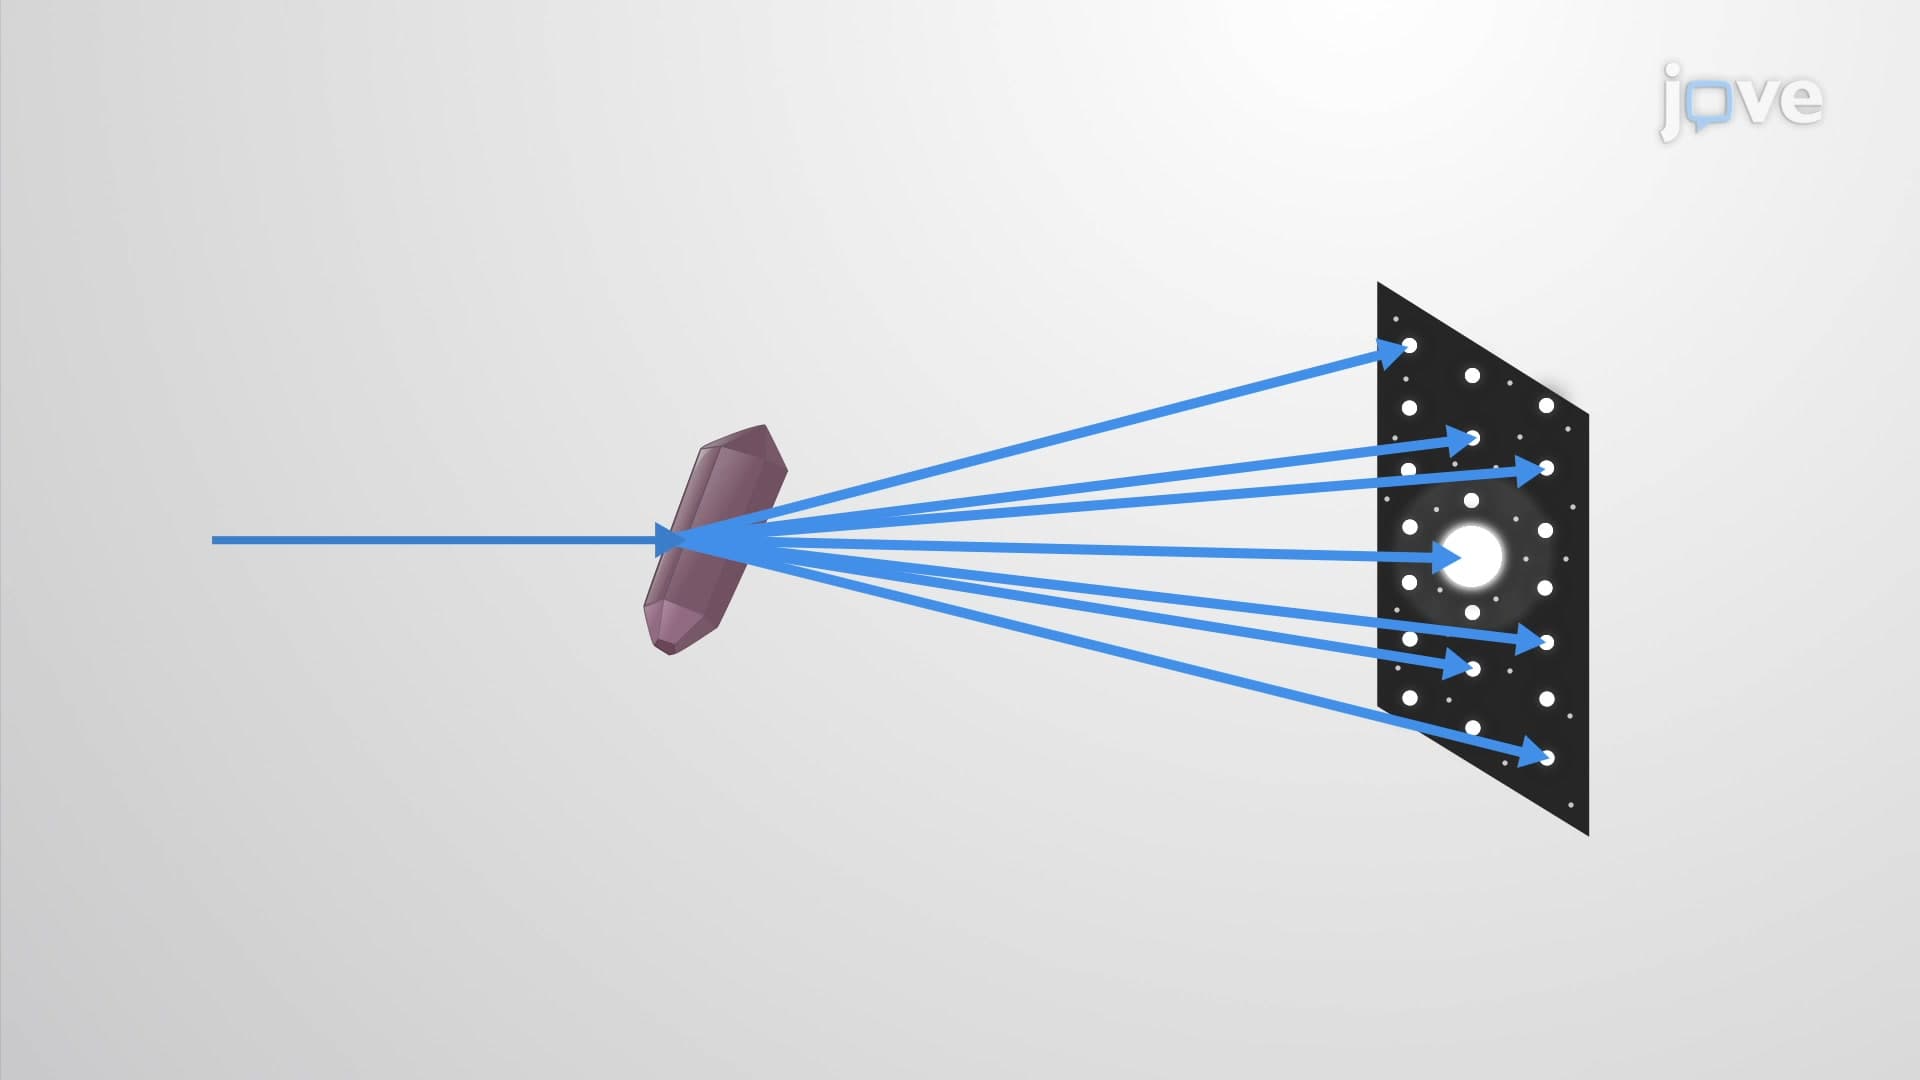
X-ray Crystallography
Diffraction
Diffraction is the change in the direction of travel experienced by an electromagnetic wave when it encounters a physical barrier whose dimensions are comparable to those of the wavelength of the light. X-rays are electromagnetic radiation with wavelengths about as long as the distance between neighboring...