Related Experiment Video
Updated: Nov 7, 2025

Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
The atomic structure and dynamics at the CaCO3 vaterite-water interface: A classical molecular dynamics study
Alicia Schuitemaker1, Paolo Raiteri1, Raffaella Demichelis1
1Curtin Institute for Computation, The Institute for Geoscience Research (TIGeR), School of Molecular and Life Sciences, Curtin University, GPO Box U1987, 6845 Perth, Western Australia, Australia.

Abstract:
Classical molecular and lattice dynamics were applied to explore the structure and dynamics of water on different surfaces of vaterite, the least abundant calcium carbonate polymorph. Surfaces were generated starting from the three possible structural models for vaterite (monoclinic, hexagonal/trigonal, and triclinic) and pre-screened using their surface energies in an implicit solvent. Surfaces with energies lower than 0.55 J/m2 were then run in explicit water. The majority of these surfaces dissolve in less than 100 ns, highlighting the low stability of this phase in abiotic environments. Three stable surfaces were identified; they exhibited only minor structural changes when in contact with explicit water and did not show any tendency to dissolve during 1 µs molecular dynamics simulations. The computed water density profiles show that all these surfaces have two distinct hydration layers. The water residence time at the various calcium sites was computed to be within 0.7 and 20.5 ns, which suggests that specific Ca ions will be more readily available to bind with organic molecules present in solution. This analysis is a step forward in understanding the structure of this complex mineral and its role in biomineralization, as it provides a solid theoretical background to explore its surface chemistry. In particular, this study provides realistic surface models and predicts the effect of water exchange at the surface active sites on the adsorption of other molecules.
Related Concept Videos
 01:09
Theories of Dissolution: The Danckwerts' Model and Interfacial Barrier Model
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:24
Van der Waals Interactions
 02:54
Molecular and Ionic Solids
Molecular Solids
Molecular crystalline solids, such as ice, sucrose (table sugar), and iodine, are solids that are composed of neutral molecules as their constituent units. These molecules are held together by weak intermolecular forces such as London dispersion forces, dipole-dipole interactions, or hydrogen bonds, which...
 01:10
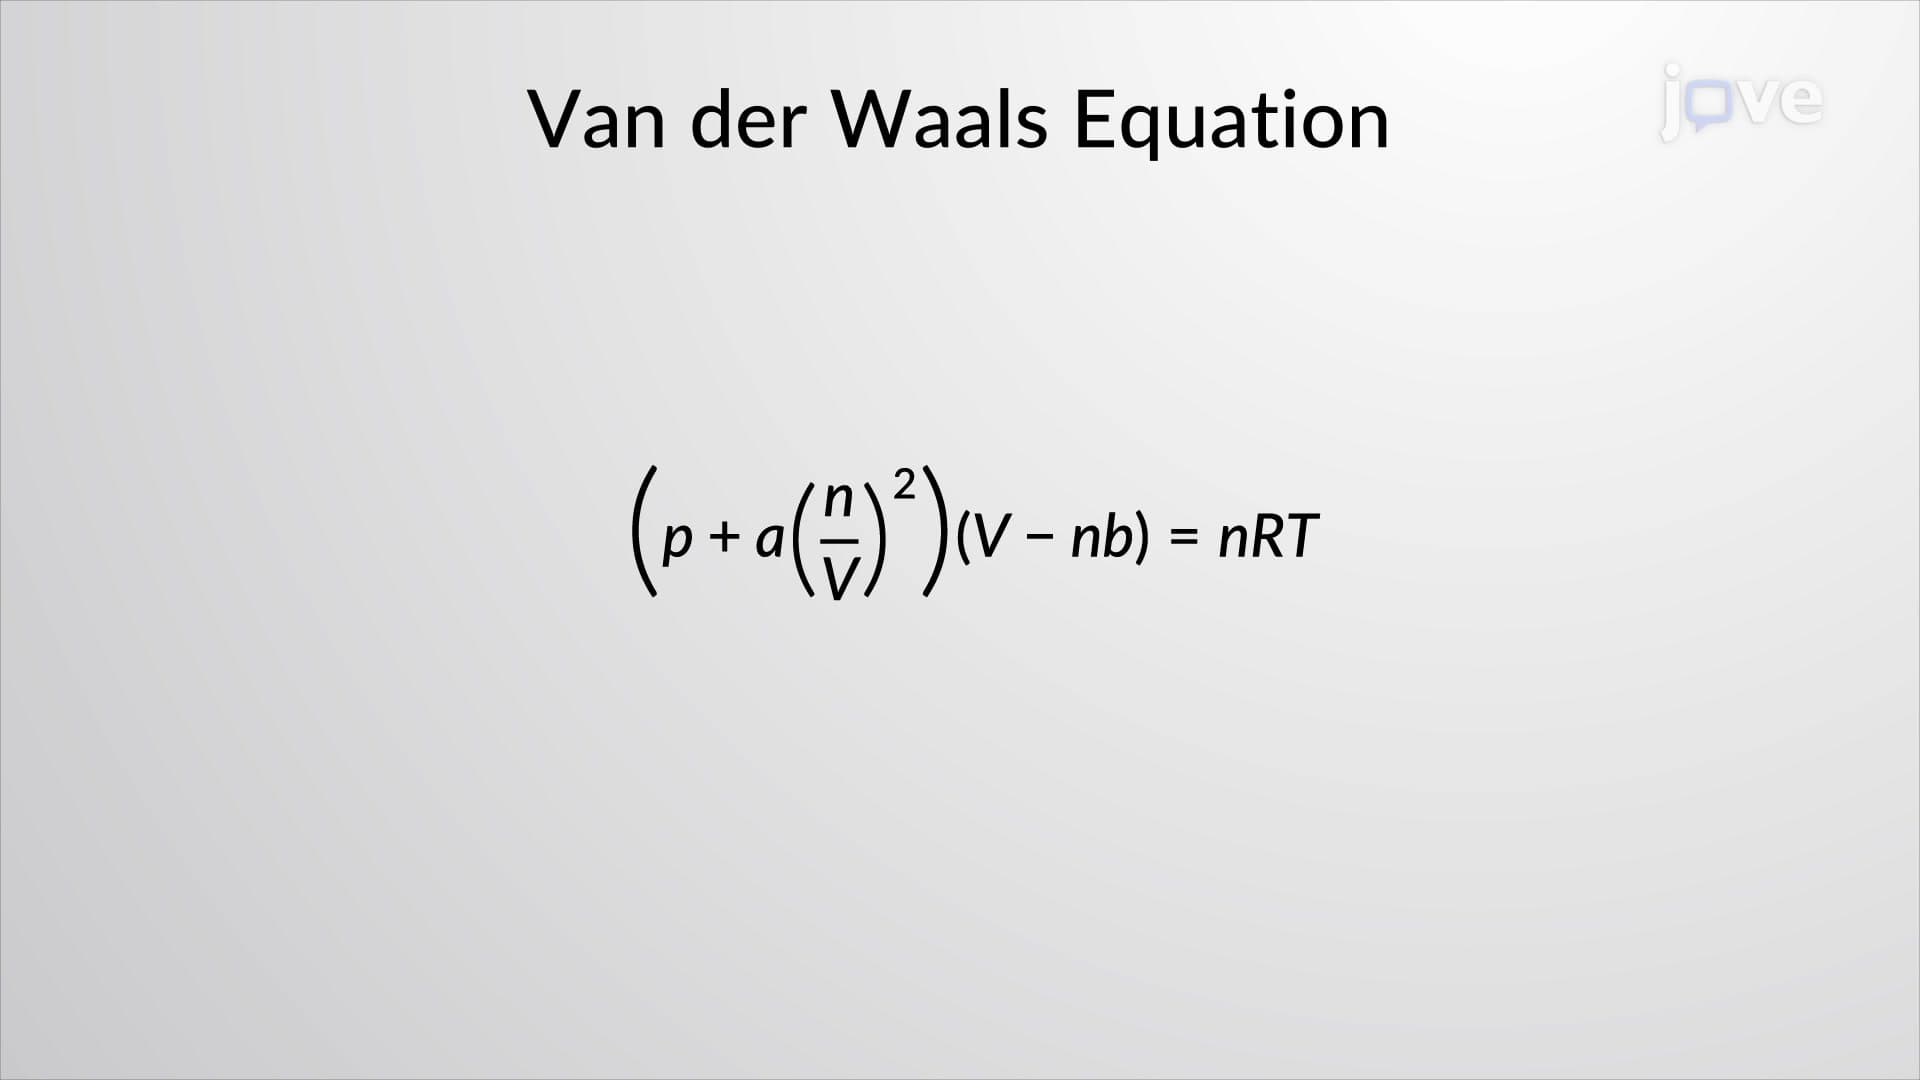
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 02:28
Intermolecular Forces in Solutions
When the strengths of the intermolecular forces of attraction between solute and solvent species in a solution are no different than those present in the separated components, the solution is formed with no accompanying energy change. Such a solution is called an ideal solution. A mixture of ideal gases (or gases such as helium and argon,...