Increasing LRP4 diminishes neuromuscular deficits in a mouse model of Duchenne muscular dystrophy
Related Experiment Video
Updated: Nov 5, 2025
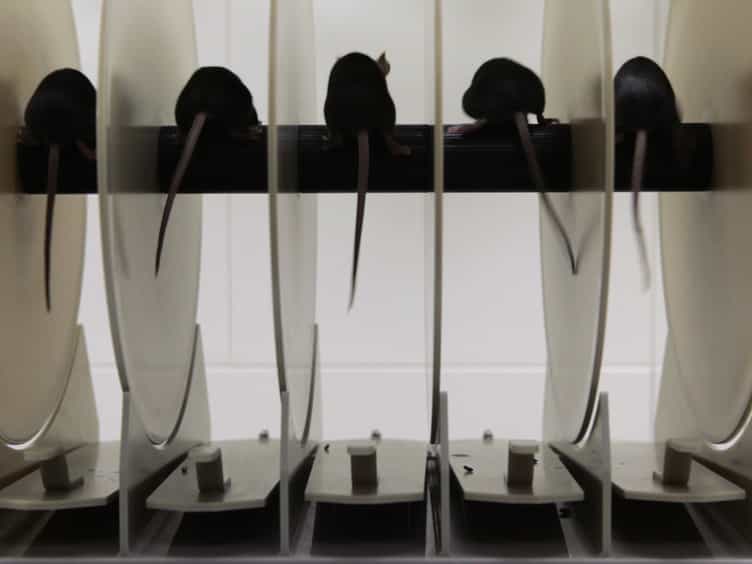
Assessing Functional Performance in the Mdx Mouse Model
Published on: March 27, 2014
Tiankun Hui1,2, Hongyang Jing1,2, Tian Zhou3
1School of Life Science, Nanchang University, Nanchang, Jiangxi, China.

Abstract:
Duchenne muscular dystrophy (DMD) is an X-linked neuromuscular disease characterized by progressive wasting of skeletal muscles. The neuromuscular junction (NMJ) is a synapse between motor neurons and skeletal muscle fibers, critical for the control of muscle contraction. The NMJ decline is observed in DMD patients, but the mechanism is unclear. LRP4 serves as a receptor for agrin, a proteoglycan secreted by motor neurons to induce NMJ, and plays a critical role in NMJ formation and maintenance. Interestingly, we found that protein levels of LRP4 were reduced both in muscles of the DMD patients and DMD model mdx mice. We explored whether increasing LRP4 is beneficial for DMD and crossed muscle-specific LRP4 transgenic mice with mdx mice (mdx; HSA-LRP4). The LRP4 transgene increased muscle strength, together with improved neuromuscular transmission in mdx mice. Furthermore, we found the LRP4 expression mitigated NMJ fragments and denervation in mdx mice. Mechanically, we showed that overexpression of LRP4 increased the activity of MuSK and expression of dystrophin-associated glycoprotein complex proteins in the mdx mice. Overall, our findings suggest that increasing LRP4 improves both function and structure of NMJ in the mdx mice and Agrin signaling might serve as a new therapeutic strategy in DMD.