Related Experiment Video
Updated: Nov 3, 2025
Phenotyping Mouse Pulmonary Function In Vivo with the Lung Diffusing Capacity
Published on: January 6, 2015
Quantification of Phenotypic Variability of Lung Disease in Children with Cystic Fibrosis
Mirjam Stahl1,2, Eva Steinke1,2, Marcus A Mall1,2
1Department of Pediatric Respiratory Medicine, Immunology and Critical Care Medicine, Charité-Universitätsmedizin Berlin, Corporate Member of Freie Universität Berlin and Humboldt-Universität zu Berlin, 13353 Berlin, Germany.

Abstract:
Cystic fibrosis (CF) lung disease has the greatest impact on the morbidity and mortality of patients suffering from this autosomal-recessive multiorgan disorder. Although CF is a monogenic disorder, considerable phenotypic variability of lung disease is observed in patients with CF, even in those carrying the same mutations in the cystic fibrosis transmembrane conductance regulator (CFTR) gene or CFTR mutations with comparable functional consequences. In most patients with CF, lung disease progresses from childhood to adulthood, but is already present in infants soon after birth. In addition to the CFTR genotype, the variability of early CF lung disease can be influenced by several factors, including modifier genes, age at diagnosis (following newborn screening vs. clinical symptoms) and environmental factors. The early onset of CF lung disease requires sensitive, noninvasive measures to detect and monitor changes in lung structure and function. In this context, we review recent progress with using multiple-breath washout (MBW) and lung magnetic resonance imaging (MRI) to detect and quantify CF lung disease from infancy to adulthood. Further, we discuss emerging data on the impact of variability of lung disease severity in the first years of life on long-term outcomes and the potential use of this information to improve personalized medicine for patients with CF.
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic...
 01:27
Chronic Obstructive Pulmonary Disease-IV: Assessement and Diagnostic Studies
Medical History
 01:11
Variability: Analysis
The range is a simple measure of variability, indicating the difference between the highest and...
 01:24
Respiratory Capacities
One key metric is the Inspiratory Capacity (IC), which represents the maximum amount of air that can be inhaled with full effort. IC is calculated by summing the tidal volume and inspiratory reserve volume, typically ranging from 2.4 to 3.6 liters.
The Functional Residual Capacity (FRC) represents the air in the...
 01:15
Respiratory Volumes
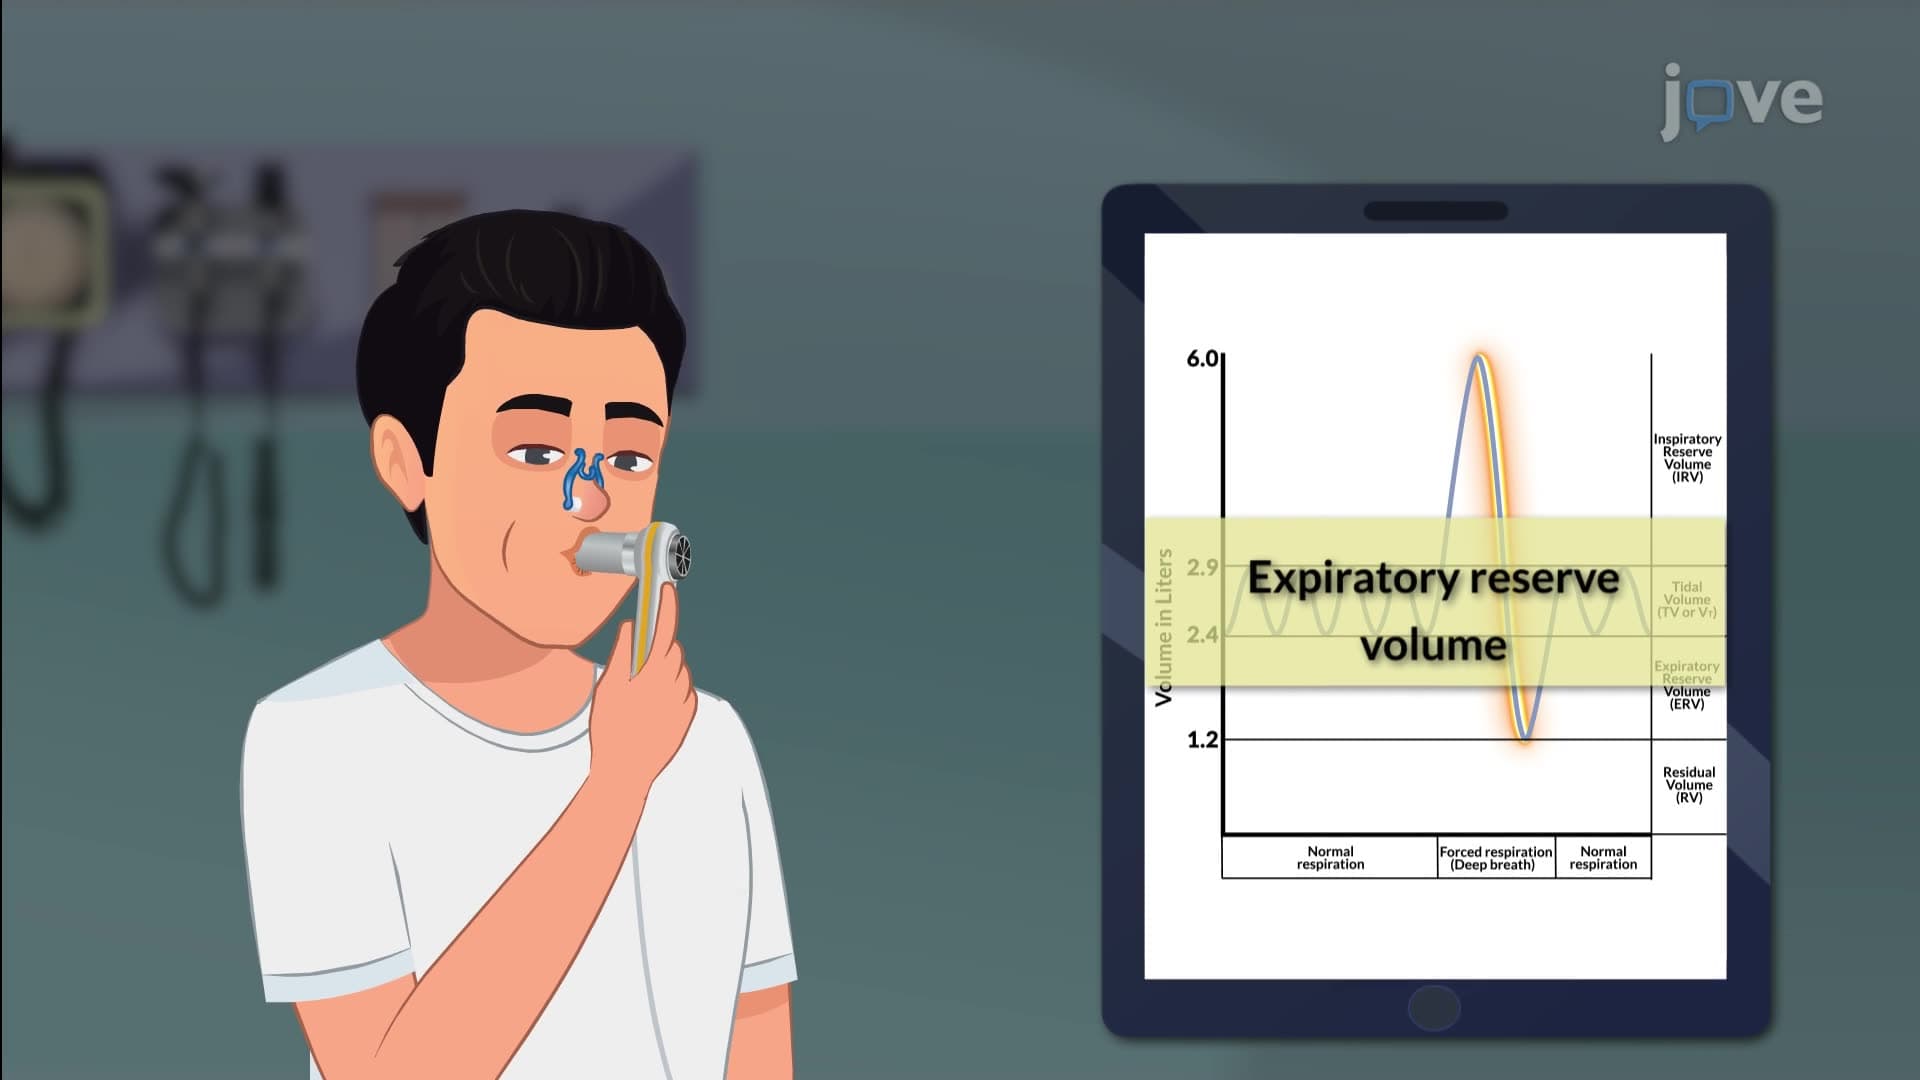
Tidal Volume (TV) Tidal volume (TV) is the air inhaled or exhaled in a...