Related Experiment Video
Updated: Nov 3, 2025


Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Efficient Band Structure Calculation of Two-Dimensional Materials from Semilocal Density Functionals
Abhilash Patra1, Subrata Jana1, Prasanjit Samal1
1School of Physical Sciences, National Institute of Science Education and Research, HBNI, Bhubaneswar 752050, India.

Abstract:
The experimental and theoretical realization of two-dimensional (2D) materials is of utmost importance in semiconducting applications. Computational modeling of these systems with satisfactory accuracy and computational efficiency is only feasible with semilocal density functional theory methods. In the search for the most useful method in predicting the band gap of 2D materials, we assess the accuracy of recently developed semilocal exchange-correlation (XC) energy functionals and potentials. Though the explicit forms of exchange-correlation (XC) potentials are very effective against XC energy functionals for the band gap of bulk solids, their performance needs to be investigated for 2D materials. In particular, the LMBJ [J. Chem. Theory Comput.2020, 16, 2654] and GLLB-SC [Phys. Rev. B82, 2010, 115106] potentials are considered for their dominance in bulk band gap calculation. The performance of recently developed meta generalized gradient approximations, like TASK [Phys. Rev. Res.1, 2019, 033082] and MGGAC [Phys. Rev. B. 100, 2019, 155140], is also assessed. We find that the LMBJ potential constructed for 2D materials is not as successful as its parent functional, i.e., MBJ [Phys. Rev. Lett.102, 2009, 226401] in bulk solids. Due to a contribution from the derivative discontinuity, the band gaps obtained with GLLB-SC are in a certain number of cases, albeit not systematically, larger than those obtained with other methods, which leads to better agreement with the quasi-particle band gap obtained from the GW method. The band gaps obtained with TASK and MGGAC can also be quite accurate.
More Related Videos


Related Concept Videos
 02:35
Band Theory
The energy difference between these bands is known as the band gap.
Conductor, Semiconductor,...
 01:01
Energy Bands in Solids
Band Formation:
When atoms are brought close together, as in a solid, these discrete energy levels begin to split due to the overlap of electron orbitals from adjacent atoms. This split occurs because of the Pauli exclusion principle, which states...
 01:18
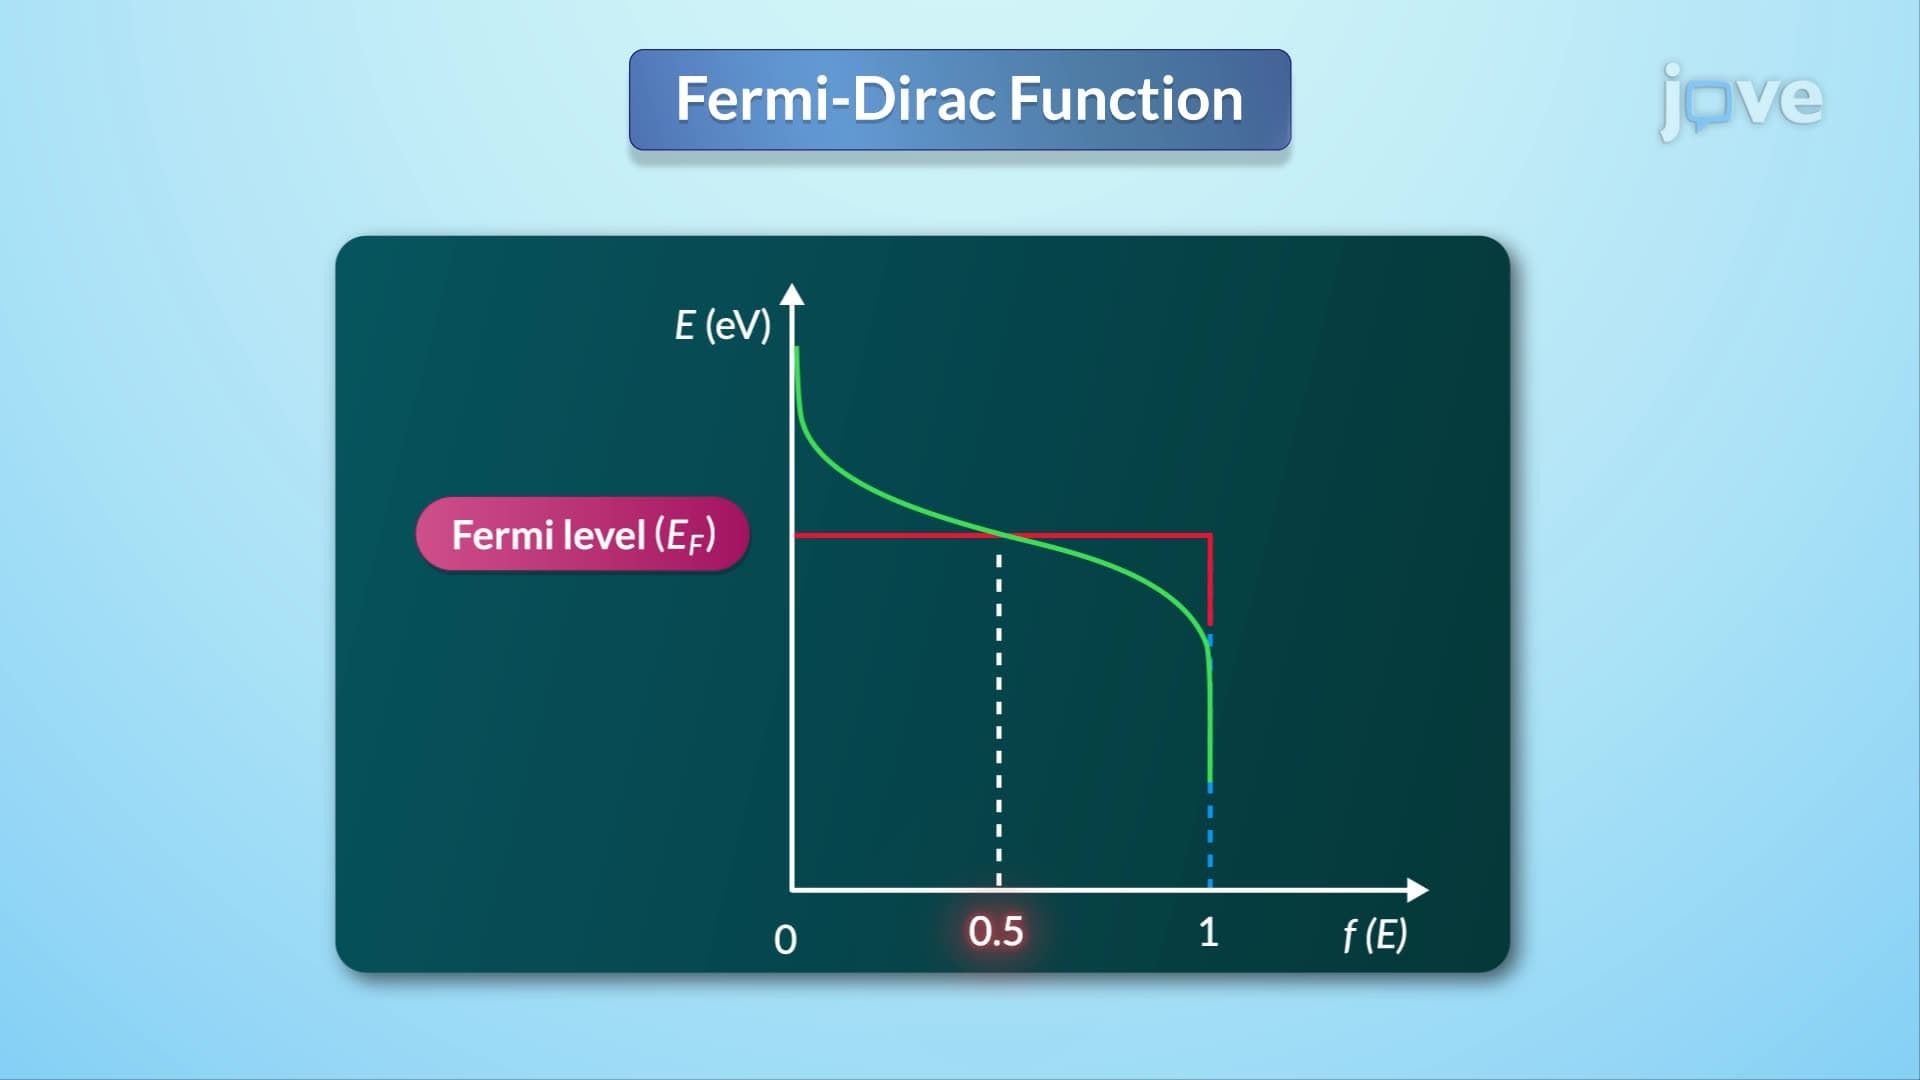
Fermi Level
At absolute zero temperature, electrons fill all energy states up to the Fermi level, leaving upper states empty. As the temperature rises,...
 01:12
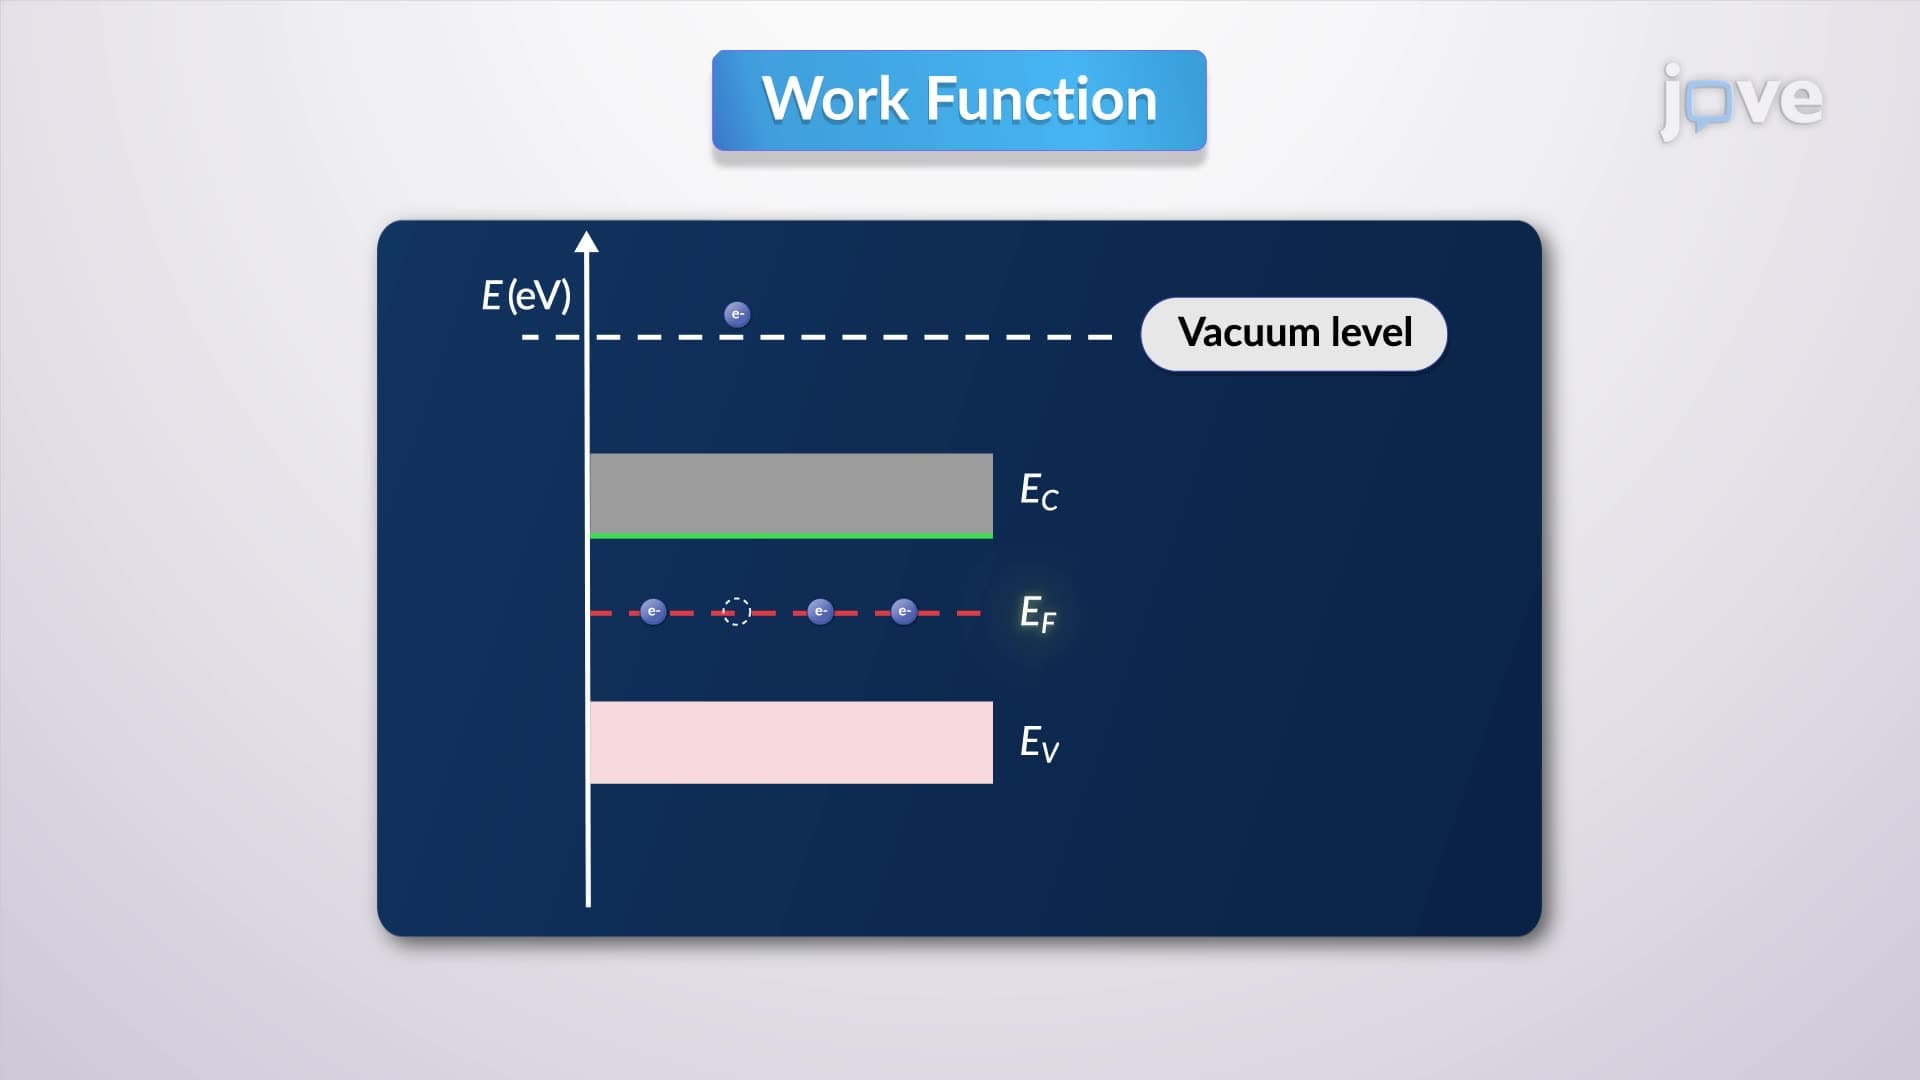
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...