Related Experiment Video
Updated: Nov 2, 2025

Author Spotlight: Integrating BRET-Based Assays and Rare Mutation Analysis to Decipher RAF Kinase Regulation in Live Cells
Published on: March 1, 2024
Exploring the Binding Interaction of Raf Kinase Inhibitory Protein With the N-Terminal of C-Raf Through Molecular
Shraddha Parate1, Shailima Rampogu2, Gihwan Lee1
1Division of Life Sciences, Division of Applied Life Science (BK21 Plus), Plant Molecular Biology and Biotechnology Research Center (PMBBRC), Research Institute of Natural Science (RINS), Gyeongsang National University (GNU), Jinju, Korea.

Abstract:
Protein-protein interactions are indispensable physiological processes regulating several biological functions. Despite the availability of structural information on protein-protein complexes, deciphering their complex topology remains an outstanding challenge. Raf kinase inhibitory protein (RKIP) has gained substantial attention as a favorable molecular target for numerous pathologies including cancer and Alzheimer's disease. RKIP interferes with the RAF/MEK/ERK signaling cascade by endogenously binding with C-Raf (Raf-1 kinase) and preventing its activation. In the current investigation, the binding of RKIP with C-Raf was explored by knowledge-based protein-protein docking web-servers including HADDOCK and ZDOCK and a consensus binding mode of C-Raf/RKIP structural complex was obtained. Molecular dynamics (MD) simulations were further performed in an explicit solvent to sample the conformations for when RKIP binds to C-Raf. Some of the conserved interface residues were mutated to alanine, phenylalanine and leucine and the impact of mutations was estimated by additional MD simulations and MM/PBSA analysis for the wild-type (WT) and constructed mutant complexes. Substantial decrease in binding free energy was observed for the mutant complexes as compared to the binding free energy of WT C-Raf/RKIP structural complex. Furthermore, a considerable increase in average backbone root mean square deviation and fluctuation was perceived for the mutant complexes. Moreover, per-residue energy contribution analysis of the equilibrated simulation trajectory by HawkDock and ANCHOR web-servers was conducted to characterize the key residues for the complex formation. One residue each from C-Raf (Arg398) and RKIP (Lys80) were identified as the druggable "hot spots" constituting the core of the binding interface and corroborated by additional long-time scale (300 ns) MD simulation of Arg398Ala mutant complex. A notable conformational change in Arg398Ala mutant occurred near the mutation site as compared to the equilibrated C-Raf/RKIP native state conformation and an essential hydrogen bonding interaction was lost. The thirteen binding sites assimilated from the overall analysis were mapped onto the complex as surface and divided into active and allosteric binding sites, depending on their location at the interface. The acquired information on the predicted 3D structural complex and the detected sites aid as promising targets in designing novel inhibitors to block the C-Raf/RKIP interaction.
Insights
Researchers identified key "hot spots" in the C-Raf/RKIP protein complex, revealing critical residues like Arg398 and Lys80. Mutations at these sites significantly reduced binding energy, offering new targets for developing inhibitors against cancer and Alzheimer's disease.
Area of Science:
- Structural Biology
- Computational Biophysics
- Molecular Pharmacology
Background:
- Protein-protein interactions are crucial for biological functions, but their complex structures are challenging to decipher.
- The Raf kinase inhibitory protein (RKIP) is a significant target for diseases like cancer and Alzheimer's, as it regulates the RAF/MEK/ERK pathway.
- RKIP inhibits C-Raf (Raf-1 kinase) activation, making the C-Raf/RKIP interaction a key area for therapeutic intervention.
Purpose of the Study:
- To elucidate the binding mode and structural complex of RKIP with C-Raf using computational methods.
- To identify critical residues and potential druggable sites within the C-Raf/RKIP interface.
- To assess the impact of mutations on the stability and binding energy of the C-Raf/RKIP complex.
Main Methods:
- Knowledge-based protein-protein docking (HADDOCK, ZDOCK) to predict the consensus binding mode of the C-Raf/RKIP complex.
- Molecular dynamics (MD) simulations in explicit solvent to analyze conformational dynamics.
- Site-directed mutagenesis of conserved interface residues, followed by MD simulations and MM/PBSA analysis to evaluate binding free energy and complex stability.
- Per-residue energy contribution analysis using HawkDock and ANCHOR web servers to identify key residues.
Main Results:
- A consensus structural complex of C-Raf/RKIP was obtained, and MD simulations provided insights into binding conformations.
- Mutant complexes showed a substantial decrease in binding free energy and increased backbone fluctuations compared to wild-type.
- Residues Arg398 (C-Raf) and Lys80 (RKIP) were identified as druggable 'hot spots' at the binding interface.
- A specific Arg398Ala mutation led to conformational changes and loss of a key hydrogen bond, confirming its importance.
Conclusions:
- The study successfully predicted the 3D structure of the C-Raf/RKIP complex and identified critical binding interface residues.
- The identified 'hot spots' (Arg398 and Lys80) represent promising targets for designing novel inhibitors.
- Blocking the C-Raf/RKIP interaction through these identified sites could offer new therapeutic strategies for cancer and Alzheimer's disease.
Related Concept Videos
 01:07
MAPK Signaling Cascades
 01:30
Assembly of Signaling Complexes
Interaction domains in cell signaling
Interaction domains recognize exposed features of their binding partners containing post-translationally modified sequences,...
 02:04
Protein-protein Interfaces
 01:24
Small GTPases - Ras and Rho
Three regulatory proteins control their activity:
 01:26
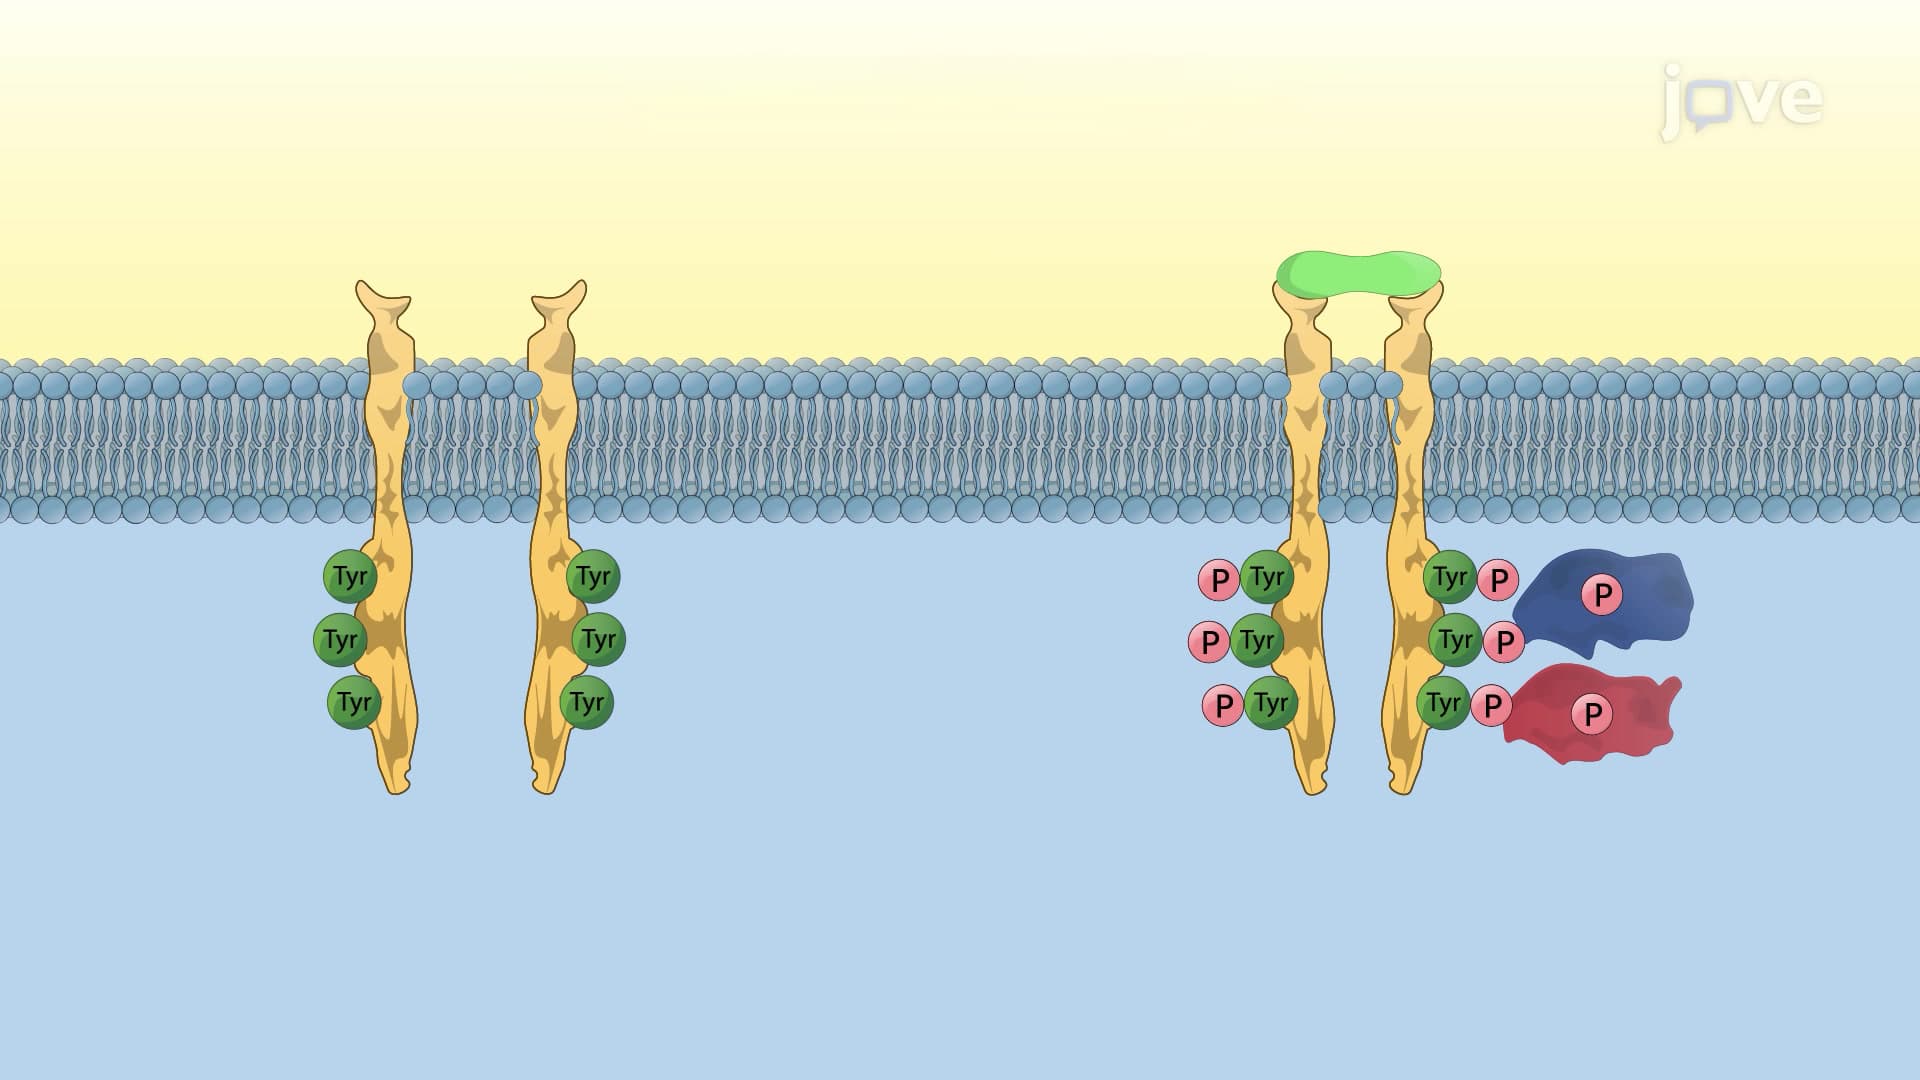
Receptor Tyrosine Kinases
 01:19
Interactions Between Signaling Pathways
Convergence and divergence, and cross-talk between signaling pathways
Two distinct signaling pathways can converge on a single functional unit, which may either be a single protein or a complex of proteins. The response is either functionally distinct or synergistic between the two pathways but different from the response...