Related Experiment Video
Updated: Nov 2, 2025
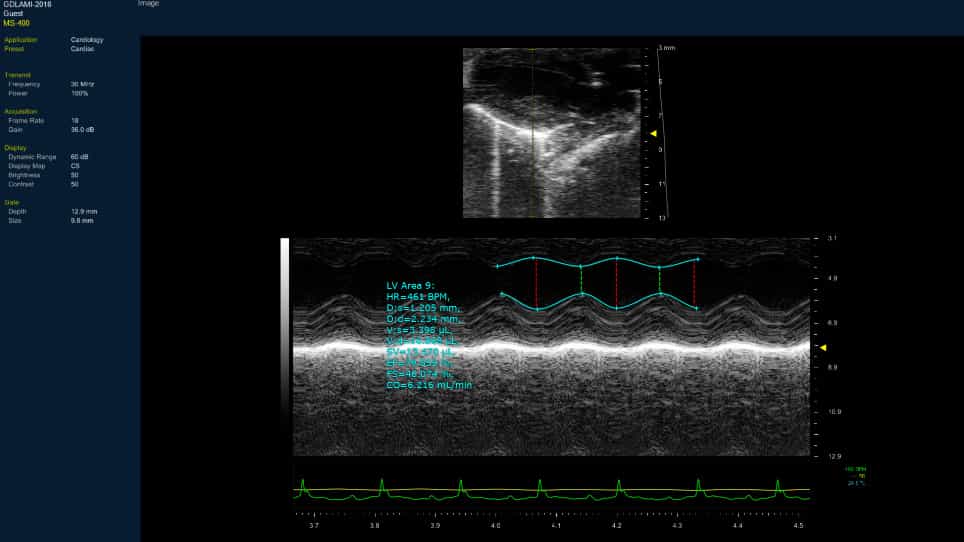
Investigating the Pathogenesis of MYH7 Mutation Gly823Glu in Familial Hypertrophic Cardiomyopathy using a Mouse Model
Published on: August 8, 2022
DIAPH1 Variants in Non-East Asian Patients With Sporadic Moyamoya Disease
Adam J Kundishora1, Samuel T Peters2, Amélie Pinard1
1Department of Internal Medicine, McGovern Medical School, University of Texas Health Science Center, Houston.

Importance:
Moyamoya disease (MMD), a progressive vasculopathy leading to narrowing and ultimate occlusion of the intracranial internal carotid arteries, is a cause of childhood stroke. The cause of MMD is poorly understood, but genetic factors play a role. Several familial forms of MMD have been identified, but the cause of most cases remains elusive, especially among non-East Asian individuals.
Objective:
To assess whether ultrarare de novo and rare, damaging transmitted variants with large effect sizes are associated with MMD risk.
Design, Setting, And Participants:
A genetic association study was conducted using whole-exome sequencing case-parent MMD trios in a small discovery cohort collected over 3.5 years (2016-2019); data were analyzed in 2020. Medical records from US hospitals spanning a range of 1 month to 1.5 years were reviewed for phenotyping. Exomes from a larger validation cohort were analyzed to identify additional rare, large-effect variants in the top candidate gene. Participants included patients with MMD and, when available, their parents. All participants who met criteria and were presented with the option to join the study agreed to do so; none were excluded. Twenty-four probands (22 trios and 2 singletons) composed the discovery cohort, and 84 probands (29 trios and 55 singletons) composed the validation cohort.
Main Outcomes And Measures:
Gene variants were identified and filtered using stringent criteria. Enrichment and case-control tests assessed gene-level variant burden. In silico modeling estimated the probability of variant association with protein structure. Integrative genomics assessed expression patterns of MMD risk genes derived from single-cell RNA sequencing data of human and mouse brain tissue.
Results:
Of the 24 patients in the discovery cohort, 14 (58.3%) were men and 18 (75.0%) were of European ancestry. Three of 24 discovery cohort probands contained 2 do novo (1-tailed Poisson P = 1.1 × 10⁻⁶) and 1 rare, transmitted damaging variant (12.5% of cases) in DIAPH1 (mammalian diaphanous-1), a key regulator of actin remodeling in vascular cells and platelets. Four additional ultrarare damaging heterozygous DIAPH1 variants (3 unphased) were identified in 3 other patients in an 84-proband validation cohort (73.8% female, 77.4% European). All 6 patients were non-East Asian. Compound heterozygous variants were identified in ena/vasodilator-stimulated phosphoproteinlike protein EVL, a mammalian diaphanous-1 interactor that regulates actin polymerization. DIAPH1 and EVL mutant probands had severe, bilateral MMD associated with transfusion-dependent thrombocytopenia. DIAPH1 and other MMD risk genes are enriched in mural cells of midgestational human brain. The DIAPH1 coexpression network converges in vascular cell actin cytoskeleton regulatory pathways.
Conclusions And Relevance:
These findings provide the largest collection to date of non-East Asian individuals with sporadic MMD harboring pathogenic variants in the same gene. The results suggest that DIAPH1 is a novel MMD risk gene and impaired vascular cell actin remodeling in MMD pathogenesis, with diagnostic and therapeutic ramifications.
Insights
Genetic variants in DIAPH1 are linked to Moyamoya disease (MMD) risk in non-East Asian individuals. This discovery offers new insights into MMD pathogenesis and potential diagnostic approaches.
Area of Science:
- Genetics
- Neurology
- Vascular Biology
Background:
- Moyamoya disease (MMD) is a progressive vasculopathy causing childhood stroke, characterized by intracranial artery narrowing.
- The genetic underpinnings of MMD, particularly in non-East Asian populations, remain largely unknown.
- Previous studies identified familial forms, but the etiology of most sporadic cases is elusive.
Purpose of the Study:
- To investigate the association of ultrarare de novo and rare, damaging transmitted genetic variants with Moyamoya disease risk.
- To identify novel genetic factors contributing to MMD pathogenesis, especially in underrepresented ethnic groups.
Main Methods:
- Whole-exome sequencing was performed on MMD patient-parent trios in a discovery cohort.
- A validation cohort was used to identify additional rare, large-effect variants in candidate genes.
- Integrative genomics and in silico modeling were employed to assess variant effects and gene expression patterns.
Main Results:
- Pathogenic variants in DIAPH1 (mammalian diaphanous-1) were identified in non-East Asian MMD patients, including de novo and rare transmitted variants.
- Compound heterozygous variants in EVL, a DIAPH1 interactor, were also found.
- Patients with DIAPH1 or EVL variants exhibited severe MMD and transfusion-dependent thrombocytopenia, with DIAPH1 implicated in vascular cell actin remodeling.
Conclusions:
- DIAPH1 is identified as a novel risk gene for Moyamoya disease, particularly in sporadic, non-East Asian cases.
- Impaired vascular cell actin remodeling due to DIAPH1 dysfunction is implicated in MMD pathogenesis.
- These findings have significant diagnostic and therapeutic implications for Moyamoya disease.
More Related Videos


Related Concept Videos
 01:29
Cardiomyopathy III: Hypertrophic Cardiomyopathy
 01:21
Aneurysm II: Clinical Manifestations and Diagnostic Studies
 01:30
Coronary Artery Disease III: Clinical Manifestations
 01:23
Mitral Stenosis II: Clinical features and Diagnostic Tests
 01:26
Coronary Artery Disease II: Pathophysiology
 01:25
Cardiomyopathy I: Introduction and Classification