Related Experiment Video
Updated: Nov 2, 2025
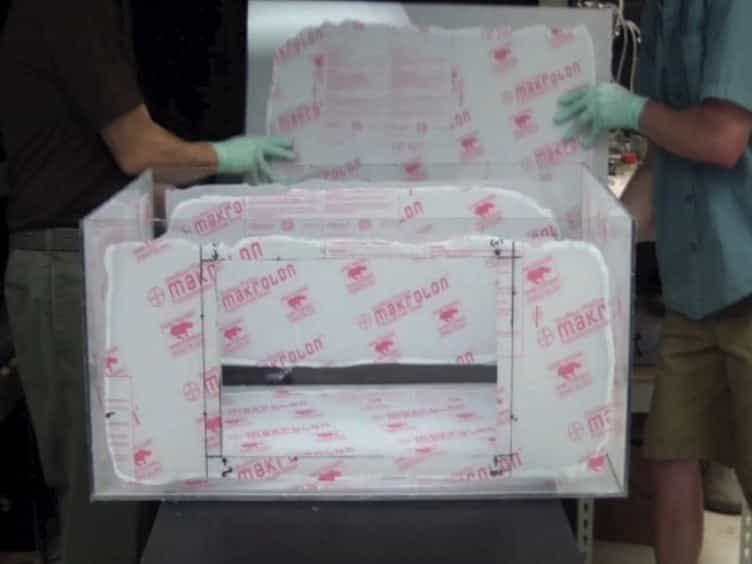

Construction of Vapor Chambers Used to Expose Mice to Alcohol During the Equivalent of all Three Trimesters of Human Development
Published on: July 13, 2014
Programmed suppression of oxidative phosphorylation and mitochondrial function by gestational alcohol exposure
Richard C Chang1,2, Kara N Thomas1, Nicole A Mehta1
1Department of Veterinary Physiology & Pharmacology, College of Veterinary Medicine and Biomedical Sciences, Texas A&M University, 4466 TAMU, College Station, TX, 77843-4466, USA.

Background:
A critical question emerging in the field of developmental toxicology is whether alterations in chromatin structure induced by toxicant exposure control patterns of gene expression or, instead, are structural changes that are part of a nuclear stress response. Previously, we used a mouse model to conduct a three-way comparison between control offspring, alcohol-exposed but phenotypically normal animals, and alcohol-exposed offspring exhibiting craniofacial and central nervous system structural defects. In the cerebral cortex of animals exhibiting alcohol-induced dysgenesis, we identified a dramatic increase in the enrichment of dimethylated histone H3, lysine 9 (H3K9me2) within the regulatory regions of key developmental factors driving histogenesis in the brain. However, whether this change in chromatin structure is causally involved in the development of structural defects remains unknown.
Results:
Deep-sequencing analysis of the cortex transcriptome reveals that the emergence of alcohol-induced structural defects correlates with disruptions in the genetic pathways controlling oxidative phosphorylation and mitochondrial function. The majority of the affected pathways are downstream targets of the mammalian target of rapamycin complex 2 (mTORC2), indicating that this stress-responsive complex plays a role in propagating the epigenetic memory of alcohol exposure through gestation. Importantly, transcriptional disruptions of the pathways regulating oxidative homeostasis correlate with the emergence of increased H3K9me2 across genic, repetitive, and non-transcribed regions of the genome. However, although associated with gene silencing, none of the candidate genes displaying increased H3K9me2 become transcriptionally repressed, nor do they exhibit increased markers of canonical heterochromatin. Similar to studies in C. elegans, disruptions in oxidative homeostasis induce the chromatin looping factor SATB2, but in mammals, this protein does not appear to drive increased H3K9me2 or altered patterns of gene expression.
Conclusions:
Our studies demonstrate that changes in H3K9me2 associate with alcohol-induced congenital defects, but that this epigenetic change does not correlate with transcriptional suppression. We speculate that the mobilization of SATB2 and increased enrichment of H3K9me2 may be components of a nuclear stress response that preserve chromatin integrity and interactions under prolonged oxidative stress. Further, we postulate that while this response may stabilize chromatin structure, it compromises the nuclear plasticity required for normal differentiation.
Related Concept Videos
 02:30
Genomic Imprinting and Inheritance
The expression of some genes depends on which parent passed the gene to the offspring, through a phenomenon known as...
 01:07
Teratogenicity
 01:37
Epigenetic Regulation
X-chromosome...
 01:46
Epigenetic Regulation
 02:59
Animal Mitochondrial Genetics