
The ortho effect in directed C-H activation
Balázs L Tóth1, Anna Monory1, Orsolya Egyed2
1ELTE "Lendület" Catalysis and Organic Synthesis Research Group, Faculty of Science, Institute of Chemistry, Eötvös Loránd University Pázmány Péter Sétány. 1/A H-1117 Budapest Hungary tothb@zng.elte.hu novakz@elte.hu.

Abstract:
The success of transition metal-catalysed ortho-directed C-H activation is often plagued by the effects of undesirable interactions between the directing group (DG) and other groups introduced into the aromatic core of the substrate. In particular, when these groups are in neighbouring positions, their interactions can affect profoundly the efficacy of the C-H activation by transition metals. In this work we introduce a simple substrate-only-based model to interpret the influence of steric hindrance of a group in ortho position to the DG in directed ortho-C-H bond activation reactions, and coined the term Ortho Effect (OE) for such situations. We consider simple descriptors such as torsion angle and torsional energy to predict and explain the reactivity of a given substrate in directed C-H activation reactions. More than 250 examples have been invoked for the model, and the nature of the ortho effect was demonstrated on a wide variety of structures. In order to guide organic chemists, we set structural and energetic criteria to evaluate a priori the efficiency of the metalation step which is usually the rate-determining event in C-H activations, i.e. we provide a simple and general protocol to estimate the reactivity of a potential substrate in C-H activation. For borderline cases these criteria help set the minimum reaction temperature to obtain reasonable reaction rates. As an example for the practical applicability of the model, we performed synthetic validations via palladium-catalysed 2,2,2-trifluoroethylation reactions in our lab. Furthermore, we give predictions for the necessary reaction conditions for several selected DGs.
Related Concept Videos
 01:11
ortho–para-Directing Activators: –CH3, –OH, –NH2, –OCH3
 02:36
Regioselectivity and Stereochemistry of Hydroboration
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn stereochemistry.
 01:26
Photochemical Electrocyclic Reactions: Stereochemistry
Selection Rules: Photochemical Activation
 01:17
Thermal Electrocyclic Reactions: Stereochemistry
Selection Rules: Thermal Activation
Conjugated systems containing an even number of π-electron pairs undergo a conrotatory ring closure. For example, thermal electrocyclization of (2E,4E)-2,4-hexadiene, a conjugated diene containing two π-electron pairs, gives trans-3,4-dimethylcyclobutene.
 01:24
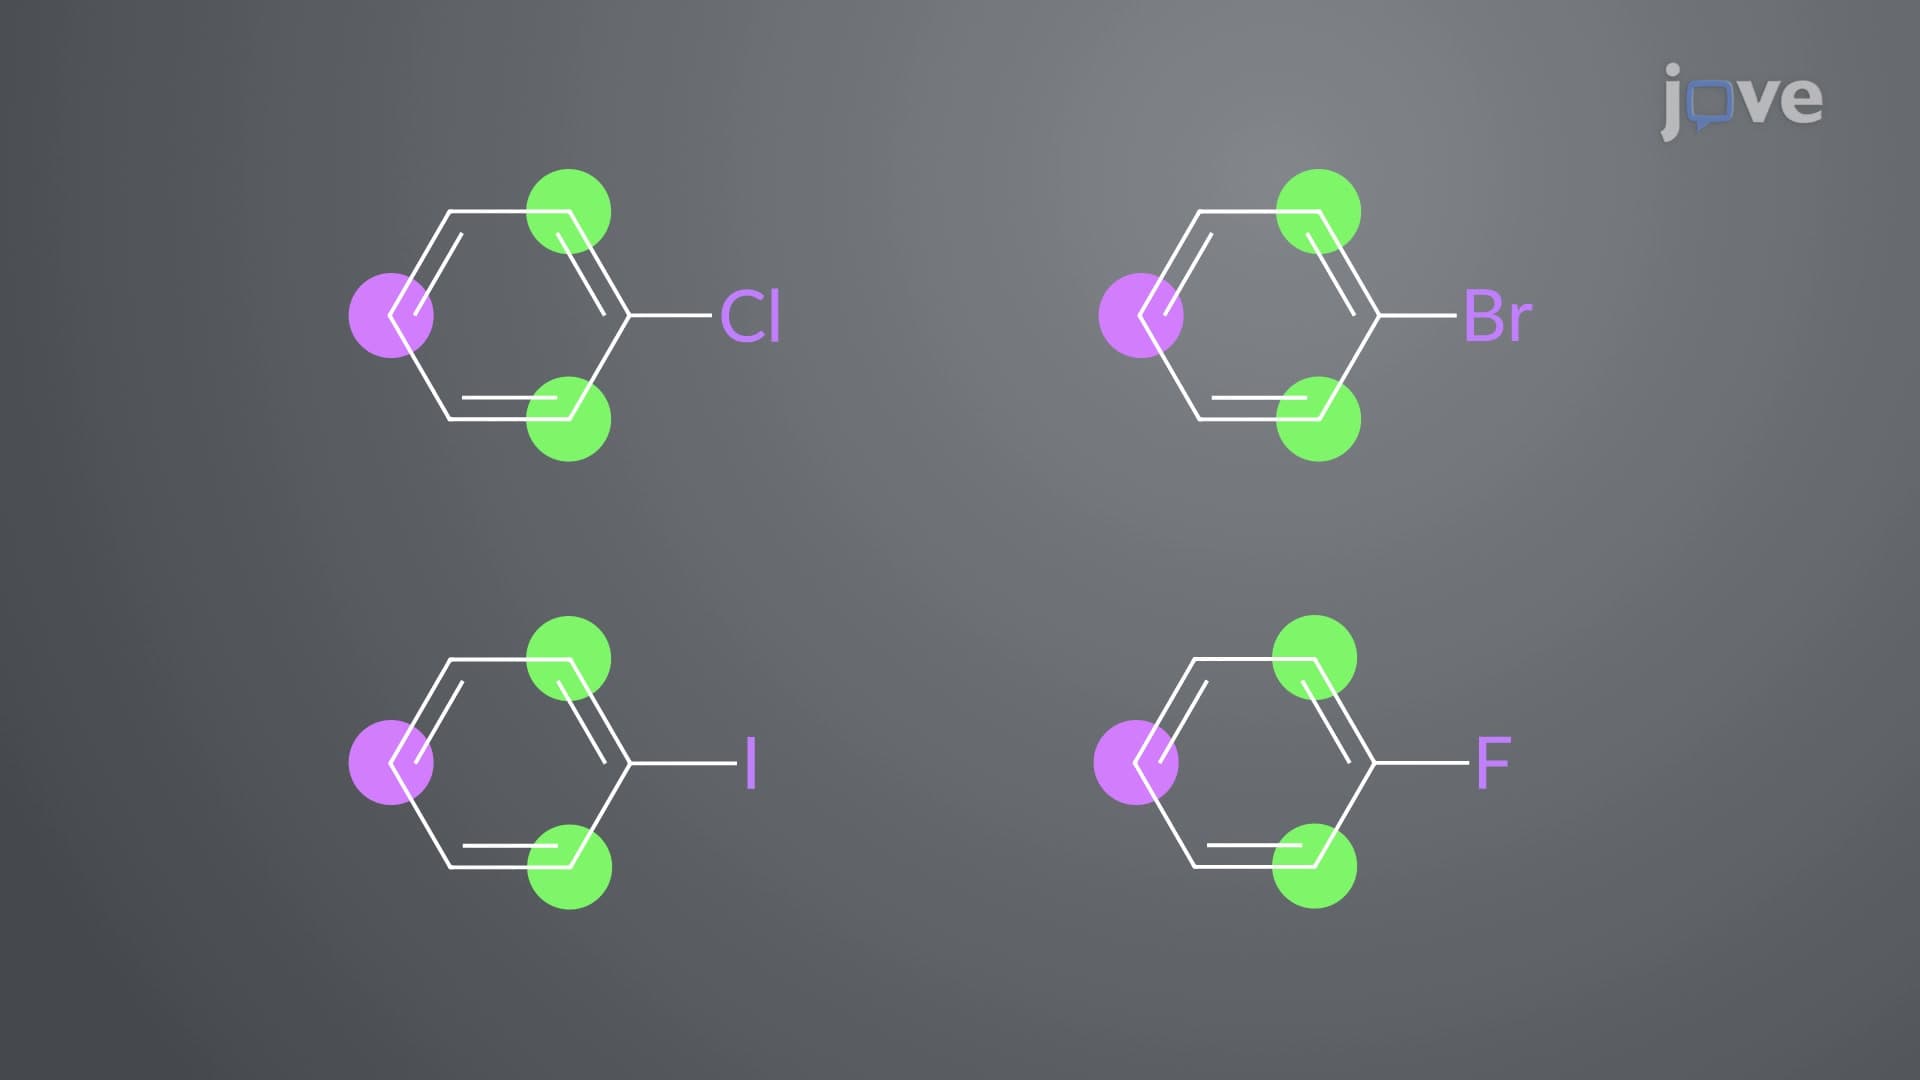
ortho–para-Directing Deactivators: Halogens
 02:34
Regioselectivity and Stereochemistry of Acid-Catalyzed Hydration