Related Experiment Video
Updated: Oct 31, 2025

Pharmacophore Modeling for Targets with Extensive Ligand Libraries: A Case Study on SARS-CoV-2 Mpro
Published on: September 26, 2025
Exploring Dual Agonists for PPARα/γ Receptors using Pharmacophore Modeling, Docking Analysis and Molecule Dynamics
Ting-Ting Ding1, Ya-Ya Liu1, Li-Ming Zhang1
1Tianjin Key Laboratory on Technologies Enabling Development of Clinical Therapeutics and Diagnostics (Theranostics), School of Pharmacy, Tianjin Medical University, Tianjin 300070, China.

Background:
The Peroxisome Proliferator-Activated Receptors (PPARs) are ligandactivated transcription factors belonging to the nuclear receptor family. The roles of PPARα in fatty acid oxidation and PPARγ in adipocyte differentiation and lipid storage have been widely characterized. Compounds with dual PPARα/γ activity have been proposed, combining the benefits of insulin sensitization and lipid lowering into one drug, allowing a single drug to reduce hyperglycemia and hyperlipidemia while preventing the development of cardiovascular complications.
Methods:
The new PPARα/γ agonists were screened through virtual screening of pharmacophores and molecular dynamics simulations. First, in the article, the constructed pharmacophore was used to screen the Ligand Expo Components-pub database to obtain the common structural characteristics of representative PPARα/γ agonist ligands. Then, the accepted ligand structure was modified and replaced to obtain 12 new compounds. Using molecular docking, ADMET and molecular dynamics simulation methods to screen the designed 12 ligands, analyze their docking scores when they bind to the PPARα/γ dual targets, their stability and pharmacological properties when they bind to the PPARα/γ dual targets.
Results:
We performed pharmacophore-based virtual screening for 22949 molecules in Ligand Expo Components-pub database. The compounds that were superior to the original ligand were performed structural analysis and modification, and a series of compounds with novel structures were designed. Using precise docking, ADMET prediction and molecular dynamics methods to screen and verify newly designed compounds, and the above compounds show higher docking scores and lower side effects.
Conclusion:
9 new PPARα/γ agonists were obtained by pharmacophore modeling, docking analysis and molecular dynamics simulation.
More Related Videos
 11:06
Network Pharmacology Prediction and Metabolomics Validation of the Mechanism of Fructus Phyllanthi against Hyperlipidemia
Published on: April 7, 2023
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Related Concept Videos
 01:25
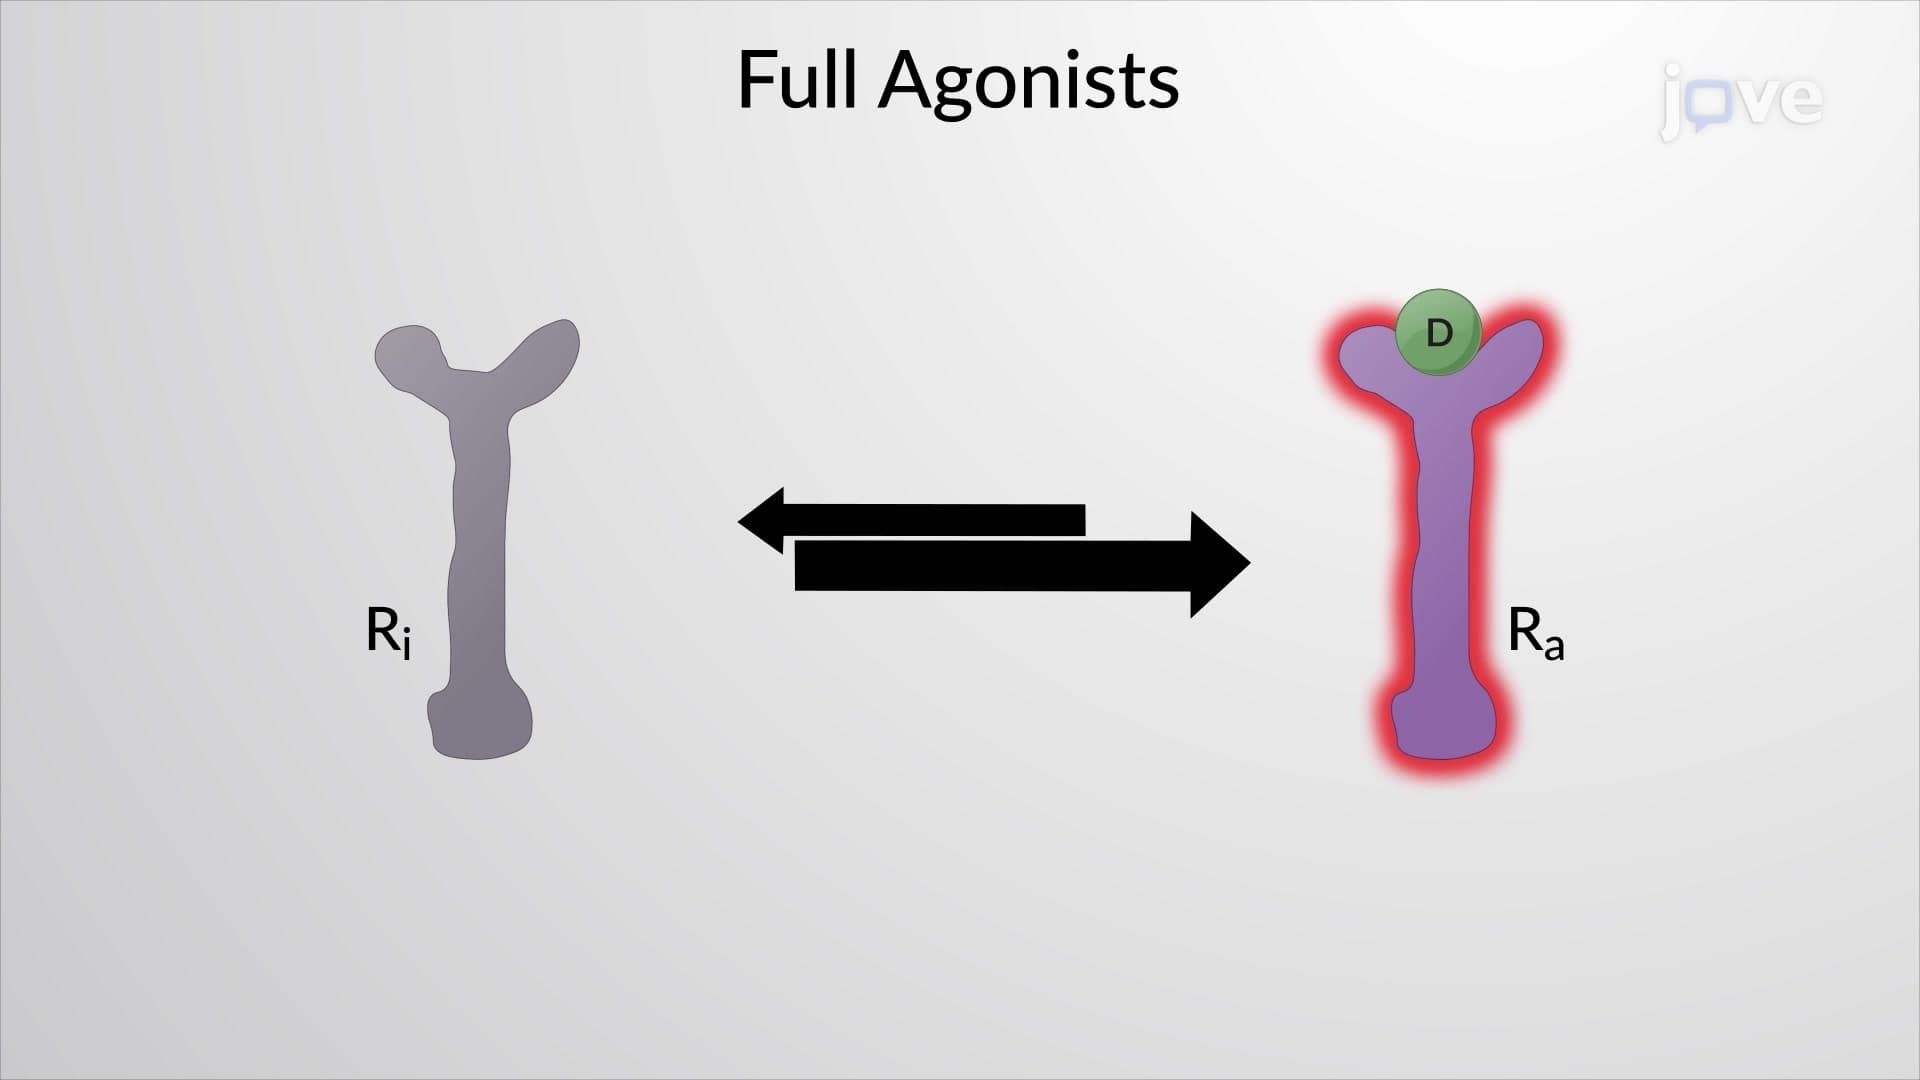
Drug-Receptor Interaction: Agonist
Agonists can bind to receptors in different ways. Some agonists bind directly to the receptor's active site, mimicking the endogenous...
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with...
 01:29
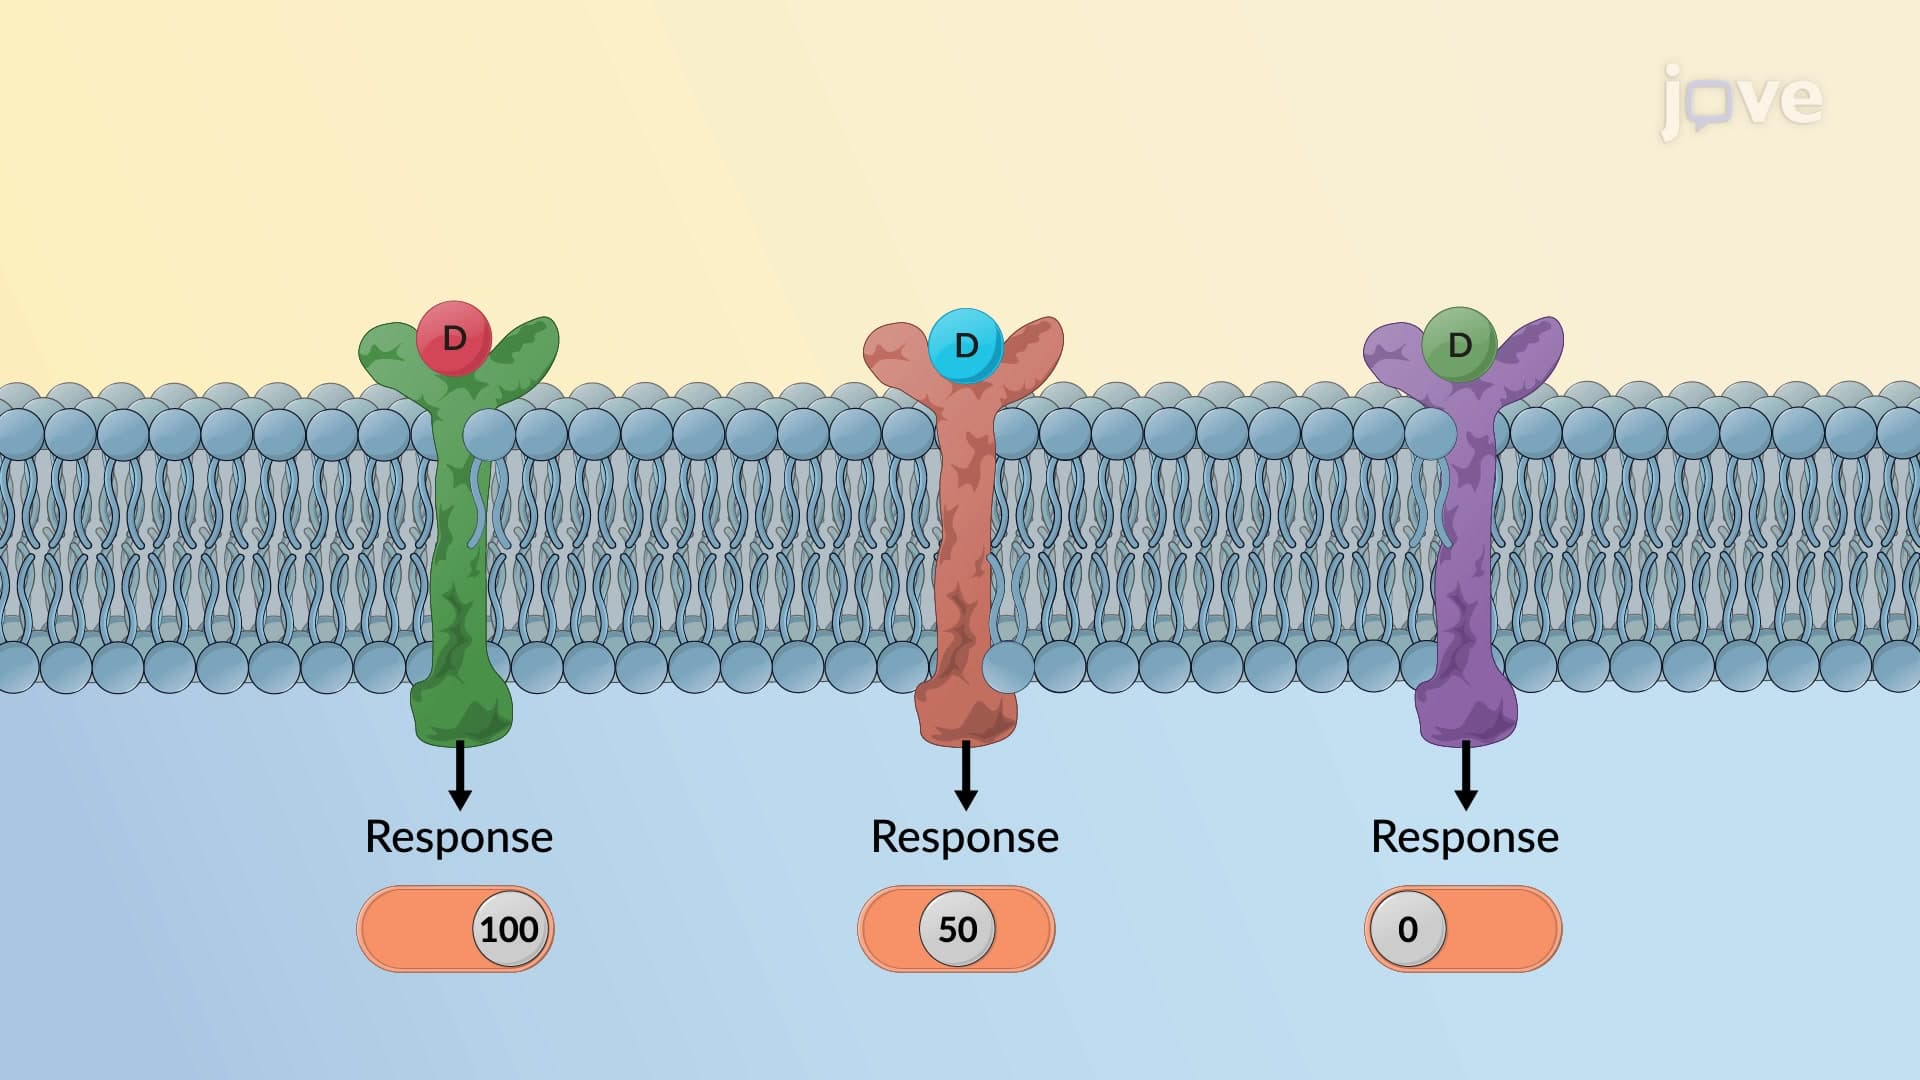
Drug-Receptor Interactions
Several parameters, such as the drug's affinity for its receptor and its efficacy, which is its ability to activate the receptor, determine the drug's effect on the tissue....
 01:15
G Protein-coupled Receptors
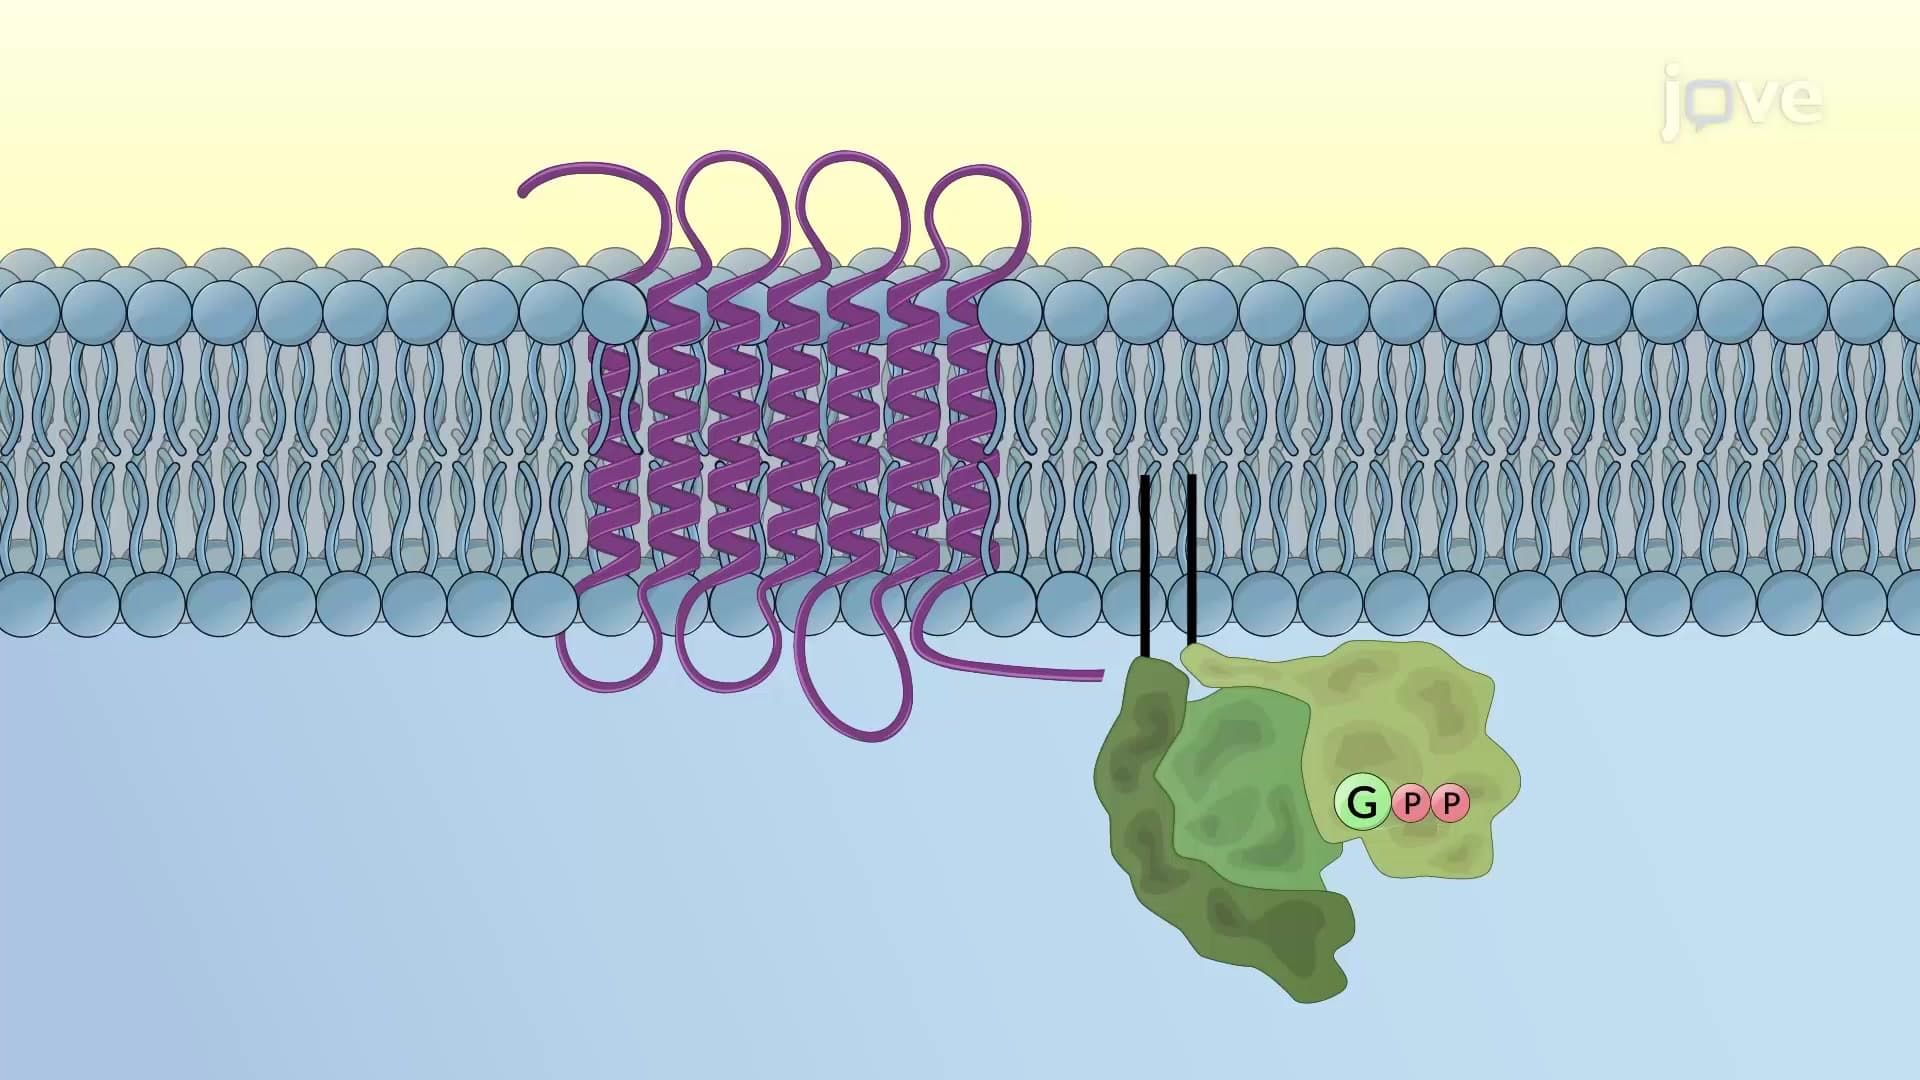
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...
 01:26
Drug Discovery: Overview
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...