Related Experiment Video
Updated: Oct 30, 2025

Fractionation for Resolution of Soluble and Insoluble Huntingtin Species
Published on: February 27, 2018
Huntingtin and the Synapse
Jessica C Barron1, Emily P Hurley1, Matthew P Parsons1
1Division of Biomedical Sciences, Faculty of Medicine, Memorial University, St. John's, NL, Canada.

Insights
Huntington disease (HD) therapies targeting mutant huntingtin (mHTT) may lower essential wild-type huntingtin (wtHTT). This review argues that wtHTT loss impairs synaptic function, preceding neurodegeneration in HD.
Area of Science:
- Neuroscience
- Genetics
- Molecular Biology
Background:
- Huntington disease (HD) is a genetic disorder caused by a CAG repeat expansion in the huntingtin (HTT) gene, leading to mutant HTT (mHTT) protein production.
- Current RNA-targeting therapies for HD, such as antisense oligonucleotides (ASOs) and RNA interference (RNAi), aim to reduce mHTT levels.
- These therapies often lack selectivity, potentially lowering beneficial wild-type HTT (wtHTT) levels, which are already reduced in HD patients.
Purpose of the Study:
- To review the critical role of wtHTT at the synapse.
- To discuss the consequences of wtHTT reduction on synaptic function in the context of HD.
- To argue that wtHTT loss is detrimental to synaptic health and may precede neurodegeneration.
Main Methods:
- Literature review focusing on the function of wtHTT in synaptic transmission.
- Analysis of the impact of reduced wtHTT levels on pre- and postsynaptic mechanisms.
- Synthesis of evidence linking synaptic dysfunction to neurodegenerative processes in HD.
Main Results:
- Wild-type HTT (wtHTT) plays a crucial role in multiple aspects of synaptic neurotransmission.
- wtHTT is implicated in protein transport, neurotransmitter release, and vesicle recycling at synapses.
- Reduced wtHTT levels negatively impact both pre- and postsynaptic functions, leading to synaptic dysfunction.
Conclusions:
- Wild-type HTT (wtHTT) is essential for maintaining normal synaptic function.
- Synaptic dysfunction resulting from wtHTT loss is a sensitive indicator of neuronal health and precedes neurodegeneration in HD.
- Non-selective therapies that deplete wtHTT may exacerbate HD pathology by compromising synaptic integrity.
Abstract:
Huntington disease (HD) is a monogenic disease that results in a combination of motor, psychiatric and cognitive symptoms. HD is caused by a CAG trinucleotide repeat expansion in the huntingtin (HTT) gene, which results in the production of a pathogenic mutant HTT protein (mHTT). Although there is no cure at present for HD, a number of RNA-targeting therapies have recently entered clinical trials which aim to lower mHTT production through the use of antisense oligonucleotides (ASOs) and RNAi. However, many of these treatment strategies are non-selective in that they cannot differentiate between non-pathogenic wild type HTT (wtHTT) and the mHTT variant. As HD patients are already born with decreased levels of wtHTT, these genetic therapies may result in critically low levels of wtHTT. The consequence of wtHTT reduction in the adult brain is currently under debate, and here we argue that wtHTT loss is not well-tolerated at the synaptic level. Synaptic dysfunction is an extremely sensitive measure of subsequent cell death, and is known to precede neurodegeneration in numerous brain diseases including HD. The present review focuses on the prominent role of wtHTT at the synapse and considers the consequences of wtHTT loss on both pre- and postsynaptic function. We discuss how wtHTT is implicated in virtually all major facets of synaptic neurotransmission including anterograde and retrograde transport of proteins to/from terminal buttons and dendrites, neurotransmitter release, endocytic vesicle recycling, and postsynaptic receptor localization and recycling. We conclude that wtHTT presence is essential for proper synaptic function.
More Related Videos
 10:52
Efficient and Scalable Production of Full-length Human Huntingtin Variants in Mammalian Cells using a Transient Expression System
Published on: December 10, 2021
 11:22
Generation of Native, Untagged Huntingtin Exon1 Monomer and Fibrils Using a SUMO Fusion Strategy
Published on: June 27, 2018
Related Concept Videos
 02:47
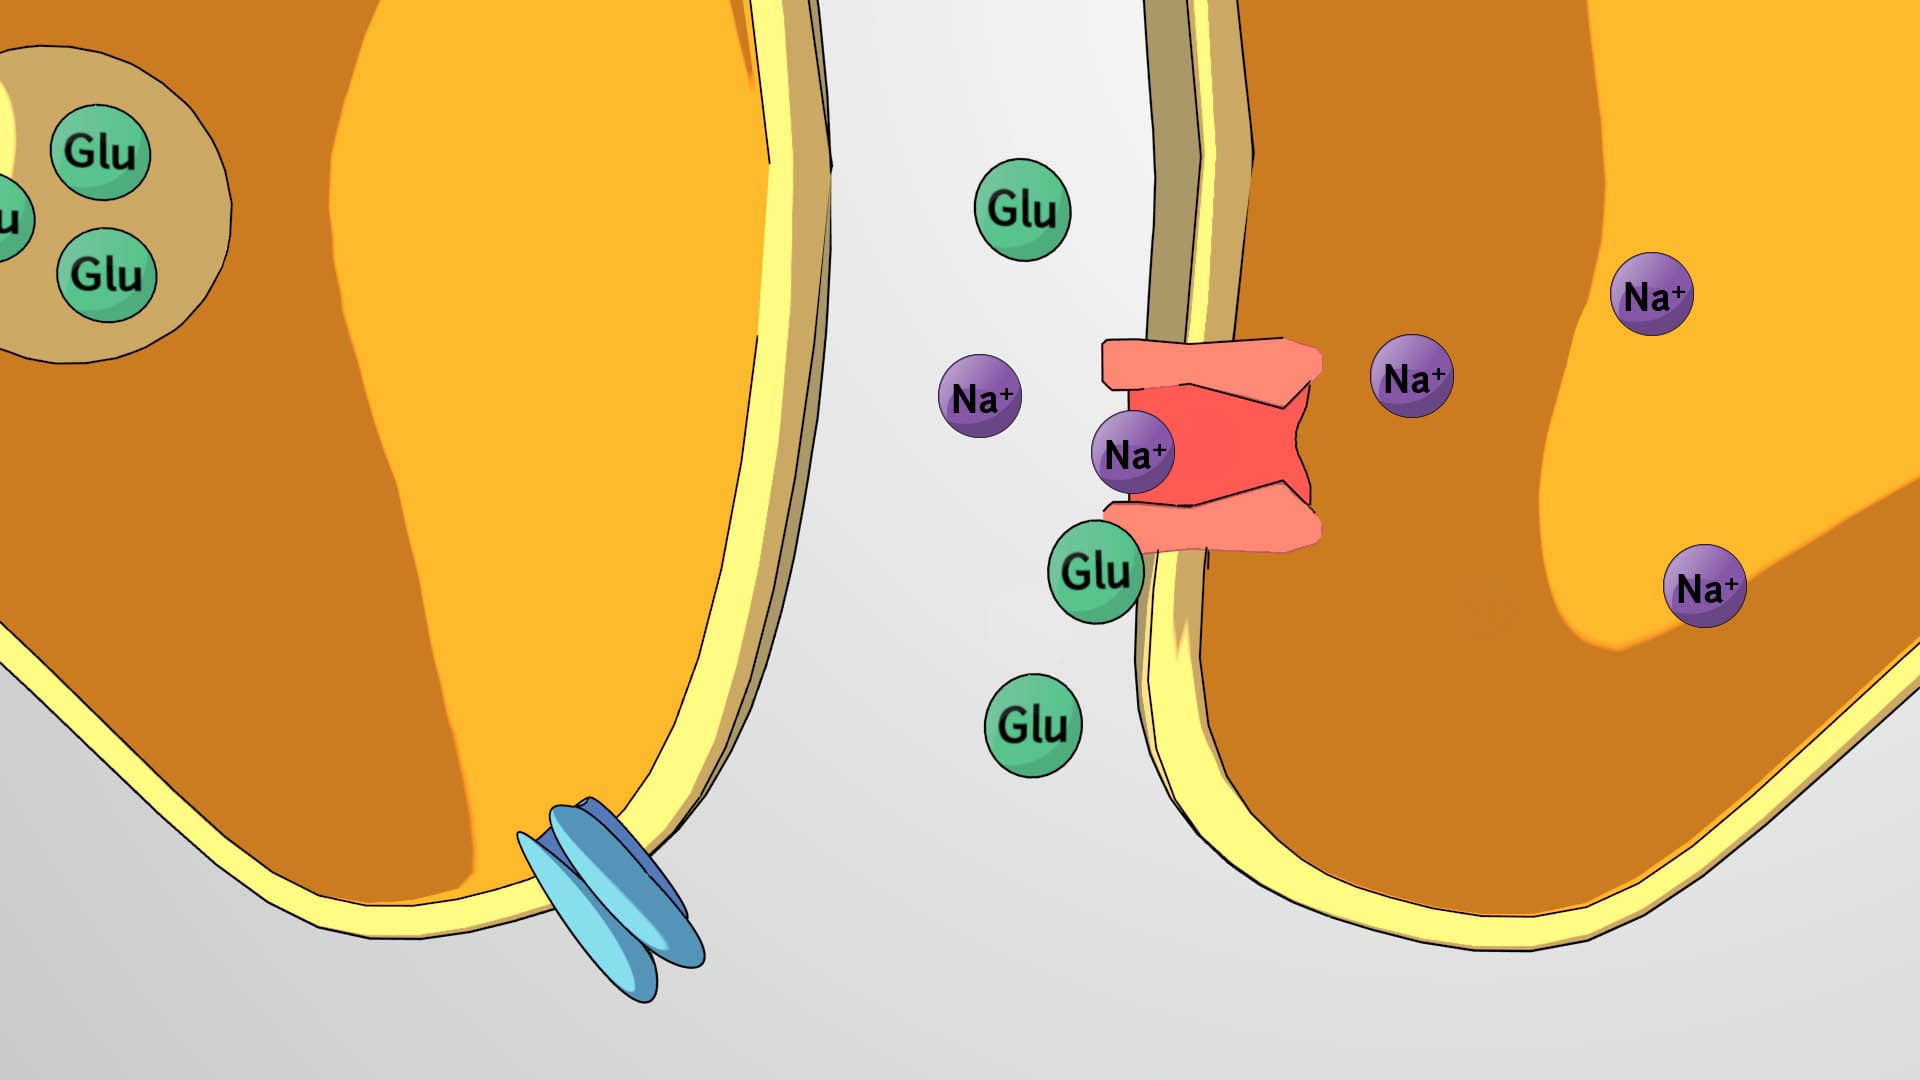
The Synapse
 01:26
Fusion of Secretory Vesicles with the Plasma Membrane
In 1993, Jim Rothman proposed that the antiparallel pairing of vesicular and transmembrane SNAREs, or...
 01:23
Catenins
Catenins in Cell Junctions
Catenins bind to cell adhesion molecules such as cadherins and link them to different cytoskeletal proteins depending on the type of cell junction. At the...
 01:26
Chemical Synapses
Because chemical synapses depend on the release of neurotransmitter molecules from synaptic vesicles to pass on their signal, there is an approximately one millisecond delay between when the axon potential reaches the presynaptic terminal and when the neurotransmitter leads to opening of postsynaptic ion channels. Additionally, this signaling is...
01:26Chemical Synapses
Because chemical synapses depend on the release of neurotransmitter molecules from synaptic vesicles to pass on their signal, there is an approximately one millisecond delay between when the axon potential reaches the presynaptic terminal and when the neurotransmitter leads to opening of postsynaptic ion channels. Additionally, this signaling is...
 01:32
Assembly of Complex Microtubule Structures