Related Experiment Video
Updated: Oct 29, 2025
15N CPMG Relaxation Dispersion for the Investigation of Protein Conformational Dynamics on the µs-ms Timescale
Published on: April 19, 2021
Model protein excited states: MRCI calculations with large active spaces vs CC2 method
Valérie Brenner1, Thibaut Véry1, Michael W Schmidt2
1LIDYL, CEA, CNRS, Université Paris-Saclay, 91191 Gif-sur-Yvette, France.

Abstract:
Benchmarking calculations on excited states of models of phenylalanine protein chains are presented to assess the ability of alternative methods to the standard and most commonly used multiconfigurational wave function-based method, the complete active space self-consistent field (CASSCF), in recovering the non-dynamical correlation for systems that become not affordable by the CASSCF. The exploration of larger active spaces beyond the CASSCF limit is benchmarked through three strategies based on the reduction in the number of determinants: the restricted active space self-consistent field, the generalized active space self-consistent field (GASSCF), and the occupation-restricted multiple active space (ORMAS) schemes. The remaining dynamic correlation effects are then added by the complete active space second-order perturbation theory and by the multireference difference dedicated configuration interaction methods. In parallel, the approximate second-order coupled cluster (CC2), already proven to be successful for small building blocks of model proteins in one of our previous works [Ben Amor et al., J. Chem. Phys. 148, 184105 (2018)], is investigated to assess its performances for larger systems. Among the different alternative strategies to CASSCF, our results highlight the greatest efficiency of the GASSCF and ORMAS schemes in the systematic reduction of the configuration interaction expansion without loss of accuracy in both nature and excitation energies of both singlet ππ* and nπ* CO excited states with respect to the equivalent CASSCF calculations. Guidelines for an optimum applicability of this scheme to systems requiring active spaces beyond the complete active space limit are then proposed. Finally, the extension of the CC2 method to such large systems without loss of accuracy is demonstrated, highlighting the great potential of this method to treat accurately excited states, mainly single reference, of very large systems.
Related Concept Videos
 02:00
Molecular Models
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...
 02:36
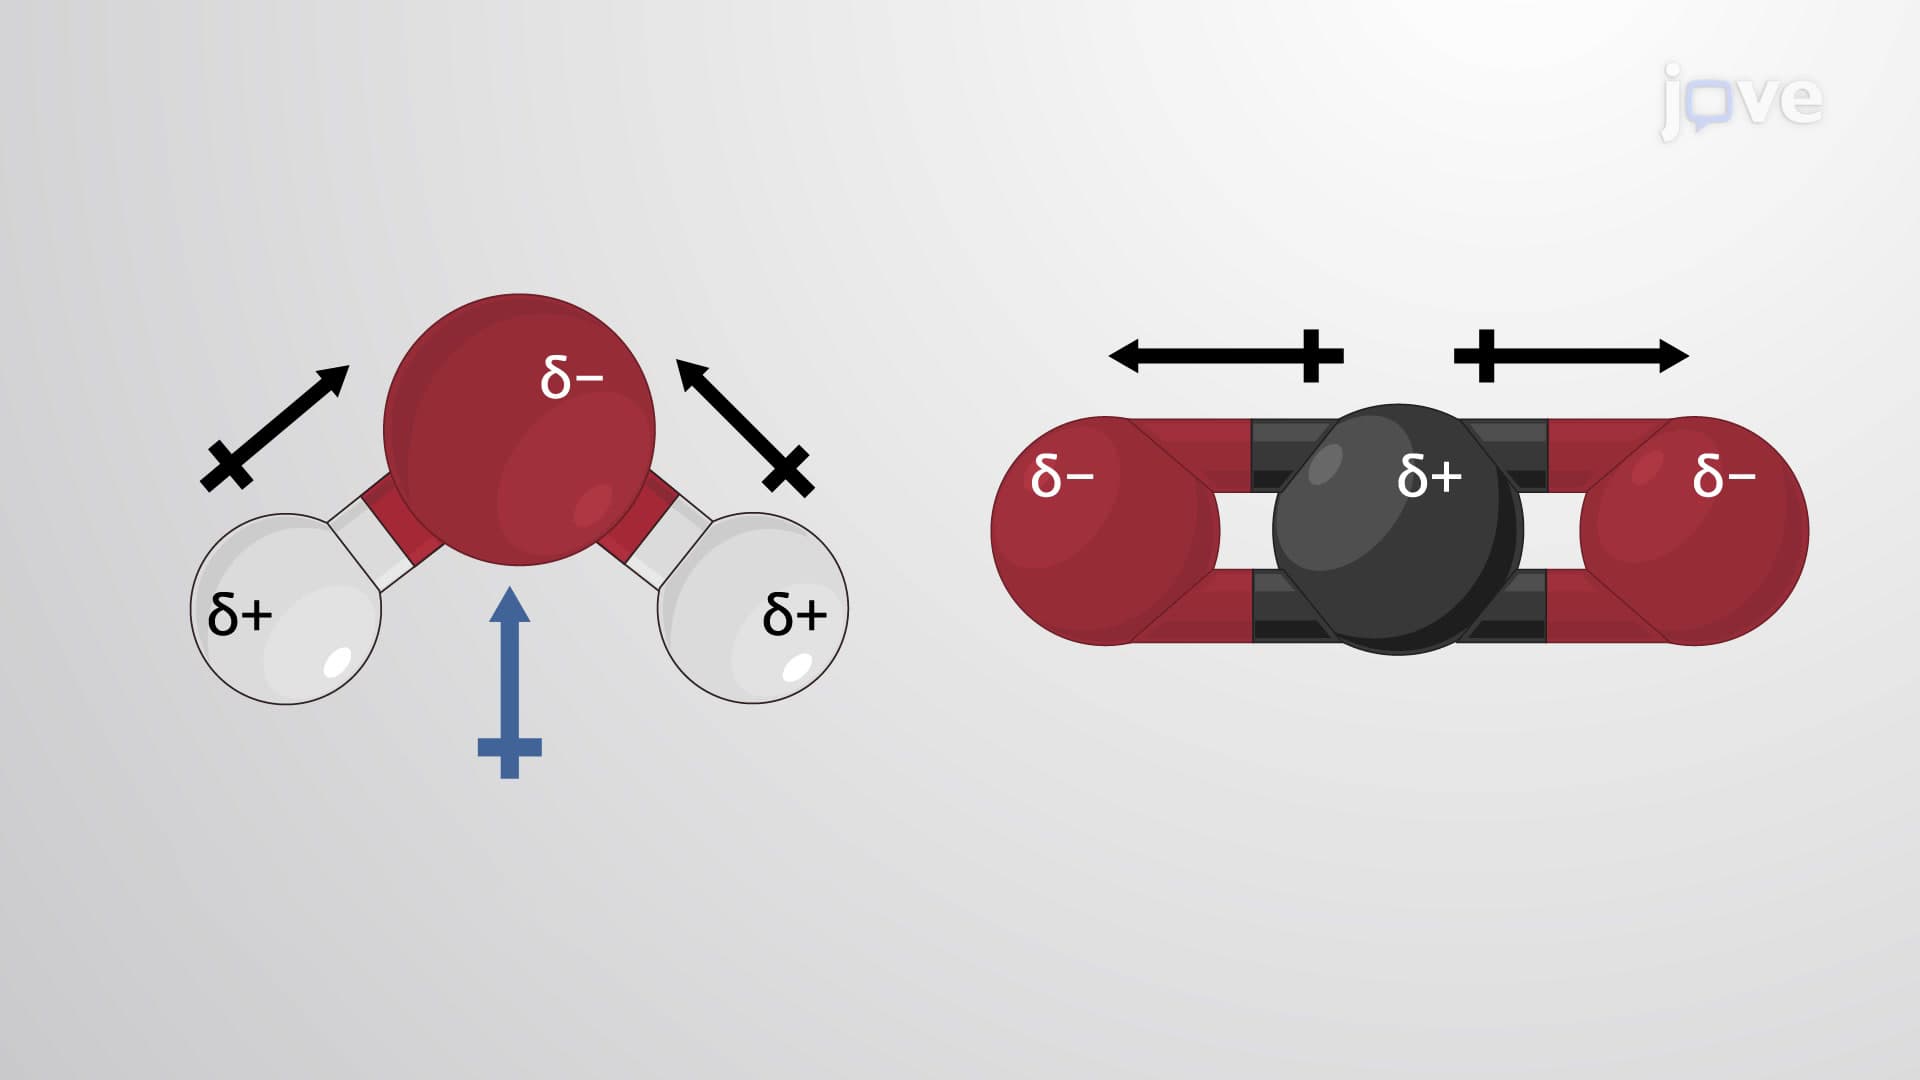
Molecular Geometry and Dipole Moments
 01:27
π Electron Effects on Chemical Shift: Overview
 03:51
Molecular Orbital Theory II
 02:35
Molecular Orbital Theory I