Related Experiment Video
Updated: Oct 29, 2025

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Approximations of density matrices in N-electron valence state second-order perturbation theory (NEVPT2). II. The
Yang Guo1, Kantharuban Sivalingam2, Christian Kollmar2
1Qingdao Institute for Theoretical and Computational Sciences, Shandong University, Qingdao, Shandong 266237, China.

Abstract:
In Paper I, the performances of pre-screening (PS), extended PS (EPS), and cumulant (CU) approximations to the fourth-order density matrix were examined in the context of second-order N-electron valence state perturbation theory (NEVPT2). It has been found that the CU, PS, and even EPS approximations with loose thresholds may introduce intruder states. In the present work, the origin of these "false intruder" states introduced by approximated density matrices is discussed. Canonical NEVPT2 implementations employ a rank reduction trick. By analyzing its residual error, we find that the omission of the rank reduction leads to a more stable multireference perturbation theory for incomplete active space reference wave functions. Such a full rank (FR)-NEVPT2 formulation is equivalent to the conventional NEVPT2 method for the complete active space self-consistent field/complete active space configuration interaction reference wave function. A major drawback of the FR-NEVPT2 formulation is the necessity of the fifth-order density matrix. To avoid the construction of the high-order density matrices, the combination of the FR-NEVPT2 with the CU approximation is studied. However, we find that the CU approximation remains problematic as it still introduces intruder states. The question of how to robustly and efficiently perform internally contracted multireference perturbation theories with approximate densities remains a challenging field of investigation.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 02:45
The Quantum-Mechanical Model of an Atom
 01:10
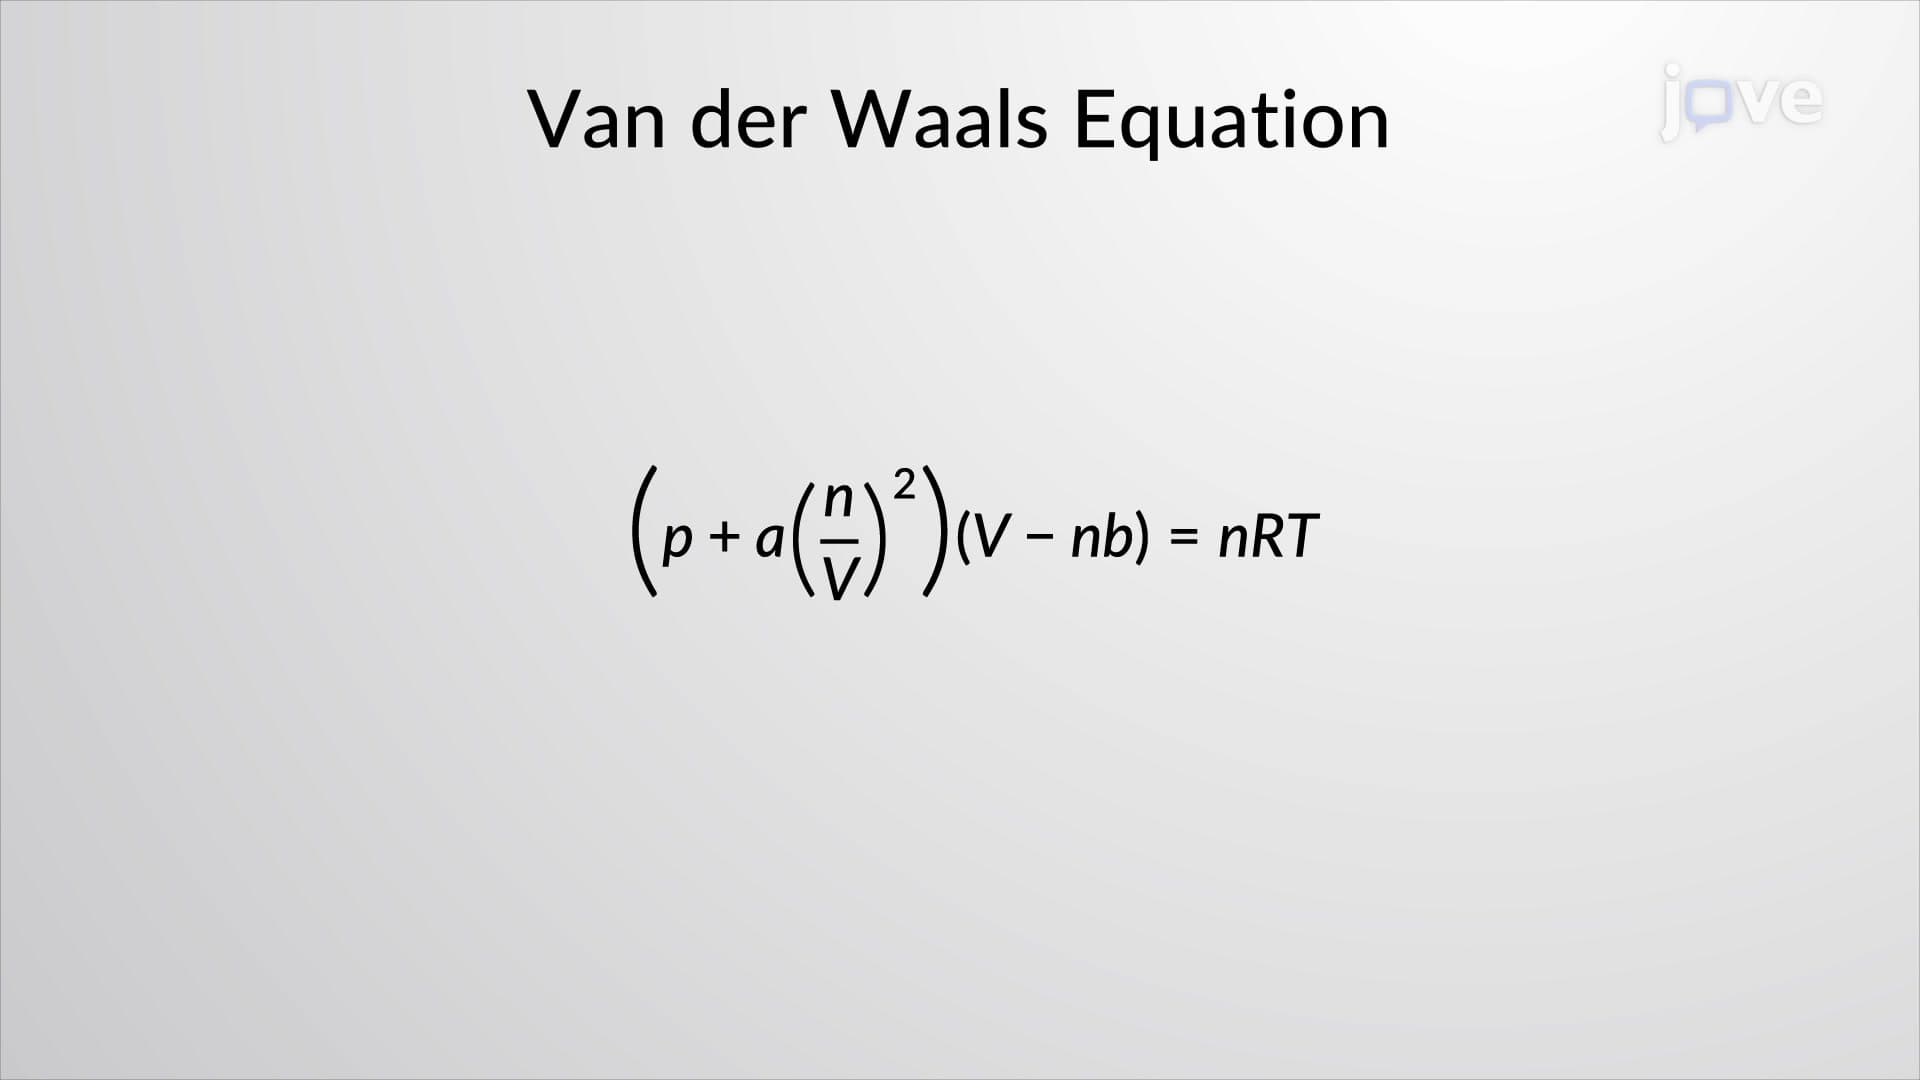
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 03:06
The Pauli Exclusion Principle
 03:02
The Aufbau Principle and Hund's Rule
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation
 03:51
Molecular Orbital Theory II