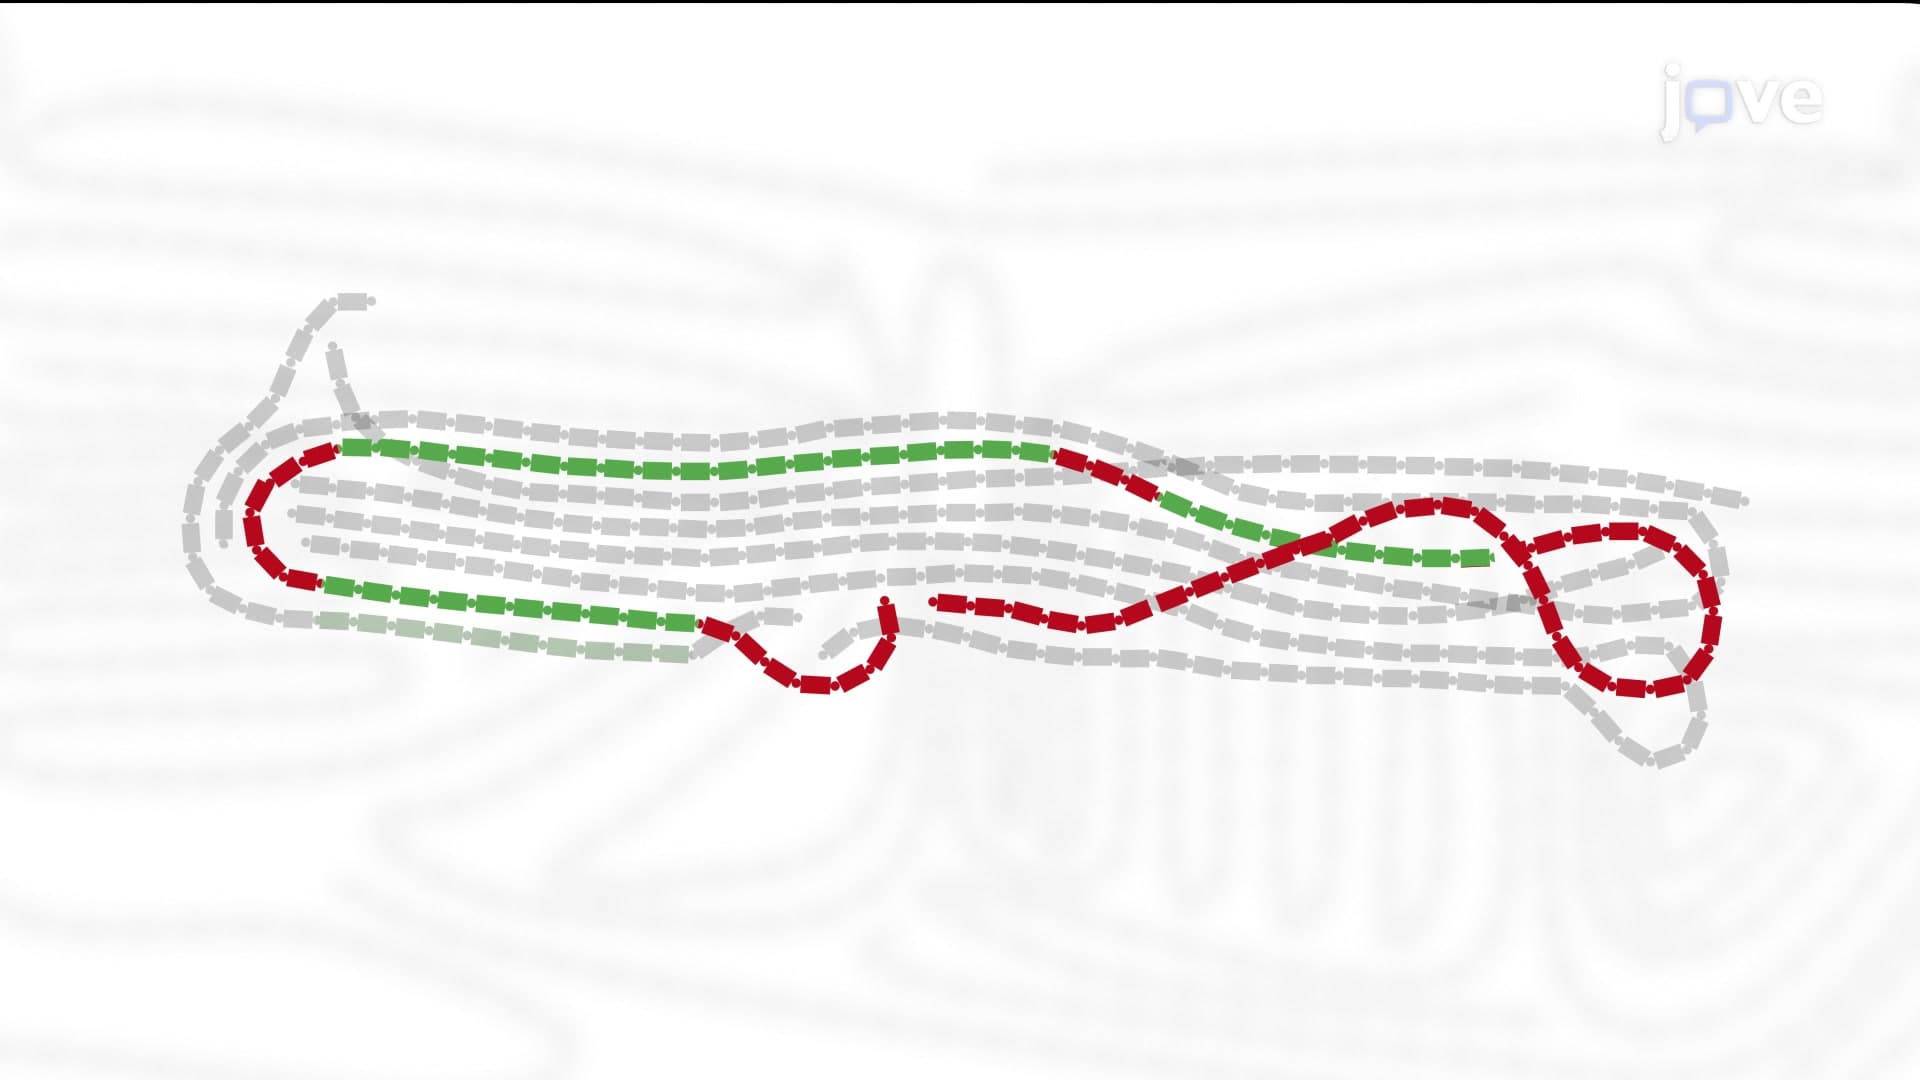
A molecular dynamics simulation of crystalline alpha-cyclodextrin hexahydrate
Related Experiment Video
Updated: Oct 2, 2026
Iterative Optimization of DNA Duplexes for Crystallization of SeqA-DNA Complexes
Published on: November 1, 2012
J E Koehler1, W Saenger, W F van Gunsteren
1Institut für Kristallographie, Freie Universität Berlin, Germany.

Abstract:
The structure of crystalline alpha-cyclodextrin (alpha-CD) hexahydrate, form I (C36H60O30.6H2O, space group P2(1)2(1)2(1)) is experimentally so well determined by X-ray and by neutron diffraction analyses that the positions of all the hydrogen atoms are available. This provides an opportunity for testing an empirical force field that is currently used in simulations of protein and nucleic acid structures by performing molecular dynamics studies employing the GROMOS program package on a system of 4 unit cells containing 16 alpha-CD molecules and 96 water molecules. A detailed comparison of the simulated and experimentally determined crystal structures shows that the experimental positions of the alpha-CD atoms are reproduced within 0.025 nm, well within the overall experimental accuracy of 0.036 nm; that the water molecules are on average within 0.072 nm from their experimental sites, with two thirds reproduced within experimental accuracy by the calculations; that high correlation is produced between the occurrence of simulated and experimentally observed hydrogen bonds. The good agreement between simulated and experimental results suggests that the tested force field is reliable.
More Related Videos
 11:27
Studying Soft-matter and Biological Systems over a Wide Length-scale from Nanometer and Micrometer Sizes at the Small-angle Neutron Diffractometer KWS-2
Published on: December 8, 2016
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020