Related Experiment Video
Updated: Oct 25, 2025

An Intestine/Liver Microphysiological System for Drug Pharmacokinetic and Toxicological Assessment
Published on: December 3, 2020
Comparison between lab variability and in silico prediction errors for the unbound fraction of drugs in human plasma
Urban Fagerholm1, Ola Spjuth1,2, Sven Hellberg1
1Prosilico AB, Huddinge, Sweden.

Abstract:
Variability of the unbound fraction in plasma (fu) between labs, methods and conditions is known to exist. Variability and uncertainty of this parameter influence predictions of the overall pharmacokinetics of drug candidates and might jeopardise safety in early clinical trials. Objectives of this study were to evaluate the variability of human in vitro fu-estimates between labs for a range of different drugs, and to develop and validate an in silico fu-prediction method and compare the results to the lab variability.A new in silico method with prediction accuracy (Q2) of 0.69 for log fu was developed. The median and maximum prediction errors were 1.9- and 92-fold, respectively. Corresponding estimates for lab variability (ratio between max and min fu for each compound) were 2.0- and 185-fold, respectively. Greater than 10-fold lab variability was found for 14 of 117 selected compounds.Comparisons demonstrate that in silico predictions were about as reliable as lab estimates when these have been generated during different conditions. Results propose that the new validated in silico prediction method is valuable not only for predictions at the drug design stage, but also for reducing uncertainties of fu-estimations and improving safety of drug candidates entering the clinical phase.
More Related Videos
Related Concept Videos
 01:29
Drug Distribution: Plasma Protein Binding
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
 01:24
Dosage Regimen: Individualization
 01:01
Dosage Regimens: Partial Pharmacokinetic Parameters
 01:20
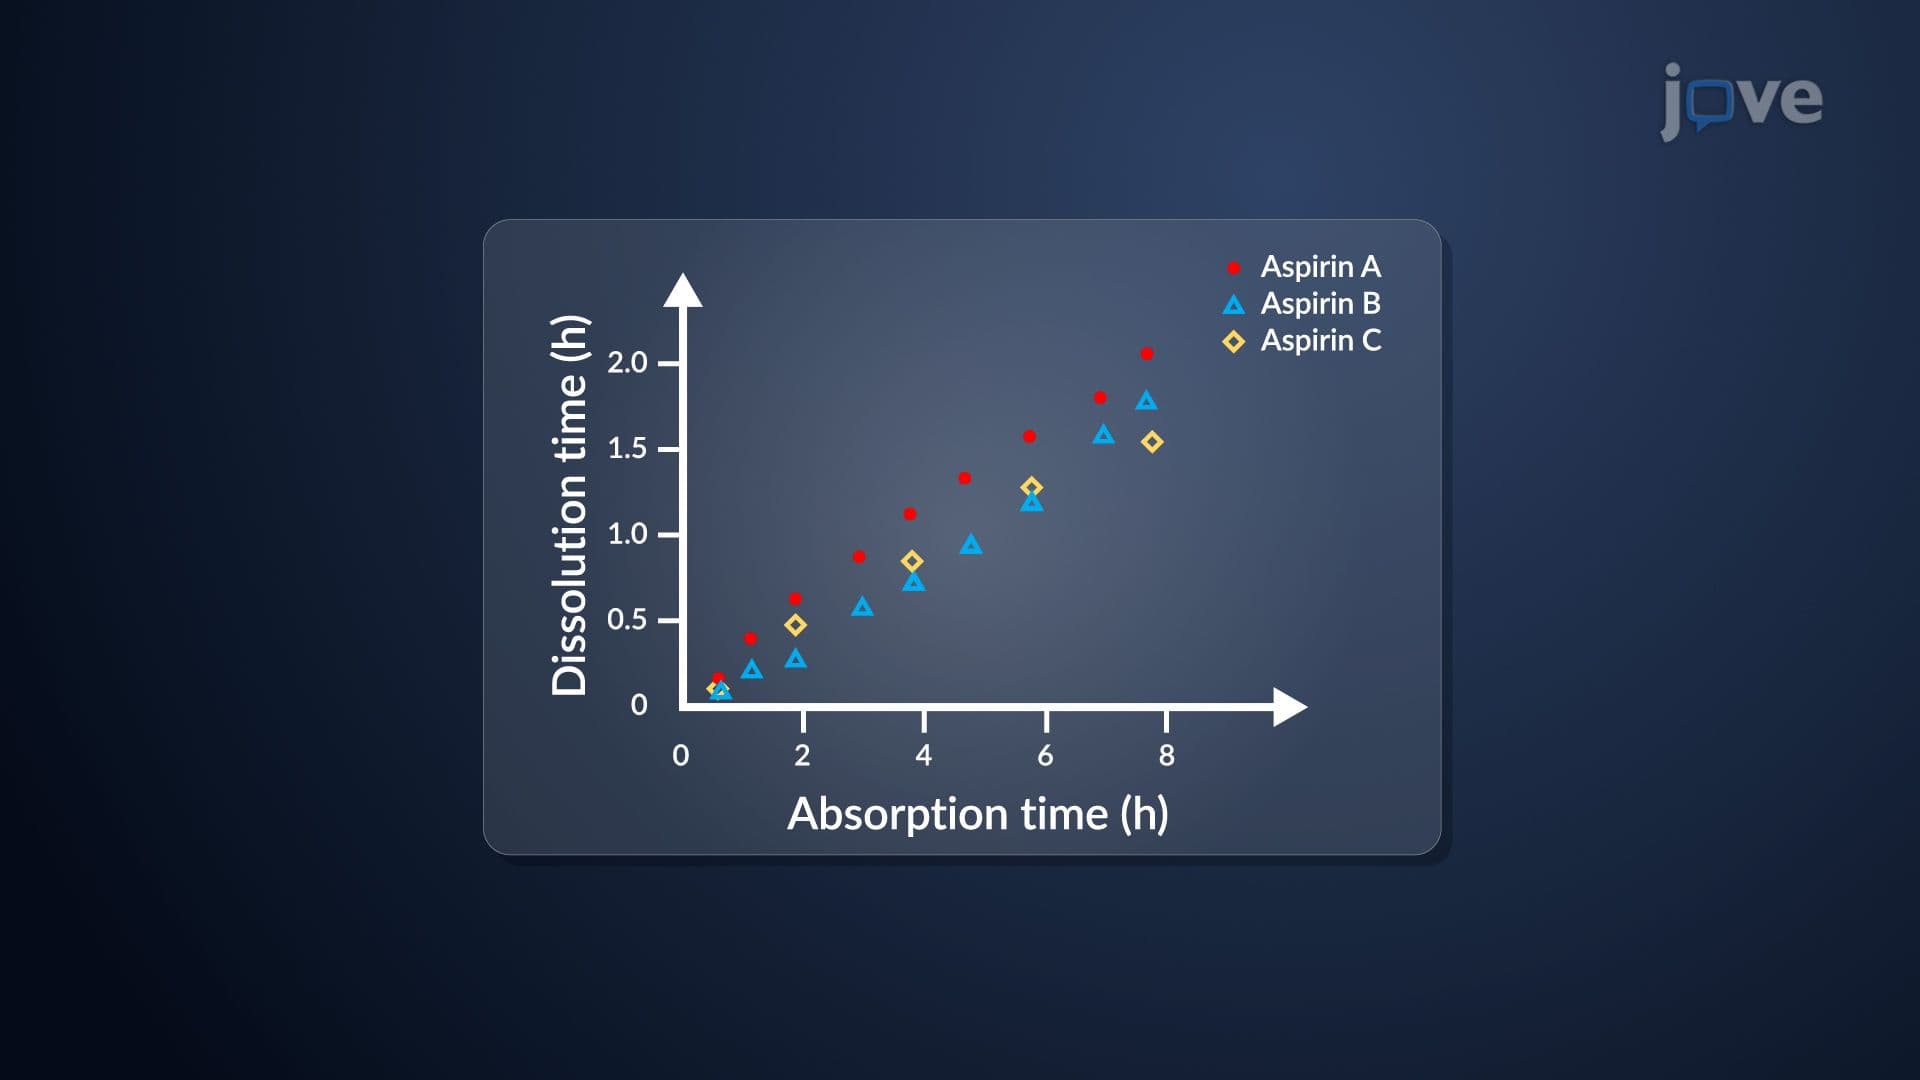
Drug Product Performance: In Vitro–In Vivo Correlation
 01:26
Pharmacokinetic Models: Comparison and Selection Criterion
Physiological models take a detailed approach by considering specific molecular processes. They can predict drug distribution, metabolism, and elimination changes, providing a comprehensive understanding of how drugs interact with the body.