Related Experiment Video
Updated: Oct 25, 2025
Facile Preparation of 2Z,4E-Dienamides by the Olefination of Electron-deficient Alkenes with Allyl Acetate
Published on: June 21, 2017
Experimental and computational studies of the mechanism of iron-catalysed C-H activation/functionalisation with allyl
Joshua C DeMuth1, Zhihui Song2, Stephanie H Carpenter1
1Department of Chemistry, University of Rochester Rochester New York 14627 USA neidig@chem.rochester.edu.

Abstract:
Synthetic methods that utilise iron to facilitate C-H bond activation to yield new C-C and C-heteroatom bonds continue to attract significant interest. However, the development of these systems is still hampered by a limited molecular-level understanding of the key iron intermediates and reaction pathways that enable selective product formation. While recent studies have established the mechanism for iron-catalysed C-H arylation from aryl-nucleophiles, the underlying mechanistic pathway of iron-catalysed C-H activation/functionalisation systems which utilise electrophiles to establish C-C and C-heteroatom bonds has not been determined. The present study focuses on an iron-catalysed C-H allylation system, which utilises allyl chlorides as electrophiles to establish a C-allyl bond. Freeze-trapped inorganic spectroscopic methods (57Fe Mössbauer, EPR, and MCD) are combined with correlated reaction studies and kinetic analyses to reveal a unique and rapid reaction pathway by which the allyl electrophile reacts with a C-H activated iron intermediate. Supporting computational analysis defines this novel reaction coordinate as an inner-sphere radical process which features a partial iron-bisphosphine dissociation. Highlighting the role of the bisphosphine in this reaction pathway, a complementary study performed on the reaction of allyl electrophile with an analogous C-H activated intermediate bearing a more rigid bisphosphine ligand exhibits stifled yield and selectivity towards allylated product. An additional spectroscopic analysis of an iron-catalysed C-H amination system, which incorporates N-chloromorpholine as the C-N bond-forming electrophile, reveals a rapid reaction of electrophile with an analogous C-H activated iron intermediate consistent with the inner-sphere radical process defined for the C-H allylation system, demonstrating the prevalence of this novel reaction coordinate in this sub-class of iron-catalysed C-H functionalisation systems. Overall, these results provide a critical mechanistic foundation for the rational design and development of improved systems that are efficient, selective, and useful across a broad range of C-H functionalisations.
Related Concept Videos
 02:17
Reduction of Alkenes: Asymmetric Catalytic Hydrogenation
The metal catalyst used can be either heterogeneous or homogeneous. When hydrogenation of an alkene generates a chiral center, a pair of enantiomeric products is expected to form. However, an enantiomeric excess of one of the products can be facilitated using an enantioselective reaction or an...
 02:24
Reduction of Alkynes to cis-Alkenes: Catalytic Hydrogenation
Like alkenes, alkynes can be reduced to alkanes in the presence of transition metal catalysts such as Pt, Pd, or Ni. The reaction involves two sequential syn additions of hydrogen via a cis-alkene intermediate.
 01:18
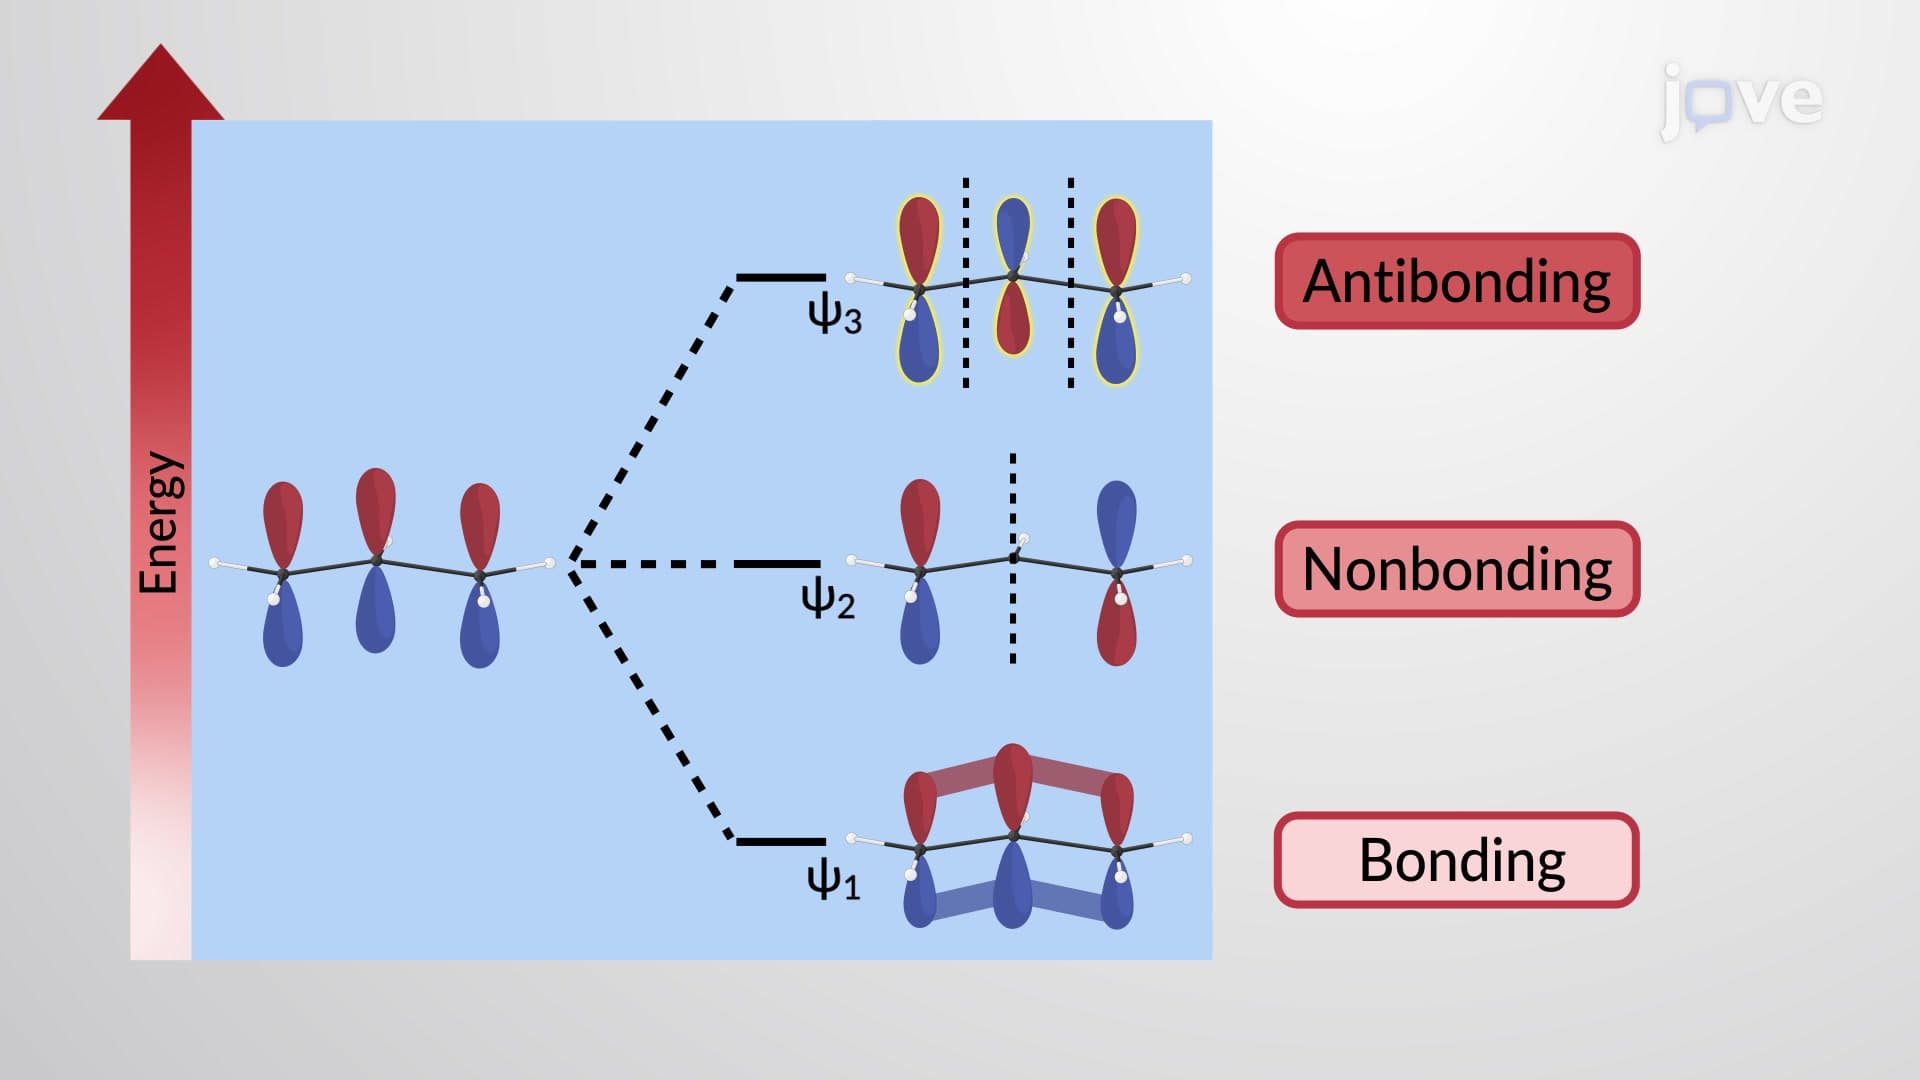
π Molecular Orbitals of the Allyl Cation and Anion
 02:13
Reduction of Alkenes: Catalytic Hydrogenation
Metals like palladium, platinum, and nickel are commonly used in their solid forms — fine powder on an inert surface. As these catalysts remain insoluble in the reaction mixture, they are referred to as heterogeneous catalysts.
The hydrogenation process takes place on the...
 02:17
Regioselectivity of Electrophilic Additions to Alkenes: Markovnikov's Rule
The hydrohalogenation of an unsymmetrical alkene can yield two haloalkane products, depending on which vinylic carbon takes up the halogen. However, one product usually predominates, where hydrogen adds to the vinylic carbon bearing the...
 01:21
Acid-Catalyzed α-Halogenation of Aldehydes and Ketones
In the first step of the mechanism, the acid protonates the carbonyl oxygen resulting in a resonance-stabilized cation, which subsequently loses an α-hydrogen to form an enol tautomer. The C=C bond in an enol is highly nucleophilic because of the electron-donating nature of the –OH group. Consequently, the double bond attacks an electrophilic halogen to form a...