Related Experiment Video
Updated: Oct 23, 2025

Functional Reconstitution and Channel Activity Measurements of Purified Wildtype and Mutant CFTR Protein
Published on: March 9, 2015
Functional stability of CFTR depends on tight binding of ATP at its degenerate ATP-binding site
Han-I Yeh1,2, Ying-Chun Yu1, Pei-Lun Kuo3
1Dalton Cardiovascular Research Center and Department of Medical Pharmacology and Physiology, University of Missouri-Columbia, Columbia, MO, 65211, USA.

Abstract:
Opening of the cystic fibrosis transmembrane conductance regulator (CFTR) channel is coupled to the motion of its two nucleotide-binding domains: they form a heterodimer sandwiching two functionally distinct ATP-binding sites (sites 1 and 2). While active ATP hydrolysis in site 2 triggers rapid channel closure, the functional role of stable ATP binding in the catalysis-incompetent (or degenerate) site 1, a feature conserved in many other ATP-binding cassette (ABC) transporter proteins, remains elusive. Here, we found that CFTR loses its prompt responsiveness to ATP after the channel is devoid of ATP for tens to hundreds of seconds. Mutants with weakened ATP binding in site 1 and the most prevalent disease-causing mutation, F508del, are more vulnerable to ATP depletion. In contrast, strengthening ligand binding in site 1 with N6 -(2-phenylethyl)-ATP, a high-affinity ATP analogue, or abolishing ATP hydrolysis in site 2 by the mutation D1370N, helps sustain a durable function of the otherwise unstable mutant channels. Thus, tight binding of ATP in the degenerate ATP-binding site is crucial to the functional stability of CFTR. Small molecules targeting site 1 may bear therapeutic potential to overcome the membrane instability of F508del-CFTR. KEY POINTS: During evolution, many ATP-binding cassette transporters - including the cystic fibrosis transmembrane conductance regulator (CFTR) chloride channel, whose dysfunction causes cystic fibrosis (CF) - lose the ability to hydrolyse ATP in one of the two ATP-binding sites. Here we show that tight ATP binding at this degenerate site in CFTR is central for maintaining the stable, robust function of normal CFTR. We also demonstrate that membrane instability of the most common CF-causing mutant, F508del-CFTR, can be rescued by strengthening ATP binding at CFTR's degenerate site. Our data thus explain an evolutionary puzzle and offer a potential therapeutic strategy for CF.
Related Concept Videos
 01:19
Allosteric Proteins-ATCase
Aspartate transcarbamoylase (ATCase) is a cytosolic enzyme that catalyzes the condensation of L-aspartate and carbamoyl phosphate to N-carbamoyl-L-aspartate. This reaction is the first step in pyrimidine biosynthesis. UTP and CTP, the end products of the pyrimidine synthesis...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 01:48
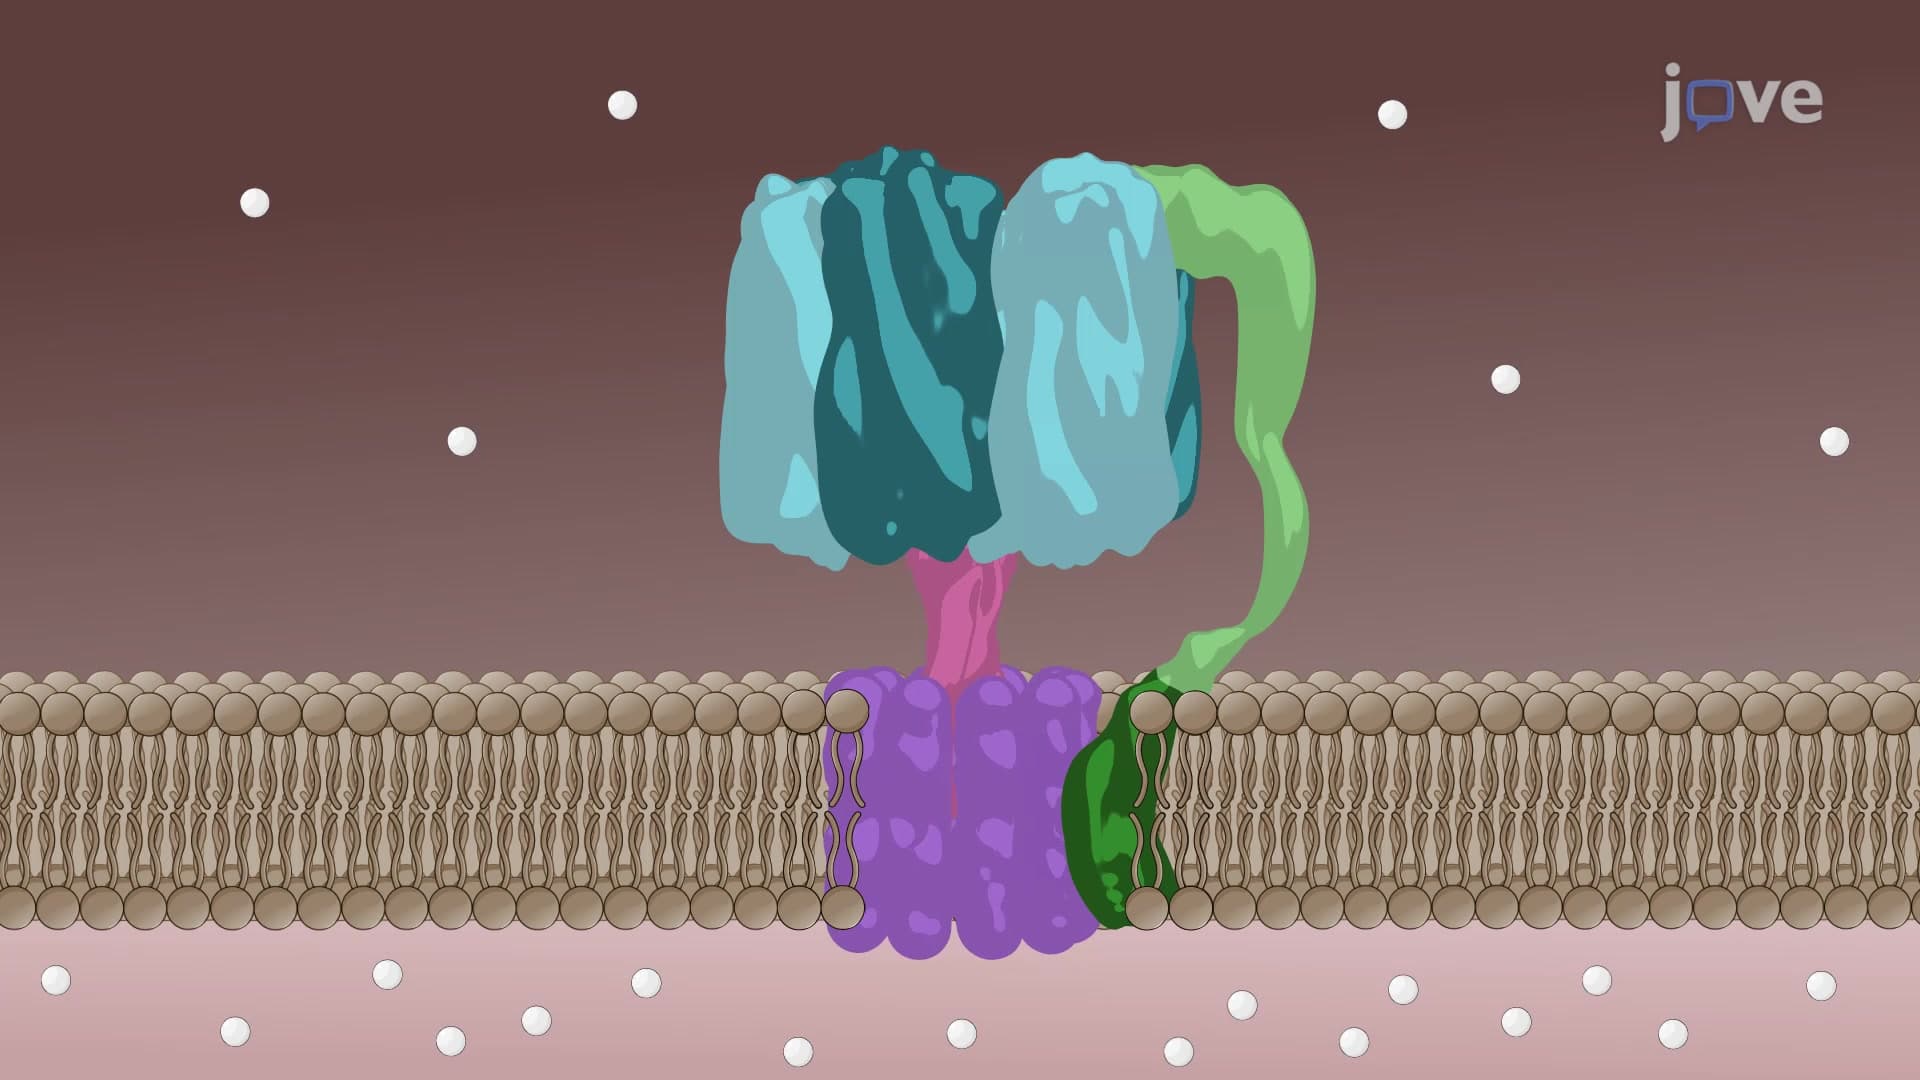
ATP Synthase: Mechanism
 02:13
Cooperative Binding of Transcription Regulators
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:18
ATP Synthase: Structure